Related Experiment Video
Updated: Aug 11, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Ab initio molecular dynamics simulation of the structure and proton transport dynamics of methanol-water solutions
Joseph A Morrone1, Kiryn E Haslinger, Mark E Tuckerman
1Department of Chemistry, Princeton University, Princeton, New Jersey 08544, USA.

Abstract:
Ab initio molecular dynamics simulations are employed to study the structural and proton transport properties of methanol-water mixtures. Structural characteristics analyzed at two different methanol mole fractions (X(M) = 0.25 and X(M) = 0.5) reveal enhanced structuring of water as the methanol mole fraction increases in agreement with recent neutron diffraction experiments. The simulations reveal the existence of separate hydrogen-bonded water and methanol networks, also in agreement with the neutron diffraction data. The addition of a single proton to the X(M) = 0.5 mixture leads to an anomalous structural or Grotthuss-type diffusion mechanism of the charge defect in which water-to-water, methanol-to-water, and water-to-methanol proton transfer reactions play the dominant role with methanol-to-methanol transfers being much less significant. Unlike in bulk water, where coordination number fluctuations drive the proton transport process, suppression of the coordination number of waters in the first solvation shell of the defect diminish the importance of coordination number fluctuations as a driving force in the structural diffusion process. The charge defect is found to reside preferentially at the interface between water and methanol networks. The length of the ab initio molecular dynamics run (approximately 120 ps), allowed the diffusion constant of the charge defect to be computed, yielding a value of D = 4.2 x 10(-5) cm2/s when deuterium masses are assigned to all protons in the system. The relation of this value to excess proton diffusion in bulk water is discussed. Finally, a kinetic theory is introduced to identify the relevant time scales in the proton transfer/transport process.

Related Concept Videos
 01:10
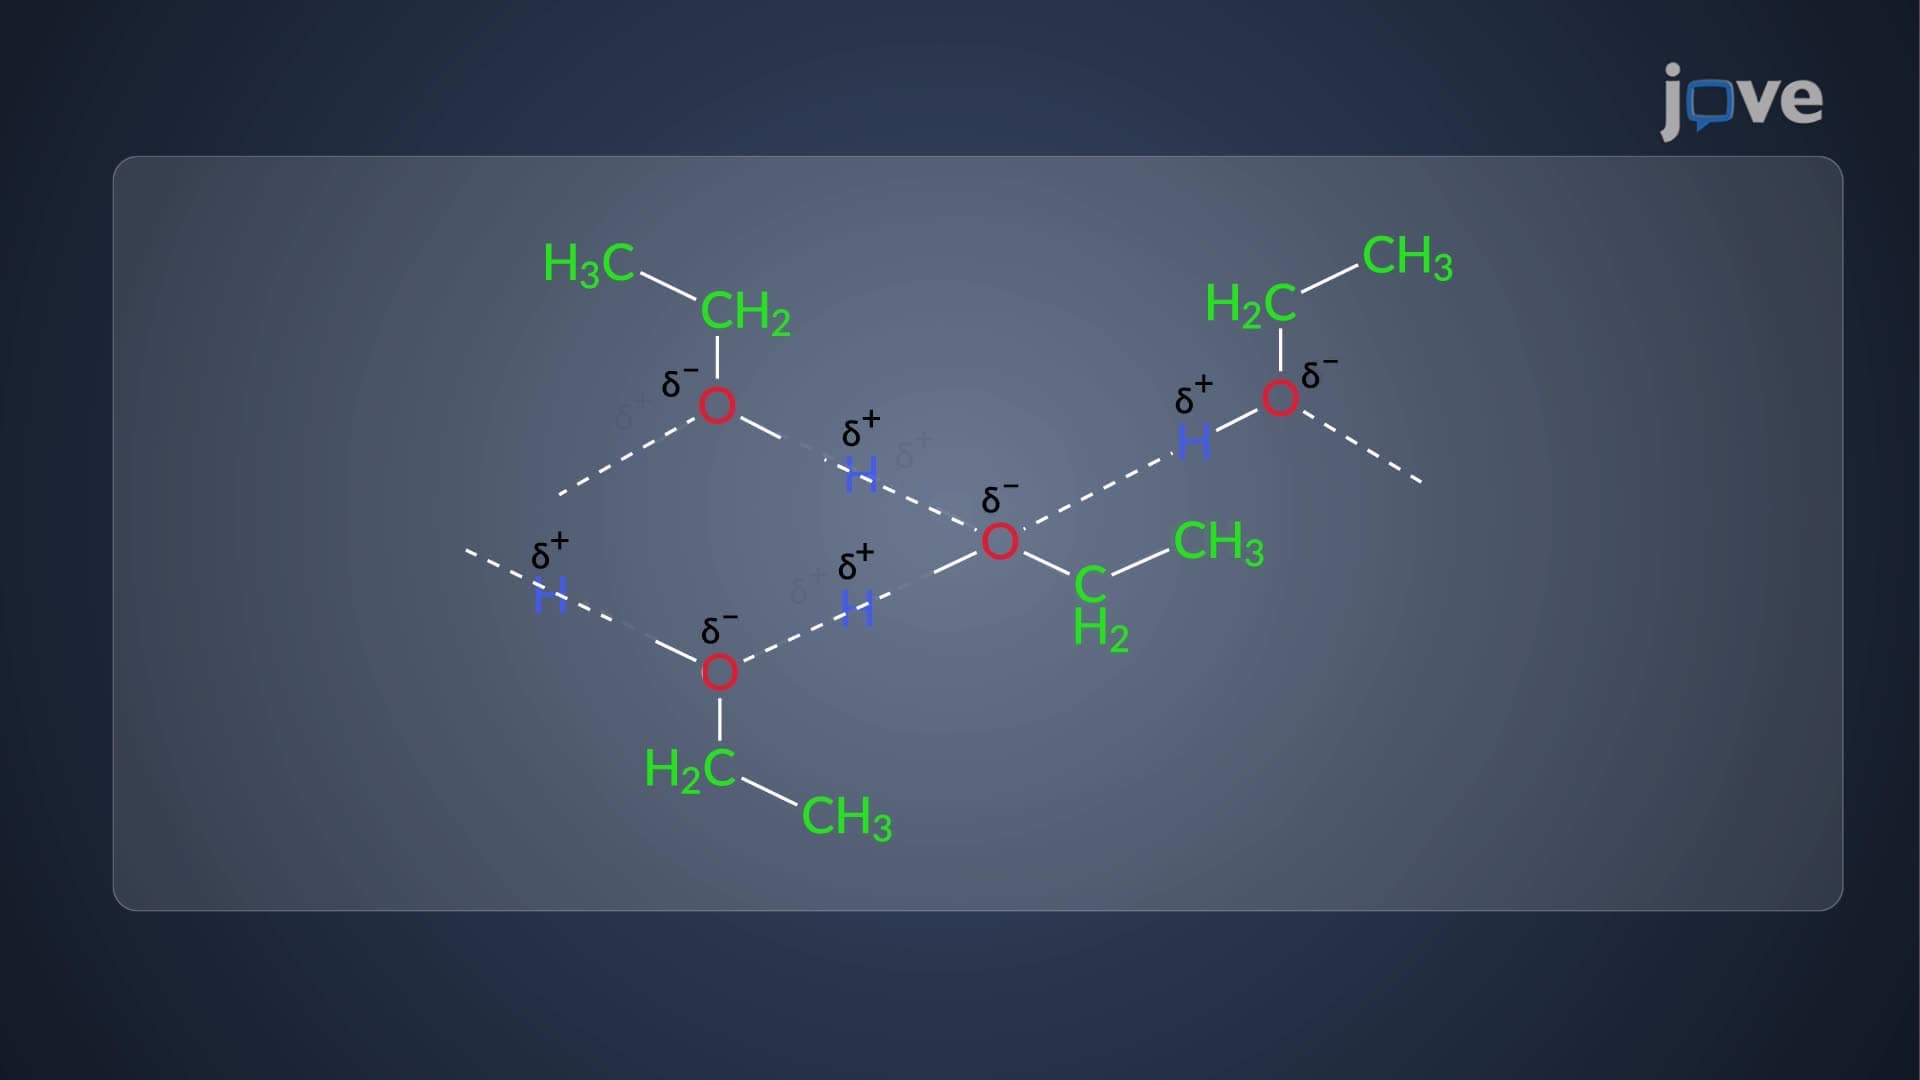
¹H NMR of Labile Protons: Temporal Resolution
The –OH proton in alcohols typically appears in the range of δ 2 to 5 ppm but can vary depending on the specific...
 00:48
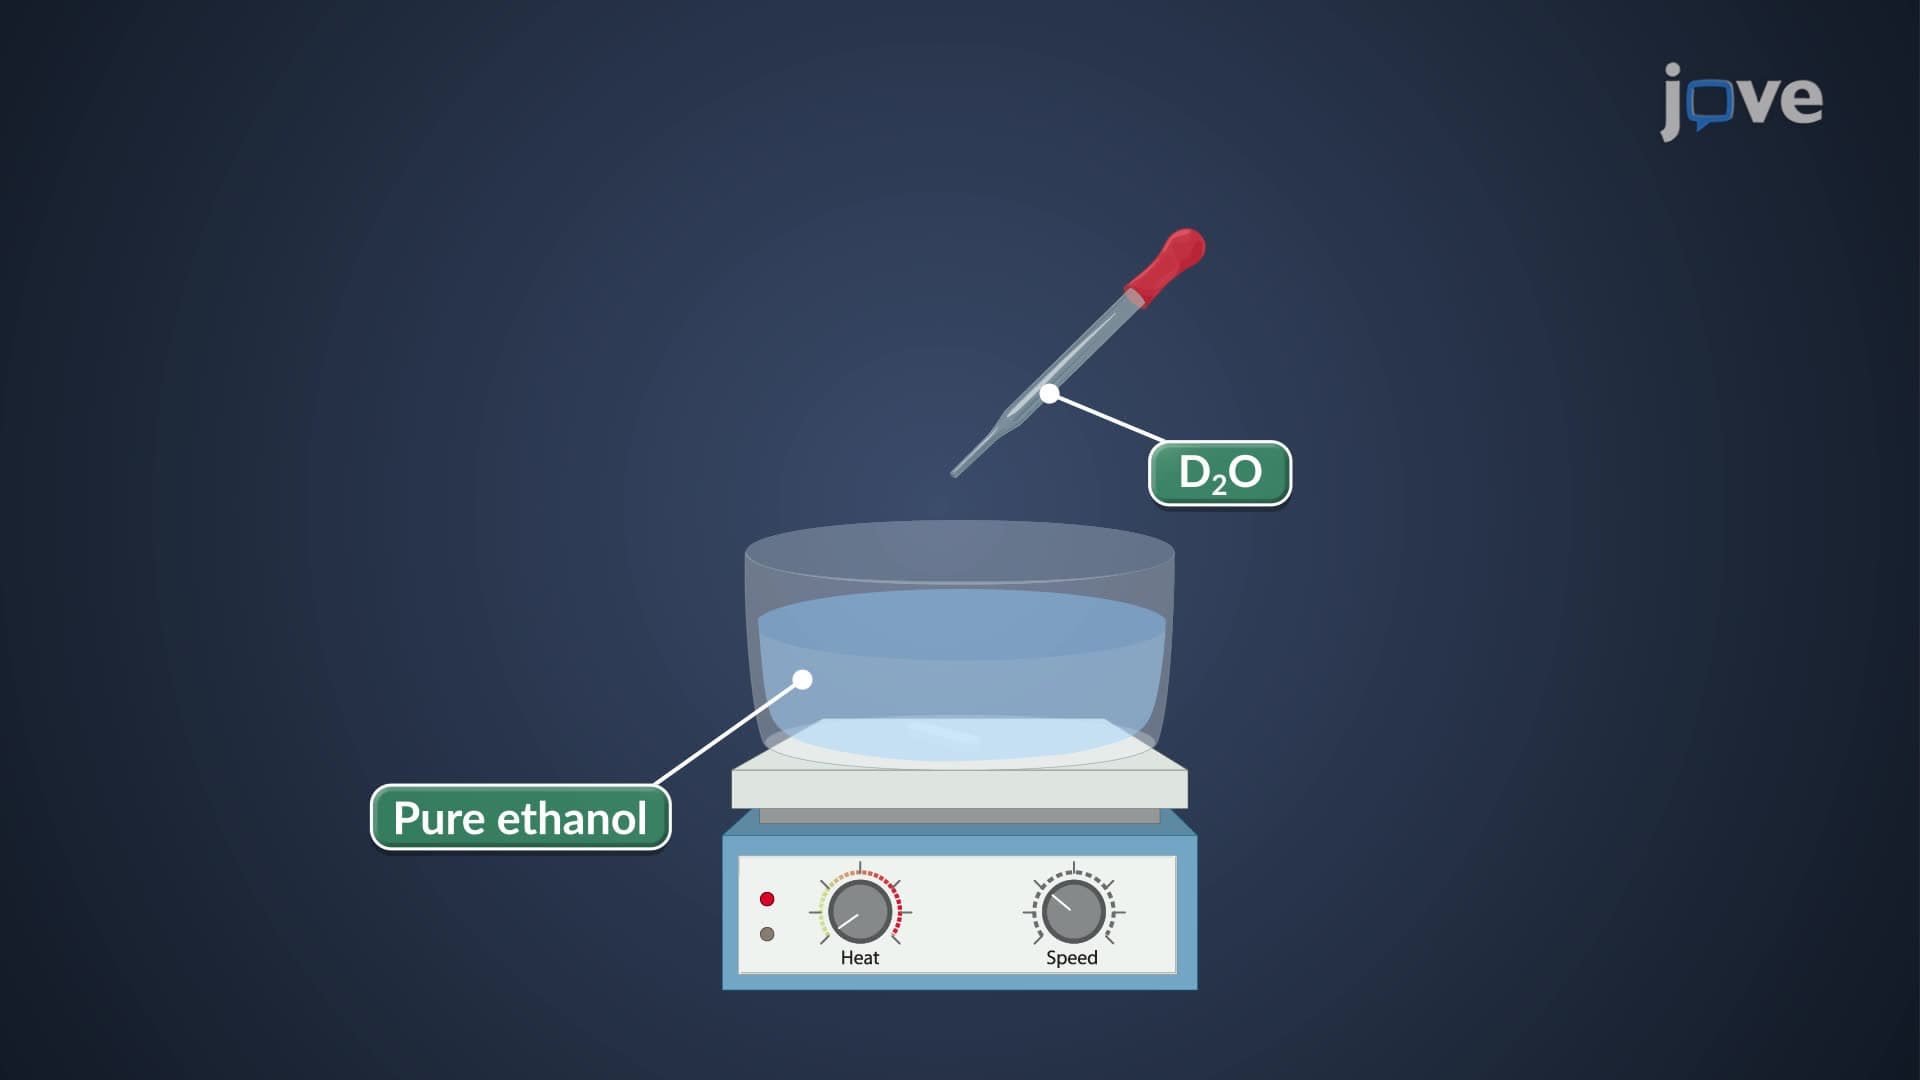
¹H NMR of Labile Protons: Deuterium (²H) Substitution
 01:20
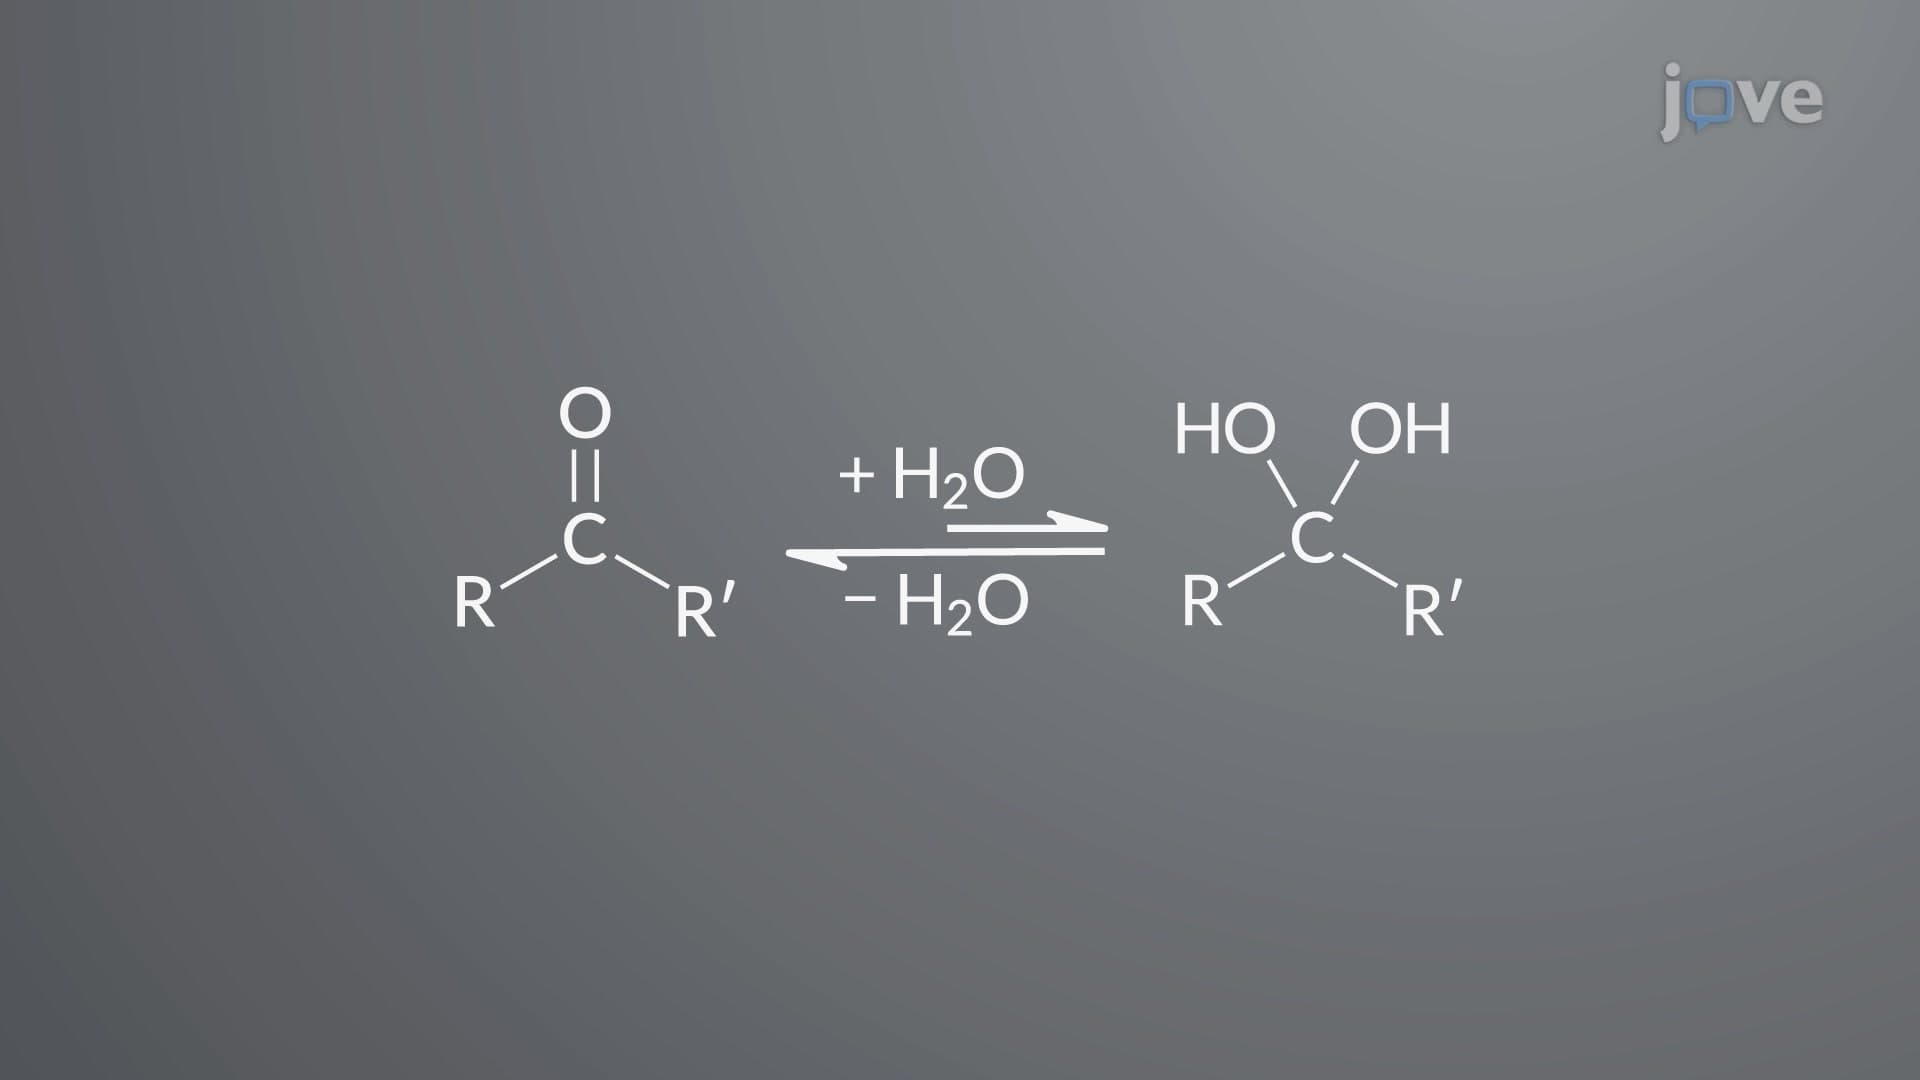
Aldehydes and Ketones with Water: Hydrate Formation
The formation of hydrates is a reversible reaction. Hydrate formation is influenced by steric and electronic factors accompanying the alkyl substituents on the carbonyl group: The rate of hydrate formation increases with a decrease in the number of alkyl groups attached to the carbonyl carbon. Hence,...
 02:34
Regioselectivity and Stereochemistry of Acid-Catalyzed Hydration
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 02:15
Preparation of Alcohols via Addition Reactions
The acid-catalyzed addition of water to the double bond of alkenes is a large-scale industrial method used to synthesize low-molecular-weight alcohols. An acidic atmosphere is required to allow the hydrogen in the water molecule to act as an electrophile and attack the double bond in an alkene. The addition of a proton to the double bond creates a carbocation intermediate. The proton preferentially bonds to the less substituted end of the double bond to create a more stable carbocation...