Related Experiment Videos
Valence- and dipole-bound anions of the thymine-water complex: ab initio characterization of the potential energy
Tomaso Frigato1, Daniel Svozil, Pavel Jungwirth
1Institute of Organic Chemistry and Biochemistry, Academy of Sciences of the Czech Republic.

Abstract:
The potential energy surfaces of the neutral and anionic thymine-water complexes are investigated using high-level ab initio calculations. Both dipole-bound (DB) and valence-bound (VB) anionic forms are considered. Four minima and three first-order stationary points are located, and binding energies are computed. All minima, for both anions, are found to be vertically and adiabatically stable. The binding energies are much higher for valence-bound than for dipole-bound anions. Adiabatic electron affinities are in the 66-287 meV range for VB anions and the 4-60 meV range for DB anions, and vertical detachment energies are in the 698-977 meV and 10-70 meV range for VB and DB anions, respectively. For cases where literature data are available, the computed values are in good agreement with previous experimental and theoretical studies. It is observed that electron attachment modifies the shape of the potential energy surfaces of the systems, especially for the valence-bound anions. Moreover, for both anions the size of the energy barrier between the two lowest energy minima is strongly reduced, rendering the coexistence of different structures more probable.
Related Concept Videos
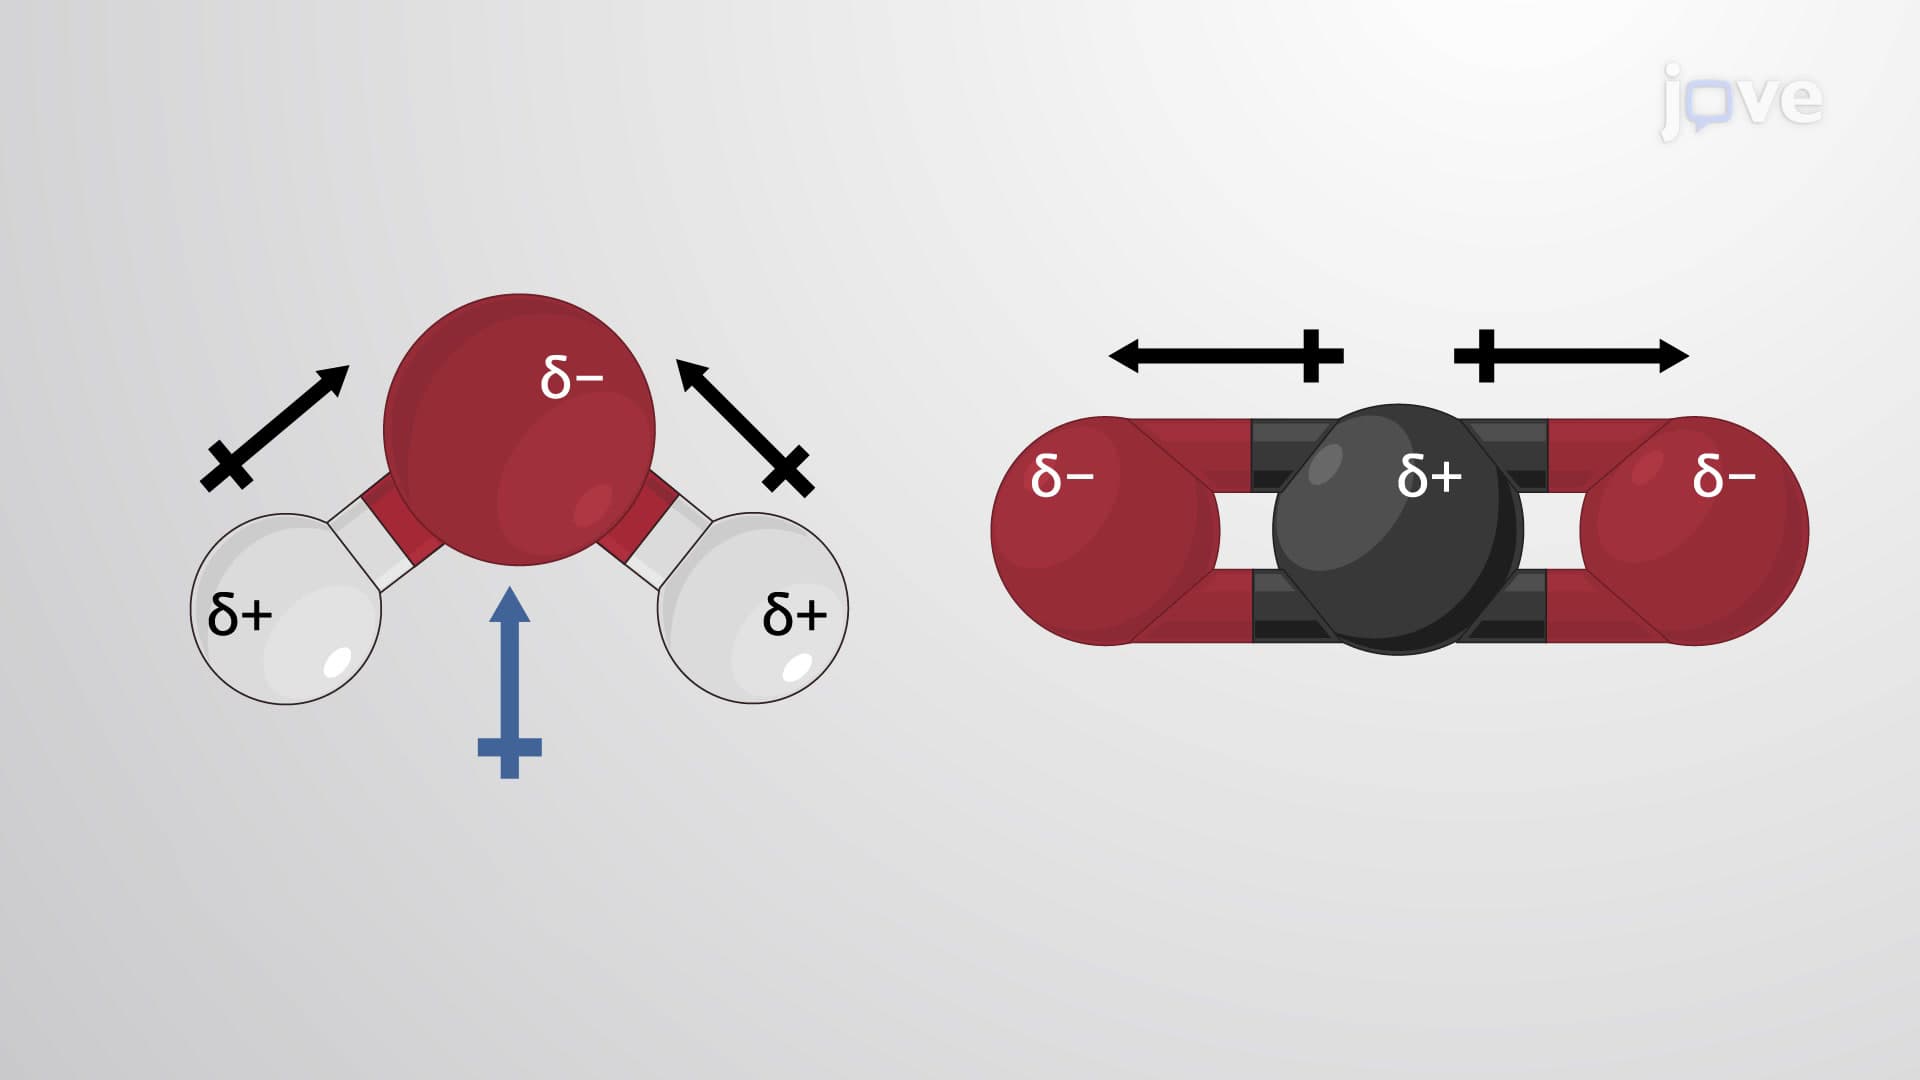 02:36
Molecular Geometry and Dipole Moments
 02:42
Valence Bond Theory
 02:45
Valence Bond Theory
 03:13
Intermolecular Forces
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:27
Predicting Molecular Geometry