Related Experiment Video
Updated: Aug 10, 2026
High Throughput, Real-time, Dual-readout Testing of Intracellular Antimicrobial Activity and Eukaryotic Cell Cytotoxicity
Published on: November 16, 2016
Pharmacodynamics of moxifloxacin and levofloxacin simulating human serum and lung concentrations
S Schubert1, A Dalhoff, H Stass
1Institute for Infection Medicine, University Hospital Schleswig-Holstein, Campus Kiel, Brunswiker Str. 4, 24105 Kiel, Germany.

Abstract:
Fluoroquinolones are known to penetrate well into the infectious foci such as lung mucosa, epithelial lining fluid and alveolar macrophages achieving higher target site concentrations than the corresponding serum levels. In order to integrate the in vitro antibacterial activity and pharmacokinetics of moxifloxacin and levofloxacin, their bactericidal efficacy was assessed by simulating human serum and lung tissue concentrations using Streptococcus pneumoniae, Staphylococcus aureus and Klebsiella pneumoniae as indicator organisms. The bacteria were exposed to fluctuating moxifloxacin and levofloxacin concentrations simulating the drug levels in serum, lung mucosa, epithelial lining fluid and alveolar macrophages. The following parameters were deduced from the kill curves: area under the bactericidal kill curve normalized to the initial inoculum (AUBKC norm), the time needed to reduce the inoculum by 3 log(10) titers, and the initial bactericidal activity. In general, all these three parameters were for all the bacterial isolates having been exposed to moxifloxacin concentration dependent. In contrast, beyond a levofloxacin concentration of optimal bactericidal effect, higher drug concentrations did not further augment the bactericidal activity of levofloxacin. These data demonstrate that not all fluoroquinolones share the same pharmacodynamic targets needed to maximize their antibacterial effect.
Related Concept Videos
 01:15
Drug Concentration Versus Time Correlation
Two pivotal parameters are the minimum effective concentration (MEC) and the minimum toxic concentration (MTC). The MEC is the lowest drug...
 01:30
Measurement of Bioavailability: Pharmacokinetic Methods
 01:20
Measurement of Bioavailability: Pharmacodynamic Methods
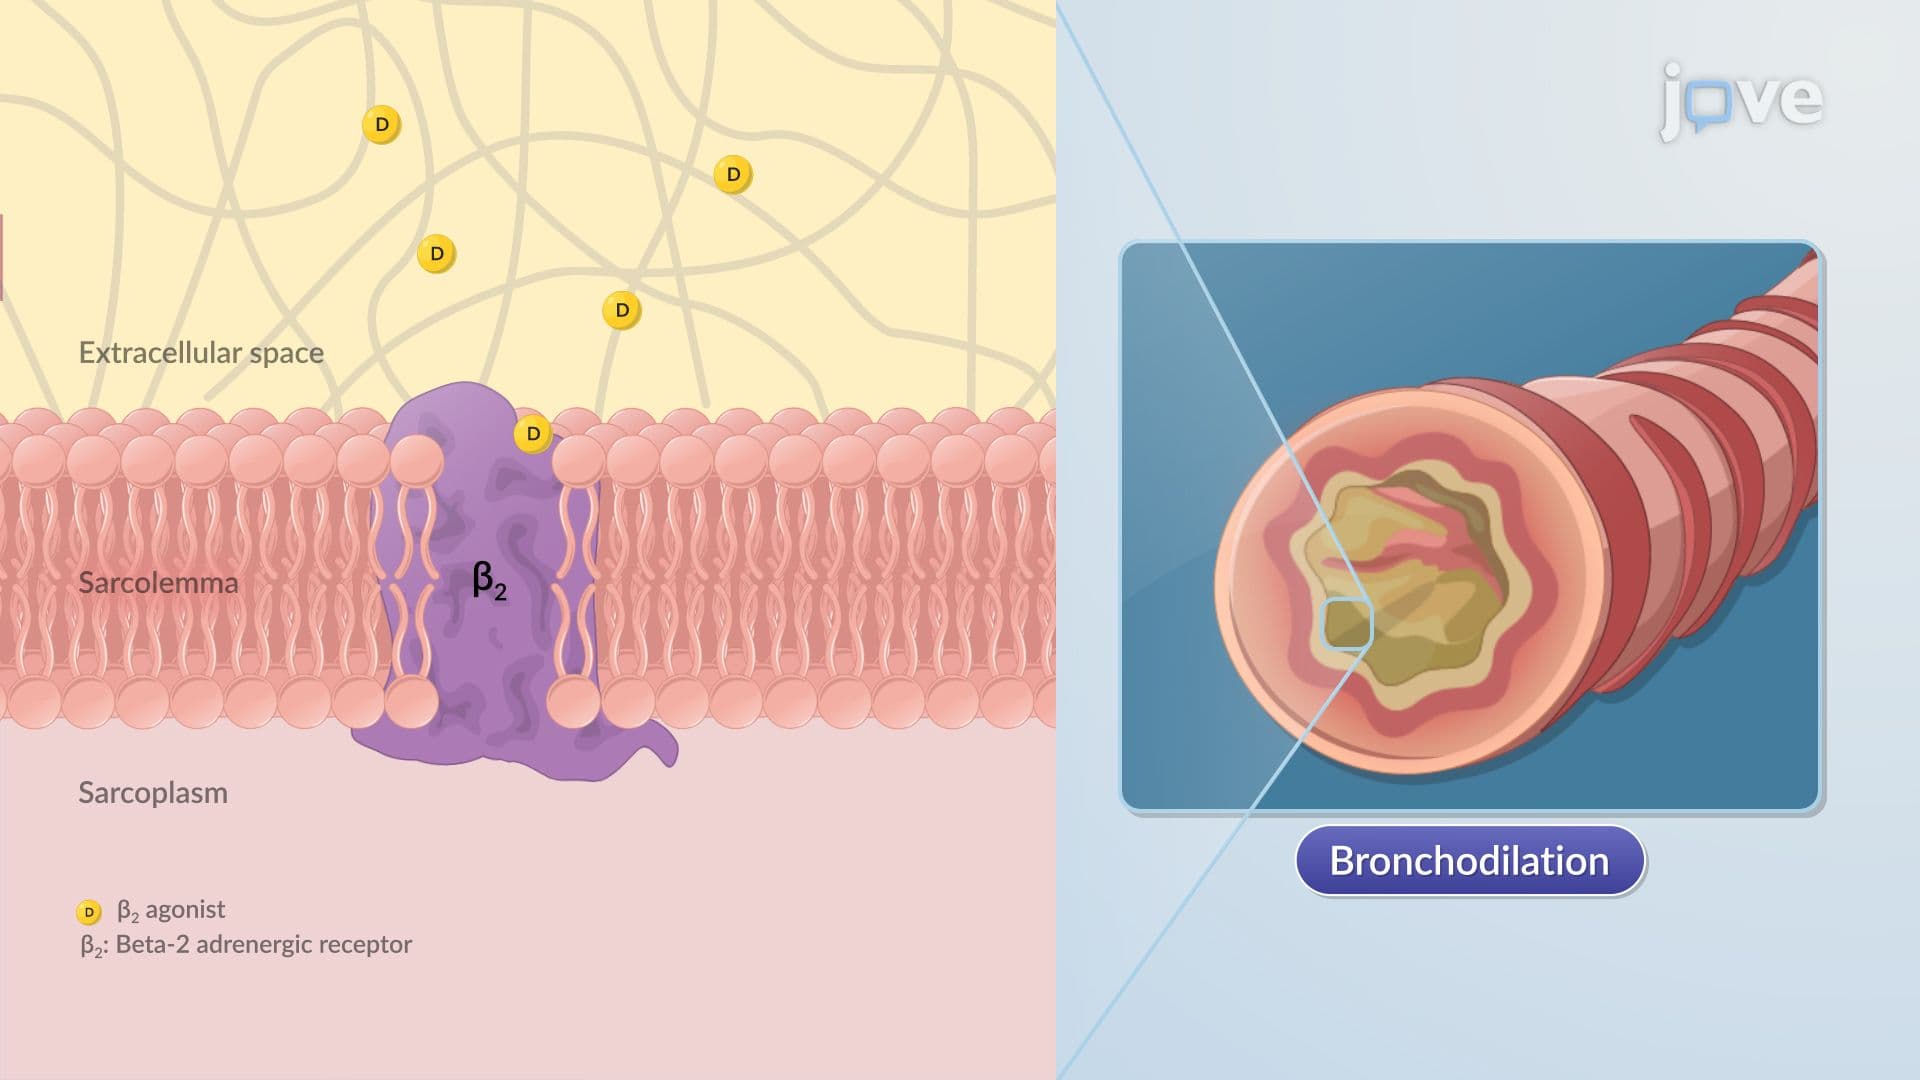 01:27
Pharmacodynamic Models: Overview
 01:15
Pharmacodynamic Models: Linear Concentration–Effect Model
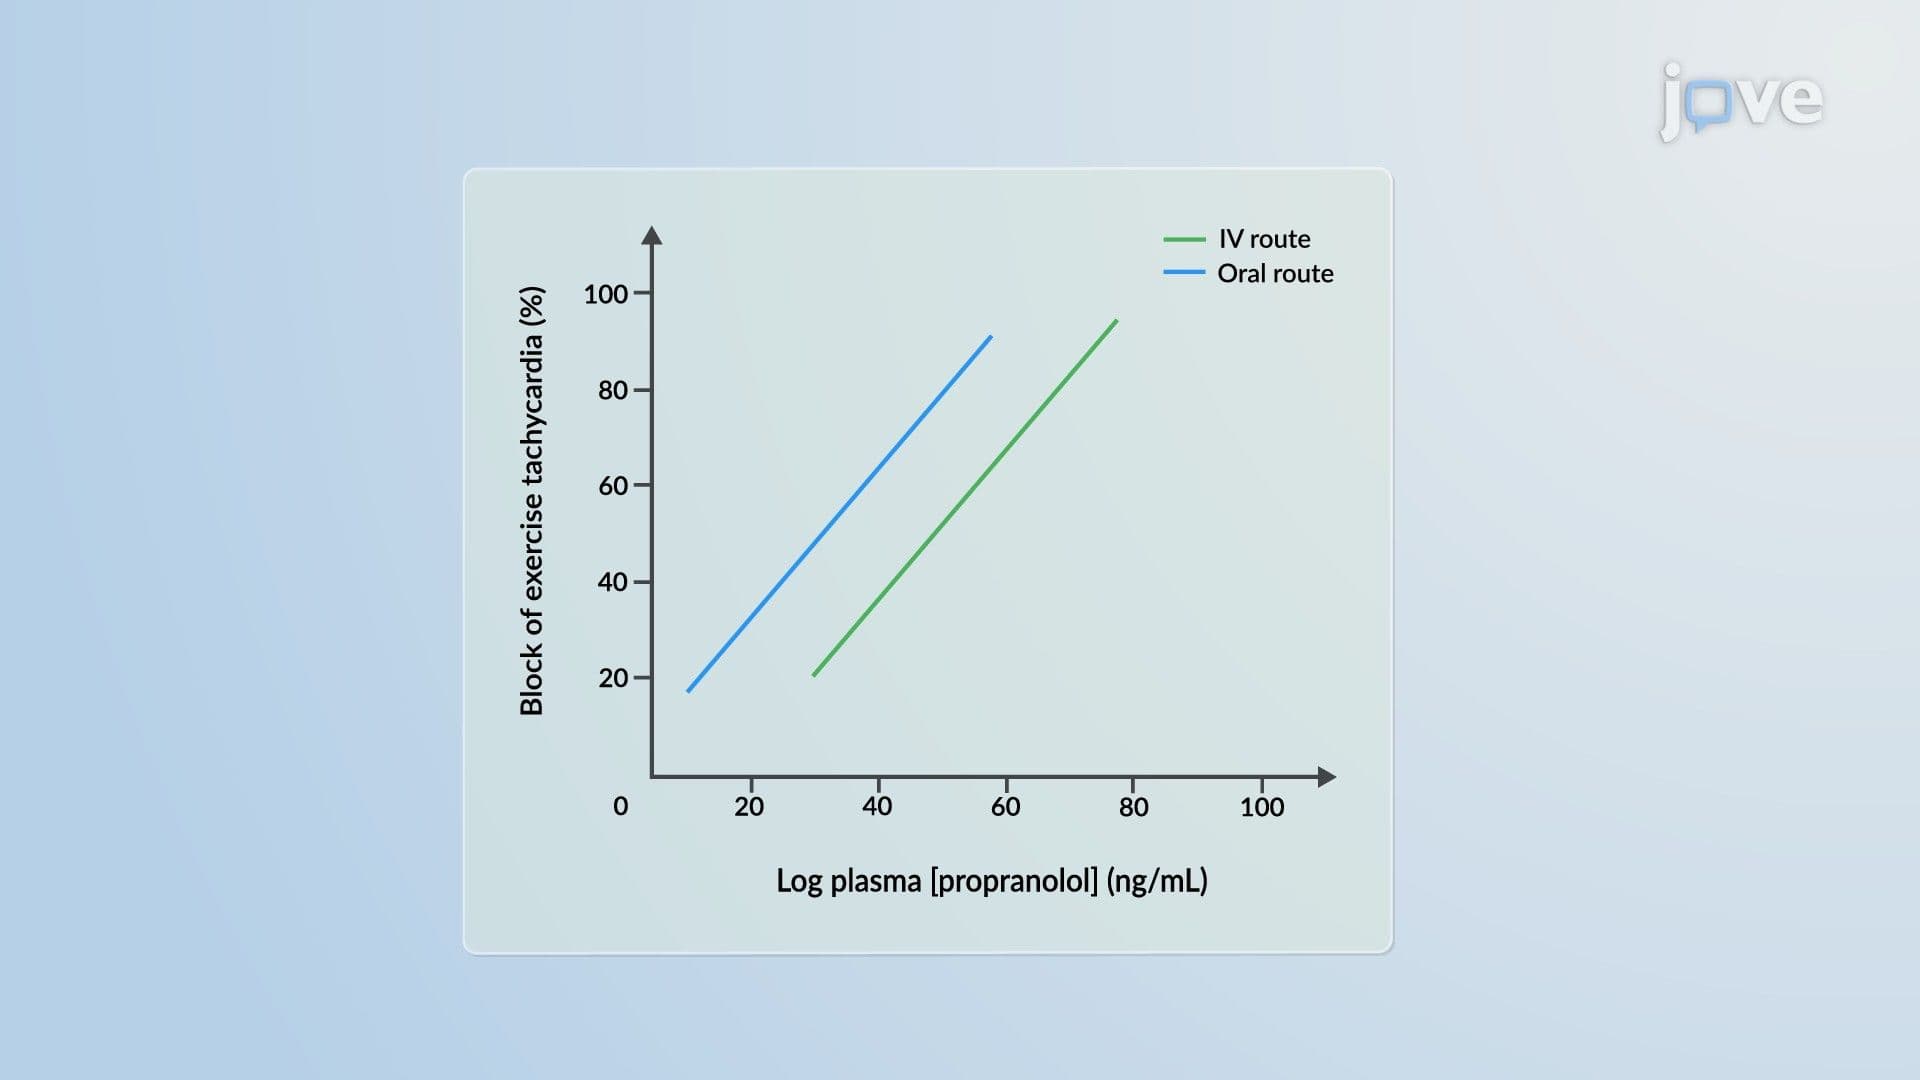 01:15
Pharmacodynamic Models: Logarithmic Concentration–Effect Model