Related Experiment Video
Updated: Aug 10, 2026
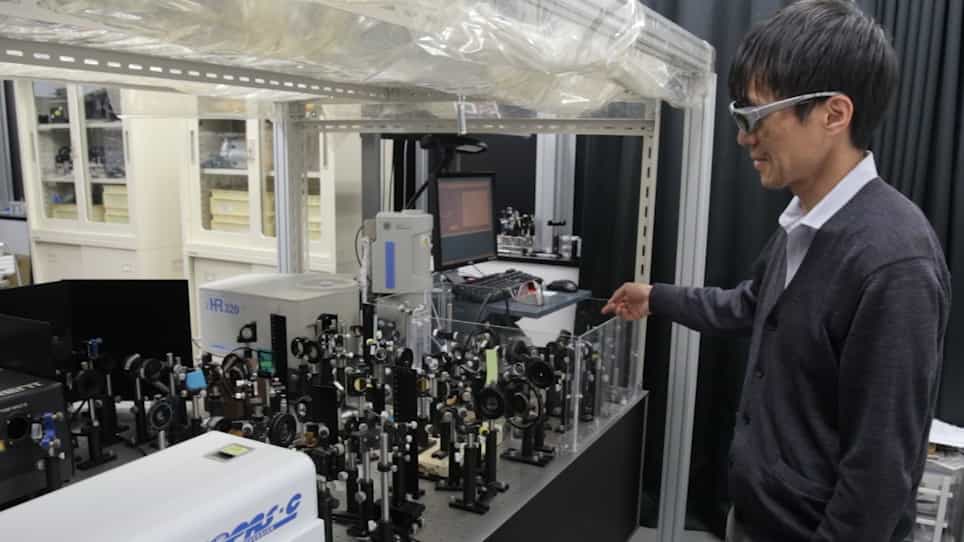
Ultrafast Time-resolved Near-IR Stimulated Raman Measurements of Functional π-conjugate Systems
Published on: February 10, 2020
Large Raman-scattering activities for the low-frequency modes of substituted benzenes: induced polarizability and
Yoshihiro Yamakita1, Yumiko Isogai, Koichi Ohno
1Department of Chemistry, Graduate School of Science, Tohoku University, Aramaki, Sendai 980-8578, Japan. yy@qpcrkk.chem.tohoku.ac.jp

Abstract:
The large nonresonant Raman-scattering activities of the out-of-plane bending and torsional modes of monosubstituted benzene analogs are studied by low-frequency Raman experiments and B3LYP6-31++G(d,p) calculations. Electronic interactions between the sigma orbitals of the substituent and the pi orbitals of the ring are found to enhance the Raman activities, depending on the substituent and its conformation. In the case of tert-butylbenzene [C6H5C(CH3)3] and trimethylphenylsilane [C6H5Si(CH3)3], three single bonds which are linked to the alpha atom of the substituent have low rotational barriers around the joint bond. Nearly free rotation of the substituents leads to a significant probability for one of the single bonds to occupy a conformation close to the vertical configuration with respect to the ring at room temperature. The resultant sigma-pi electronic interaction gives rise to the large Raman activities. In contrast, those possessing a single bond in a coplanar (or nearly coplanar) configuration at the most stable equilibrium state, i.e., anisole (C6H5OCH3), thioanisole (C6H5SCH3), and N-methylaniline (C6H5NHCH3), display no prominent Raman bands for the low-frequency vibrational modes. In these molecules, the sigma-pi conjugation does not take place due to the orthogonal orientation of the orbitals. Strong conformational dependence of the sigma-pi Raman enhancement is clearly obtained for the metastable vertical conformer of thioanisole, for which Raman activities are one-order magnitude greater than those of the coplanar conformer.
More Related Videos
 09:11
Raman and IR Spectroelectrochemical Methods as Tools to Analyze Conjugated Organic Compounds
Published on: October 12, 2018
 03:33
Fabrication of polydimethylsiloxane (PDMS)-Based Flexible Surface-Enhanced Raman Scattering (SERS) Substrate for Ultrasensitive Detection
Published on: November 17, 2023
Related Concept Videos
 01:37
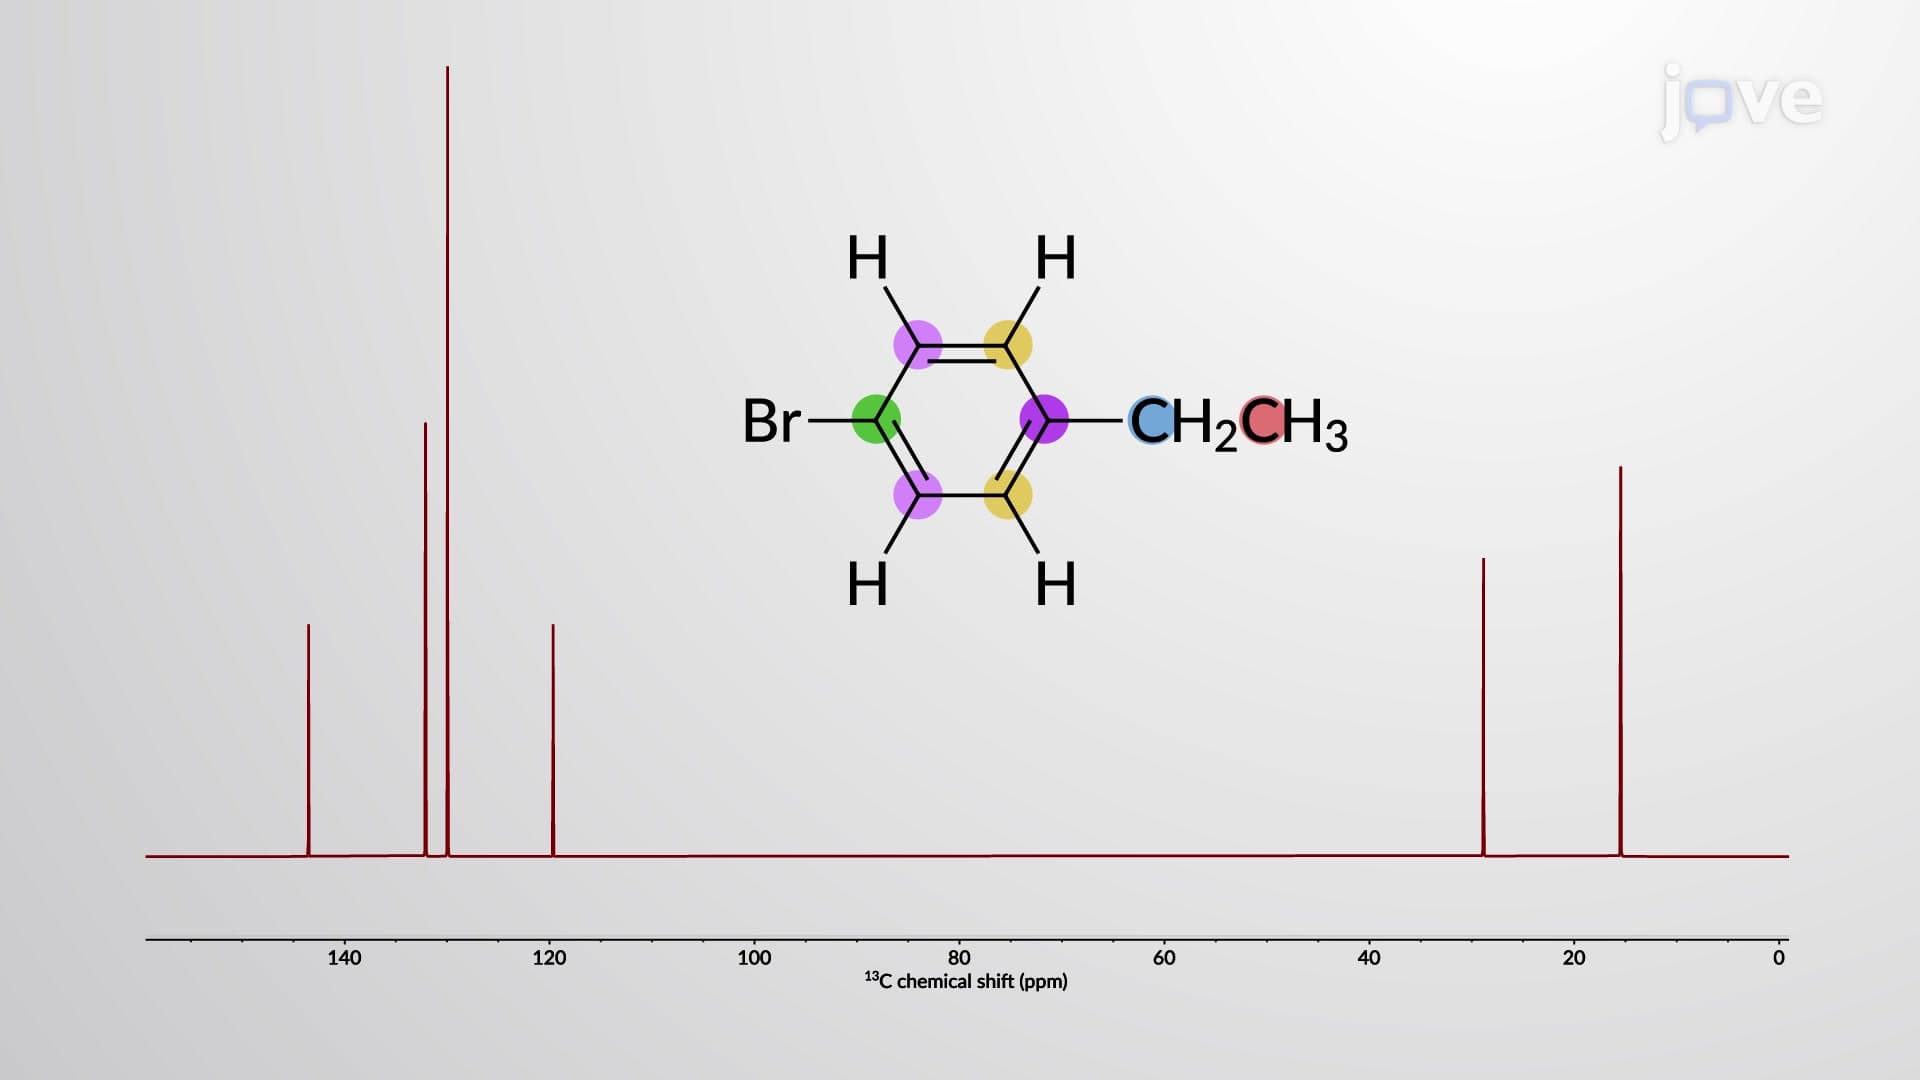
NMR Spectroscopy of Benzene Derivatives
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:20
Raman Spectroscopy: Overview
However, a small fraction of the scattered light exhibits a frequency shift due to the exchange of energy between the incident photons and the...
 01:18
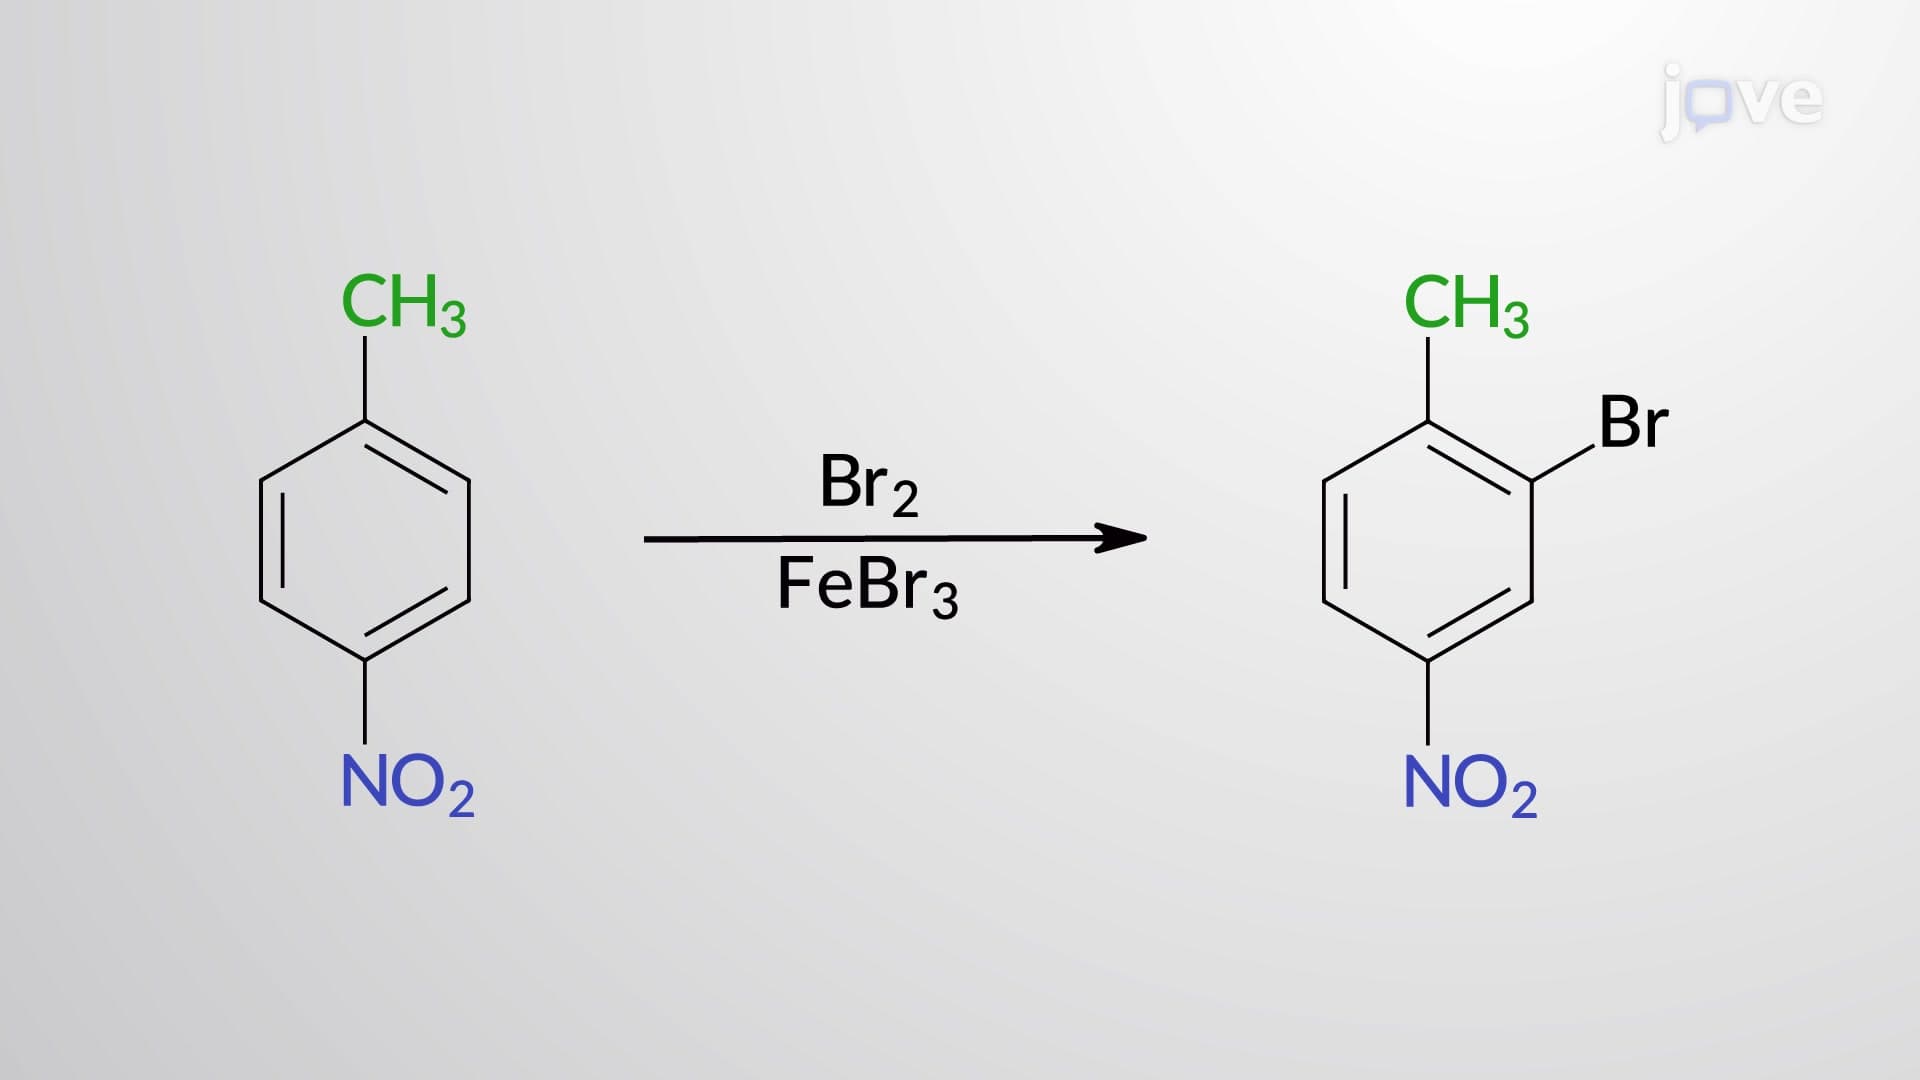
Directing and Steric Effects in Disubstituted Benzene Derivatives
 01:14
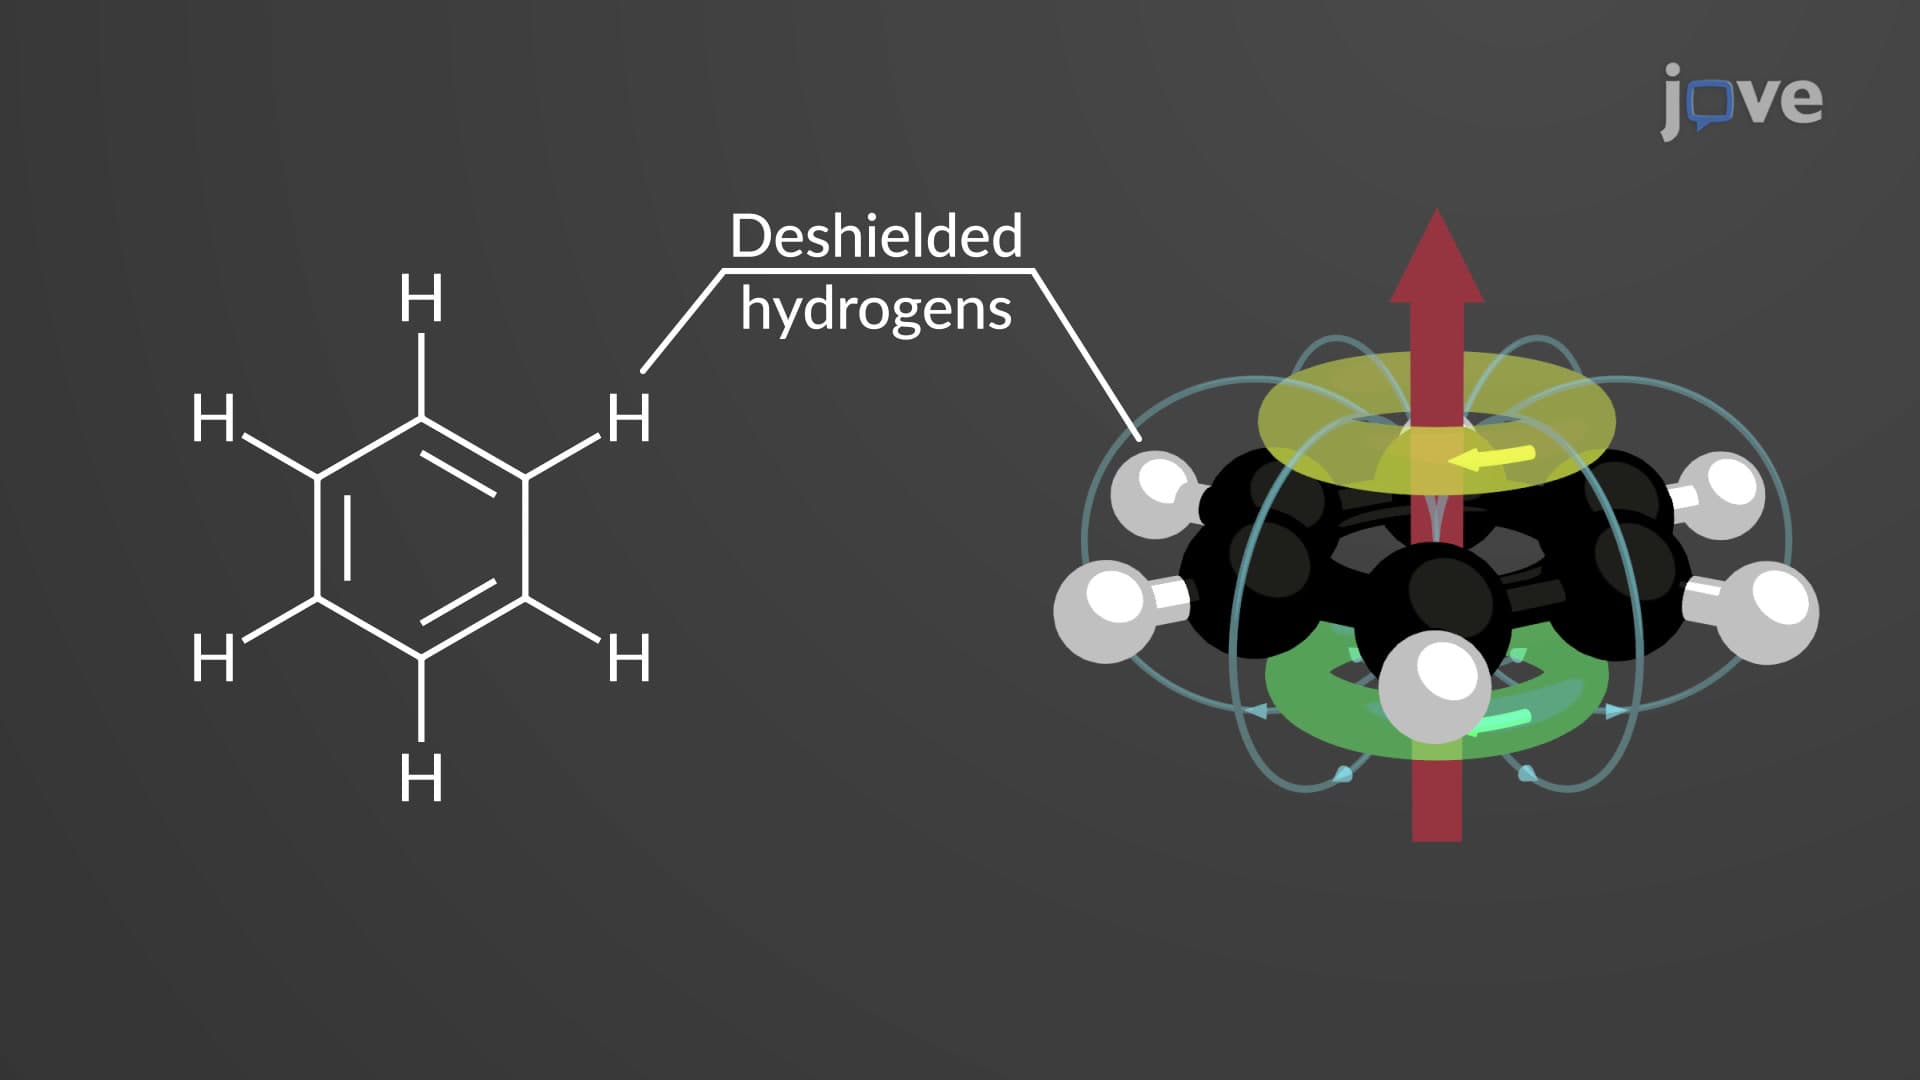
NMR Spectroscopy of Aromatic Compounds
 01:13
meta-Directing Deactivators: –NO2, –CN, –CHO, –CO2R, –COR, –CO2H