Related Experiment Video
Updated: Aug 10, 2026

Separation and Fractionation of Cell Wall and Cell Membrane Proteins from Mycobacterium tuberculosis for Downstream Protein Analysis
Published on: September 26, 2025
Computational identification of beta-barrel outer-membrane proteins in Mycobacterium tuberculosis predicted proteomes
1Meningococcal Research Department, Vaccine Division, CIGB, Cubanacán, Playa. Havana City, Cuba. rolando.pajon@cigb.edu.cu

Abstract:
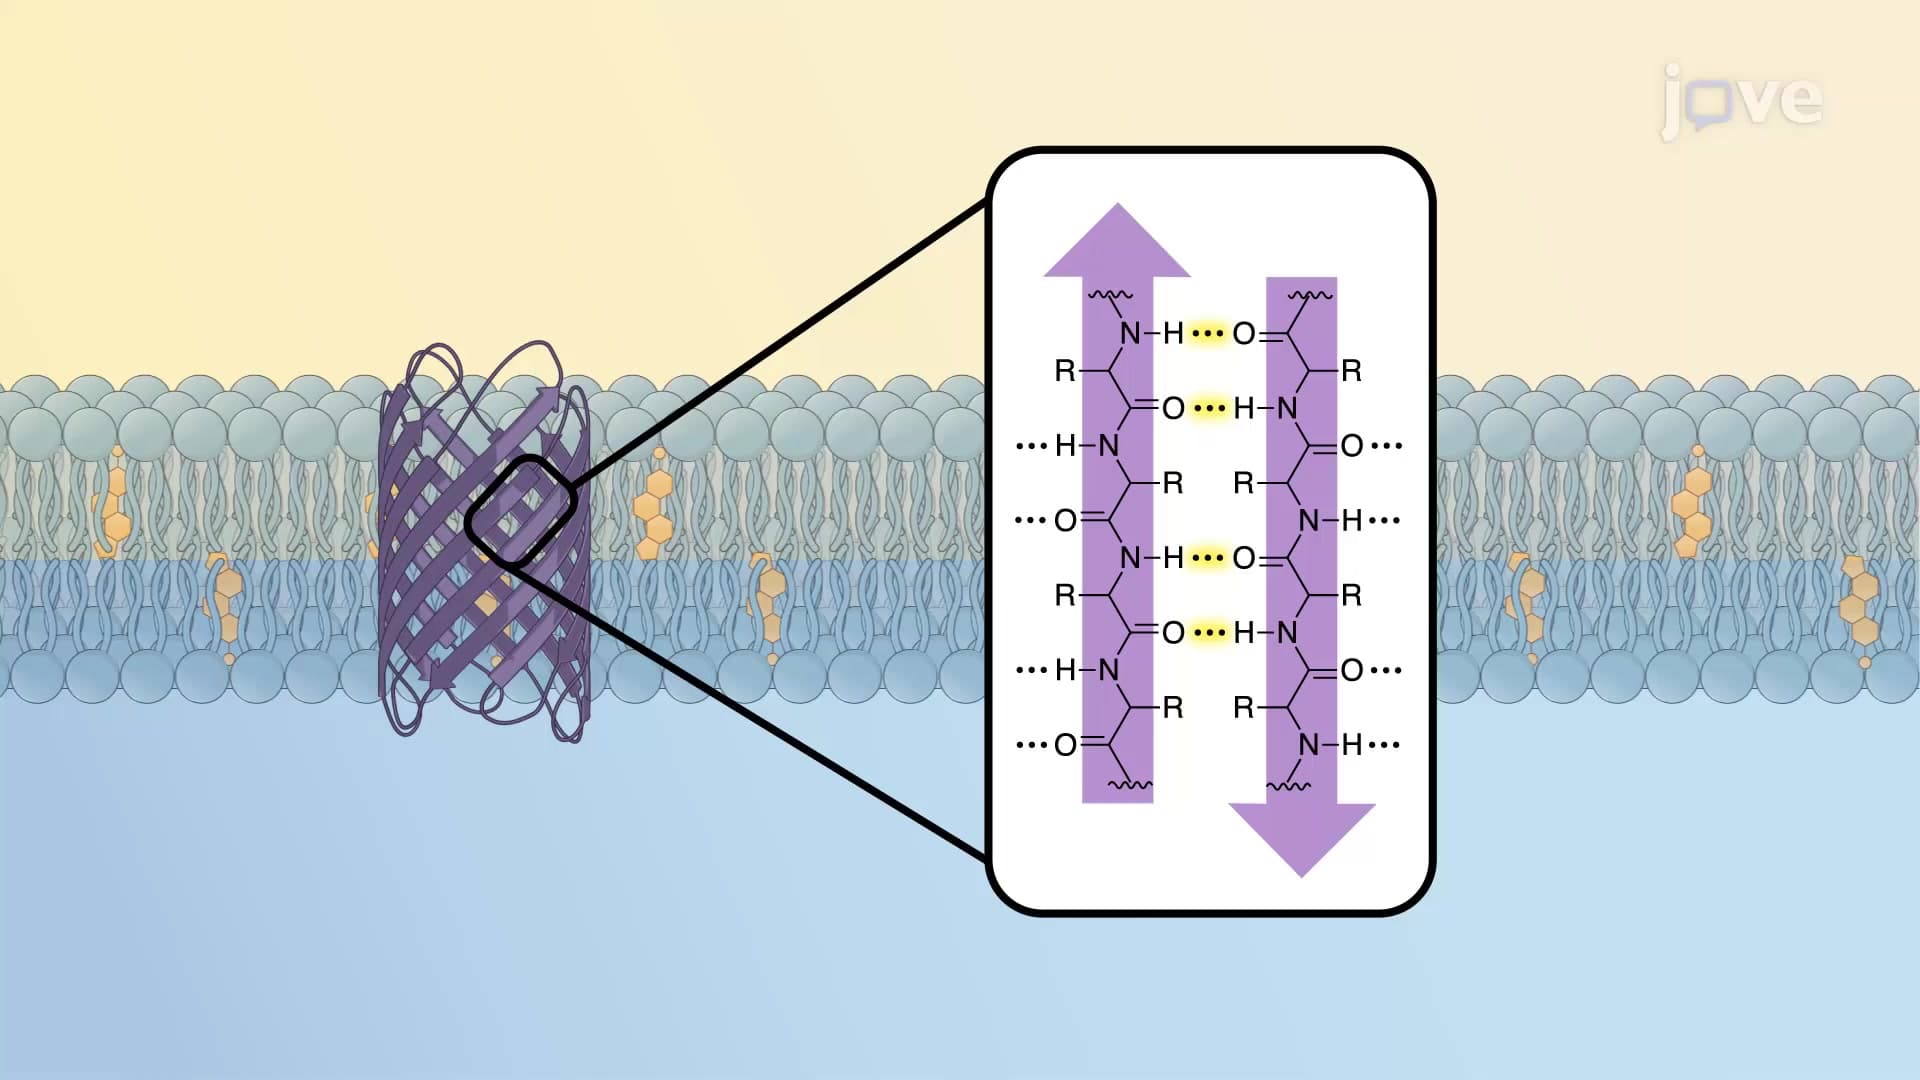
Mycobacterial porins and other beta-barrel outer-membrane proteins are represented by the structure of Mycobacterium smegmatis porin MspA. On the basis of existing knowledge of beta-barrel outer-membrane proteins, several state of the art prediction methods, as well as a new in-house program (PROB) were employed for the systematic exploration of Mycobacterium tuberculosis predicted proteomes for potential beta-barrel structures. PROB allowed parameter optimization while functioning with an adaptive algorithm for the detection of outer-membrane beta-barrel proteins in highly divergent proteomes. As a result of the predictions, 114 proteins in total were predicted to be beta-barrel structures; of these, 40 were PE-PPE proteins, 8 Mce proteins, 24 hypothetical, 11 probable membrane proteins, 10 transporters, 4 lipoproteins, and 14 classified as other. The congruence among three of the predictors, PROB, TMB-Hunt, and BOMP, was low with only three proteins (MT0318, MT0356, and MT2423) predicted by the three. Overall, 79 new proteins for which no previous experimental work has been performed are reported. At least 10 of these have high potential of being not only surface-exposed but also served as putative vaccine candidates as determined by in silico predictions of CD4T cell MHC-II restricted epitopes.
Insights
Researchers identified 114 potential beta-barrel outer-membrane proteins in Mycobacterium tuberculosis, including novel vaccine candidates. This study advances understanding of mycobacterial outer membranes and potential drug targets.
Area of Science:
- Microbiology
- Structural Biology
- Bioinformatics
Background:
- Mycobacterial outer membrane proteins, particularly beta-barrels, are crucial for cellular structure and function.
- The structure of Mycobacterium smegmatis porin MspA serves as a reference for understanding these proteins.
Purpose of the Study:
- To systematically identify potential beta-barrel outer-membrane proteins within the Mycobacterium tuberculosis proteome.
- To evaluate the efficacy of prediction methods, including a novel in-house program (PROB), for detecting these structures.
Main Methods:
- Utilized established prediction methods and a new adaptive algorithm program (PROB) for analyzing Mycobacterium tuberculosis proteomes.
- Employed in silico prediction of CD4+ T cell MHC-II restricted epitopes to assess potential vaccine candidates.
Main Results:
- Predicted a total of 114 beta-barrel structures, comprising various protein classes including PE-PPE and Mce proteins.
- Identified 79 novel proteins with no prior experimental data, with at least 10 showing potential as surface-exposed vaccine candidates.
- Observed low congruence among prediction tools (PROB, TMB-Hunt, BOMP), with only three proteins identified by all three.
Conclusions:
- The study expands the known repertoire of mycobacterial outer-membrane beta-barrel proteins.
- Identified novel potential vaccine candidates for Mycobacterium tuberculosis, warranting further experimental validation.
- Highlights the challenges and potential of computational methods in predicting membrane protein structures in divergent proteomes.