Late onset axonal Charcot-Marie-Tooth phenotype caused by a novel myelin protein zero mutation

H M E Bienfait1, C G Faber, F Baas

  • 1Department of Neurology, Academic Medical Centre, University of Amsterdam, The Netherlands.

Related Concept Videos

Multiple Sclerosis l: Introduction01:19

Multiple Sclerosis l: Introduction

Multiple sclerosis is a chronic autoimmune disease of the central nervous system (CNS) that affects the brain, spinal cord, and optic nerves. It is an inflammatory demyelinating disorder and a leading cause of neurological disability in young adults.EpidemiologyMS commonly begins between 20 and 40 years of age and is twice as common in women. Its exact cause remains unclear, but genetic susceptibility contributes, with higher risk in first-degree relatives and identical twins. A greater...
Pleiotropy01:33

Pleiotropy

Pleiotropy is the phenomenon in which a single gene impacts multiple, seemingly unrelated phenotypic traits. For example, defects in the SOX10 gene cause Waardenburg Syndrome Type 4, or WS4, which can cause defects in pigmentation, hearing impairments, and an absence of intestinal contractions necessary for elimination. This diversity of phenotypes results from the expression pattern of SOX10 in early embryonic and fetal development. SOX10 is found in neural crest cells that form melanocytes,...
Botulism01:22

Botulism

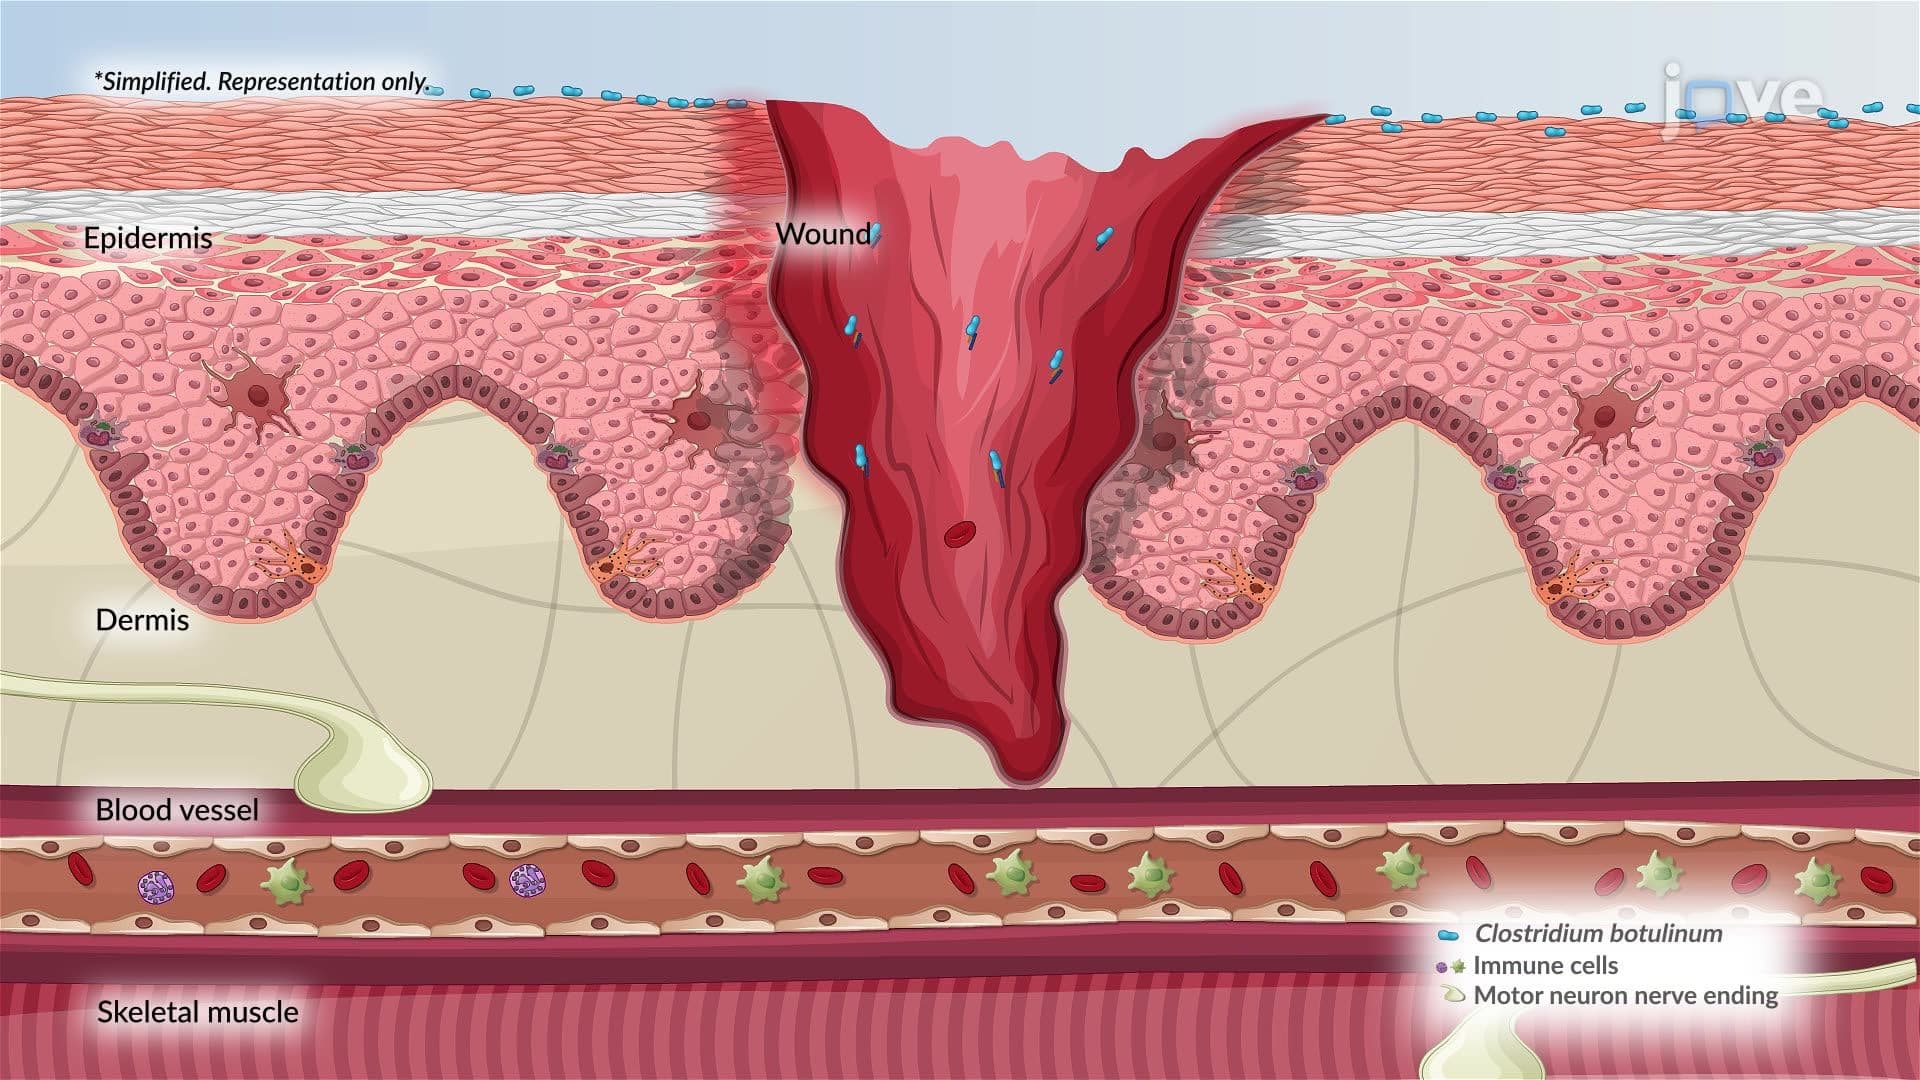
Botulism is a life-threatening neuroparalytic condition caused by botulinum neurotoxin, which is produced by the bacterium Clostridium botulinum, a Gram-positive, spore-forming, obligate anaerobe.In adults, the toxin enters the body in different ways: in foodborne botulism, the preformed toxin is absorbed in the intestine. In wound botulism, spores grow in injured tissue and release the toxin into the blood. Infant botulism differs mechanistically from adult forms. In infants, botulism commonly...
Amyloid Fibrils03:03

Amyloid Fibrils

Amyloid fibrils are aggregates of misfolded proteins.  Under most circumstances, misfolded proteins are either refolded by chaperone proteins or degraded by the proteasome. However, in the case of a mutation or a disease, these proteins can accumulate to form large clusters and often further assemble to form elongated fibers, called fibrils. 
Amyloid deposits were observed as early as 1639 in the liver and the spleen.   In 1854, Rudolph Virchow performed iodine staining, normally used to...
Myasthenia Gravis ll: Pathophysiology01:22

Myasthenia Gravis ll: Pathophysiology

The disease process of myasthenia gravis begins at the neuromuscular junction, where antibodies attack key proteins needed for muscle activation. This immune reaction weakens signal transmission, leading to the characteristic muscle fatigue and weakness that define the condition.Immune-Mediated DamageIn most individuals, antibodies target acetylcholine receptors (AChRs) on the postsynaptic membrane of muscle cells. By blocking acetylcholine binding, these antibodies prevent the nerve signal...
Huntington Disease l: Introduction01:21

Huntington Disease l: Introduction

Huntington disease or HD is a progressive, fatal neurodegenerative disorder inherited in an autosomal dominant pattern.PathophysiologyIt is caused by expansion of the CAG trinucleotide repeat in the HTT gene on chromosome 4 (4p16.3), producing an abnormal huntingtin protein with an expanded polyglutamine tract. This misfolded protein disrupts cellular function, leading to neuronal death. Normal alleles have ≤26 repeats, 27–35 are intermediate (risk of expansion), 36–39 show reduced penetrance,...