Related Experiment Video
Updated: Aug 9, 2026

Preparation and Reactivity of a Triphosphenium Bromide Salt: A Convenient and Stable Source of Phosphorus(I)
Published on: November 22, 2016
DFT calculations of structure and IR spectra of the phoshorus-containing G'0v generation dendron
V L Furer1, I I Vandyukova, J P Majoral
1Kazan State Architect and Civil Engineering University, Zelenaya, 1, Kazan 420043, Russia. furer@ksaba.ru

Abstract:
FT-IR spectra of the phosphorus-containing dendron G'(0v) generation with terminal aldehyde groups have been recorded. The structural optimization and normal mode analysis are performed for the G'(0v) on the basis of the ab initio density functional theory. This calculations gave vibrational frequencies and infrared intensities for the t,g,g- and t,-g,g-conformers of the G'(0v). The t,-g,g-conformer is 0.71 kcal/mol less stable compared to t,g,g-conformer. Relying on DFT calculations a complete vibrational assignment is proposed for different parts of the dendron. The influence of encirclement on band frequencies and intensity is studied and the information usually inaccessible is obtained. The global and local reactivity descriptors have been used to characterize the reactivity pattern of the core function and terminal groups. Our study reveals that the most reactive site in the dendron is the core function and the CH(2) side of the vinyl group is preferred for nucleophilic attack. In the dendrimer the most reactive are the terminal groups.
More Related Videos
Related Concept Videos
 02:27
Predicting Molecular Geometry
 03:35
Hybridization of Atomic Orbitals II
 02:36
Molecular Geometry and Dipole Moments
 02:52
VSEPR Theory and the Basic Shapes
 01:16
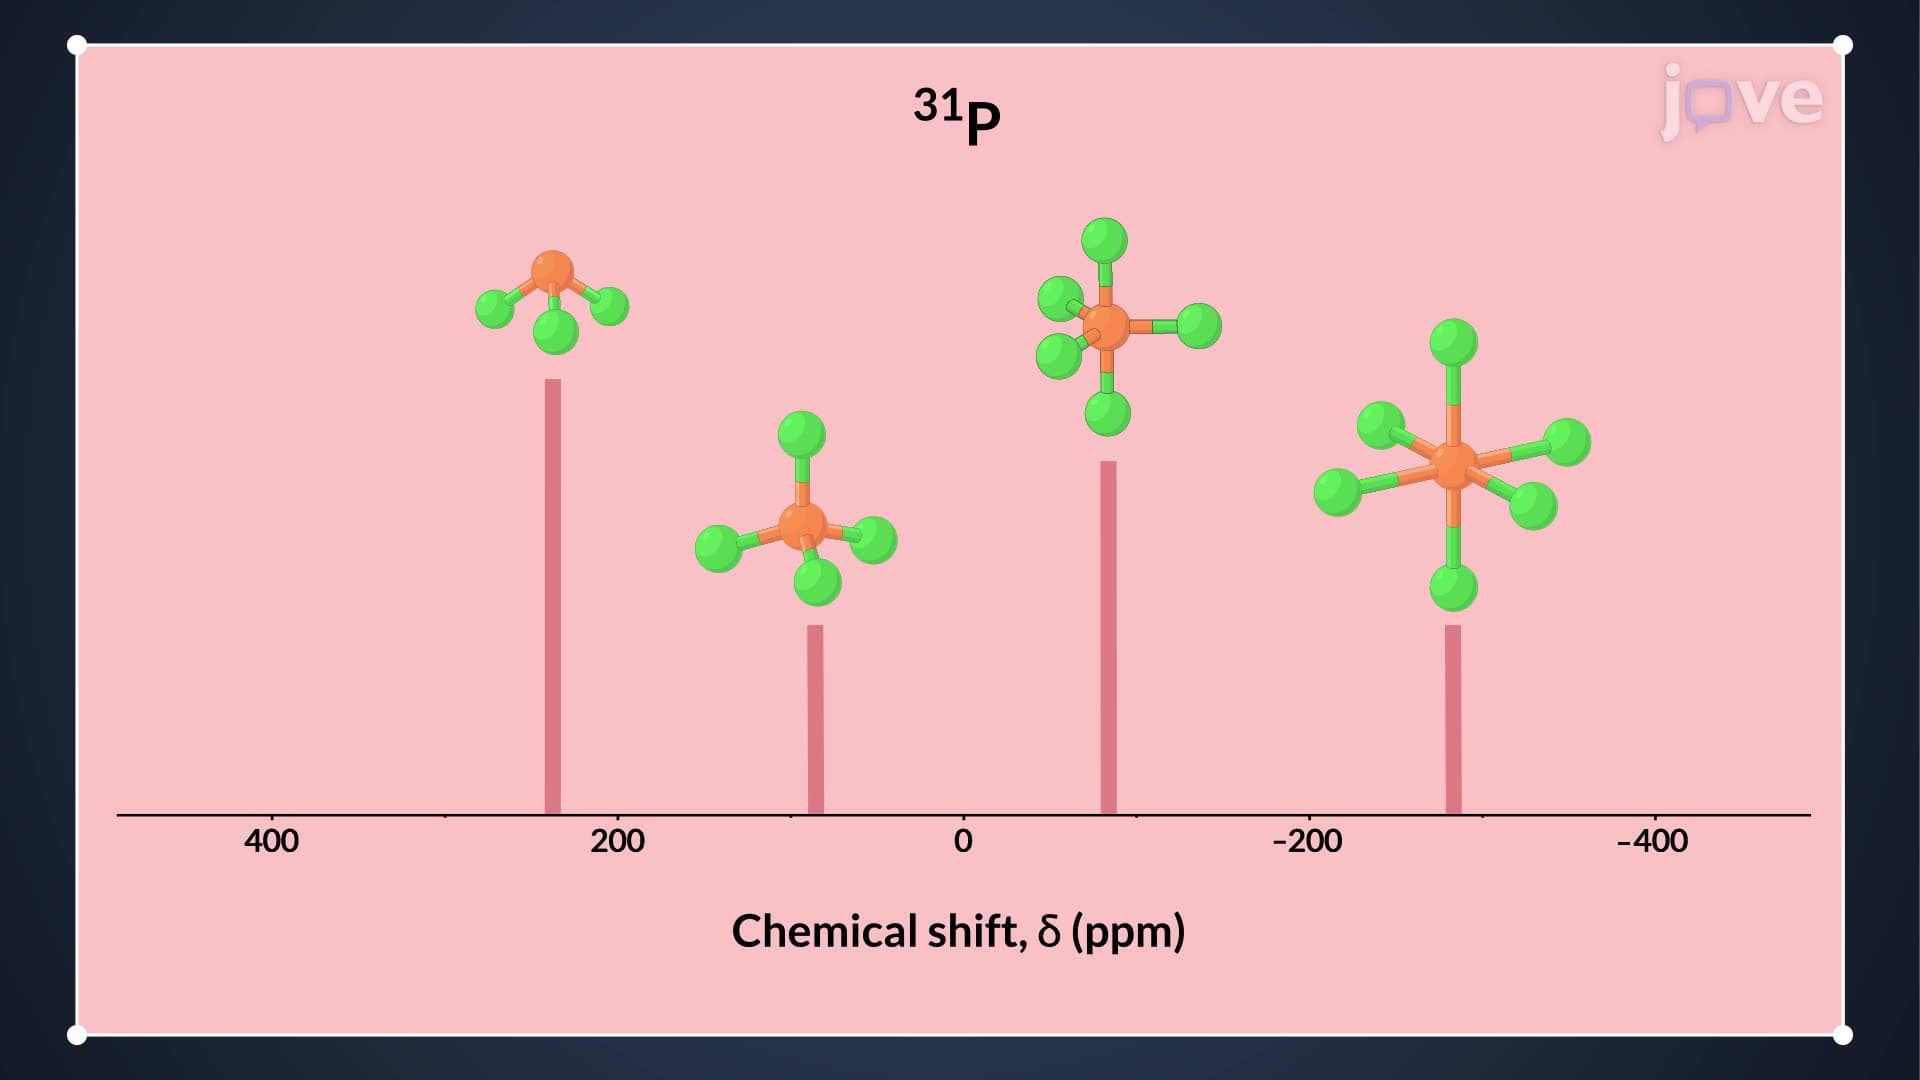
Other Nuclides: 31P, 19F, 15N NMR
While fluorine-19 and phosphorous-31 have high natural abundances (100%) and positive gyromagnetic ratios, nitrogen-15 has a low natural abundance and a negative gyromagnetic ratio. However, nitrogen-15 is still preferred over nitrogen-14 (which has a high...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...