Related Experiment Video
Updated: Aug 9, 2026
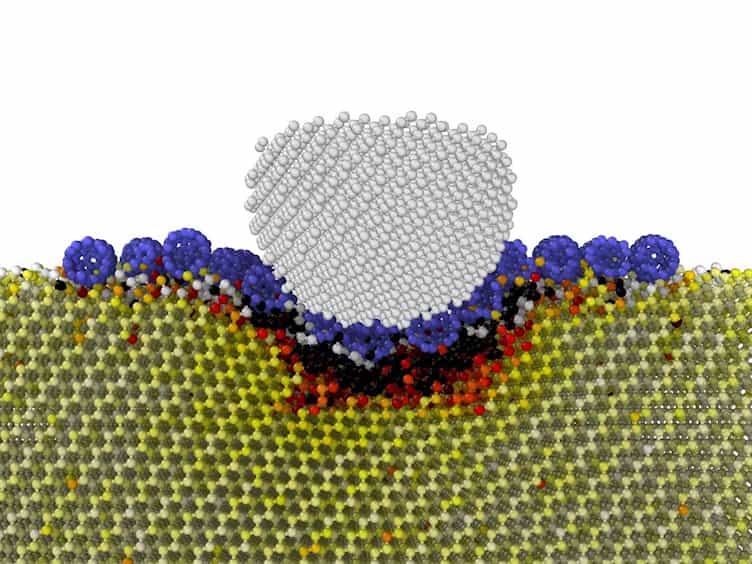

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Surface structure of (10(-)10) and (11(-)20) surfaces of ZnO with density functional theory and atomistic simulation
David J Cooke1, Arnaud Marmier, Stephen C Parker
1Department of Chemistry, University of Bath, BATH, United Kingdom.

Abstract:
We have calculated the stability of two of the low-index surfaces known to dominate the morphology of ZnO as a function of stoichiometry. These two surfaces are (10(-)10) and (11(-)20). In each case, two terminations only are stable for a significant range of oxygen and hydrogen chemical potential: the pure stoichiometric surface and a surface covered in a monolayer of water. The mode by which the water adsorbs is however different for the two surfaces considered. On the (10(-)10) surface the close proximity of the water molecules means hydrogen bonding can occur between adjacent chemiabsorbed water molecules and hence there is little difference in the stability of the hydrated and hydroxylated surface, and in fact the most stable surface occurs with a combination of dissociated and undissociated water adsorption. In the case of the (11(-)20) surface, it is only when full dissociation has occurred that a hydrogen-bonding network can form. Our results also show good agreement between DFT and atomistic simulations, suggesting that potential based methods can usefully be applied to ZnO.
Related Concept Videos
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:18
Network Covalent Solids
To break or to melt a covalent network solid, covalent bonds must be broken. Because covalent bonds are relatively strong, covalent network solids are typically...
 01:18
Structure of Benzene: Molecular Orbital Model
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:22
Structures of Solids
 01:26
Imperfections in Crystal Structure: Stoichiometric Point Defects