Related Experiment Video
Updated: Aug 9, 2026
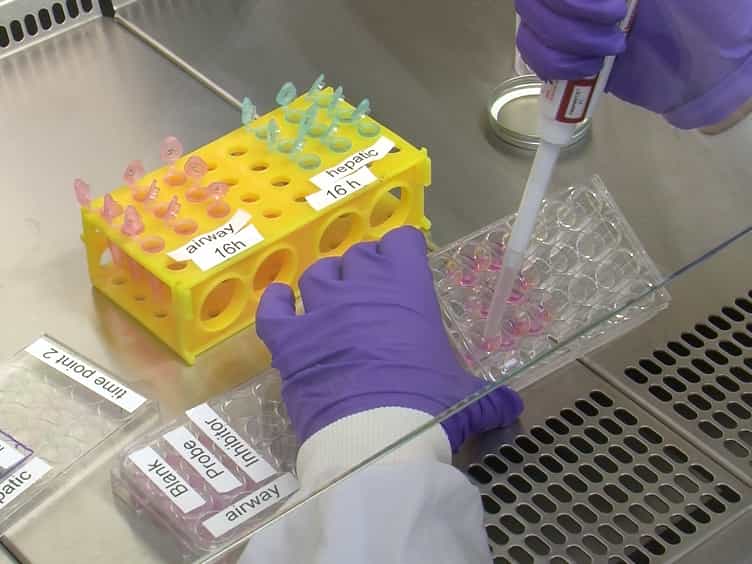

Mass Spectrometry and Luminogenic-based Approaches to Characterize Phase I Metabolic Competency of In Vitro Cell Cultures
Published on: March 28, 2017
Prediction of maximum exposure in poor metabolizers following inhibition of nonpolymorphic pathways
C Collins1, R Levy, I Ragueneau-Majlessi
1Department of Pharmaceutics, University of Washington, Seattle, 98195, USA. carolc3@u.washington.edu

Abstract:
Marked increases in exposure of some substrates have been noted in poor metabolizers given inhibitors of nonpolymorphic enzymes. Among the small number of clinical trials conducted to investigate this problem, a wide variation in the degree of maximum exposure ratios (area under the curve in poor metabolizers in the presence of inhibitor/area under the curve in extensive metabolizers) among the different substrates has been reported, with some trials reporting profound increases (> tenfold), and others demonstrating less remarkable changes (< twofold). The conduct of such trials raises safety concerns for the trial participants, in addition to other ethical and logistic concerns; therefore, the possibility was investigated that maximum exposure (area under the curve in poor metabolizers in the presence of an inhibitor) could be predicted, and that substrates susceptible to large increases in exposure could be identified. Existing clinical trials were identified by data mining the literature. A theoretical approach was developed to predict maximum exposure in poor metabolizers from studies in extensive metabolizers treated with an inhibitor of the nonpolymorphic pathway. Maximum exposure was predicted in eleven instances and the mean percentage difference between predicted and observed was 11.9%. Substrates with a fraction of substrate dose metabolized by the polymorphic enzyme (fm(POLY)) higher than 75% are at greater risk of exhibiting maximum exposure ratios of more than tenfold.
Insights
Drug exposure can significantly increase in individuals with poor drug metabolism when taking certain inhibitors. A new predictive model helps identify high-risk substrates, improving drug safety during clinical trials.
Area of Science:
- Pharmacokinetics
- Drug Metabolism
- Clinical Pharmacology
Background:
- Inhibitors of nonpolymorphic enzymes can cause substantial increases in drug exposure in poor metabolizers.
- Clinical trials show variable increases in drug exposure ratios, raising safety and ethical concerns.
Purpose of the Study:
- To develop a predictive model for maximum drug exposure in poor metabolizers when co-administered with inhibitors.
- To identify drug substrates susceptible to large increases in exposure.
Main Methods:
- Literature data mining to identify relevant clinical trials.
- Development of a theoretical approach to predict maximum exposure in poor metabolizers.
- Validation of the predictive model using existing clinical trial data.
Main Results:
- The predictive model achieved a mean percentage difference of 11.9% between predicted and observed maximum exposure.
- Substrates with a high fraction metabolized by polymorphic enzymes (fm(POLY) > 75%) are at increased risk for > tenfold exposure increases.
Conclusions:
- A theoretical model can effectively predict drug exposure in poor metabolizers.
- Identifying substrates with high fm(POLY) is crucial for assessing risks associated with enzyme inhibitors.
Related Concept Videos
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:27
Pharmacogenetics of Drug Metabolism: Overview
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:11
Pharmacogenetics of Drug Targets: β₂-Adrenergic Receptors, Apo E, Thymidylate Synthase