Related Experiment Video
Updated: Aug 8, 2026

Generating the Transcriptional Regulation View of Transcriptomic Features for Prediction Task and Dark Biomarker Detection on Small Datasets
Published on: March 1, 2024
Characterizing disease states from topological properties of transcriptional regulatory networks
David P Tuck1, Harriet M Kluger, Yuval Kluger
1Department of Pathology, Yale University School of Medicine, New Haven, Connecticut 06510, USA. david.tuck@yale.edu

Background:
High throughput gene expression experiments yield large amounts of data that can augment our understanding of disease processes, in addition to classifying samples. Here we present new paradigms of data Separation based on construction of transcriptional regulatory networks for normal and abnormal cells using sequence predictions, literature based data and gene expression studies. We analyzed expression datasets from a number of diseased and normal cells, including different types of acute leukemia, and breast cancer with variable clinical outcome.
Results:
We constructed sample-specific regulatory networks to identify links between transcription factors (TFs) and regulated genes that differentiate between healthy and diseased states. This approach carries the advantage of identifying key transcription factor-gene pairs with differential activity between healthy and diseased states rather than merely using gene expression profiles, thus alluding to processes that may be involved in gene deregulation. We then generalized this approach by studying simultaneous changes in functionality of multiple regulatory links pointing to a regulated gene or emanating from one TF (or changes in gene centrality defined by its in-degree or out-degree measures, respectively). We found that samples can often be separated based on these measures of gene centrality more robustly than using individual links. We examined distributions of distances (the number of links needed to traverse the path between each pair of genes) in the transcriptional networks for gene subsets whose collective expression profiles could best separate each dataset into predefined groups. We found that genes that optimally classify samples are concentrated in neighborhoods in the gene regulatory networks. This suggests that genes that are deregulated in diseased states exhibit a remarkable degree of connectivity.
Conclusion:
Transcription factor-regulated gene links and centrality of genes on transcriptional networks can be used to differentiate between cell types. Transcriptional network blueprints can be used as a basis for further research into gene deregulation in diseased states.
More Related Videos
 09:58
Mapping the Structure-Function Relationships of Disordered Oncogenic Transcription Factors Using Transcriptomic Analysis
Published on: June 27, 2020
 10:44
Inherent Dynamics Visualizer, an Interactive Application for Evaluating and Visualizing Outputs from a Gene Regulatory Network Inference Pipeline
Published on: December 7, 2021
Related Concept Videos
 02:23
Master Transcription Regulators
02:23Master Transcription Regulators
 01:30
General Transcription Factors
 02:02
Cis-regulatory Sequences
02:02Cis-regulatory Sequences
 01:30
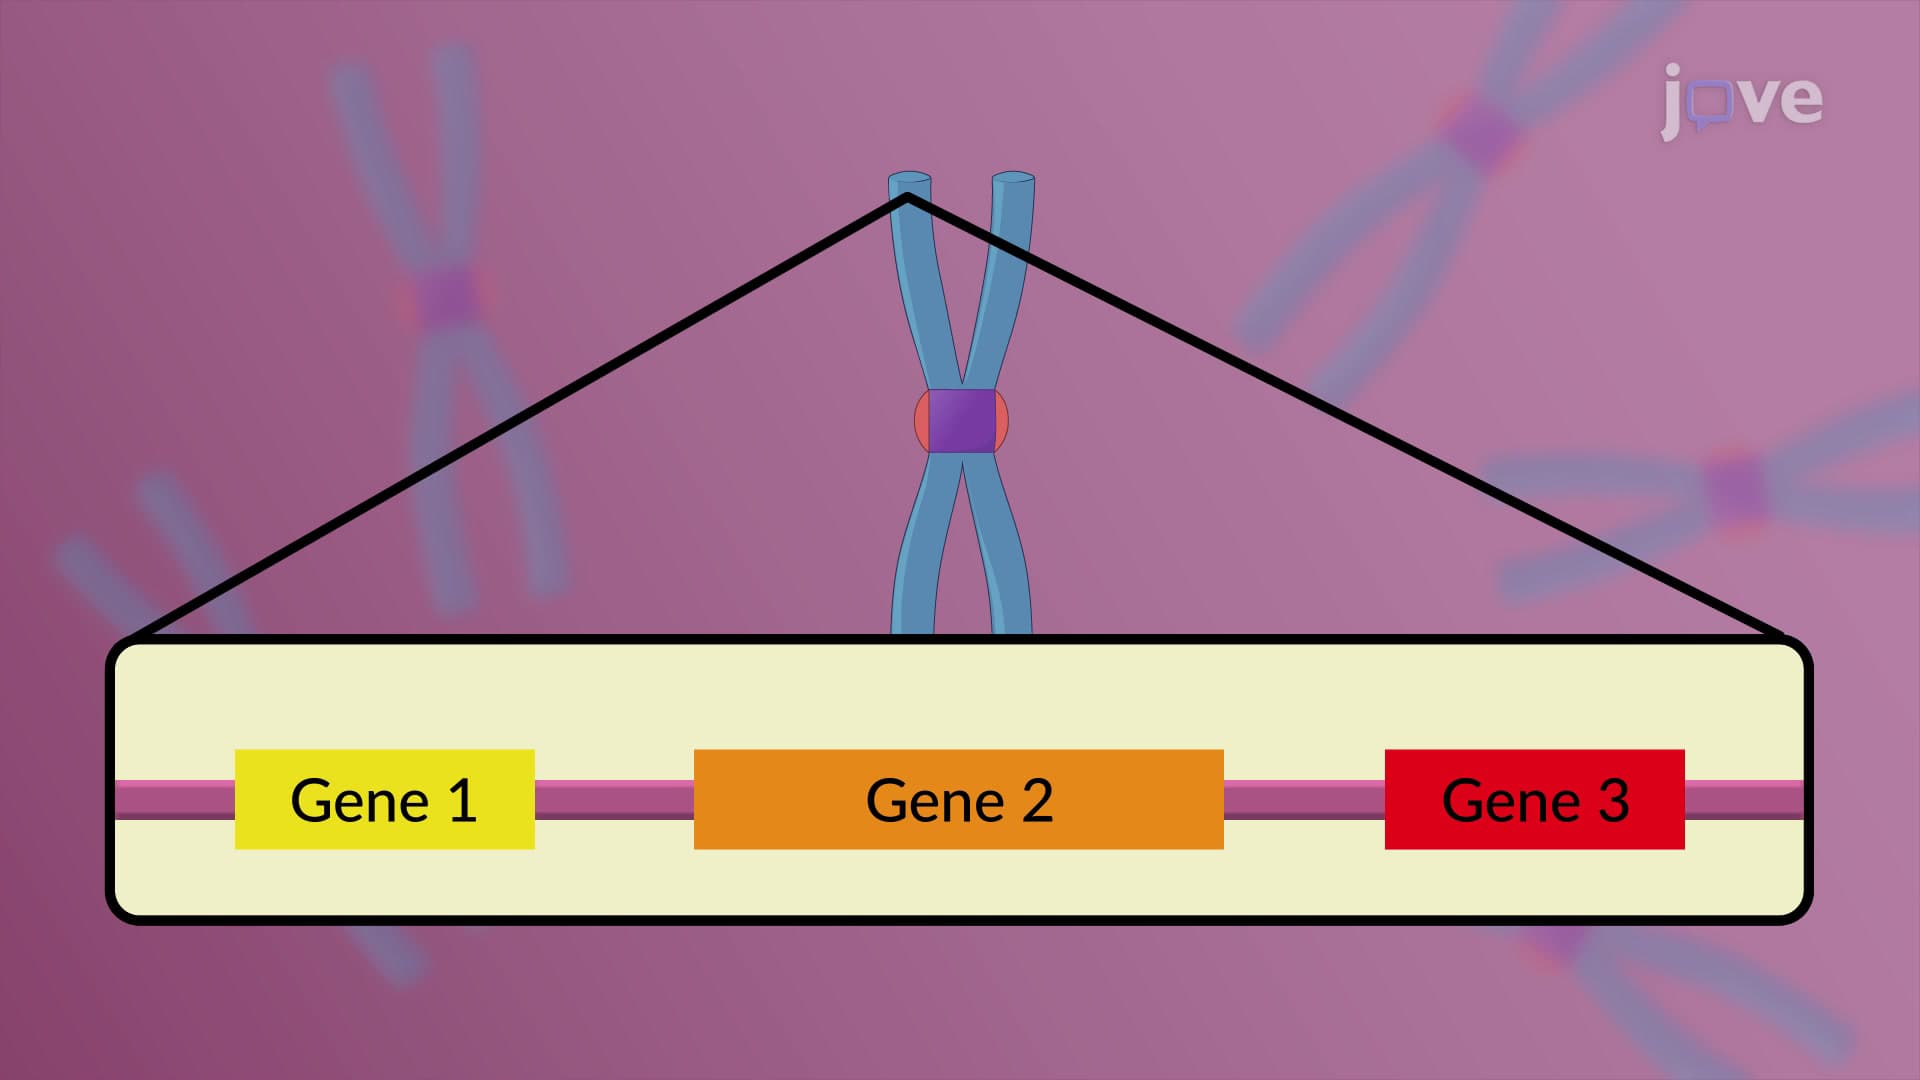
Structure of a Gene
However, only 1% of the DNA is composed of genes that encode proteins; the rest, 99% is non-coding DNA. This non-coding DNA performs...