Theoretical study of alkali cation-benzene complexes: potential energy surfaces and binding energies with improved results for rubidium and cesium
Related Experiment Video
Updated: May 1, 2026
1,3,5-Triphenylbenzene and Corannulene as Electron Receptors for Lithium Solvated Electron Solutions
Published on: October 10, 2016
Cecilia Coletti1, Nazzareno Re
1Dipartimento di Scienze del Farmaco, Università G. d'Annunzio, Via dei Vestini 31, 66100 Chieti, Italy.

Abstract:
High level ab initio quantum chemical calculations have been carried out on the binding of alkali metal to benzene with special attention to heavier metals for which the agreement between the most recent theoretical investigations and the experimental bond dissociation energies (BDEs) is not very good. We performed BSSE-corrected geometry optimizations employing the MP2 level of theory with large basis sets and a modified Stuttgart RSC 1997 basis set for rubidium and cesium and carried out single point energy calculations at the MP4 level, obtaining, also for the latter metals, BDE values in good agreement with the experimental results. Furthermore, in view of the development of empirical correction terms to force fields to describe cation-pi interactions, we evaluated the potential energy surface along the benzene symmetry axis and discussed the role of the BSSE correction on the accuracy of our results.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 16:11
Thermochemical Studies of NiII and ZnII Ternary Complexes Using Ion Mobility-Mass Spectrometry
Published on: June 8, 2022
Related Concept Videos
 02:48
Ionic Bonding and Electron Transfer
 02:45
Valence Bond Theory
 03:51
Molecular Orbital Theory II
 02:42
Valence Bond Theory
 01:07
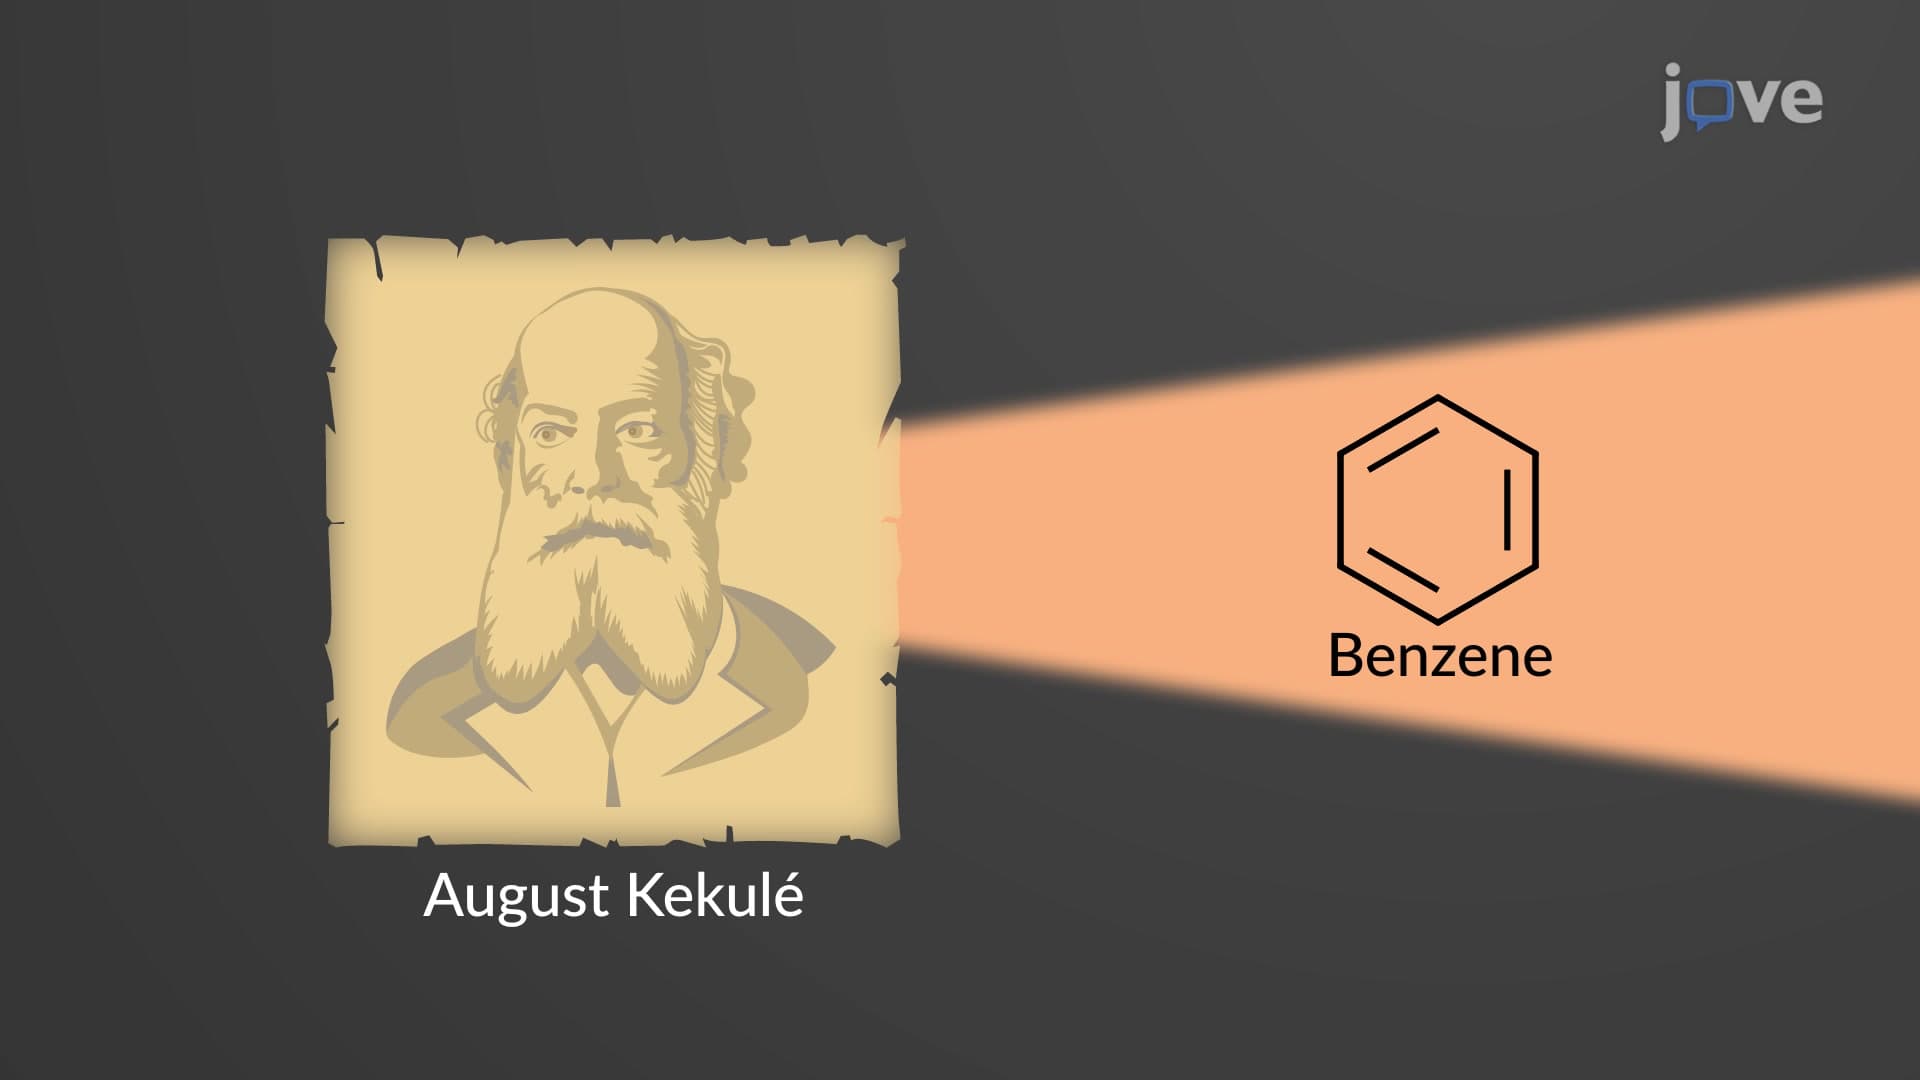
Structure of Benzene: Kekulé Model
He proposed that benzene has a cyclic structure of six carbon atoms attached to one hydrogen atom each, with three alternating pi bonds.
 01:18
Structure of Benzene: Molecular Orbital Model