Related Experiment Video
Updated: Aug 8, 2026

Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Imprint of evolutionary conservation and protein structure variation on the binding function of protein tyrosine
1Department of Pharmacology, University of California San Diego 9500 Gilman Drive, La Jolla, CA 92093-0392, USA. gverkhiv@ucsd.edu

Motivation:
According to the models of divergent molecular evolution, the evolvability of new protein function may depend on the induction of new phenotypic traits by a small number of mutations of the binding site residues. Evolutionary relationships between protein kinases are often employed to infer inhibitor binding profiles from sequence analysis. However, protein kinases binding profiles may display inhibitor selectivity within a given kinase subfamily, while exhibiting cross-activity between kinases that are phylogenetically remote from the prime target. The emerging insights into kinase function and evolution combined with a rapidly growing number of publically available crystal structures of protein kinases complexes have motivated structural bioinformatics analysis of sequence-structure relationships in determining the binding function of protein tyrosine kinases.
Results:
In silico profiling of Imatinib mesylate and PD-173955 kinase inhibitors with protein tyrosine kinases is conducted on kinome scale by using evolutionary analysis and fingerprinting inhibitor-protein interactions with the panel of all publically available protein tyrosine kinases crystal structures. We have found that sequence plasticity of the binding site residues alone may not be sufficient to enable protein tyrosine kinases to readily evolve novel binding activities with inhibitors. While evolutionary signal derived solely from the tyrosine kinase sequence conservation can not be readily translated into the ligand binding phenotype, the proposed structural bioinformatics analysis can discriminate a functionally relevant kinase binding signal from a simple phylogenetic relationship. The results of this work reveal that protein conformational diversity is intimately linked with sequence plasticity of the binding site residues in achieving functional adaptability of protein kinases towards specific drug binding. This study offers a plausible molecular rationale to the experimental binding profiles of the studied kinase inhibitors and provides a theoretical basis for constructing functionally relevant kinase binding trees.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
01:49Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:26
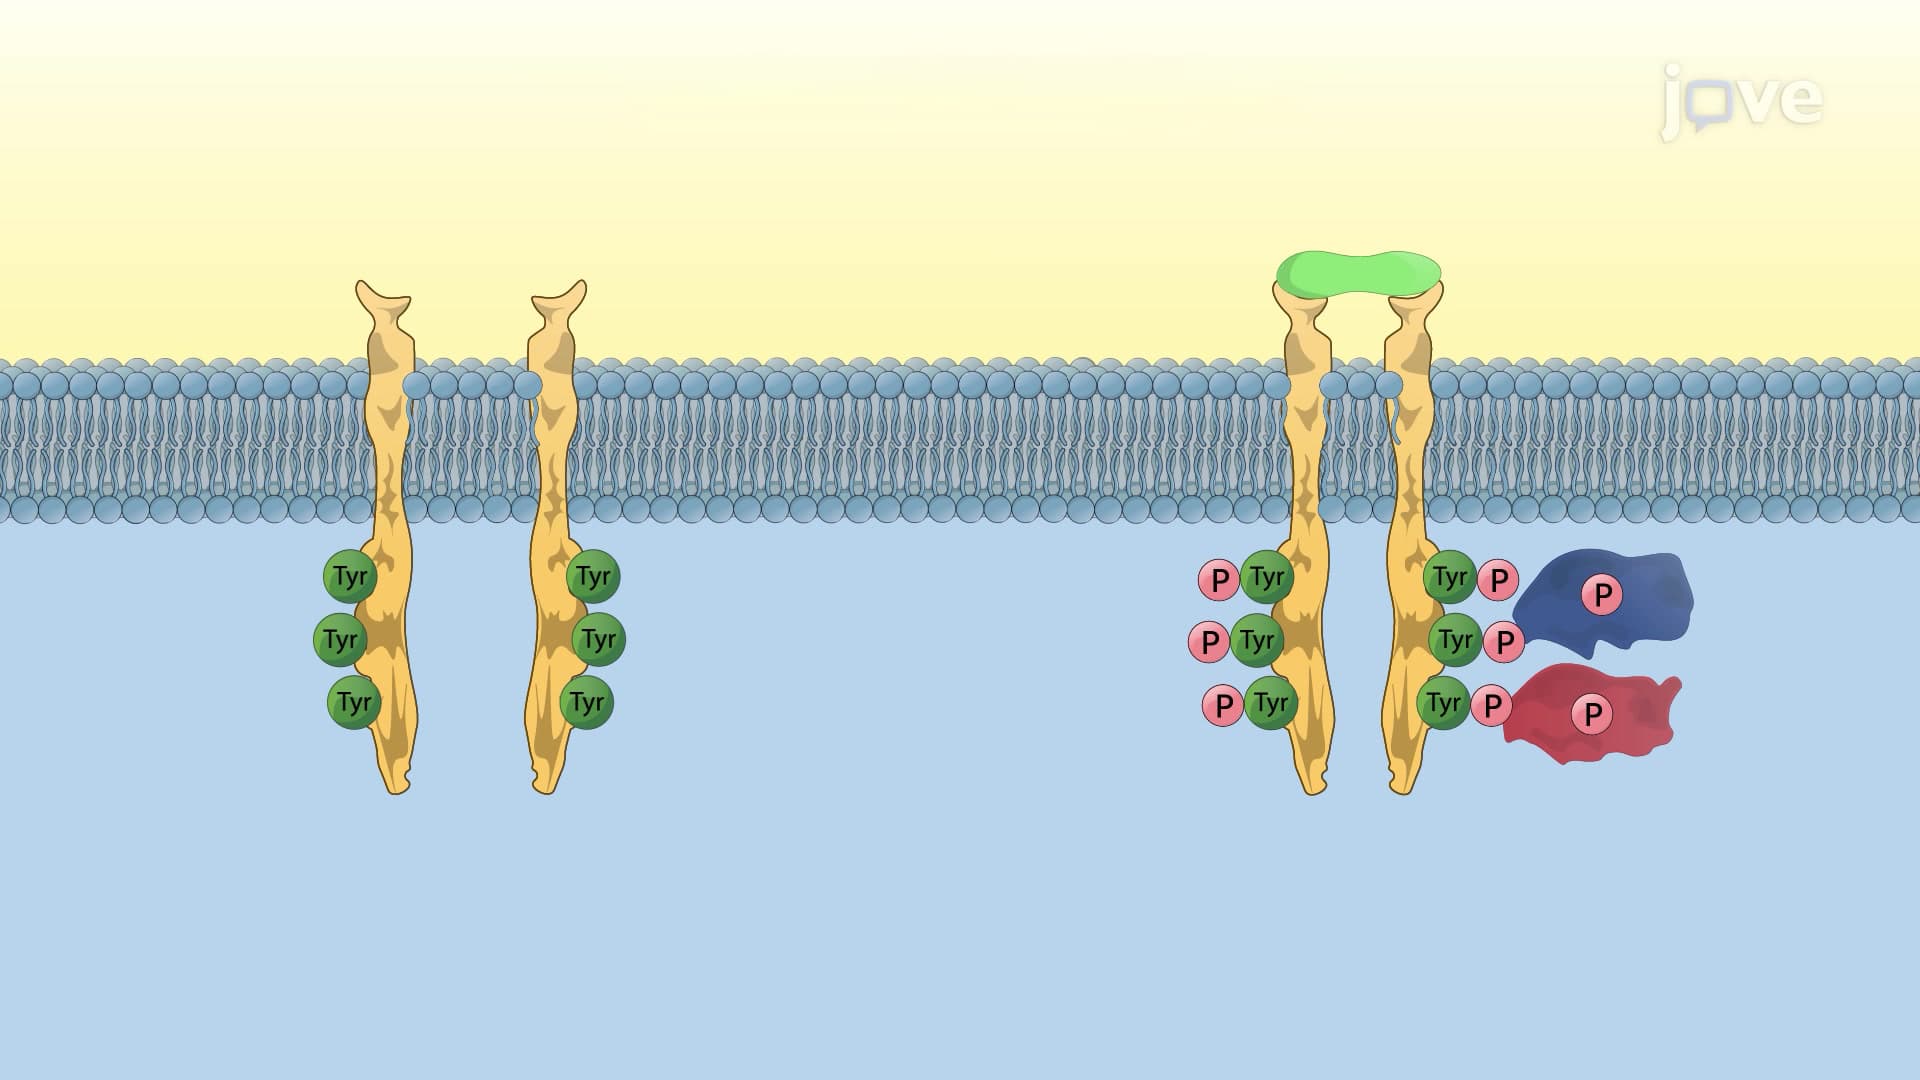
Receptor Tyrosine Kinases
 02:26
Conservation of Protein Domains Over Different Proteins
A limited set of protein domains often duplicate and recombine during evolution. These domains can be organized in different combinations to form...
 02:54
Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...
02:54Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...