

Electronic and vibrational circular dichroism of aromatic amino acids by density functional theory
Takeyuki Tanaka1, Takashi S Kodama, Hayato E Morita
1Department of Biosystems Science, Graduate School of Science and Technology, Kobe University, Kobe, Japan.

Abstract:
Structures of model compounds mimicking aromatic amino acid residues in proteins are optimized by density functional theory (DFT), assuming that the main-chain conformation was a random coil. Excitation energies and dipole and rotational strengths for the optimized structures were calculated based on time-dependent DFT (TD-DFT). The electronic circular dichroism (ECD) bands of the models were significantly affected by side-chain conformations. Hydration models of the aromatic residues were also subjected to TD-DFT calculations, and the ECD bands of these models were found to be highly perturbed by the hydration of the main-chain amide groups. In addition to calculating the random-coil conformation, we also performed TD-DFT calculations of the aromatic residue models, assuming that the main-chain conformation was an alpha-helix or beta-strand. As expected, the overall feature of the ECD bands was also perturbed by the main-chain conformations. Moreover, vibrational circular dichroism (VCD) spectra of the hydration models in a random-coil structure were simulated by DFT, which showed that the VCD spectra are more sensitive to the side-chain conformations than the ECD spectra. The present results show that analyses combining ECD and VCD spectroscopy and using DFT calculations can elucidate the main- and side-chain conformations of aromatic residues in proteins.
Related Concept Videos
 01:14
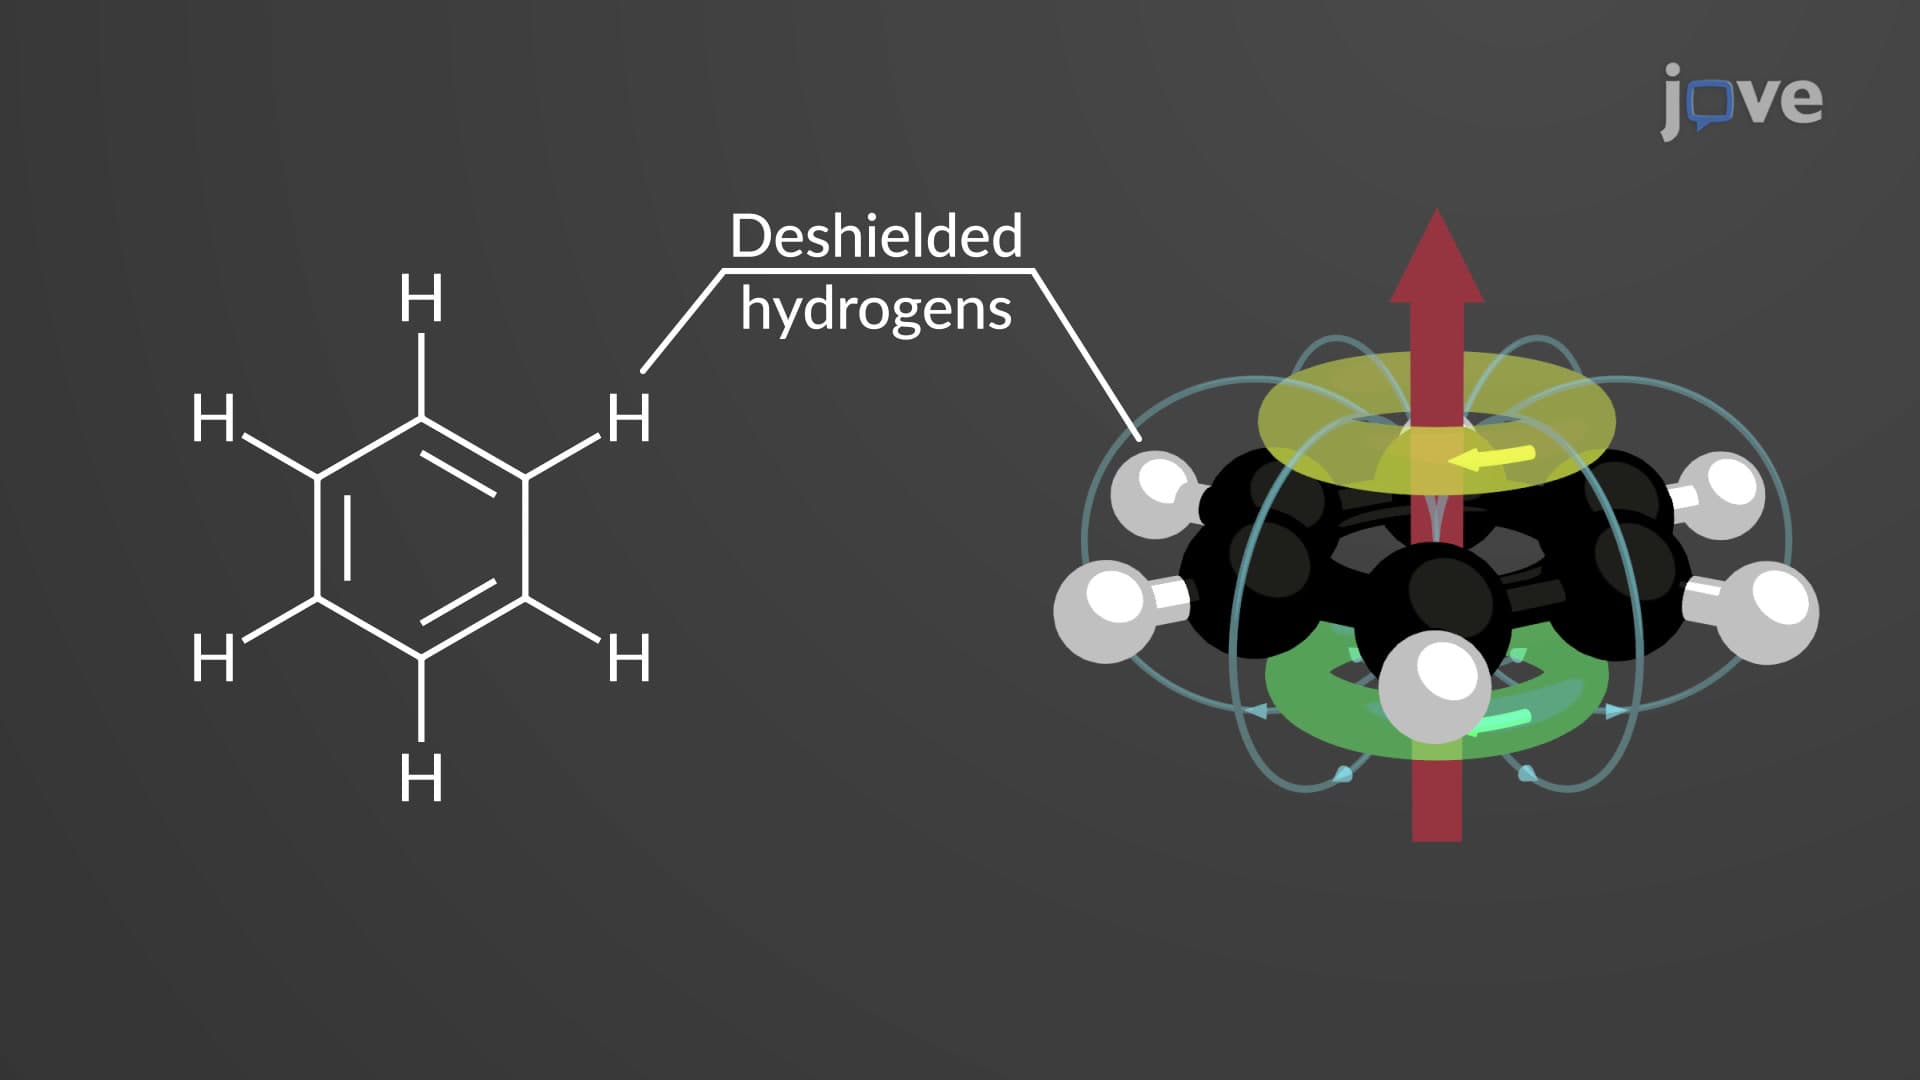
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:14
NMR Spectroscopy of Aromatic Compounds
 01:16
UV–Vis Spectroscopy: Molecular Electronic Transitions
 01:08
IR Spectrum Peak Splitting: Symmetric vs Asymmetric Vibrations
 01:19
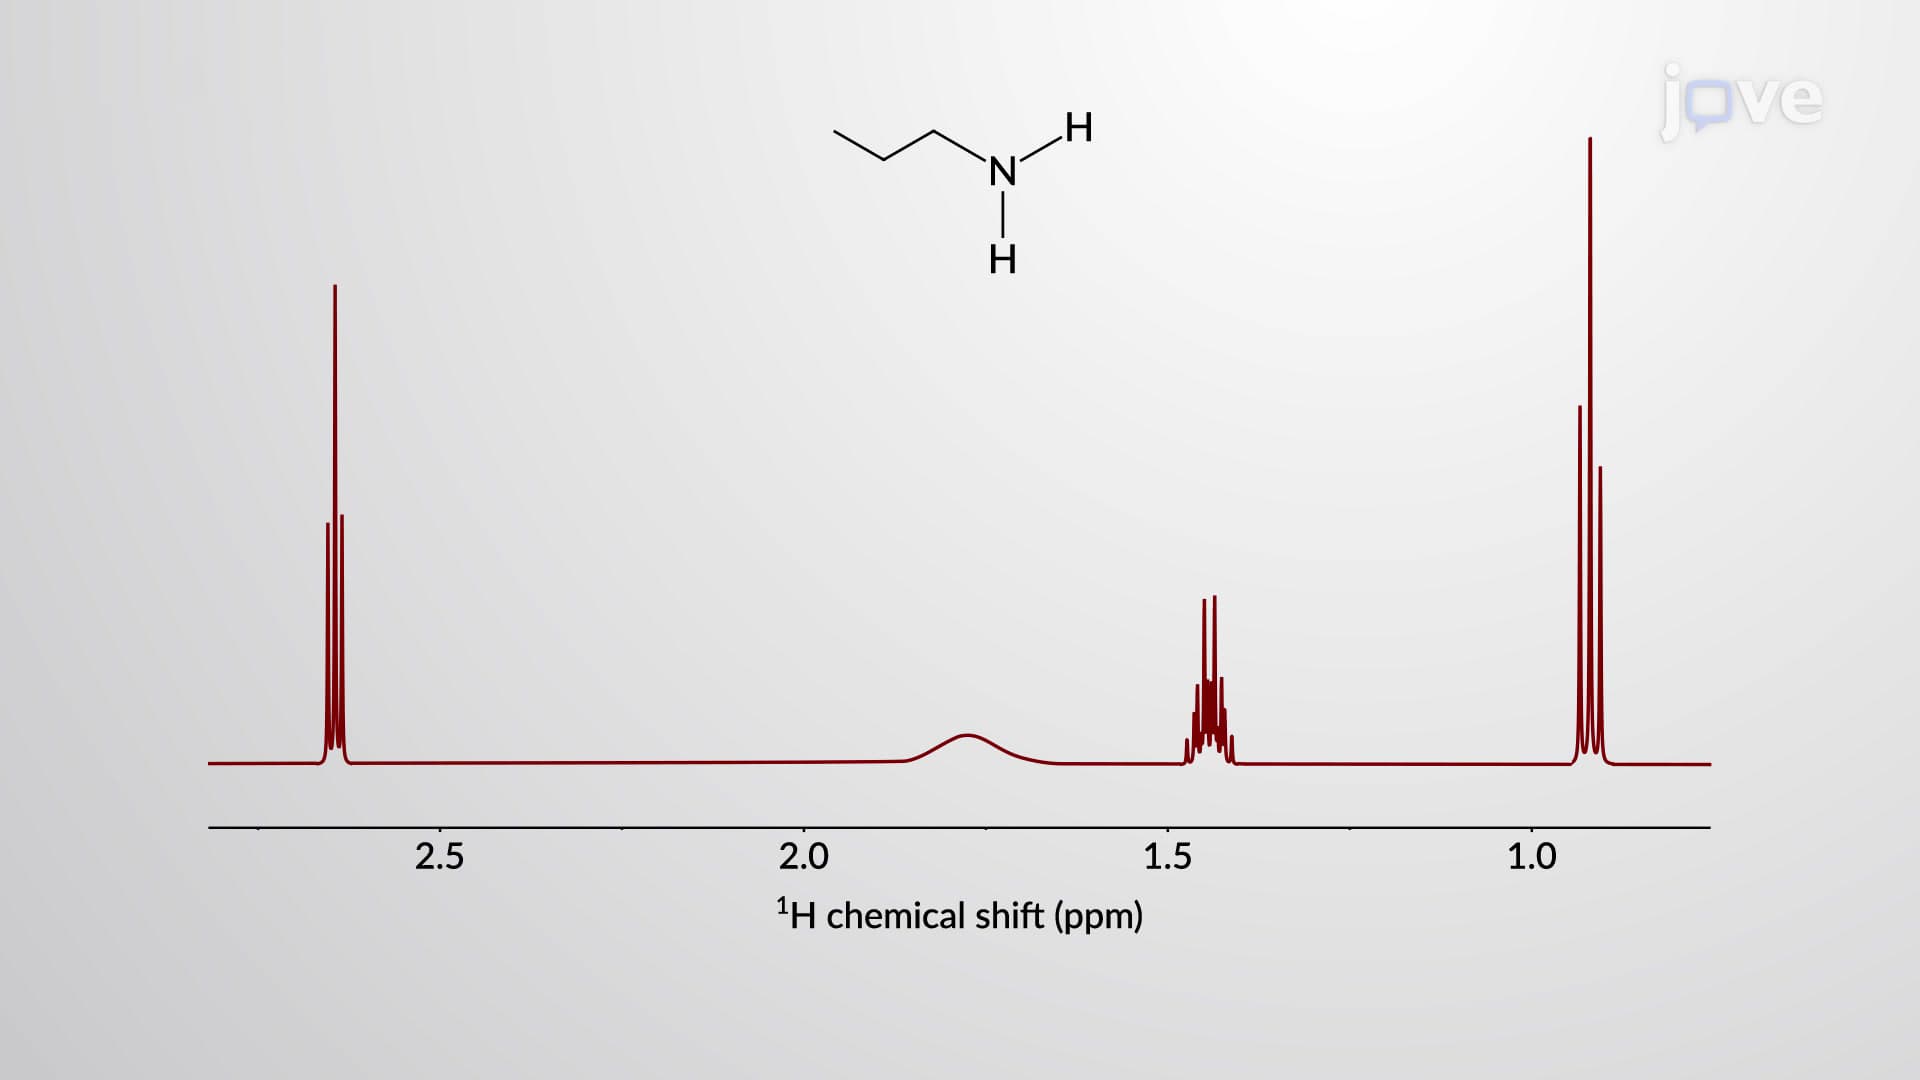
NMR Spectroscopy Of Amines
 01:18
Aromatic Hydrocarbon Anions: Structural Overview
Due to the absence of continuous overlap of p...