Related Experiment Video
Updated: Jun 10, 2026
Ethylene Polymerizations Using Parallel Pressure Reactors and a Kinetic Analysis of Chain Transfer Polymerization
Published on: November 27, 2015
Dynamic effects on the periselectivity, rate, isotope effects, and mechanism of cycloadditions of ketenes with
Bryson R Ussing1, Chao Hang, Daniel A Singleton
1Department of Chemistry, Texas A&M University, P.O. Box 30012, College Station, Texas 77842, USA.

Abstract:
The cycloadditions of cyclopentadiene with diphenylketene and dichloroketene are studied by a combination of kinetic and product studies, kinetic isotope effects, standard theoretical calculations, and trajectory calculations. In contrast to recent reports, the reaction of cyclopentadiene with diphenylketene affords both [4 + 2] and [2 + 2] cycloadducts directly. This is surprising, since there is only one low-energy transition structure for adduct formation in mPW1K calculations, but quasiclassical trajectories started from this single transition structure afford both [4 + 2] and [2 + 2] products. The dichloroketene reaction is finely balanced between [4 + 2] and [2 + 2] cycloaddition modes in mPW1K calculations, as the minimum-energy path (MEP) leads to different products depending on the basis set. The MEP is misleading in predicting a single product, as trajectory studies for the dichloroketene reaction predict that both [4 + 2] and [2 + 2] products should be formed. The periselectivity does not reflect transition state orbital interactions. The (13)C isotope effects for the dichloroketene reaction are well-predicted from the mPW1K/6-31+G** transition structure. However, the isotope effects for the diphenylketene reaction are not predictable from the cycloaddition transition structure and transition state theory. The isotope effects also appear inconsistent with kinetic observations, but the trajectory studies evince that nonstatistical recrossing can reconcile the apparently contradictory observations. B3LYP calculations predict a shallow intermediate on the energy surface, but trajectory studies suggest that the differing B3LYP and mPW1K surfaces do not result in qualitatively differing mechanisms. Overall, an understanding of the products, rates, selectivities, isotope effects, and mechanism in these reactions requires the explicit consideration of dynamic trajectories.
Related Concept Videos
 02:35
Regioselectivity of Electrophilic Additions-Peroxide Effect
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.
 01:17
Thermal Electrocyclic Reactions: Stereochemistry
Selection Rules: Thermal Activation
Conjugated systems containing an even number of π-electron pairs undergo a conrotatory ring closure. For example, thermal electrocyclization of (2E,4E)-2,4-hexadiene, a conjugated diene containing two π-electron pairs, gives trans-3,4-dimethylcyclobutene.
 00:57
Cationic Chain-Growth Polymerization: Mechanism
 01:13
Olefin Metathesis Polymerization: Overview
Ruthenium-based Grubbs catalyst is the most commonly used catalyst for olefin metathesis polymerization. Grubbs catalyst consists of a...
 01:17
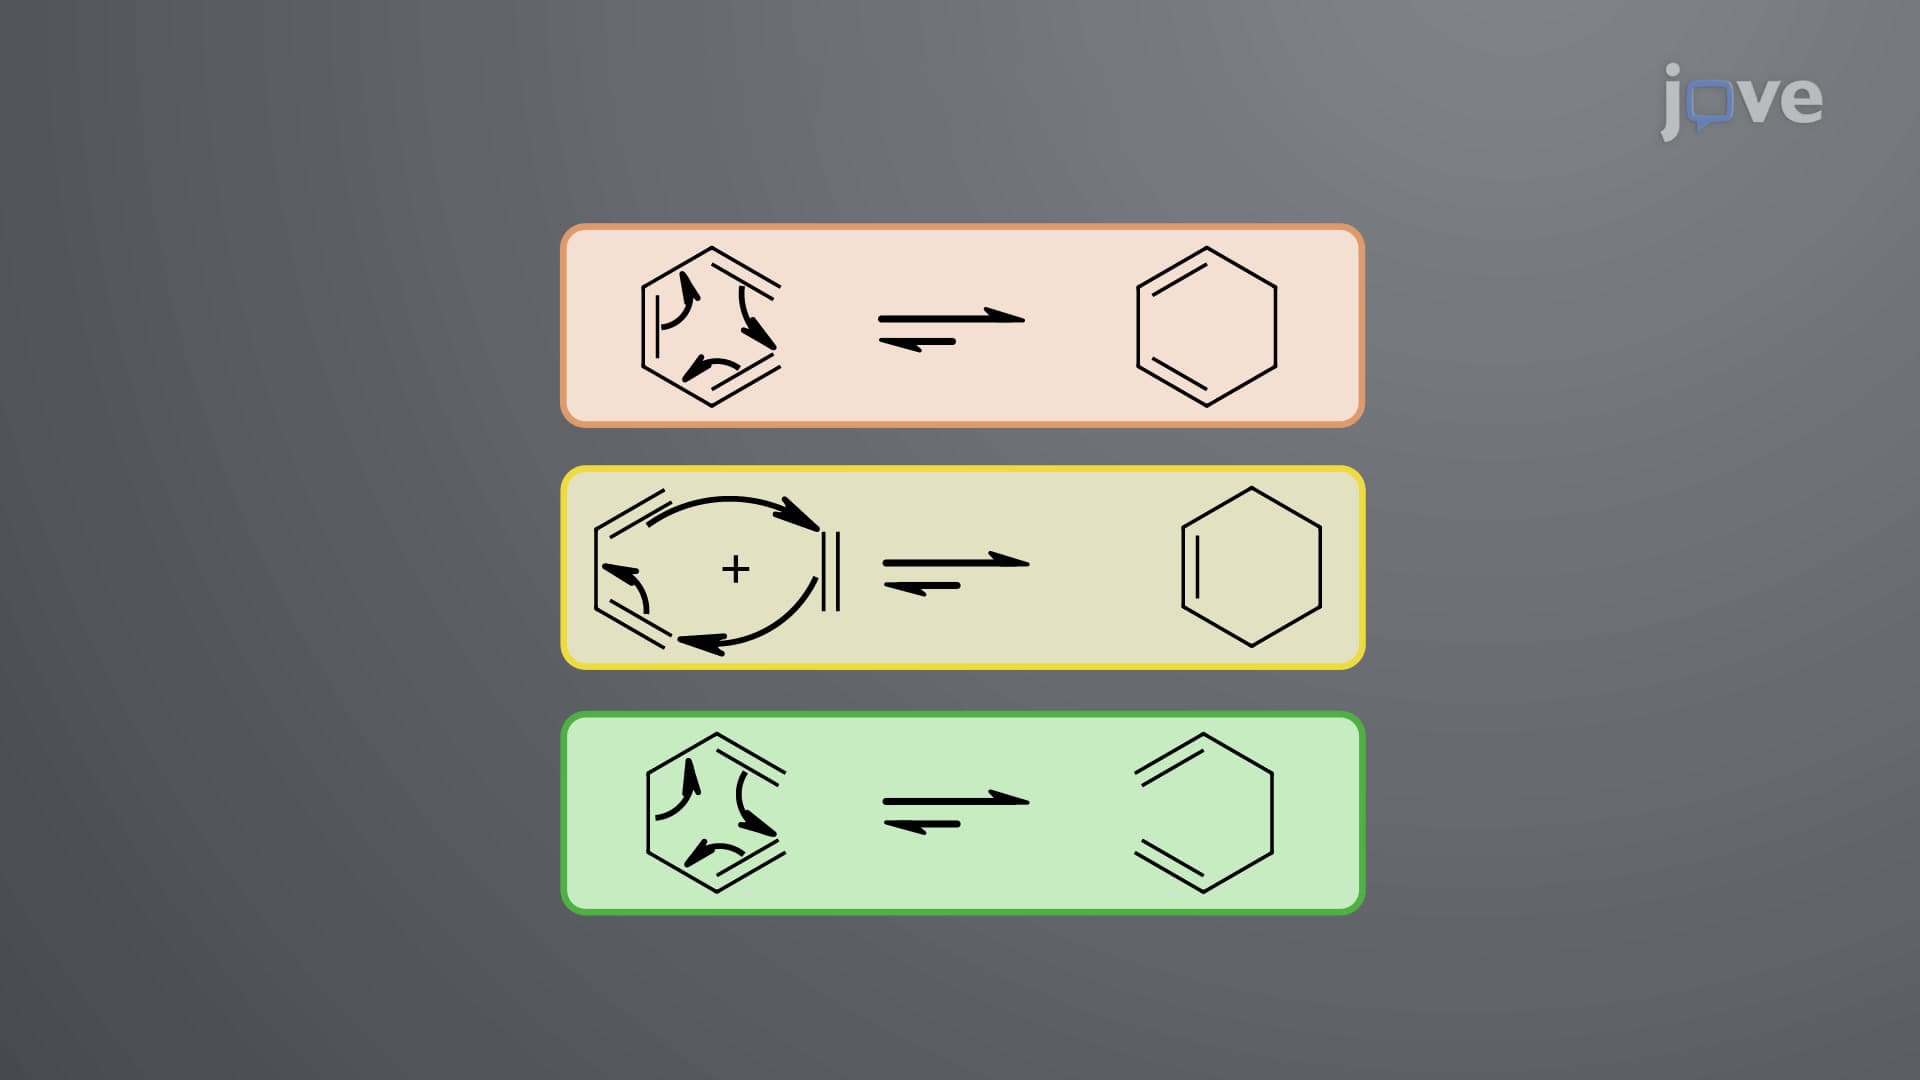
Pericyclic Reactions: Introduction
Pericyclic reactions can be classified into three categories: electrocyclic reactions, cycloaddition reactions, and sigmatropic rearrangements. Electrocyclic reactions and sigmatropic rearrangements are...