Related Experiment Video
Updated: Aug 7, 2026
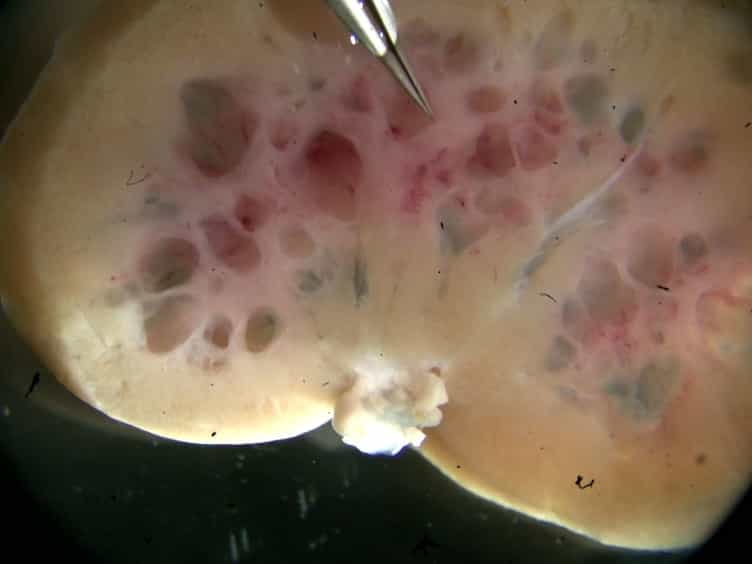

Implementing Patch Clamp and Live Fluorescence Microscopy to Monitor Functional Properties of Freshly Isolated PKD Epithelium
Published on: September 1, 2015
Molecular and cellular pathophysiology of autosomal recessive polycystic kidney disease (ARPKD)
William E Sweeney1, Ellis D Avner
1Children's Research Institute, Children's Hospital Health System of Wisconsin, Medical College of Wisconsin, Milwaukee, WI 53226, USA.

Abstract:
Autosomal recessive polycystic kidney disease (ARPKD) belongs to a group of congenital hepatorenal fibrocystic syndromes characterized by dual renal and hepatic involvement of variable severity. Despite the wide clinical spectrum of ARPKD (MIM 263200), genetic linkage studies indicate that mutations at a single locus, PKHD1 (polycystic kidney and hepatic disease 1), located on human chromosome region 6p21.1-p12, are responsible for all phenotypes of ARPKD. Identification of cystic disease genes and their encoded proteins has provided investigators with critical tools to begin to unravel the molecular and cellular mechanisms of PKD. PKD cystic epithelia share common phenotypic abnormalities despite the different genetic mutations that underlie the disease. Recent studies have shown that many cyst-causing proteins are expressed in multimeric complexes at distinct subcellular locations within epithelia. This co-expression of cystoproteins suggests that cyst formation, regardless of the underlying disease gene, results from perturbations in convergent and/or integrated signal transduction pathways. To date, no specific therapies are in clinical use for ameliorating cyst growth in ARPKD. However, studies noted in this review suggest that therapeutic targeting of the cAMP and epidermal growth factor receptor (EGFR)-axis abnormalities in cystic epithelia may translate into effective therapies for ARPKD and, by analogy, autosomal dominant polycystic kidney disease (ADPKD). A particularly promising approach appears to be the targeting of downstream intermediates of both the cAMP and EGFR axis. This review focuses on ARPKD and presents a concise summary of the current understanding of the molecular genetics and cellular pathophysiology of this disease. It also highlights phenotypic and mechanistic similarities between ARPKD and ADPKD.
Related Concept Videos
 01:29
Acute Kidney Injury II: Pathophysiology
 01:10
Nephrons
 01:25
Aquaporins
 01:22
Lysosomal Hydrolases
 01:28
Diabetic Nephropathy
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...