Related Experiment Video
Updated: Aug 7, 2026

Combining Double Fluorescence In Situ Hybridization with Immunolabelling for Detection of the Expression of Three Genes in Mouse Brain Sections
Published on: March 26, 2016
Canavan disease: a white matter disorder
Shalini Kumar1, Natalia S Mattan, Jean de Vellis
1Department of Neurobiology, Mental Retardation Research Center, Semel Institute for Neuroscience and Human Behavior, University of California, Los Angeles, California 90095-7332, USA.

Abstract:
Breakdown of oligodendrocyte-neuron interactions in white matter (WM), such as the loss of myelin, results in axonal dysfunction and hence a disruption of information processing between brain regions. The major feature of leukodystrophies is the lack of proper myelin formation during early development or the onset of myelin loss late in life. These early childhood WM diseases are described as hypomyelination or dysmyelination arising from a primary block in normal myelin synthesis because of a genetic mutation expressed in oligodendrocytes, or failure in myelination secondary to neuronal or astroglial dysfunctions (van der Knaap 2001 Dev. Med. Child Neurol. 43:705-712). Here, we describe the pathophysiological parameters of Canavan disease (CD), caused by genetic mutations of the aspartoacylase (ASPA) gene, a metabolic enzyme restricted in the central nervous system (CNS) to oligodendrocytes. CD presents pathophysiological dysfunctions similar to diseases caused by myelin gene mutations, such as Pelizaeus-Merzbacher disease (PMD) and several animal models, such as myelin deficient rat (md), jimpy (jp), shiverer (sh), and quaking (qk viable) mutant mice. These single gene mutations have pleiotropic effects, whereby the alteration of one myelin gene expression disrupts functional expression of other oligodendrocyte genes with an outcome of hypomyelination/dysmyelination. Among all of the known leukodystrophies, CD is the first disorder, which was approved and tested for the adeno-associated virus vector (AAV)-ASPA gene therapy (Leone et al. 2000 Ann. Neurol. 48:27-38; Janson et al. 2001 Trends Neurosci. 24:706-712) without much success following the first two attempts. ASPA gene delivery attempts in animal models have shown a lowering of N-acetyl L-aspartate and a change in motor functions, while sponginess of the WM, a characteristic of CD remained unchanged (Matalon et al. 2003 Mol. Ther. 7 (5, Part 1):580-587; McPhee et al. 2005 Brain Res. Mol. Brain Res. 135:112-121) even with better viral serotype and delivery of the gene during early phase of development (Klugmann et al. 2005 Mol. Ther. 11:745-753). While different approaches are being sought for the success of gene therapy, there are pivotal developmental questions to address regarding the specific regions of the CNS and cell lineages that become the target for the onset and progression of CD symptoms from early to late stages of development.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:23
Alzheimer Disease ll: Pathophysiology
 01:22
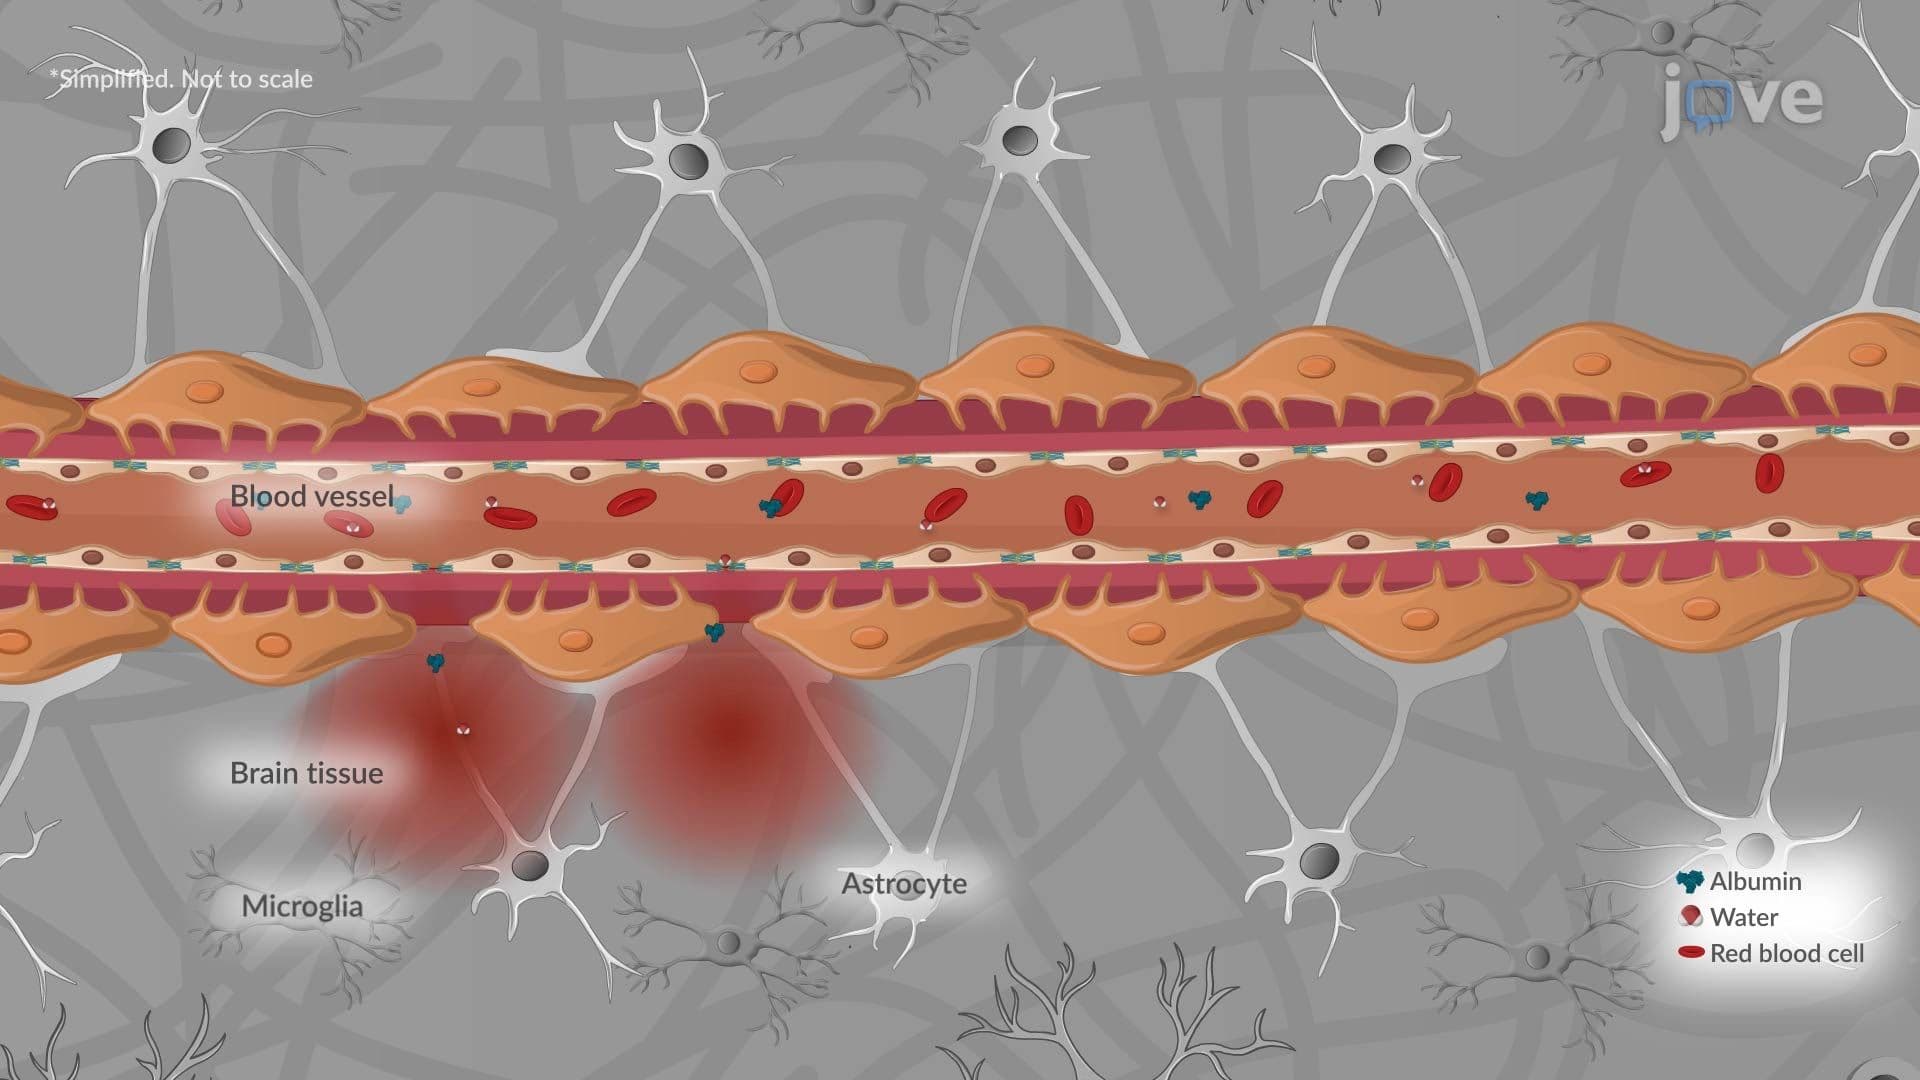
Cerebral Edema ll: Pathophysiology
 01:28
Disorders of the Nervous Tissue
Homeostatic Imbalances:
Alzheimer's disease manifests as a gradual decline in memory and cognitive abilities, attributed to the buildup of amyloid plaques and neurofibrillary tangles in the brain.
Parkinson's disease arises from the...
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:19
Multiple Sclerosis l: Introduction