Related Experiment Video
Updated: Aug 7, 2026

Atomic Scale Structural Studies of Macromolecular Assemblies by Solid-state Nuclear Magnetic Resonance Spectroscopy
Published on: September 17, 2017
Modeling NMR chemical shift: A survey of density functional theory approaches for calculating tensor properties
Travis H Sefzik1, Domenic Turco, Robbie J Iuliucci
1Washington and Jefferson College, Washington, Pennsylvania 15301, USA.

Abstract:
The NMR chemical shift, a six-parameter tensor property, is highly sensitive to the position of the atoms in a molecule. To extract structural parameters from chemical shifts, one must rely on theoretical models. Therefore, a high quality group of shift tensors that serve as benchmarks to test the validity of these models is warranted and necessary to highlight existing computational limitations. Here, a set of 102 13C chemical-shift tensors measured in single crystals, from a series of aromatic and saccharide molecules for which neutron diffraction data are available, is used to survey models based on the density functional (DFT) and Hartree-Fock (HF) theories. The quality of the models is assessed by their least-squares linear regression parameters. It is observed that in general DFT outperforms restricted HF theory. For instance, Becke's three-parameter exchange method and mpw1pw91 generally provide the best predicted shieldings for this group of tensors. However, this performance is not universal, as none of the DFT functionals can predict the saccharide tensors better than HF theory. Both the orientations of the principal axis system and the magnitude of the shielding were compared using the chemical-shift distance to evaluate the quality of the calculated individual tensor components in units of ppm. Systematic shortcomings in the prediction of the principal components were observed, but the theory predicts the corresponding isotropic value more accurately. This is because these systematic errors cancel, thereby indicating that the theoretical assessment of shielding predictions based on the isotropic shift should be avoided.
More Related Videos
 10:52
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
 08:04
Excitonic Hamiltonians for Calculating Optical Absorption Spectra and Optoelectronic Properties of Molecular Aggregates and Solids
Published on: May 27, 2020
Related Concept Videos
 01:15
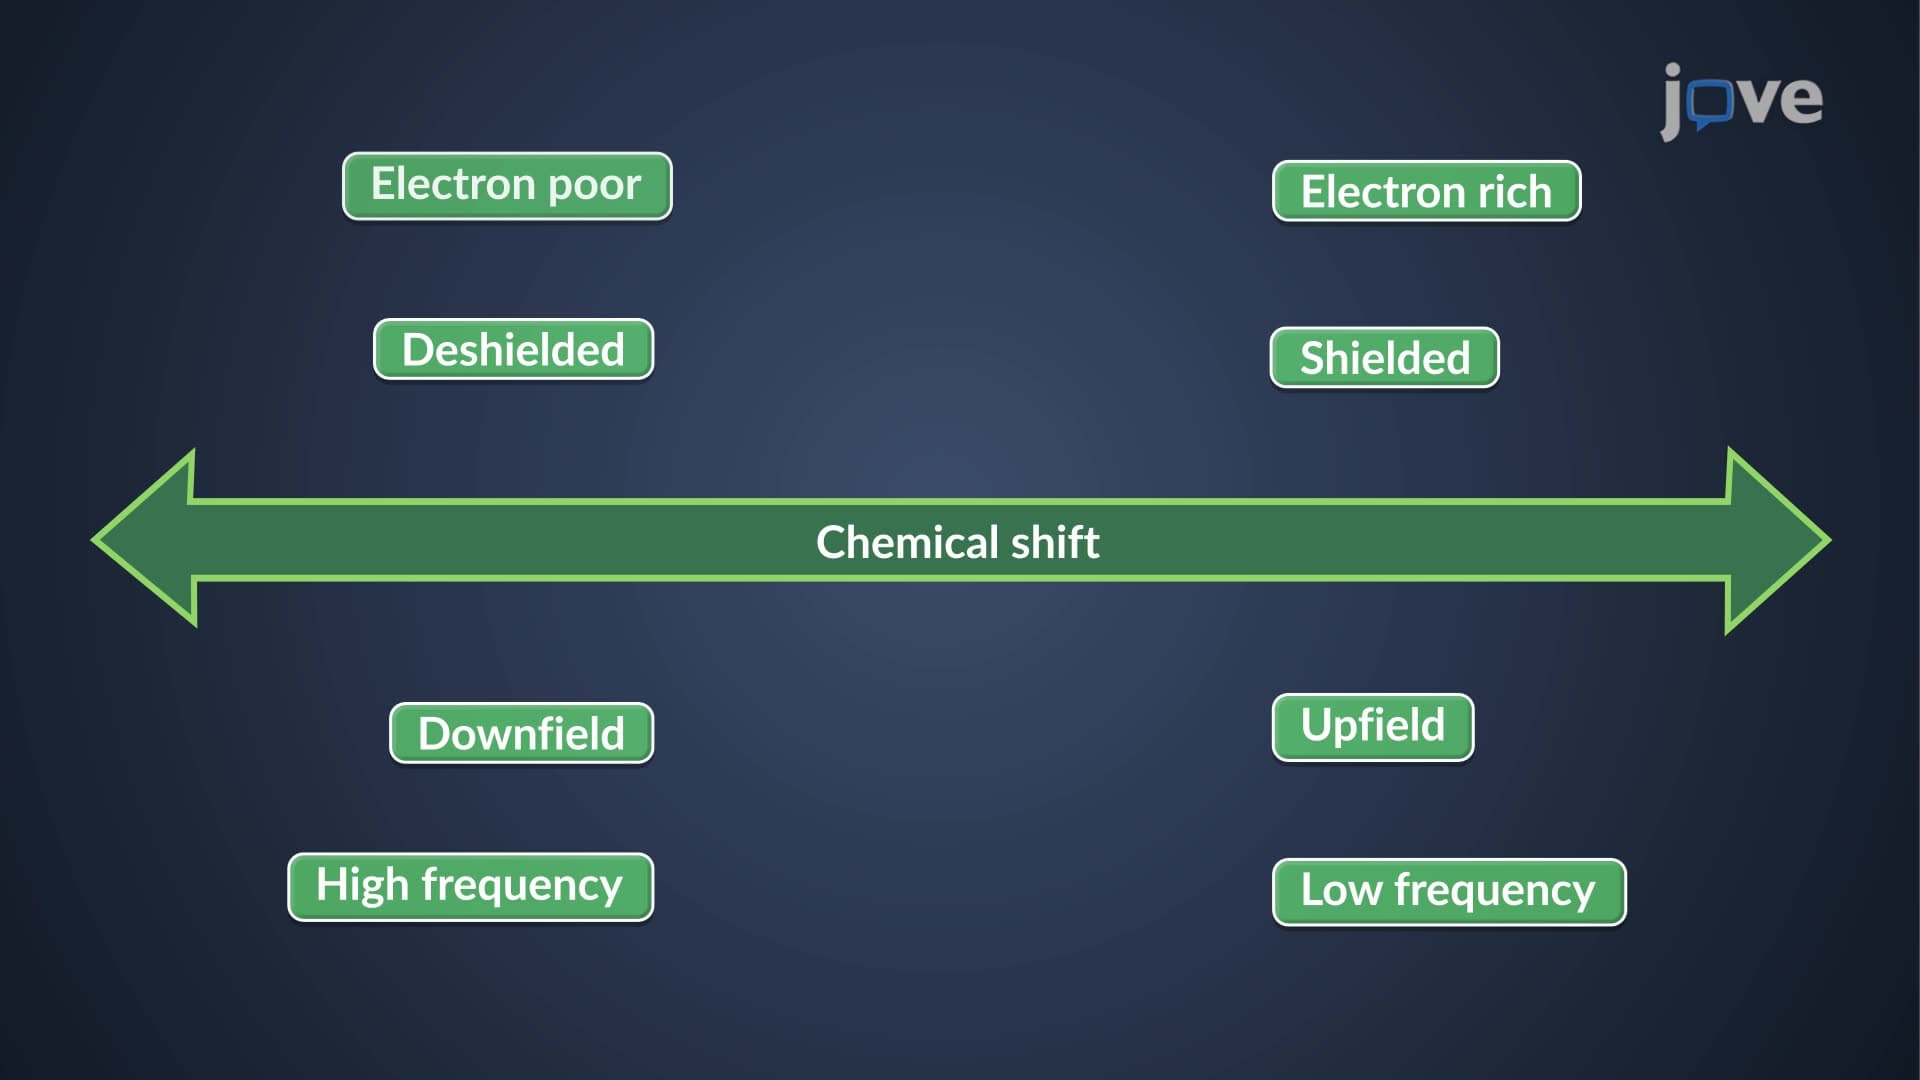
NMR Spectroscopy: Chemical Shift Overview
For instance, the proton...
 01:14
π Electron Effects on Chemical Shift: Aromatic and Antiaromatic Compounds
 01:02
¹H NMR: Interpreting Distorted and Overlapping Signals
As Δν decreases and the signals move closer, the doublets appear increasingly distorted. The intensities of the inner lines increase at the cost of those of the outer lines as the signals are slanted or...
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:07
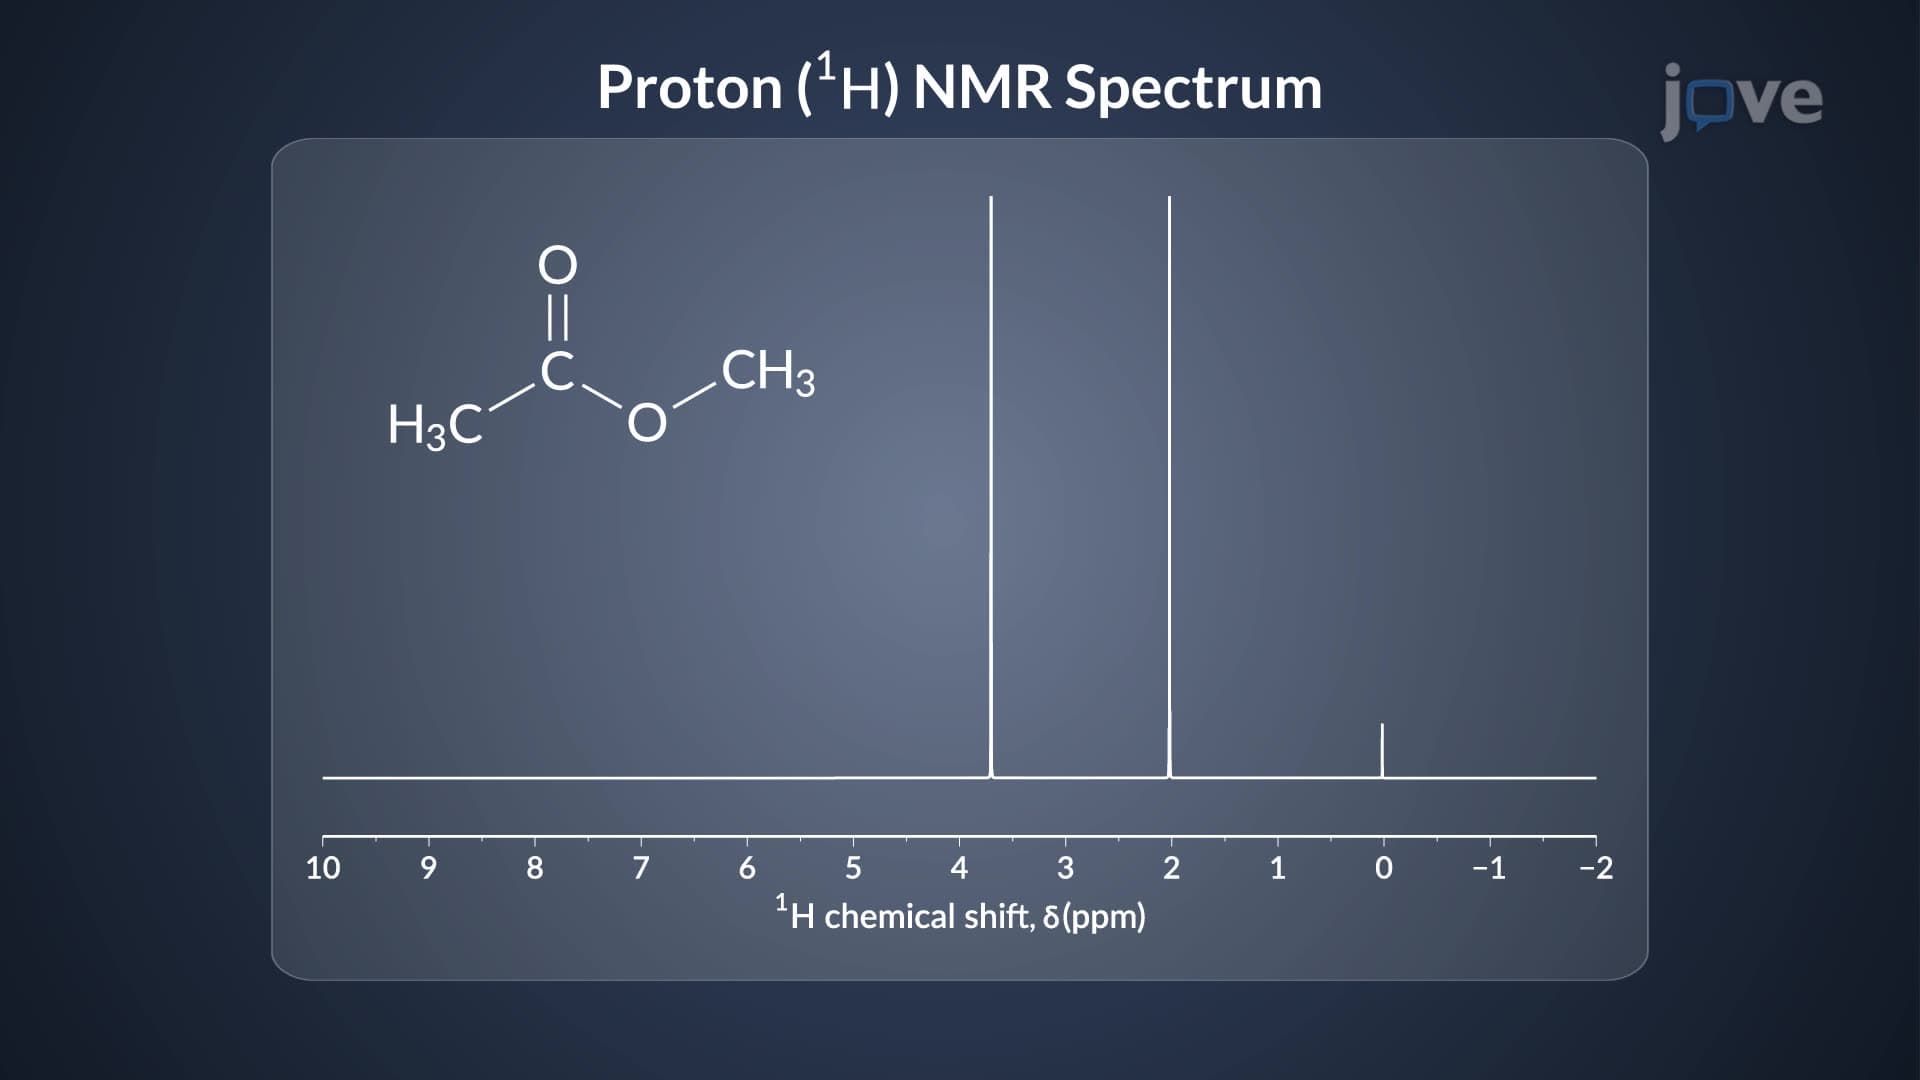
Proton (¹H) NMR: Chemical Shift
Absorption signals of all the protium nuclei in a...
 01:20
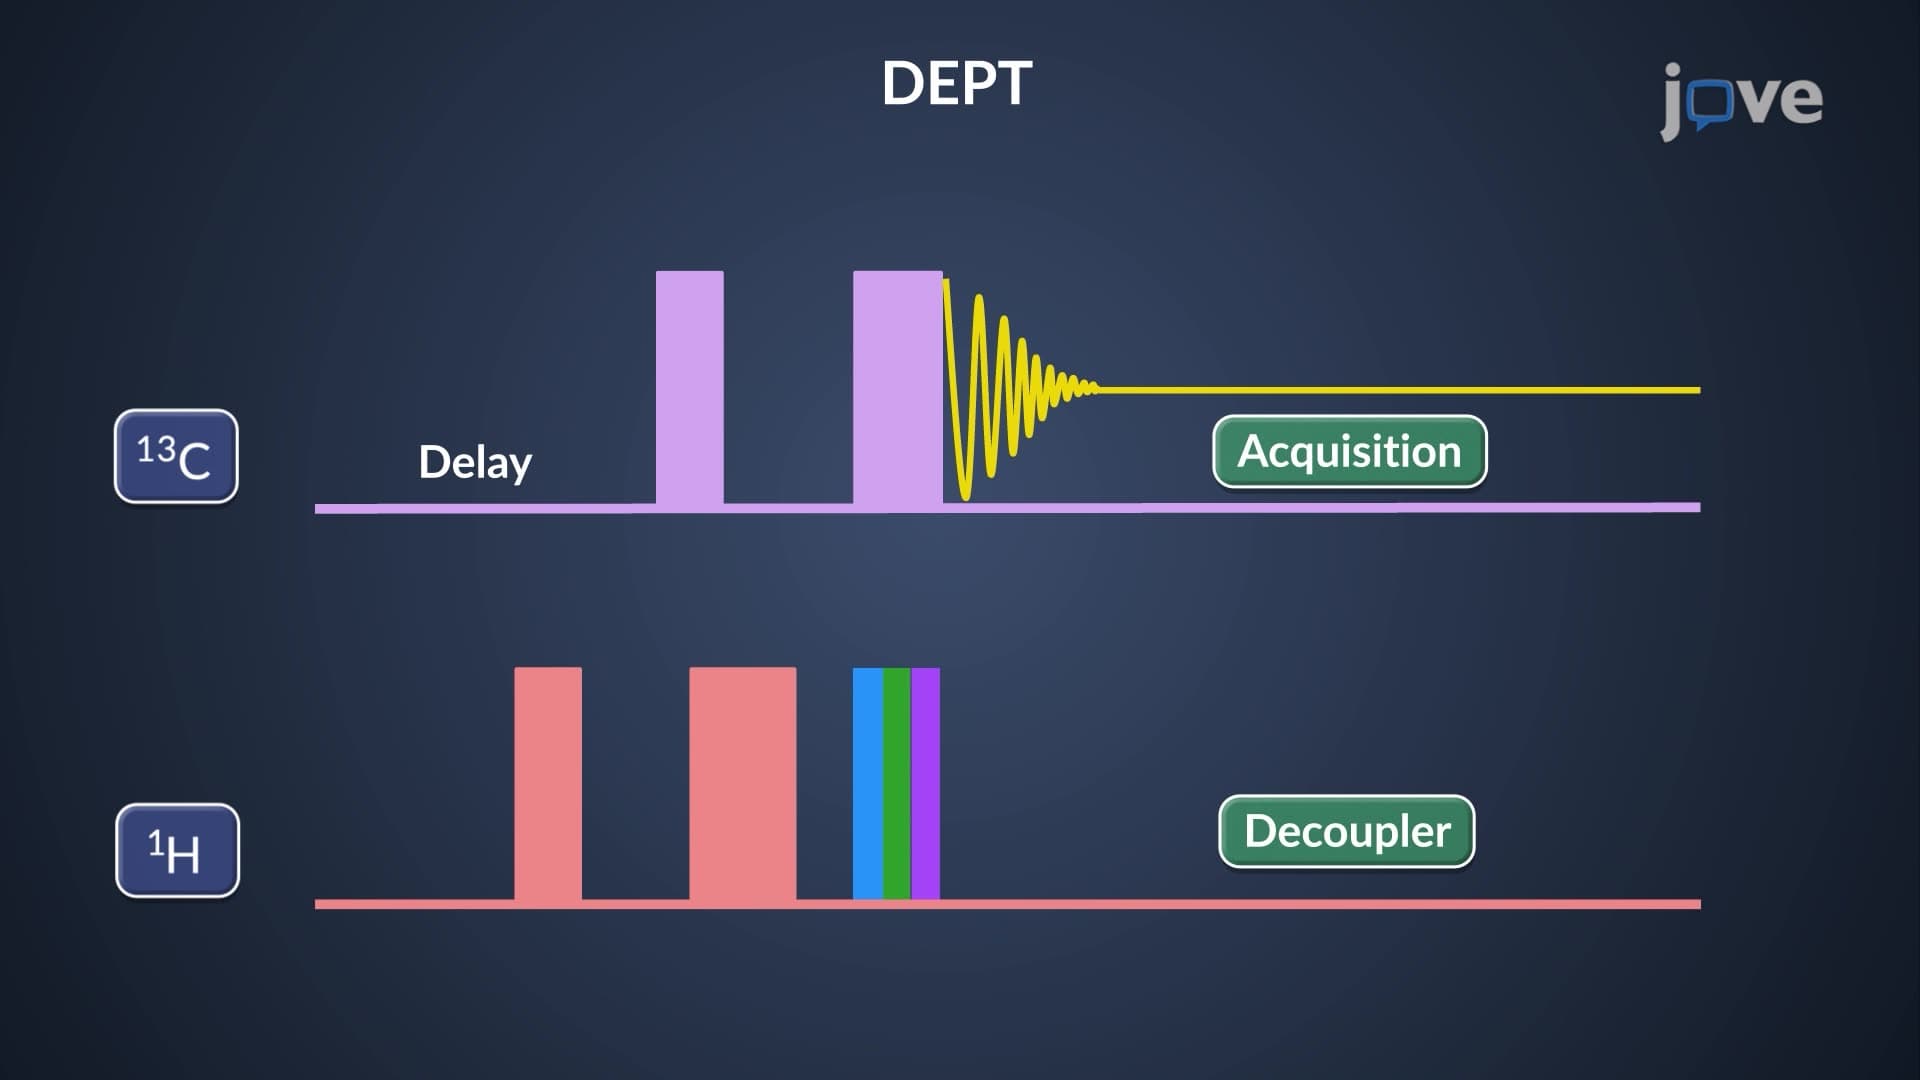
¹³C NMR: Distortionless Enhancement by Polarization Transfer (DEPT)