Related Experiment Video
Updated: Aug 7, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Quasiclassical trajectory study of the F + CH4 reaction dynamics on a dual-level interpolated potential energy
J F Castillo1, F J Aoiz, L Bañares
1Departamento de Química Física, Facultad de Química, Universidad Complutense, 28040 Madrid, Spain. jfc@legendre.quim.ucm.es

Abstract:
An ab initio interpolated potential energy surface (PES) for the F + CH4 reactive system has been constructed using the interpolation method of Collins and co-workers. The ab initio calculations have been performed using second-order Möller-Plesset (MP2) perturbation theory to build the initial PES. Scaling all correlation (SAC) methodology has been employed to improve the ab initio calculations and to construct a dual-level PES. Using this PES, a detailed quasiclassical trajectory study of integral and differential cross sections, product rovibrational populations and internal energy distributions has been carried out for the F + CH4 and F + CD4 reactions and the theoretical results have been compared with the available experimental data.


Related Concept Videos
 01:12
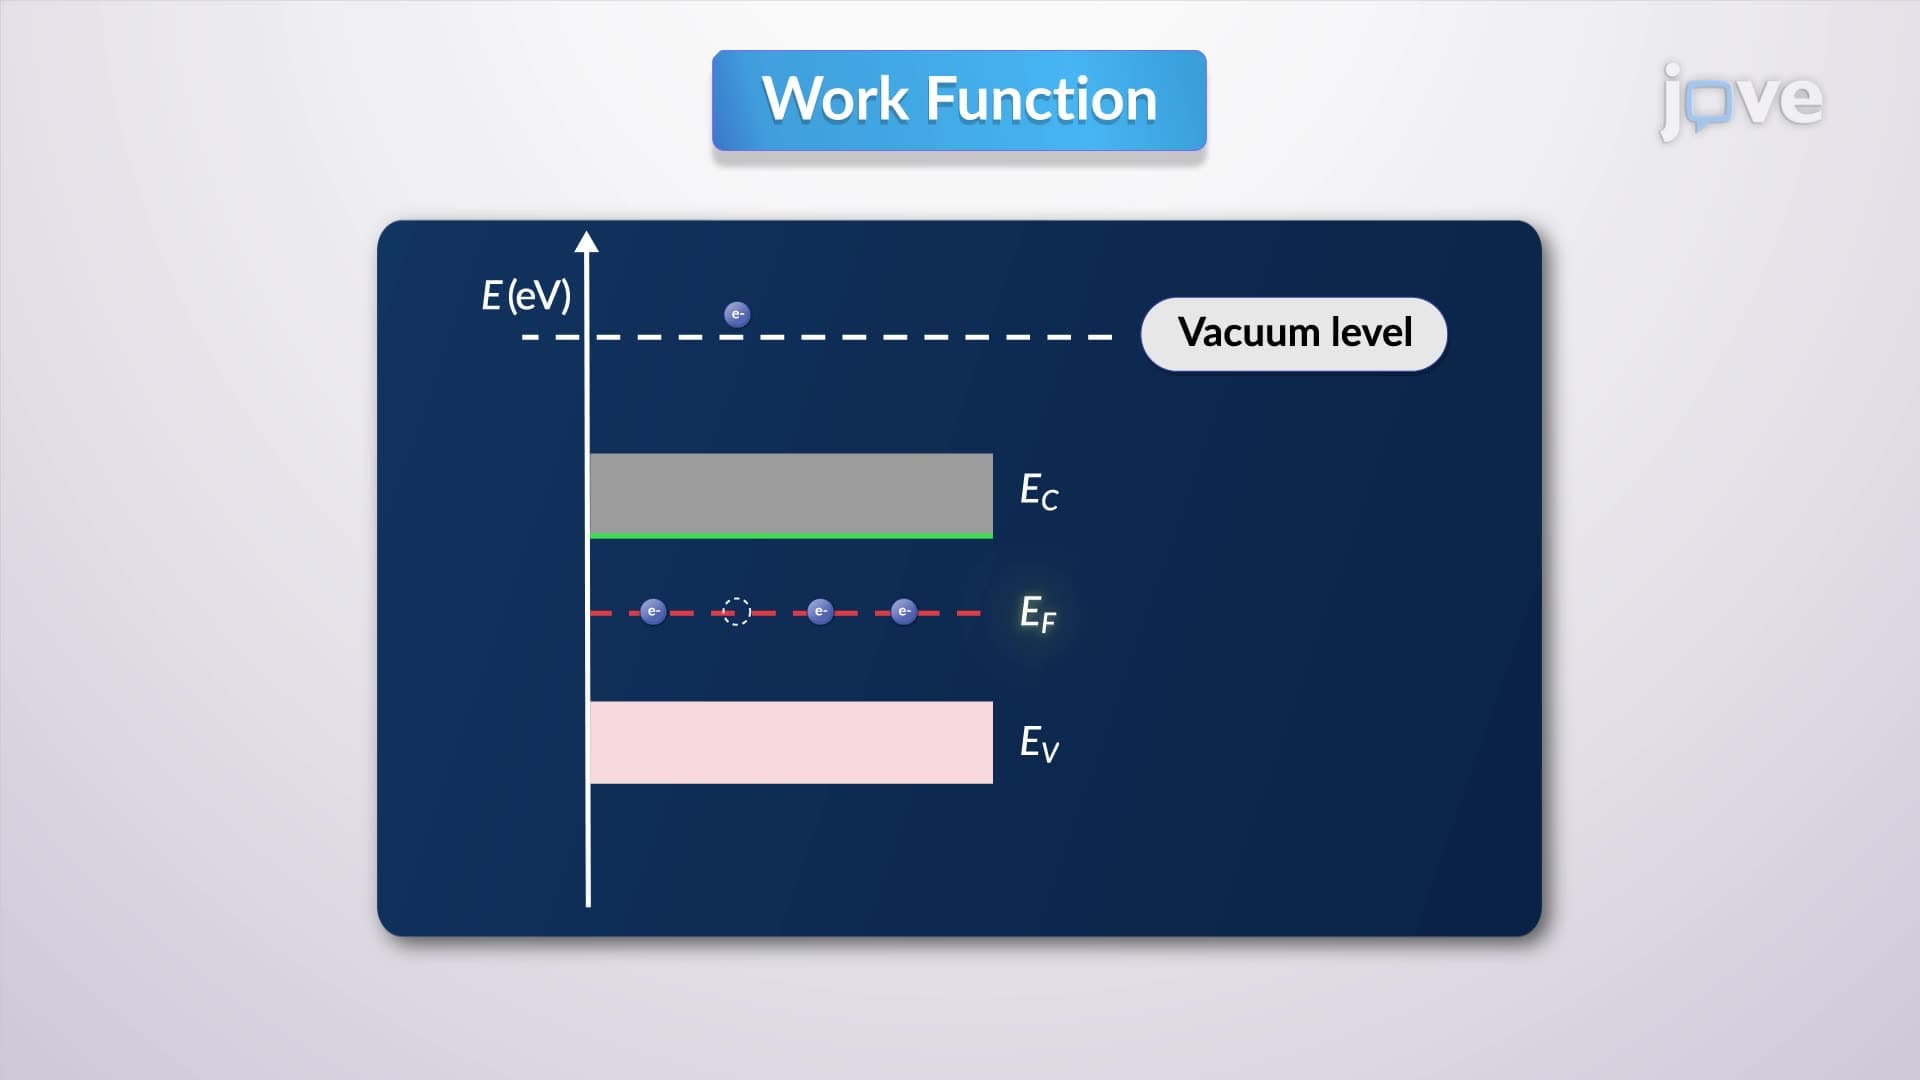
Fermi Level Dynamics
Electron affinity in semiconductors refers to the energy gap between the minimum of its conduction band and the vacuum level and it is a critical parameter in determining how easily a semiconductor can accept additional electrons.
The work...
 02:13
Energy Diagrams, Transition States, and Intermediates
 01:25
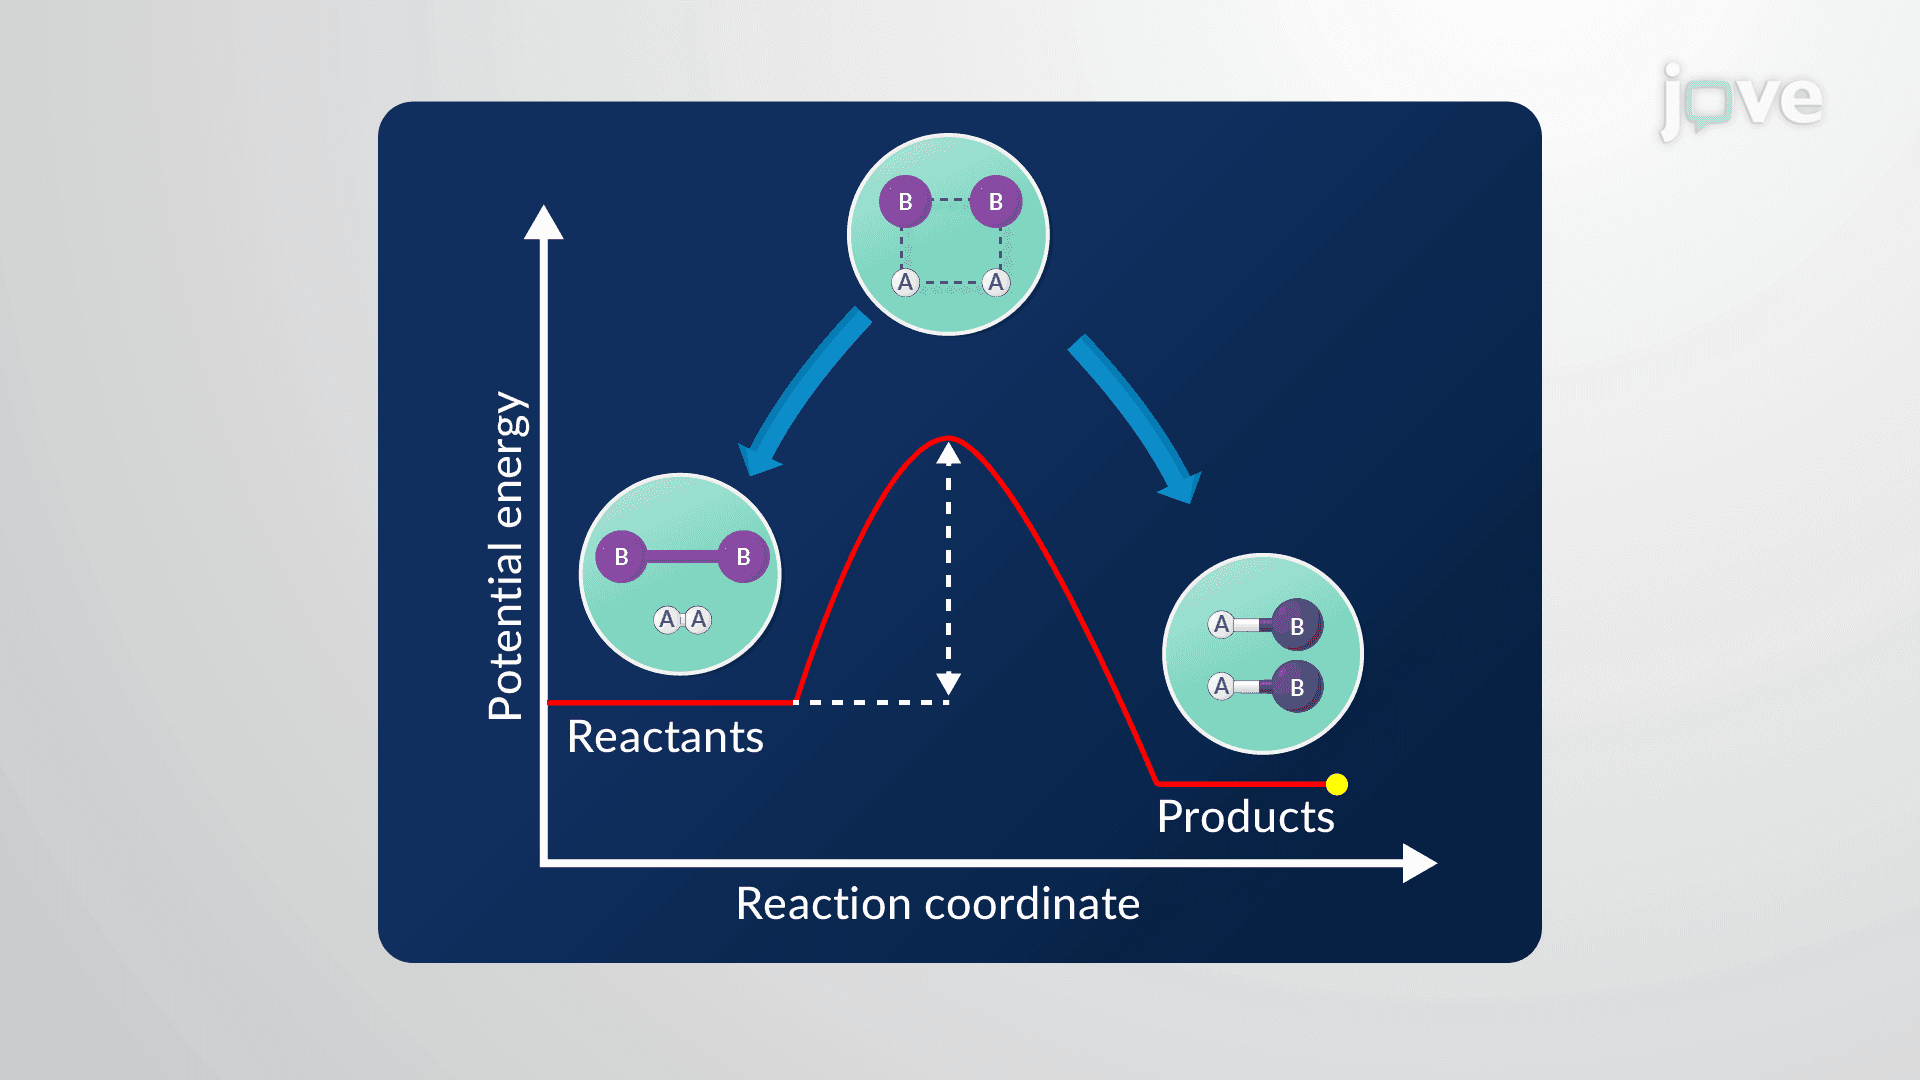
Transition State Theory
 01:26
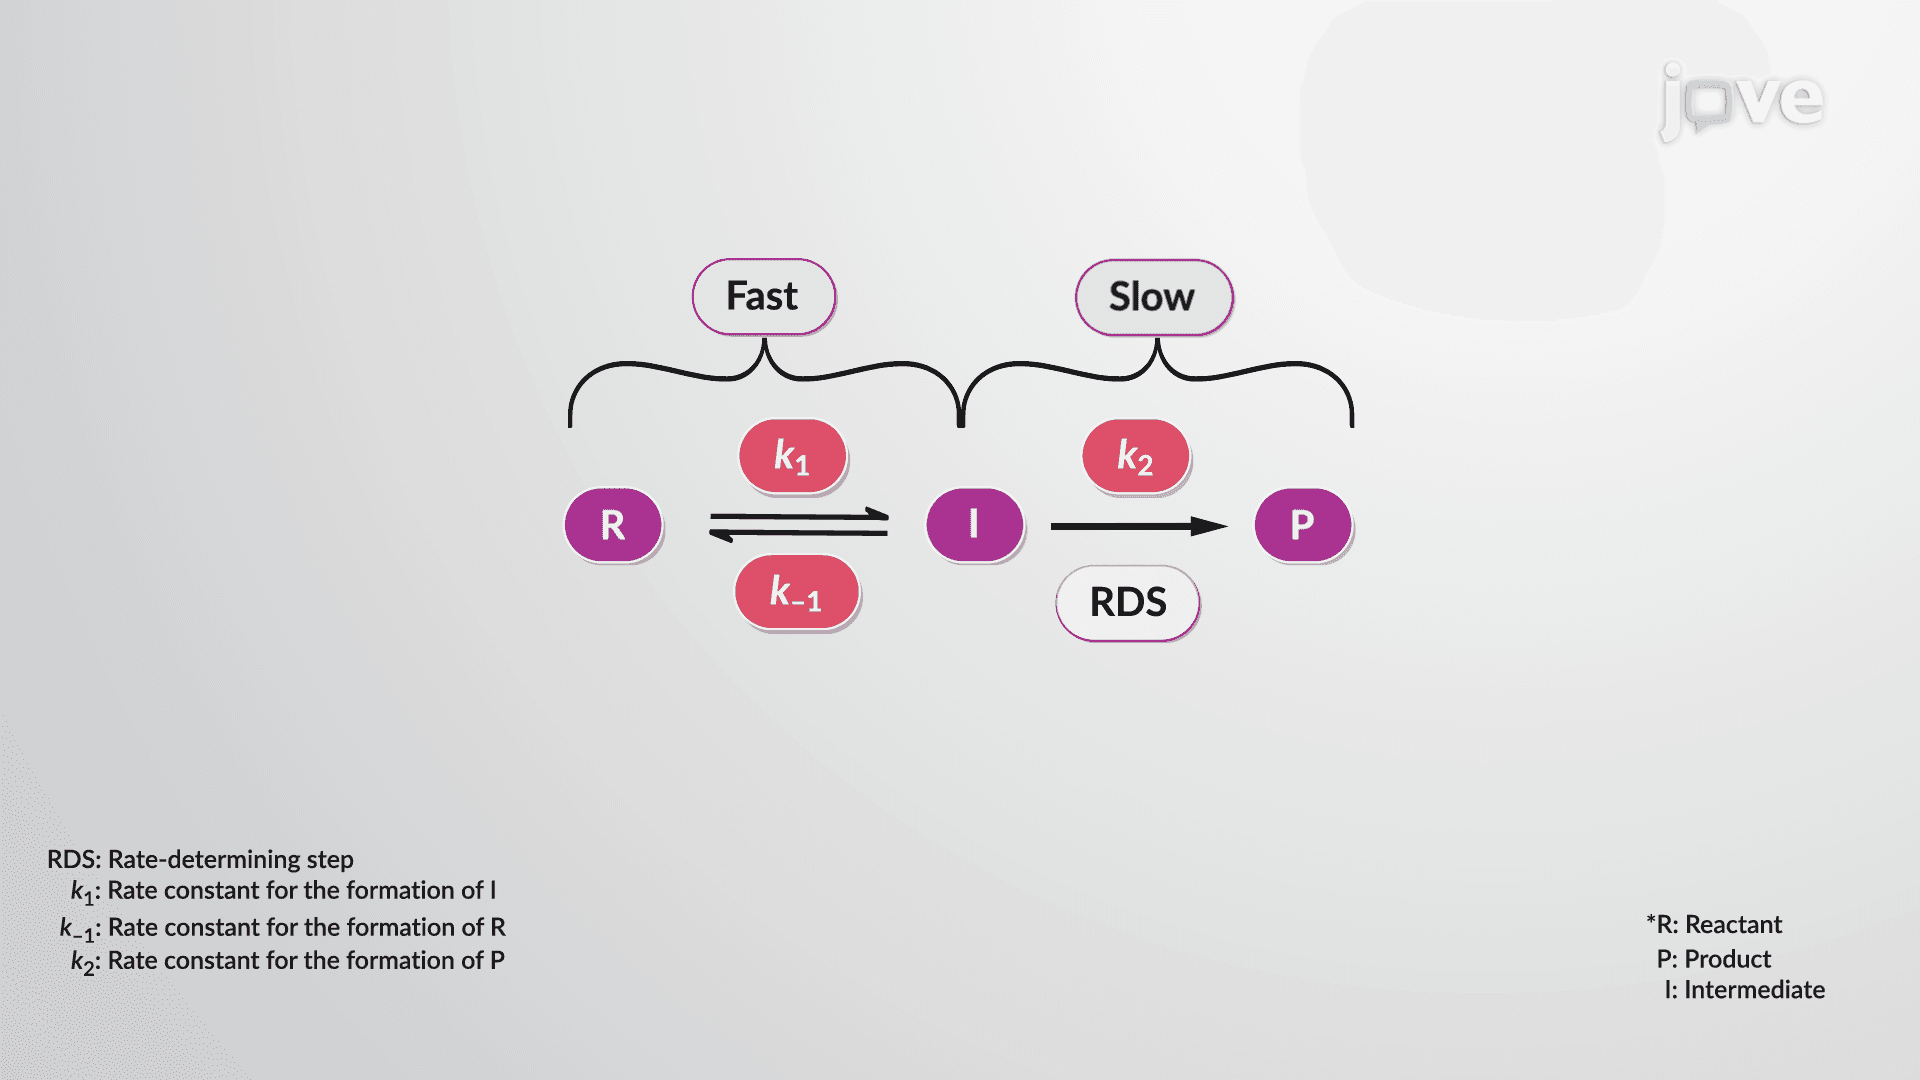
Reaction Mechanisms: The Steady-State Approximation
 03:35
Hybridization of Atomic Orbitals II
 03:51
Molecular Orbital Theory II