Related Experiment Video
Updated: Aug 7, 2026

Analysis of Complex Molecules and Their Reactions on Surfaces by Means of Cluster-Induced Desorption/Ionization Mass Spectrometry
Published on: March 1, 2020
Quantum mechanical model for the dissociative adsorption of diatomic molecules on metal surfaces
Ernst D German1, Alexander M Kuznetsov, Moshe Sheintuch
1Department of Chemical Engineering, Technion-Israel Institute of Technology, Haifa 32000, Israel. ernst_german@yahoo.com

Abstract:
A quantum mechanical nonadiabatic theory of dissociative adsorption of diatomic molecules X2 on metal surface is presented. The following reaction coordinates are used to construct crossing diabatic potential energy surfaces (PES): the distance y between the atoms of the X2 molecule, the distance x of the X2 molecular axis from the surface, the set of coordinates describing possible displacements of metal atoms under adsorption. Expression for the rate constant is derived using the model potentials describing vibrations along these coordinates. The calculated dependency of the rate constant W on the reaction heat DeltaE is compared with that in classical approximation. It is shown that quantum effects lead to a weaker dependence of W on DeltaE as compared to that for classical one.

Related Concept Videos
 01:28
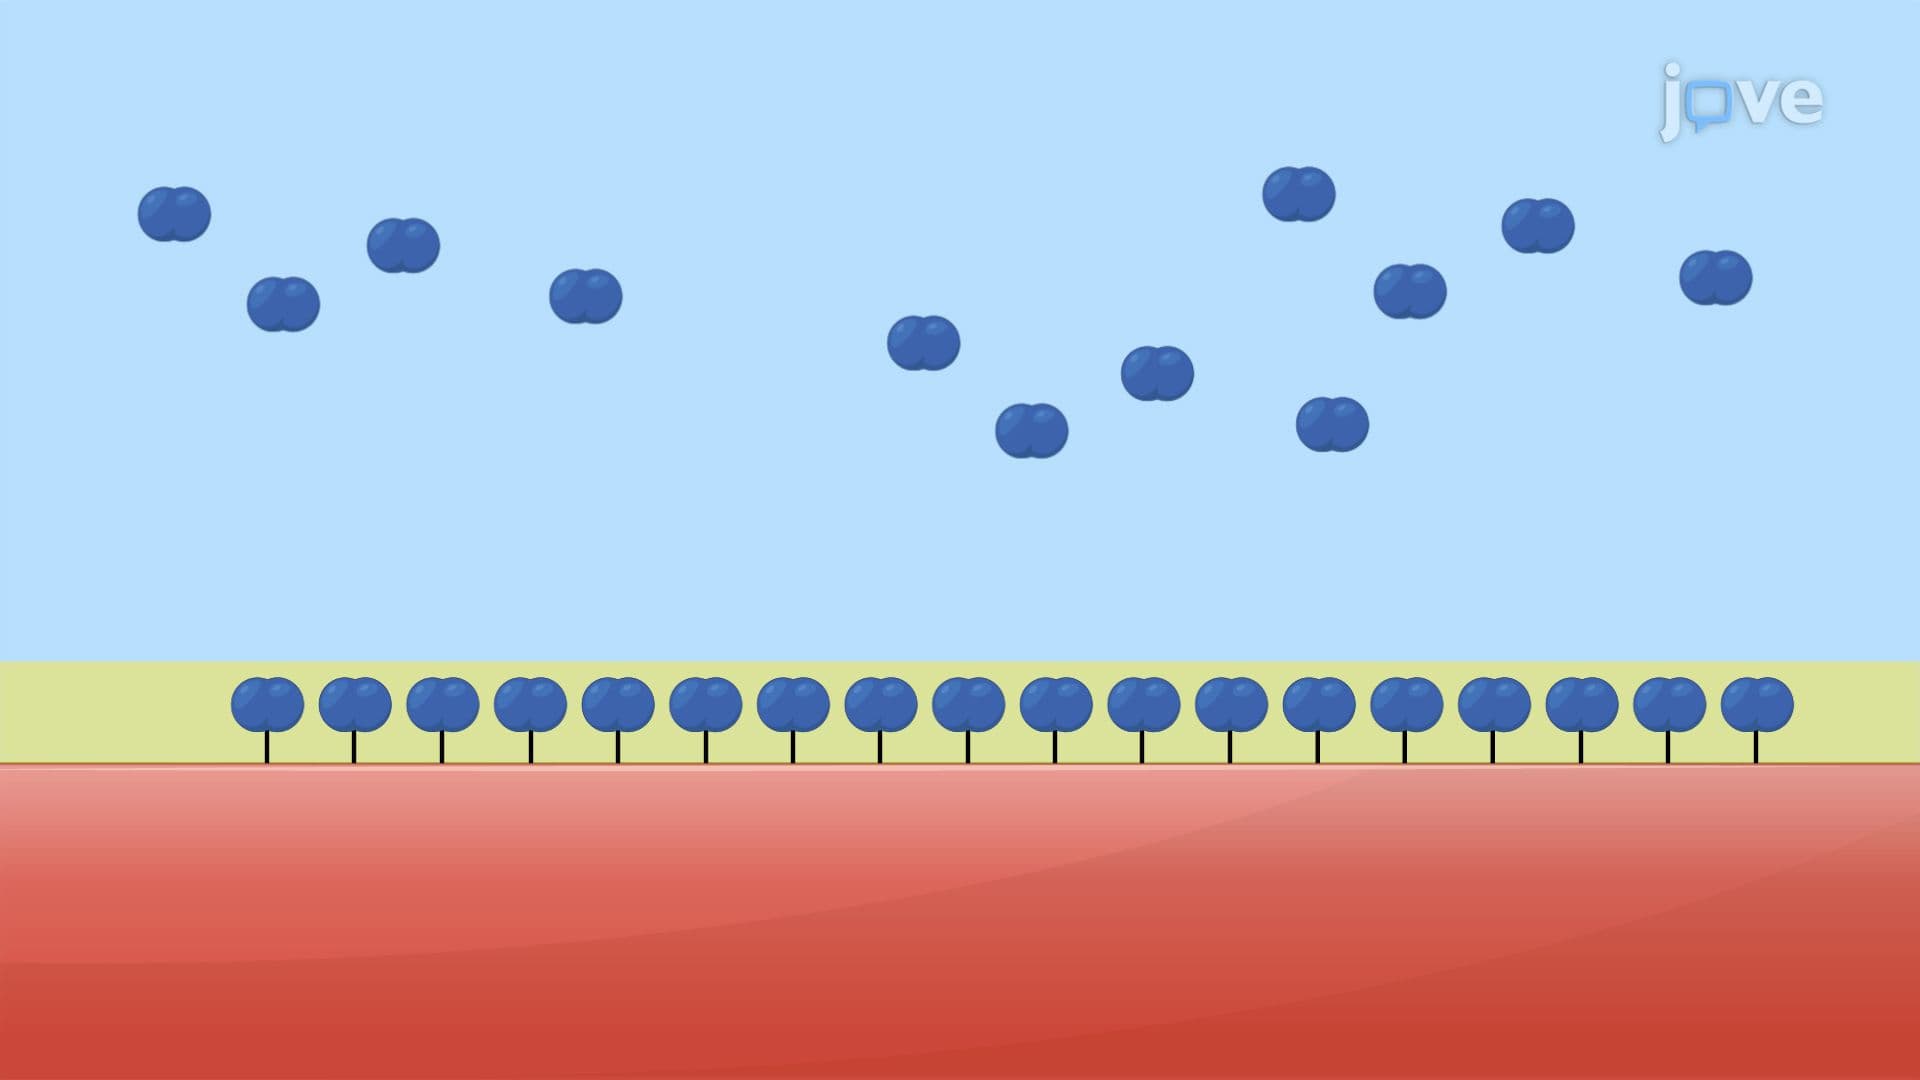
Adsorption of Gases on Solids
 01:29
Adsorption Isotherms I
 03:51
Molecular Orbital Theory II
 01:14
Molecular Spectroscopy: Absorption and Emission
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:45
The Quantum-Mechanical Model of an Atom