Related Experiment Video
Updated: Aug 7, 2026
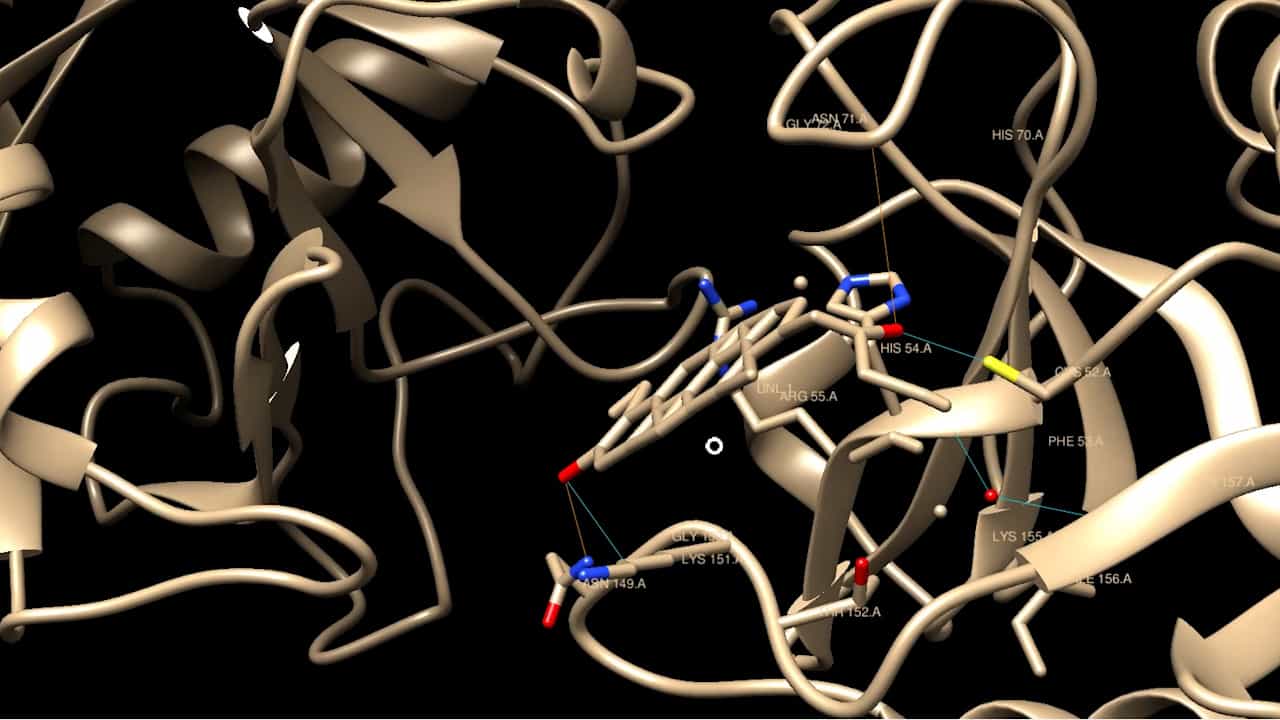

Structure-Guided Design and Development of Novel Cyclophilin A Inhibitors and Ganoderiol-F Derivatives: An In-Silico Approach
Published on: June 23, 2026
Molecular docking study and development of an empirical binding free energy model for phosphodiesterase 4 inhibitors
Fernanda G Oliveira1, Carlos M R Sant'Anna, Ernesto R Caffarena
1LASSBio, Laboratório de Avaliação e Síntese de Substãncias Bioativas, Faculdade de Farmácia and Instituto de Química, Universidade Federal do Rio de Janeiro, PO Box 68006, Rio de Janeiro, RJ 21944-910, Brazil.

Abstract:
In the present work, several computational methodologies were combined to develop a model for the prediction of PDE4B inhibitors' activity. The adequacy of applying the ligand docking approach, keeping the enzyme rigid, to the study of a series of PDE4 inhibitors was confirmed by a previous molecular dynamics analysis of the complete enzyme. An exhaustive docking procedure was performed to identify the most probable binding modes of the ligands to the enzyme, including the active site metal ions and the surrounding structural water molecules. The enzyme-inhibitor interaction enthalpies, refined by using the semiempirical molecular orbital approach, were combined with calculated solvation free energies and entropy considerations in an empirical free energy model that enabled the calculation of binding free energies that correlated very well with experimentally derived binding free energies. Our results indicate that both the inclusion of the structural water molecules close to the ions in the binding site and the use of a free energy model with a quadratic dependency on the ligand free energy of solvation are important aspects to be considered for molecular docking investigations involving the PDE4 enzyme family.
More Related Videos
 08:49
Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
 11:06
Network Pharmacology Prediction and Metabolomics Validation of the Mechanism of Fructus Phyllanthi against Hyperlipidemia
Published on: April 7, 2023
Related Concept Videos
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:27
Pharmacodynamic Models: Overview
 01:13
Physiological Pharmacokinetic Models: Assumption with Protein Binding
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 02:54
Protein Kinases and Phosphatases
Protein kinases
Many proteins in the cell are regulated by phosphorylation, the addition of a phosphate group. A family of enzymes called kinases...
 01:18
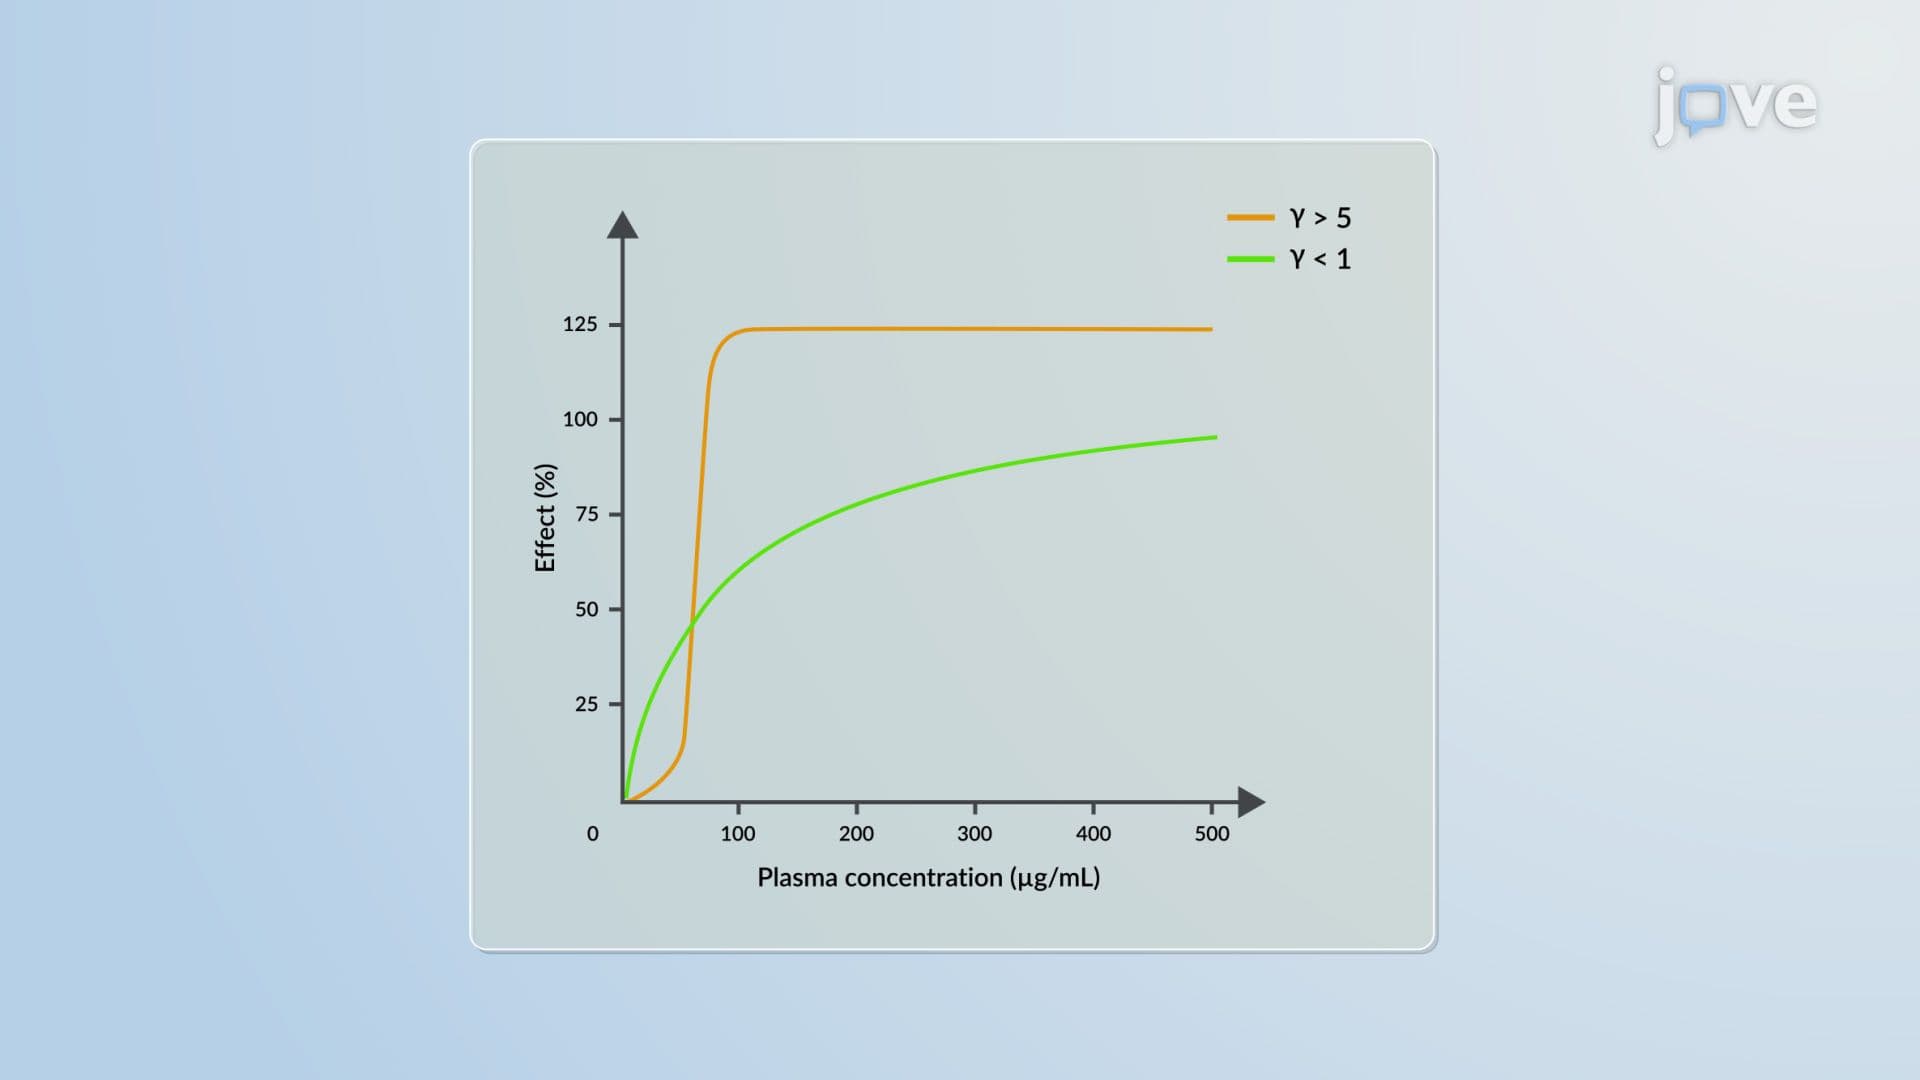
Pharmacodynamic Models: Emax Drug–Concentration Effect Model