Related Experiment Video
Updated: Aug 7, 2026
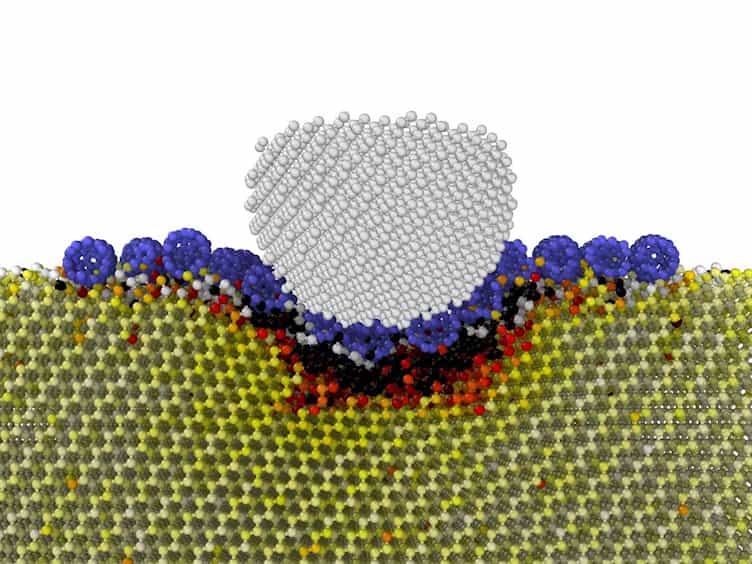

Probing C84-embedded Si Substrate Using Scanning Probe Microscopy and Molecular Dynamics
Published on: September 28, 2016
Hierarchical modeling N2 adsorption on the surface of and within a C60 crystal: from quantum mechanics to molecular
Jianwen Jiang1, Jeffery B Klauda, Stanley I Sandler
1Center for Molecular and Engineering Thermodynamics, Department of Chemical Engineering, University of Delaware, Newark, Delaware 19716, USA. jiangj@che.udel.edu

Abstract:
The adsorption of N(2) on the surface of, and within, a C(60) face-centered cubic crystal has been studied using a hierarchical approach. First, an ab initio potential between N(2) and C(60) is obtained from a recently developed quantum mechanical hybrid method, and then the adsorption behavior is predicted using Monte Carlo simulation. On the crystal surface, N(2) adsorption isotherm at 77.3 K is of type II. The adsorption simulated with the ab initio potential is slightly greater than that with the empirical Steele potential derived from experimental N(2) adsorption on planar graphite, and both are in fairly good agreement with measured results. With increasing pressure, N(2) molecules are found to sequentially occupy three favorable sites: the octahedral sites, the tetrahedral sites, and the top of C(60) molecules. Finally multiple layers form and wetting occurs as the bulk N(2) saturation pressure is reached. The isosteric heat of adsorption exhibits two maxima and finally approaches the enthalpy of vaporization of bulk N(2). Within the crystal, the N(2) adsorption isotherm at 77.3 K is of type I, and the use of ab initio potential leads to significantly greater adsorption than the Steele potential. N(2) molecules are observed to intercalate only the octahedral sites, and the isosteric heat of adsorption is nearly a constant. As in our previous work of N(2) and O(2) adsorption in the C(168) schwarzite (Jiang et al. J. Phys. Chem. B 2004, 108, 9852), this work demonstrates the importance of an accurate adsorbate-adsorbent interaction potential in the determination of gas adsorption behavior.
More Related Videos
 08:44
Measurements of Long-range Electronic Correlations During Femtosecond Diffraction Experiments Performed on Nanocrystals of Buckminsterfullerene
Published on: August 22, 2017
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Related Concept Videos
 02:00
Molecular Models
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 02:06
Newman Projections
The organic molecules rotate across the single bonds leading to numerous temporary three-dimensional structures of varying energy known as conformers.
 02:37
Metallic Solids
All metallic solids exhibit high thermal and electrical conductivity, metallic luster, and malleability. Many...
 01:18
Structure of Benzene: Molecular Orbital Model