Related Experiment Video
Updated: Aug 7, 2026

Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
A density functional theory study of molecular and dissociative adsorption of H2 on active sites in mordenite
1Institut für Materialphysik and Center for Computational Materials Science, Universität Wien, Sensengasse 8, A-1090 Wien, Austria. lubomir.benco@univie.ac.at

Abstract:
Adsorption and chemisorption of H2 in mordenite is studied using ab initio density functional theory (DFT) calculations. The geometries of the adsorption complex, the adsorption energies, stretching frequencies, and the capacity to dissociate the adsorbed molecule are compared for different active sites. The active centers include a Brønsted acid site, a three-coordinated surface Al site, and Lewis sites formed by extraframework cations: Na+, Cu+, Ag+, Zn2+, Cu2+, Ga3+, and Al3+. Adsorption properties of cations are compared for a location of the cation in the five-membered ring. This location differs from the location in the six-membered ring observed for hydrated cations. The five-membered ring, however, represents a stable location of the bare cation. In this position any cation exhibits higher reactivity compared with the location in the six-membered ring and is well accessible by molecules adsorbed in the main channel of the zeolite. Calculated adsorption energies range from 4 to 87 kJ/mol, depending on electronegativity and ionic radius of the cation and the stability of the cation-zeolite complex. The largest adsorption energy is observed for Cu+ and the lowest for Al3+ integrated into the interstitial site of the zeolite framework. A linear dependence is observed between the stretching frequency and the bond length of the adsorbed H2 molecule. The capacity of the metal-exchanged zeolite to dissociate the H2 molecule does not correlate with the adsorption energy. Dissociation is not possible on single Cu+ cation. The best performance is observed for the Ga3+, Zn2+, and Al3+ extraframework cations, in good agreement with experimental data.
More Related Videos
 11:38
In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
 09:46
Adsorption Device Based on a Langatate Crystal Microbalance for High Temperature High Pressure Gas Adsorption in Zeolite H-ZSM-5
Published on: August 25, 2016
Related Concept Videos
 01:29
Adsorption Isotherms I
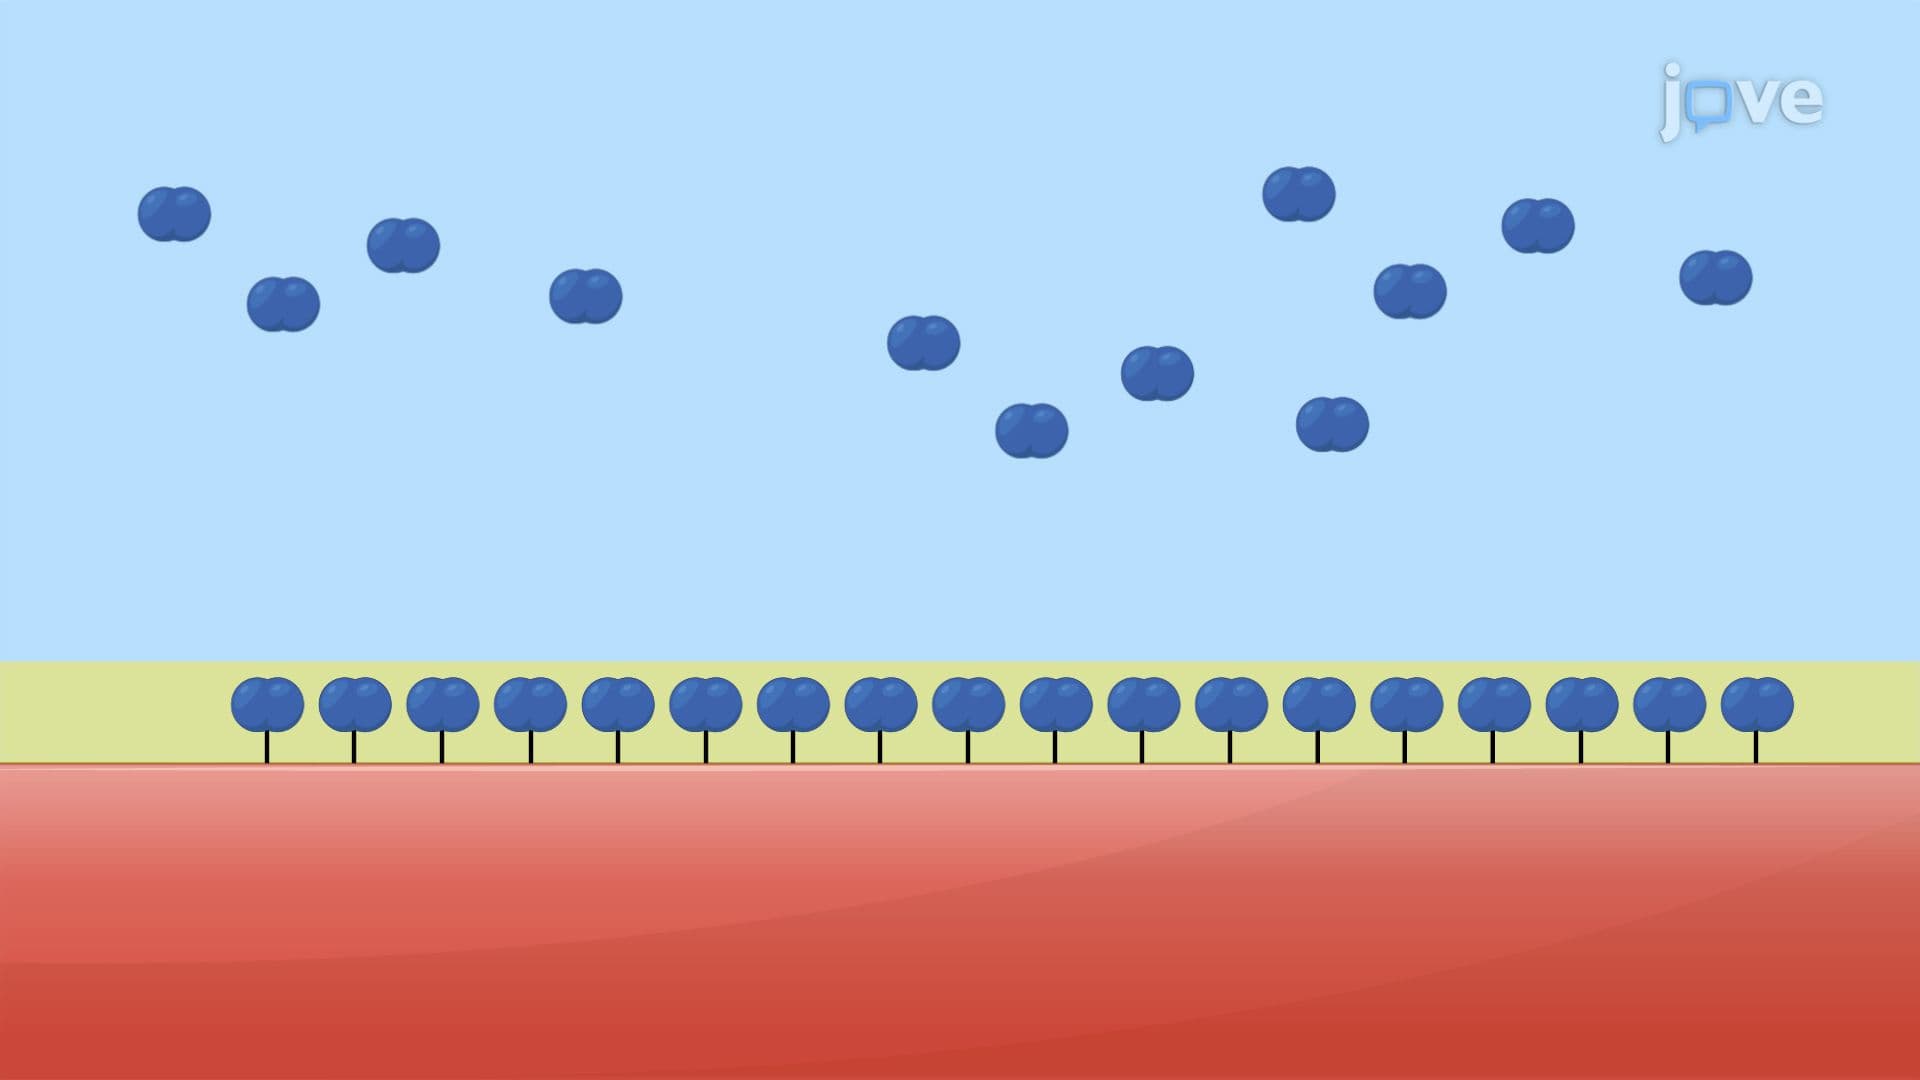 01:28
Adsorption of Gases on Solids
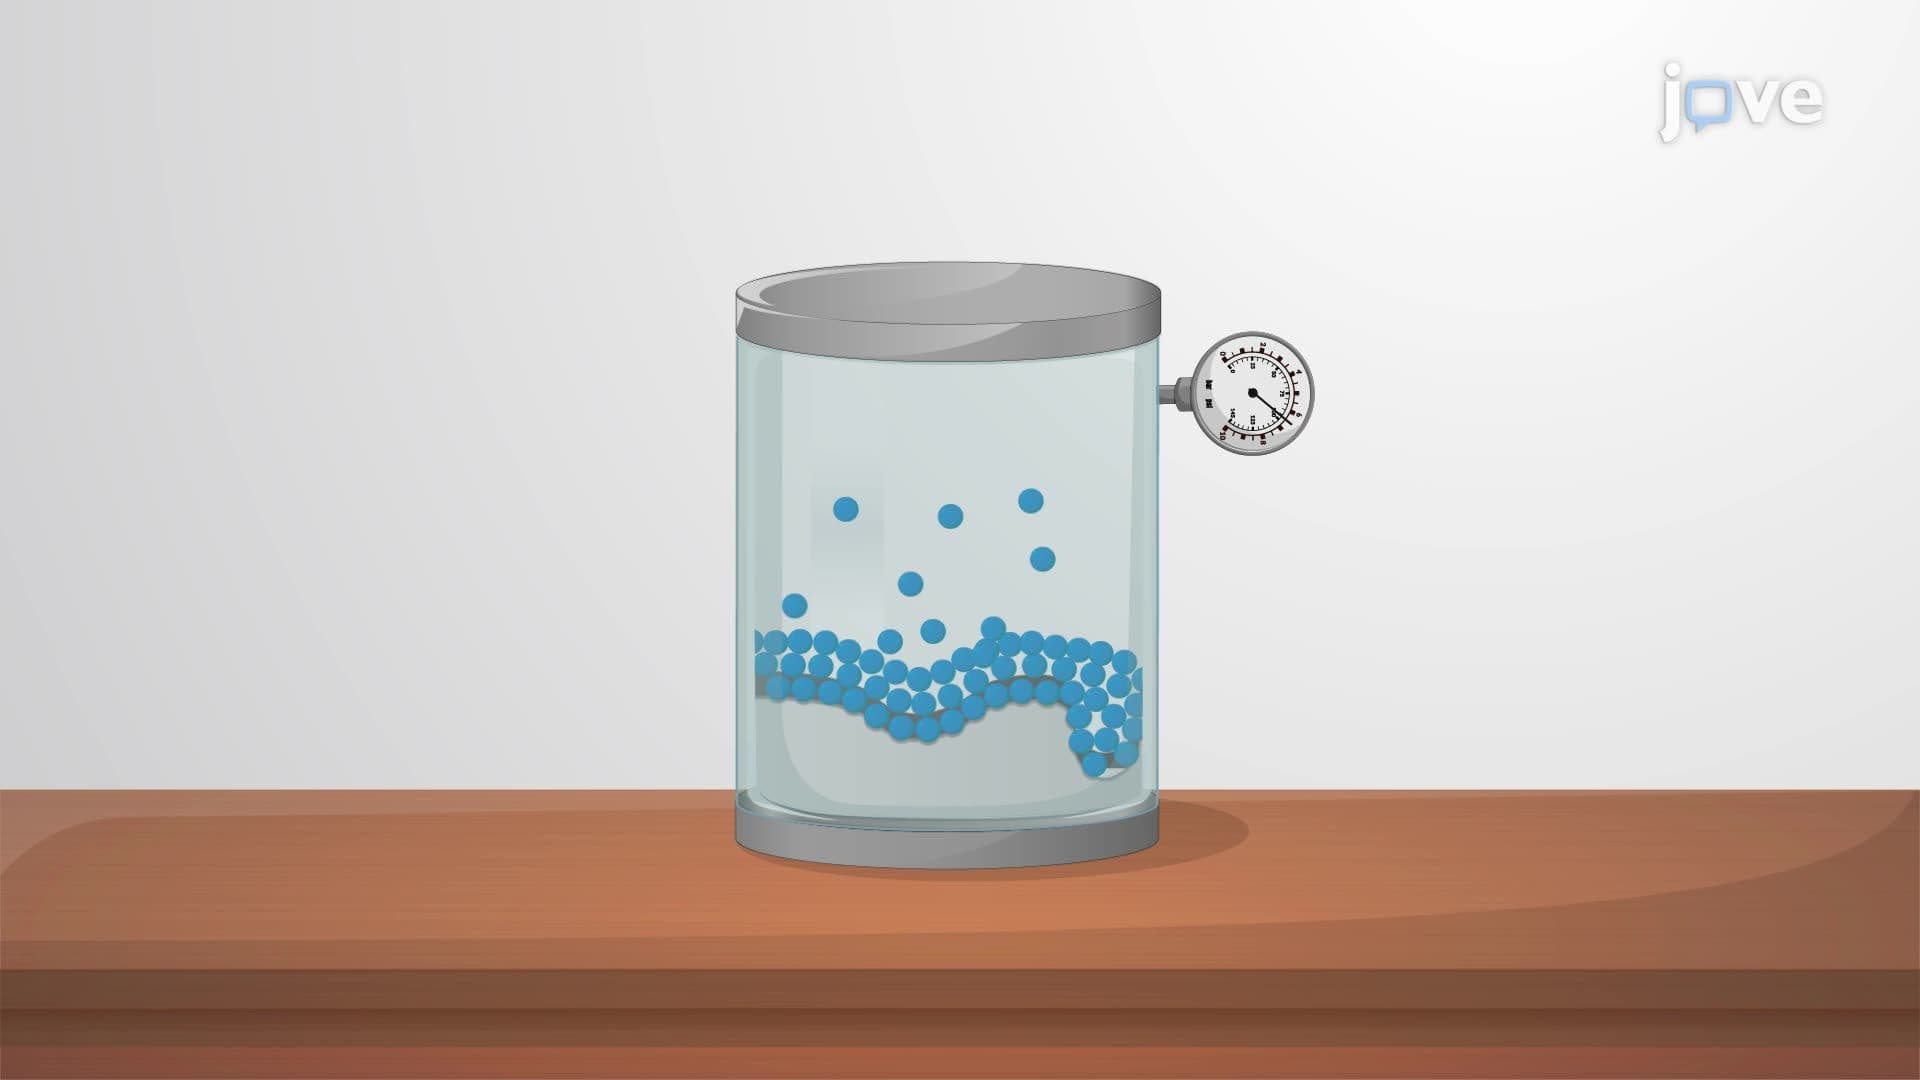 01:25
Adsorption Isotherms II
 01:22
Heterogeneous Catalysis
 03:51
Molecular Orbital Theory II
 02:58
Crystal Field Theory - Octahedral Complexes
To explain the observed behavior of transition metal complexes (such as colors), a model involving electrostatic interactions between the electrons from the ligands and the electrons in the unhybridized d orbitals of the central metal atom has been developed. This electrostatic model is crystal field theory (CFT). It helps to understand, interpret, and predict the colors, magnetic behavior, and some structures of coordination compounds of transition metals.
CFT focuses on...