Related Experiment Video
Updated: Aug 6, 2026

Prion Safety Laboratory Swipe Test
Published on: February 14, 2025
[Creutzfeldt-Jakob disease--the past or the future]
1I Klinika Neurologiczna, Instytutu Psychiatrii i Neurologii w Warszawie.

Insights
Creutzfeldt-Jakob disease (CJD) encompasses four forms: sporadic, familial, iatrogenic, and variant. While sporadic CJD has stable incidence, familial forms stem from PRNP gene mutations, and iatrogenic CJD is preventable. Variant CJD poses significant concerns due to its link with BSE.
Area of Science:
- Neurology
- Epidemiology
- Genetics
Context:
- Creutzfeldt-Jakob disease (CJD) comprises four primary forms with distinct ethiopathogenesis and epidemiological profiles.
- Understanding the incidence, genetic underpinnings, and transmission routes of CJD is crucial for public health.
- Recent advancements provide insights into the long-term outlook and challenges associated with CJD.
Purpose:
- To present current perspectives on the ethiopathogenesis and epidemiology of the four main forms of CJD.
- To discuss the stable incidence of sporadic CJD (sCJD) and the unknown reasons behind it.
- To highlight the genetic basis of familial CJD (fCJD), the preventability of iatrogenic CJD (iCJD), and the concerns surrounding variant CJD (vCJD) linked to BSE.
Summary:
- Sporadic CJD (sCJD) maintains a consistent incidence of approximately 1 case per million population annually, with its etiology remaining elusive.
- Familial CJD (fCJD) is an autosomal dominant condition linked to over 40 PRNP gene mutations, exhibiting diverse clinical and neuropathological phenotypes.
- Iatrogenic CJD (iCJD) has the most favorable epidemiological prognosis due to well-recognized causative medical errors and effective prevention strategies. Variant CJD (vCJD), associated with Bovine Spongiform Encephalopathy (BSE), raises serious global concerns due to its unknown incubation period and prion involvement of reticulo-lymphatic tissues.
Impact:
- Provides a comprehensive overview of CJD forms, aiding researchers and clinicians in understanding disease dynamics.
- Highlights the need for continued research into the causes of sCJD and the long-term implications of vCJD.
- Informs public health strategies for CJD prevention and management, particularly concerning iatrogenic and variant forms.
Abstract:
Some recent views on ethiopathogenesis and epidemiology of four main forms of CJD, based on up to-day experiences and expectations for the future, are presented. The sporadic form of the disease (sCJD) displays a stable morbidity--ca. 1 case/1 million population yearly. The reasons of its so constant appearance remain still unknown. The familial forms of CJD (fCJD) depending upon more than 40 mutations in PRNP gene known today are inherited as autosomal dominant train. The clinical and neuropathological phenotype of patients belonging to various families are usually very different. The best epidemiological prognosis is attributed to iatrogenic form of CJD (iCJD), since both the medical errors causative of the disease and methods of avoiding of them are now very good recognized. The serious fears in many countries raises variant CJD (vCJD), connected etiologically with BSE, because of unknown duration of its incubation period and regular chronic involvement by prions reticulo-lymphatic tissue in infected persons.
Related Concept Videos
 01:21
Huntington Disease l: Introduction
 01:23
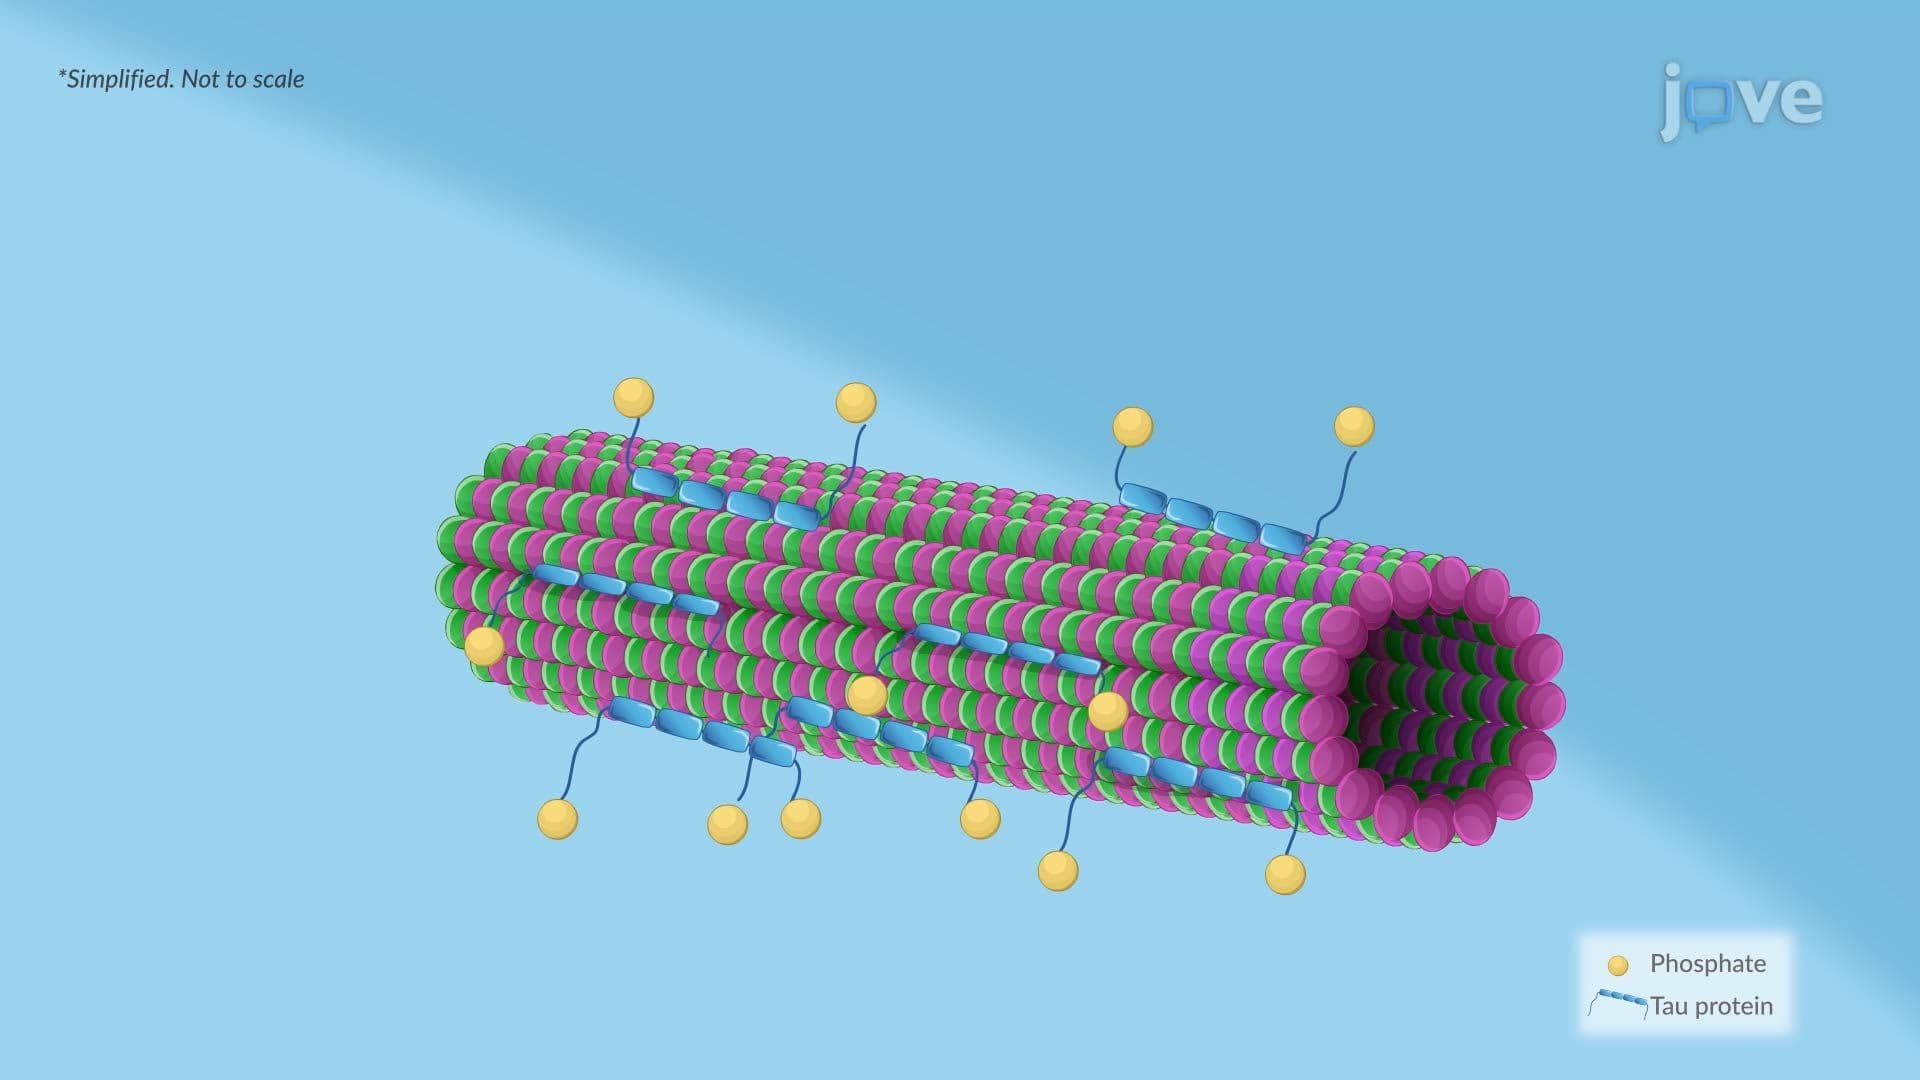
Alzheimer Disease ll: Pathophysiology
 01:26
Alzheimer's Disease: Overview
The clinical diagnosis of AD hinges on the presence of memory and other cognitive impairments. Biomarkers, such as changes in Aβ and tau...
 01:29
Alzheimer Disease l: Introduction
 03:03
Amyloid Fibrils
Amyloid deposits were observed as early as 1639 in the liver and the spleen. In 1854, Rudolph Virchow performed iodine staining, normally used to...
 01:24
Parkinson Disease ll: Pathophysiology