Related Experiment Video
Updated: Aug 6, 2026


Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
Drug transporter and metabolizing enzyme gene variants and nonnucleoside reverse-transcriptase inhibitor
Marylyn D Ritchie1, David W Haas, Alison A Motsinger
1Center for Human Genetics Research, Vanderbilt University School of Medicine, Nashville, TN 37232, USA. marylyn.ritchie@vanderbilt.edu

Abstract:
This nested case-control study examined relationships between MDR1, CYP2B6, and CYP3A4 variants and hepatotoxicity during antiretroviral therapy with either efavirenz- or nevirapine-containing regimens. Decreased risk of hepatotoxicity was associated with MDR1 3435C-->T (odds ratio, 0.254; P=.021). An interaction between MDR1 and hepatitis B surface antigen status predicted risk with 82% accuracy (P<.001).
Insights
Certain genetic variants, like MDR1, may reduce the risk of liver damage during antiretroviral therapy. An interaction between MDR1 and hepatitis B status accurately predicted this risk.
Area of Science:
- Pharmacogenetics
- Hepatology
- Infectious Diseases
Background:
- Hepatotoxicity is a significant concern during antiretroviral therapy (ART).
- Specific genetic factors and viral co-infections may influence ART-induced liver injury.
- Efavirenz and nevirapine are commonly used non-nucleoside reverse transcriptase inhibitors.
Purpose of the Study:
- To investigate the association between genetic variants in MDR1, CYP2B6, and CYP3A4 and hepatotoxicity in patients on efavirenz- or nevirapine-based ART.
- To identify genetic predictors of liver injury during antiretroviral treatment.
Main Methods:
- Nested case-control study design.
- Genotyping of MDR1, CYP2B6, and CYP3A4 gene variants.
- Analysis of clinical data including ART regimen and hepatitis B surface antigen (HBsAg) status.
Main Results:
- The MDR1 3435C-->T variant was associated with a decreased risk of hepatotoxicity (OR=0.254, P=.021).
- A significant interaction between MDR1 genotype and HBsAg status was observed, predicting hepatotoxicity risk with 82% accuracy (P<.001).
Conclusions:
- MDR1 genetic variations may play a protective role against ART-induced hepatotoxicity.
- Combined assessment of MDR1 genotype and HBsAg status can improve prediction of liver injury risk in patients receiving efavirenz or nevirapine.
Related Concept Videos
 01:27
Pharmacogenetics of Phase II Enzymes: N-acetyltransferase, Thiopurine S-methyltransferase, UDP-glucuronosyltransferase
 01:29
Pharmacogenetic Phenotypes: Alterations in Pharmacokinetics, Drug Targets and Biologic Milieu
 01:16
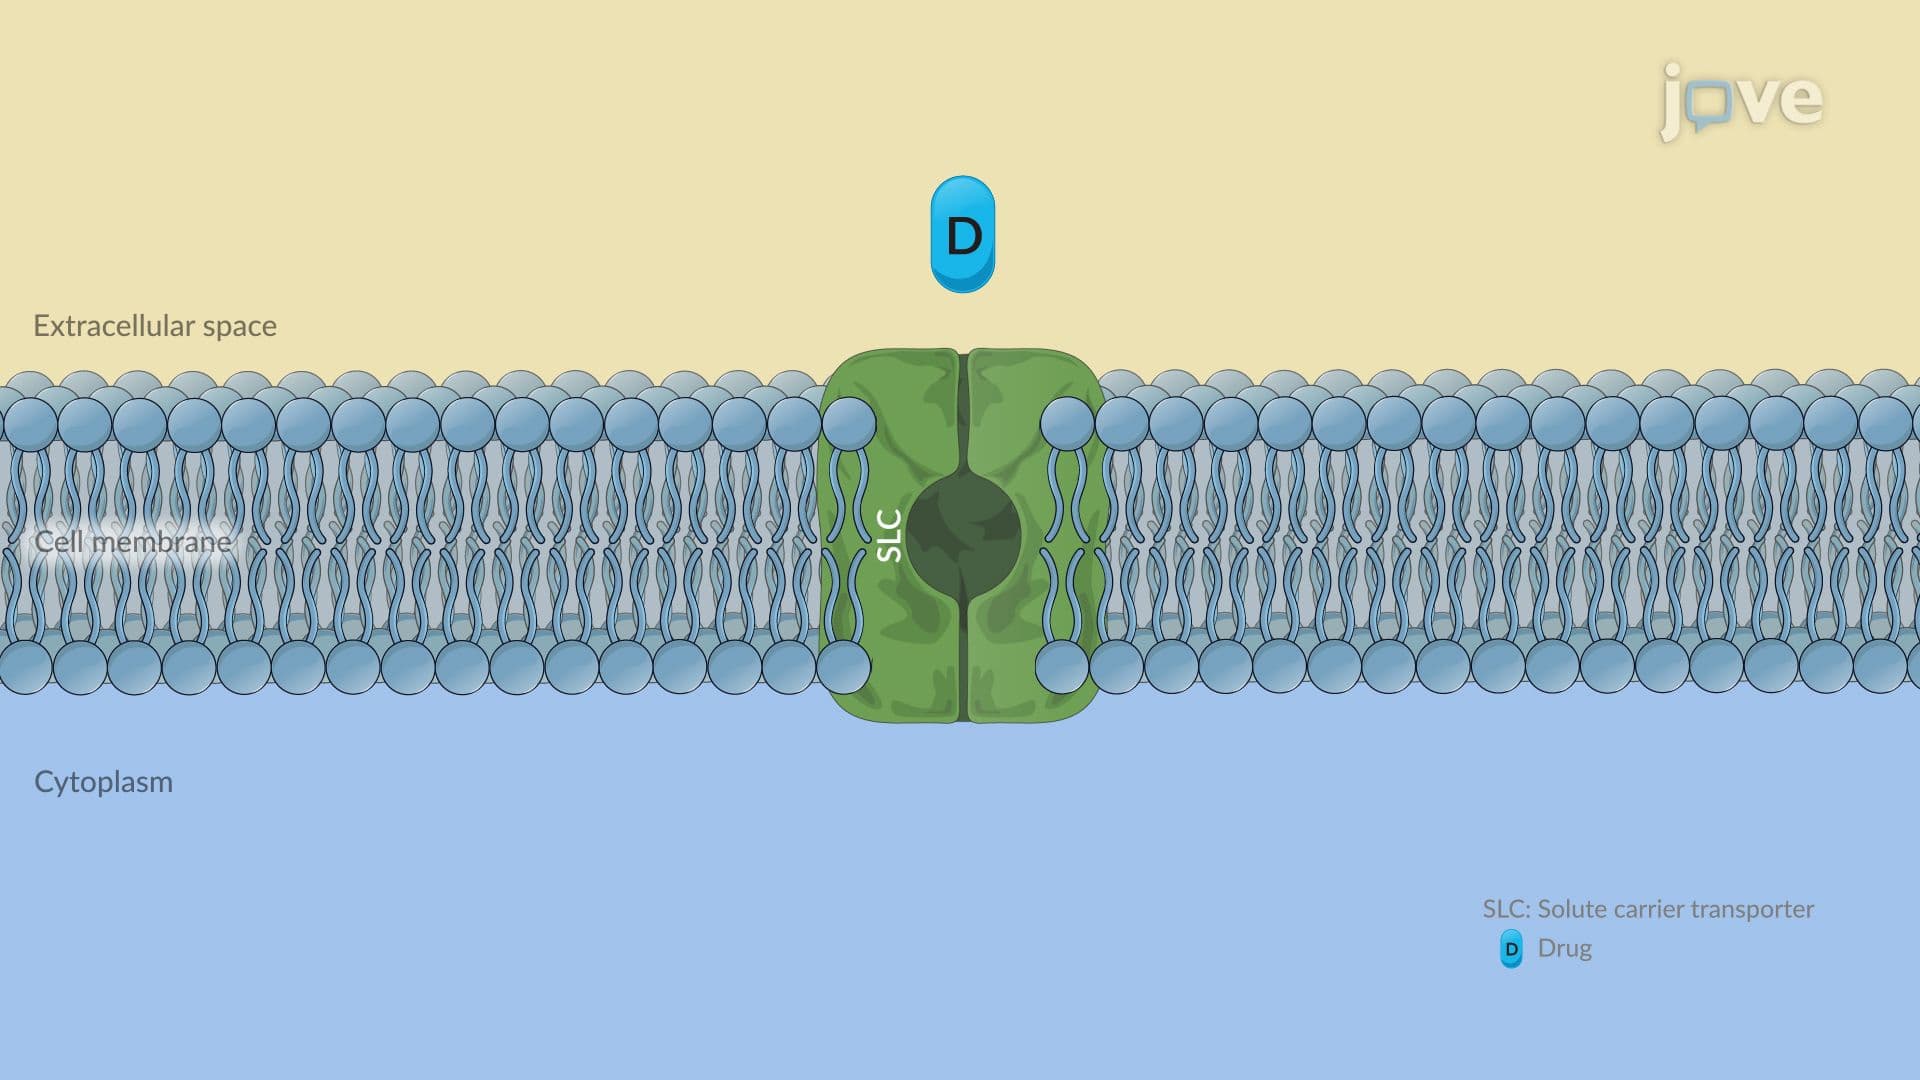
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:16
Drug toxicity: Idiosyncratic Reactions
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes
 01:27
Pharmacogenetics of Drug Metabolism: Overview