Related Experiment Video
Updated: Aug 6, 2026

The WinCF Model - An Inexpensive and Tractable Microcosm of a Mucus Plugged Bronchiole to Study the Microbiology of Lung Infections
Published on: May 8, 2017
Characterizing mucous cell remodeling in cystic fibrosis: relationship to neutrophils
1Division of Pulmonary and Critical Care Medicine, Department of Medicine, Cardiovascular Research Institute, University of California, San Francisco, San Francisco, CA 94143, USA.

Rationale:
Relatively few studies have characterized mucous cells or mucins in detail in cystic fibrosis (CF), and the relationship between mucous cell abnormalities and neutrophilic inflammation is uncertain.
Objectives:
To characterize mucous cell phenotypes and mucin profiles in CF and to determine if neutrophils accumulate around goblet cells in the epithelium and gland acini in the submucosa.
Methods:
Bronchial biopsies were collected from 7 subjects with CF and 15 control subjects, and the morphology of mucous cells was measured. Immunostains for gel-forming mucins and neutrophil elastase were quantified.
Measurements And Main Results:
Goblet cell size was increased in CF (p = 0.004), but the number of goblet cells was normal. The volume of submucosal glands was fourfold higher than normal (p = 0.031), but the proportion of mucous and serous cells in CF glands was normal. The patterns of expression of gel-forming mucins in epithelial and submucosal compartments in CF were similar to normal. Although neutrophil elastase immunostaining was intense in the epithelium in CF, neutrophils were largely absent around gland acini in the submucosa.
Conclusion:
The most prominent pathologic feature in the CF airway is an increase in submucosal gland volume, but serous cell transdifferentiation to mucous cells does not occur, nor are gland acini inflamed with neutrophils. The mechanism for increased submucosal gland volume in CF deserves further study.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:15
Differentiation of Common Myeloid Progenitor Cells
 01:24
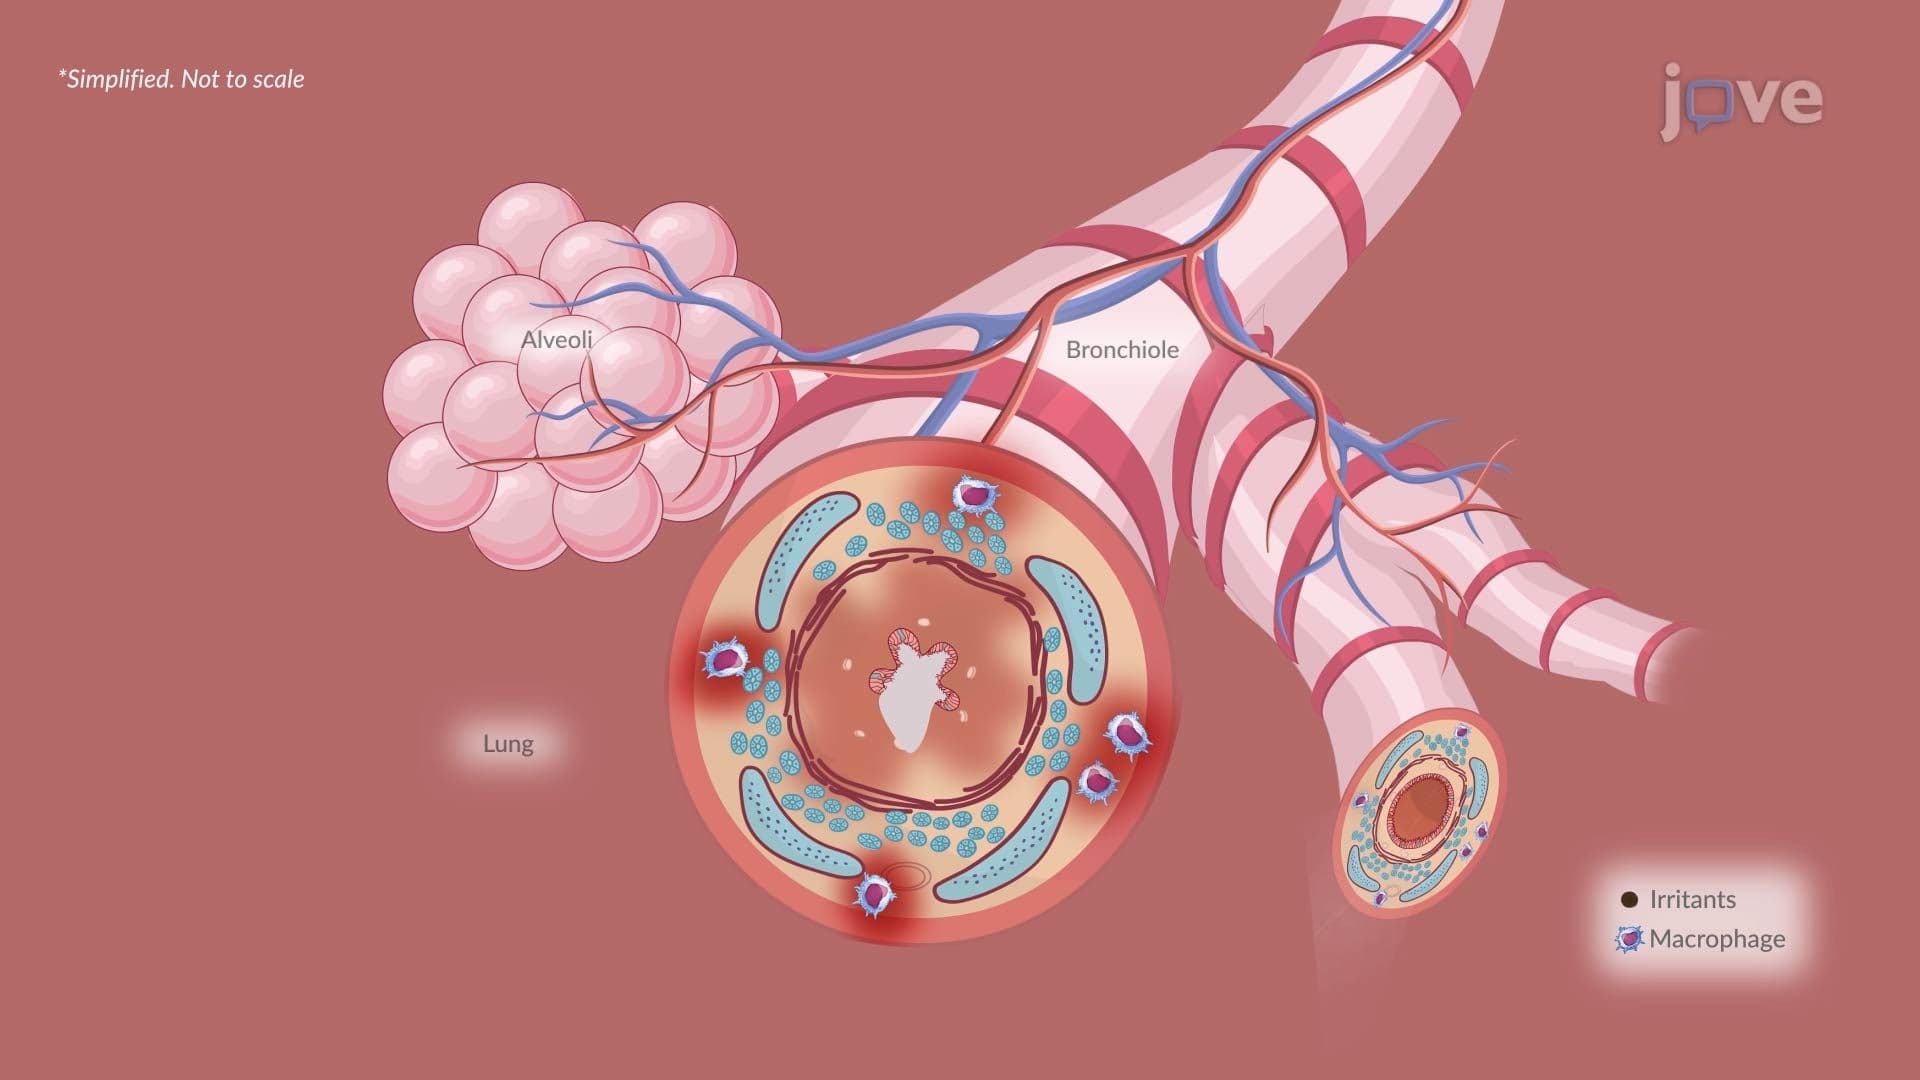
Chronic Obstructive Pulmonary Disease III: Chronic Bronchitis Features