Related Experiment Video
Updated: Jul 20, 2026
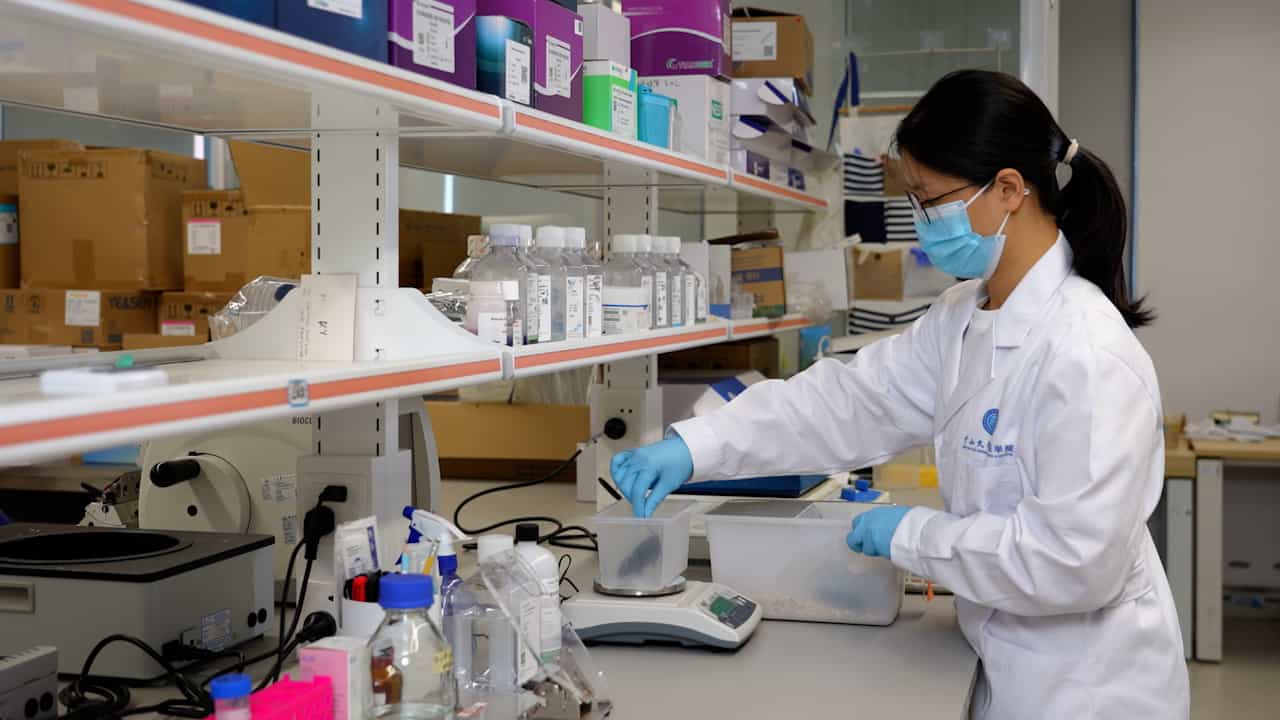

Refined Murine Model of Idiopathic Pulmonary Fibrosis
Published on: June 17, 2025
Idiopathic pulmonary fibrosis : new concepts in pathogenesis and implications for drug therapy
Jeffrey C Horowitz1, Victor J Thannickal
1Division of Pulmonary and Critical Care Medicine, Department of Medicine, University of Michigan Medical Center, Ann Arbor, Michigan, USA.

Abstract:
Idiopathic pulmonary fibrosis (IPF) is a chronic, progressive, and usually fatal pulmonary disease for which there are no proven drug therapies. Anti-inflammatory and immunosuppressive agents have been largely ineffective. The precise relationship of IPF to other idiopathic interstitial pneumonias (IIPs) is not known, despite the observation that different histopathologic patterns of IIP may coexist in the same patient. We propose that these different histopathologic 'reaction' patterns may be determined by complex interactions between host and environmental factors that alter the local alveolar milieu. Recent paradigms in IPF pathogenesis have focused on dysregulated epithelial-mesenchymal interactions, an imbalance in T(H)1/T(H)2 cytokine profile and potential roles for aberrant angiogenesis. In this review, we discuss these evolving concepts in disease pathogenesis and emerging therapies designed to target pro-fibrogenic pathways in IPF.
Related Concept Videos
 01:20
COPD: Pathogenesis and Clinical Features
The primary cause for the onset of COPD is cigarette smoking and exposure to air pollution. These hazardous factors initiate a chain reaction within the lungs, resulting in chronic inflammation, damage to the airways, and a...
 01:30
Pulmonary Hypertension: Classification and Pathogenesis
There are various classifications for PH, each relating to different underlying causes and also...
 01:20
Chronic Obstructive Pulmonary Disease-II: Pathophysiology
Chronic Inflammation
 01:29
Pharmacogenomics: Identification of New Drug Targets
 01:23
Chronic Obstructive Pulmonary Disease I: Introduction
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...