Novel aminoglycosides increase SMN levels in spinal muscular atrophy fibroblasts
Related Experiment Video
Updated: Jul 20, 2026
Evaluation of Exon Inclusion Induced by Splice Switching Antisense Oligonucleotides in SMA Patient Fibroblasts
Published on: May 11, 2018
Virginia B Mattis1, Ravi Rai, Jinhua Wang
1Department of Veterinary Pathobiology, Life Sciences Center, University of Missouri, Room 471G, Columbia, MO 65211-7310, USA.

Abstract:
Spinal muscular atrophy (SMA) is the leading genetic cause of infant mortality. SMA is caused by the homozygous absence of survival motor neuron-1 (SMN1). SMN2, a nearly identical copy gene, is retained in all SMA patients and encodes an identical protein as SMN1; however, SMN1 and SMN2 differ by a silent C to T transition which results in the production of an alternatively spliced isoform (SMNDelta7), which encodes a defective protein, demonstrating that the absence of the short peptide encoded by SMN exon 7 is critical in SMA development. Previously, we have shown that for some functions heterologous sequences can compensate for the exon 7 peptide, suggesting that the SMN C-terminus functions non-specifically. Consistent with this hypothesis, we now identify novel aminoglycosides that can induce SMN protein levels in patient fibroblasts. This hypothesis was supported, in part, by a novel fluorescent SMN read-through assay. Interestingly, however, through the development of a SMN exon 7-specific antibody, results suggested that levels of normal full-length SMN might also be elevated by aminoglycoside treatment. These results demonstrate that the compounds that promote read-through may provide an alternative platform for the discovery of compounds that induce SMN protein levels.
Insights
Novel aminoglycosides can increase levels of the full-length survival motor neuron (SMN) protein in spinal muscular atrophy (SMA) patient cells. This discovery offers a new avenue for developing SMA therapies by promoting SMN protein production.
Area of Science:
- Genetics
- Molecular Biology
- Neuroscience
Background:
- Spinal muscular atrophy (SMA) is a severe genetic disorder and the leading cause of infant mortality.
- SMA arises from the homozygous absence of the survival motor neuron-1 (SMN1) gene.
- The SMN2 gene, present in all SMA patients, produces a truncated, non-functional protein due to alternative splicing, specifically the exclusion of exon 7.
Purpose of the Study:
- To investigate novel compounds that can modulate SMN protein levels in SMA.
- To explore the potential of aminoglycosides in restoring functional SMN protein in SMA patient cells.
- To understand the mechanism by which compounds influence SMN protein production.
Main Methods:
- Utilized patient-derived fibroblasts to screen for compounds that increase SMN protein.
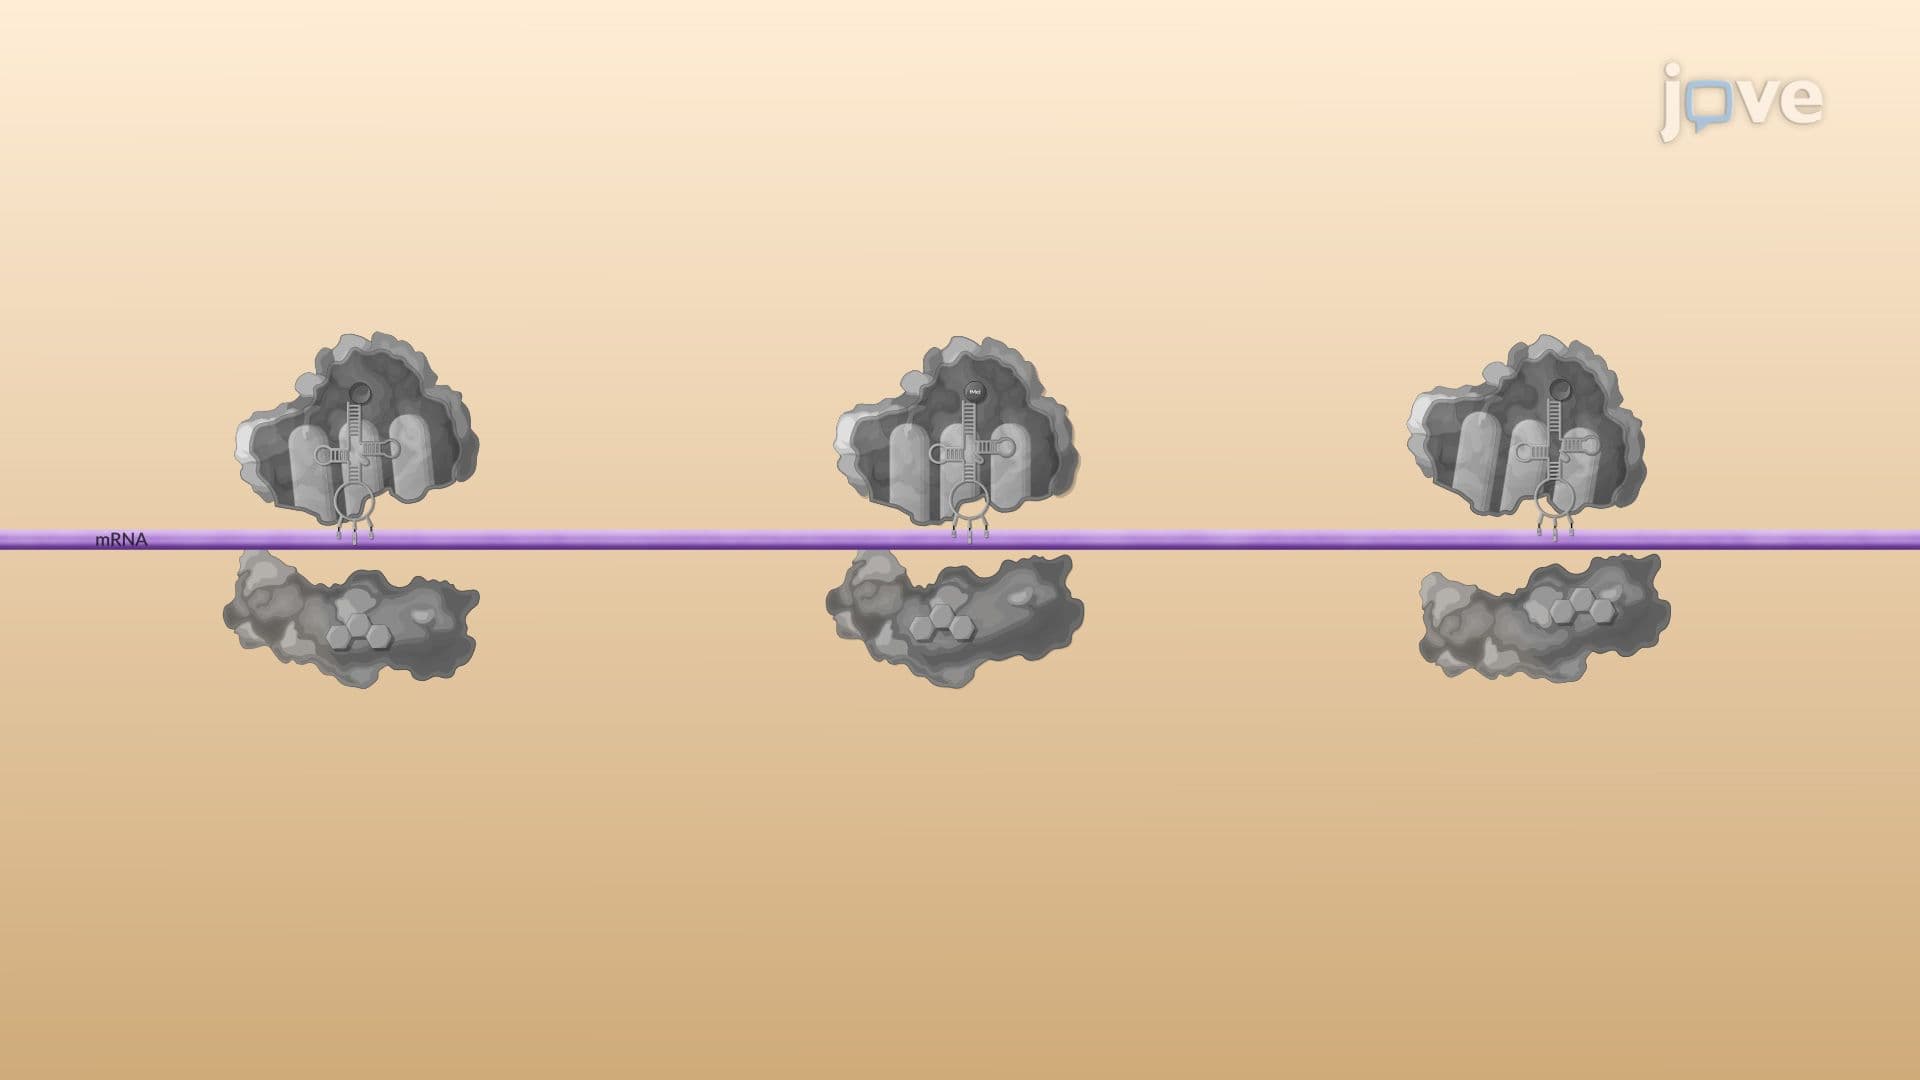
- Developed a novel fluorescent SMN read-through assay to monitor SMN protein production.
- Employed a newly developed SMN exon 7-specific antibody to differentiate between full-length and truncated SMN protein.
Main Results:
- Identified novel aminoglycosides capable of inducing SMN protein levels in SMA patient fibroblasts.
- The fluorescent SMN read-through assay provided partial support for the hypothesis that compounds can enhance SMN levels.
- Evidence suggested that aminoglycoside treatment may also elevate levels of normal, full-length SMN protein.
Conclusions:
- Aminoglycosides represent a promising class of compounds for increasing SMN protein levels in SMA.
- Compounds promoting SMN 'read-through' offer a potential therapeutic strategy for SMA.
- Further research into these compounds could lead to novel treatments for spinal muscular atrophy.