Related Experiment Video
Updated: Jul 20, 2026

Synthesis of a Borylated Ibuprofen Derivative Through Suzuki Cross-Coupling and Alkene Boracarboxylation Reactions
Published on: November 30, 2022
Structure of the boronic acid dimer and the relative stabilities of its conformers
Joseph D Larkin1, Krishna L Bhat, George D Markham
1Center for Computational Chemistry, University of Georgia, Athens, GA 30602, USA. jlarkin@ccqc.uga.edu

Abstract:
Despite the widespread use of boronic acids in materials science and as pharmaceutical agents, many aspects of their structure and reactivity are not well understood. In this research the boronic acid dimer, [HB(OH)(2)](2), was studied by second-order Møller-Plesset (MP2) perturbation theory and coupled cluster methodology with single and double excitations (CCSD). Pople split-valence 6-31+G*, 6-311G**, and 6-311++G** and Dunning-Woon correlation-consistent cc-pVDZ, aug-cc-pVDZ, cc-pVTZ, and aug-cc-pVTZ basis sets were employed for the calculations. A doubly hydrogen-bonded conformer (1) of the dimer was consistently found to be lowest in energy; the structure of 1 was planar (C(2h)) at most computational levels employed but was significantly nonplanar (C(2)) at the MP2/6-311++G** and CCSD/6-311++G** levels, the result of an intrinsic problem with Pople-type sp-diffuse basis functions on heavy atoms. The dimerization energy, enthalpy, and free energy for the formation of (1) from the exo-endo conformer of the monomer were -10.8, -9.2, and +1.2 kcal/mol, respectively, at the MP2/aug-cc-pVTZ level. Several other hydrogen-bonded conformers of the dimer were local minima on the potential energy surface (PES) and ranged from 2 to 5 kcal/mol higher in energy than 1. Nine doubly OH-bridged conformers, in which the boron atoms were tetracoordinated, were also local minima on the PES, but they were all greater than 13 kcal/mol higher in energy than 1; doubly H-bridged structures proved to be transition states. MP2 and CCSD results were compared to those from the BLYP, B3LYP, OLYP, O3LYP, PBE1PBE, and TPSS functionals with the 6-311++G** and aug-cc-pVTZ basis sets; the PBE1PBE functional performed best relative to the MP2 and CCSD results. Self-consistent reaction field (SCRF) calculations predict that boronic acid dimerization is less favorable in solution than in vacuo.
Related Concept Videos
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:20
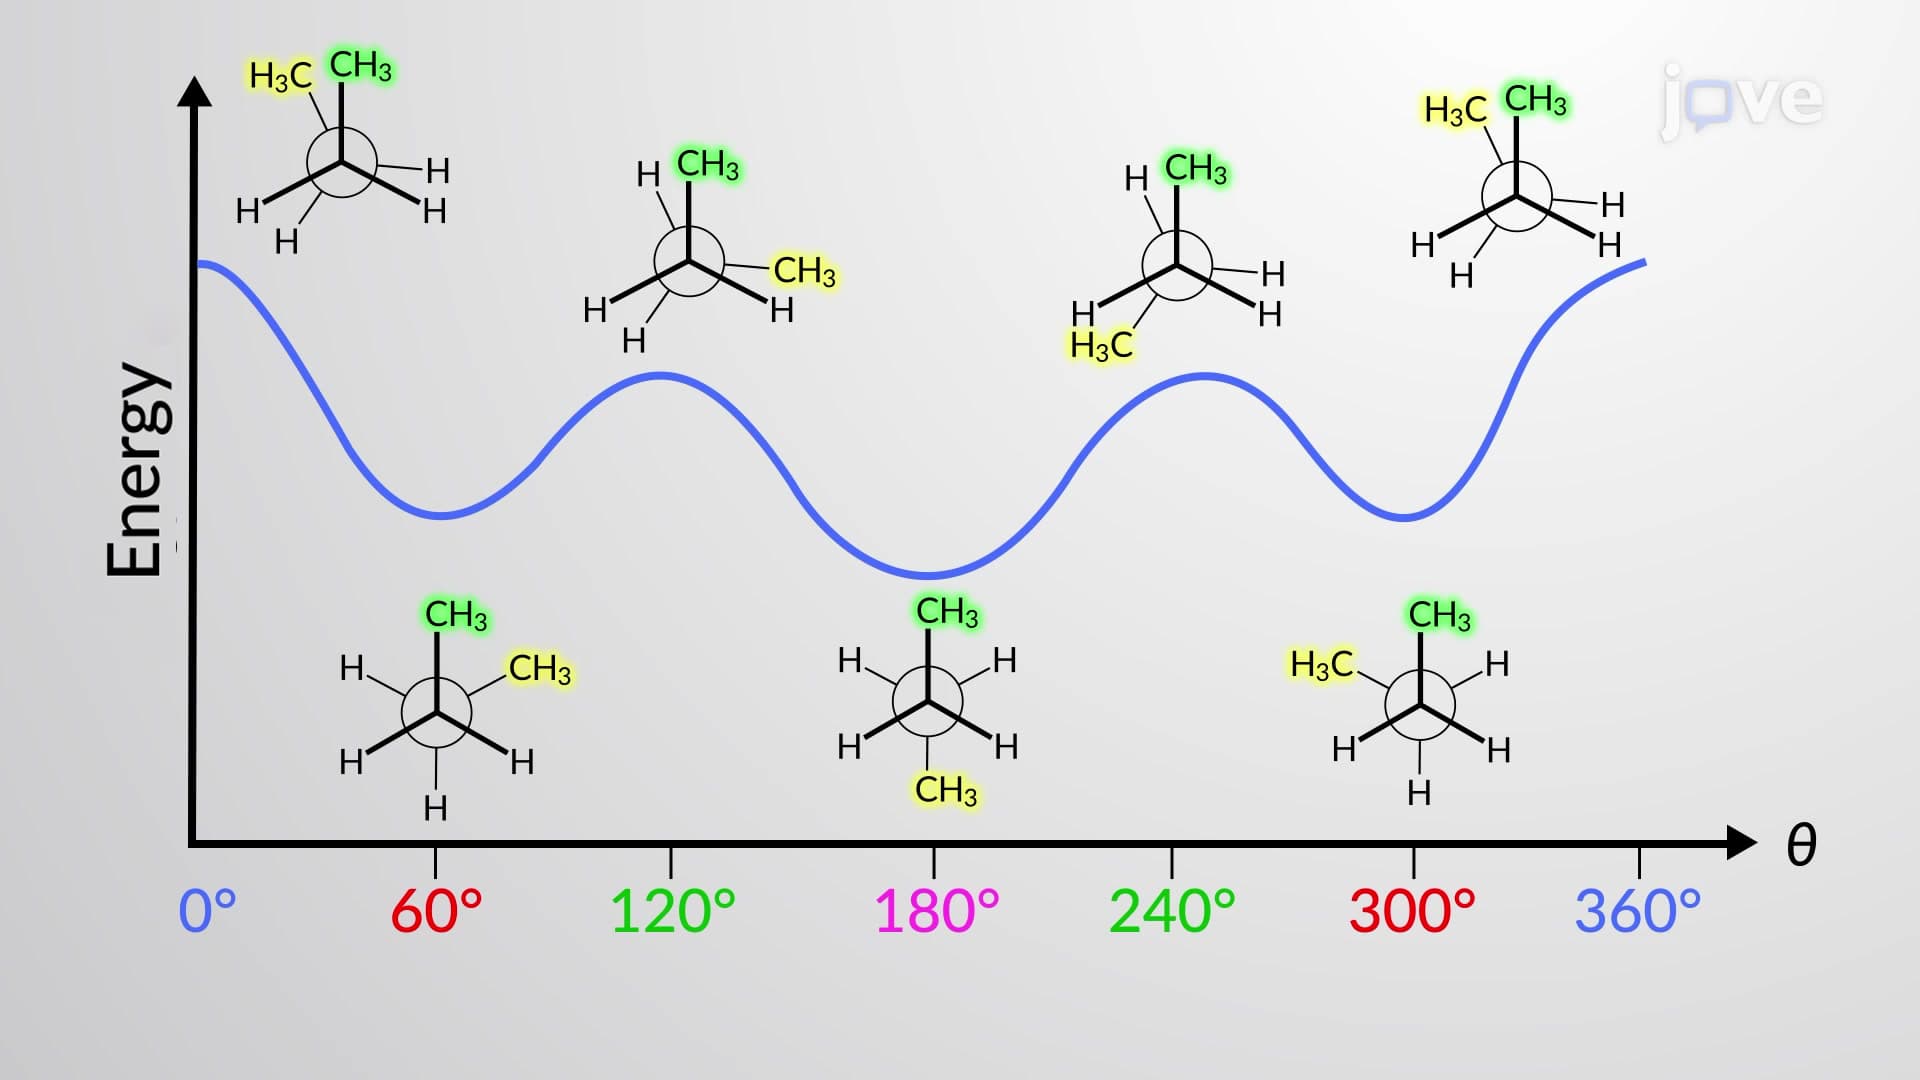
Conformations of Butane
 02:52
VSEPR Theory and the Basic Shapes
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 02:36
Regioselectivity and Stereochemistry of Hydroboration
Hydroboration proceeds in a concerted fashion with the attack of borane on the π bond, giving a cyclic four-centered transition state. The –BH2 group is bonded to the less substituted carbon and –H to the more substituted carbon. The concerted nature requires the simultaneous addition of –H and –BH2 across the same face of the alkene giving syn stereochemistry.