Related Experiment Video
Updated: Jul 20, 2026

Quantitative Structure-Activity Relationship, Activity Prediction, and Molecular Dynamics of Non-nucleotide Reverse Transcriptase Inhibitors
Published on: May 9, 2025
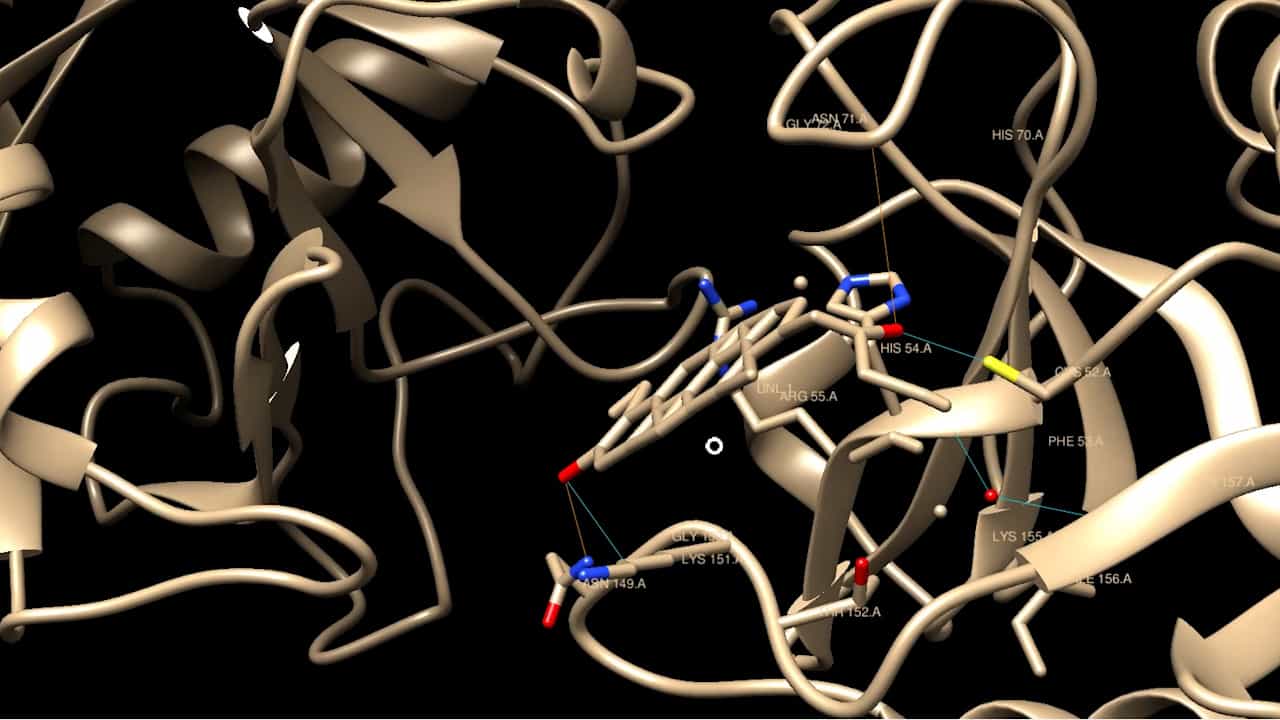
Structure modeling, ligand binding, and binding affinity calculation (LR-MM-PBSA) of human heparanase for inhibition
Zhigang Zhou1, Maria Bates, Jeffry D Madura
1Department of Chemistry and Biochemistry, Duquesne University, Pittsburgh, Pennsylvania 15213, USA. zhou@purdue.edu

Abstract:
Human heparanase is an endo-beta-D-glycosidase that cleaves heparan sulphate (HS) chains in the extracellular matrix and basement membrane. It is known that the cleavage of HS by heparanase results in cell invasion and metastasis of cancer. Therefore, heparanase is considered an important target for cancer drug development. The three-dimensional structure of heparanase would be useful in the rational design of inhibitors targeted to the enzyme; however, the three-dimensional structure has not yet been determined. In our effort to design inhibitors, we developed a three-dimensional structure of heparanase using a homology-modeling approach. The homology-built structure is consistent to previous bioinformatics and site-mutation experimental results. The heparanase features a (alpha/beta)(8) TIM-barrel fold with two glutamate residues (Glu225 and Glu343) located in the active-site cleft. This feature supports the putative mechanism of proton donor and nucleophilic sites. Docking simulations yielded 41 complex structures, which indicate that the bound inhibitor could block ligand binding into the catalytic site. A free energy of binding model was established for 25 heparanase inhibitors with a training set of 25 heparanase inhibitors using the linear response MM-PBSA approach (LR-MM-PBSA). The correlation between calculated and experimental activity was 0.79 and the reliability of the model was validated with leave-one-out cross-validation method. Its predictive capability was further validated using a test set of 16 inhibitors similar to the training set of inhibitors. The correlation between the predicted and observed activities is significantly improved by the protein "induced-fit" that accounts for the flexibility of the receptor. These interaction and pharmacophore elements provide a unique insight to the rational design of new ligands targeted to the enzyme.
More Related Videos
Related Concept Videos
 02:40
Ligand Binding Sites
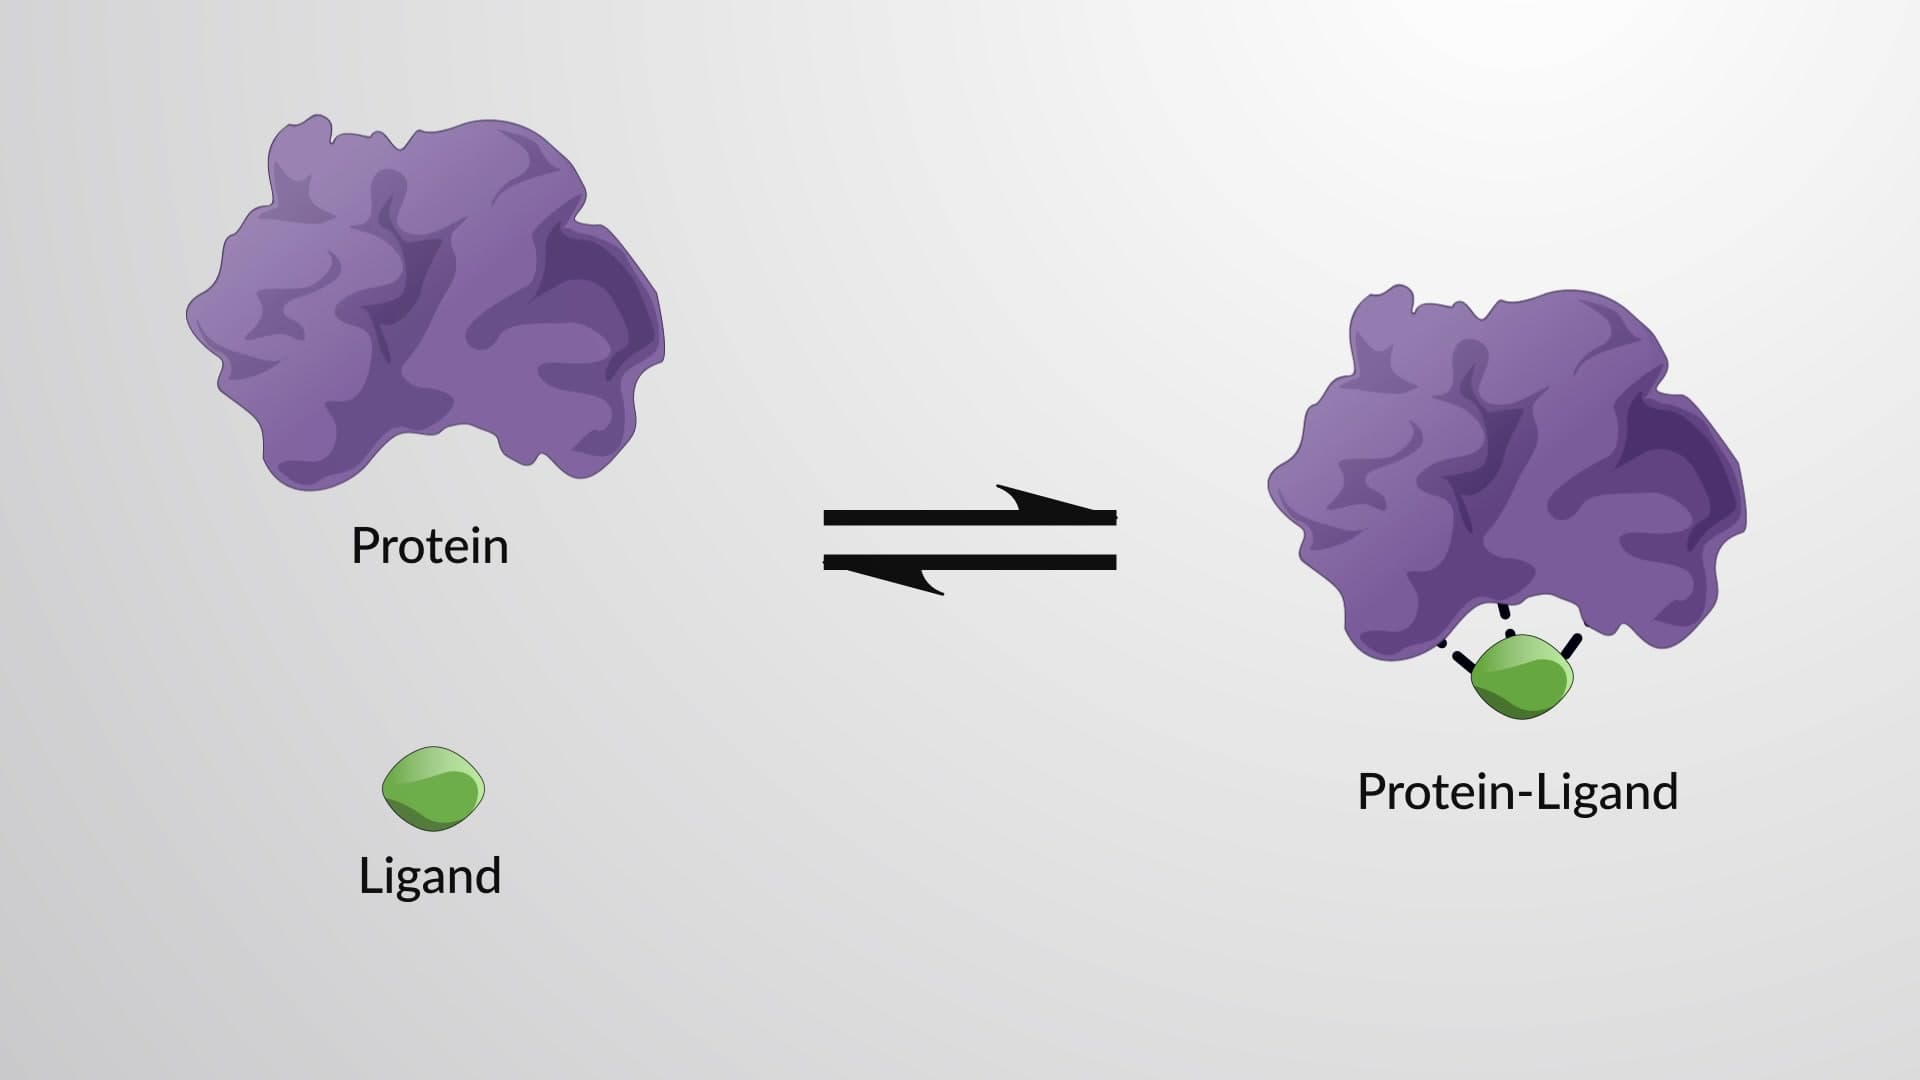
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
02:40Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 02:18
The Equilibrium Binding Constant and Binding Strength
 00:49
Ligand Binding and Linkage
00:49Ligand Binding and Linkage