Related Experiment Video
Updated: Jul 19, 2026

Identification of Orexin and Endocannabinoid Receptors in Adult Zebrafish Using Immunoperoxidase and Immunofluorescence Methods
Published on: June 25, 2019
Orexin-1 receptor-cannabinoid CB1 receptor heterodimerization results in both ligand-dependent and -independent
James Ellis1, John D Pediani, Meritxell Canals
1Molecular Pharmacology Group, Division of Biochemistry and Molecular Biology, Institute of Biomedical and Life Sciences, University of Glasgow, Glasgow G12 8QQ, Scotland, United Kingdom.

Abstract:
Following inducible expression in HEK293 cells, the human orexin-1 receptor was targeted to the cell surface but became internalized following exposure to the peptide agonist orexin A. By contrast, constitutive expression of the human cannabinoid CB1 receptor resulted in a predominantly punctate, intracellular distribution pattern consistent with spontaneous, agonist-independent internalization. Expression of the orexin-1 receptor in the presence of the CB1 receptor resulted in both receptors displaying the spontaneous internalization phenotype. Single cell fluorescence resonance energy transfer imaging indicated the two receptors were present as heterodimers/oligomers in intracellular vesicles. Addition of the CB1 receptor antagonist SR-141716A to cells expressing only the CB1 receptor resulted in re-localization of the receptor to the cell surface. Although SR-141716A has no significant affinity for the orexin-1 receptor, in cells co-expressing the CB1 receptor, the orexin-1 receptor was also re-localized to the cell surface by treatment with SR-141716A. Treatment of cells co-expressing the orexin-1 and CB1 receptors with the orexin-1 receptor antagonist SB-674042 also resulted in re-localization of both receptors to the cell surface. Treatment with SR-141716A resulted in decreased potency of orexin A to activate the mitogen-activated protein kinases ERK1/2 only in cells co-expressing the two receptors. Treatment with SB-674042 also reduced the potency of a CB1 receptor agonist to phosphorylate ERK1/2 only when the two receptors were co-expressed. These studies introduce an entirely novel pharmacological paradigm, whereby ligands modulate the function of receptors for which they have no significant inherent affinity by acting as regulators of receptor heterodimers.
Insights
Ligands can alter the function of unrelated receptors by targeting receptor heterodimers. This novel mechanism was observed with orexin-1 and cannabinoid CB1 receptors, impacting cellular signaling pathways.
Area of Science:
- Neuropharmacology
- Cell Biology
- Receptor Signaling
Background:
- The human orexin-1 receptor (OX1R) and cannabinoid CB1 receptor (CB1R) are G protein-coupled receptors involved in various physiological processes.
- Receptor trafficking and internalization are critical for regulating cellular responses to stimuli.
- Heterodimerization between different receptor types can influence their individual signaling properties.
Purpose of the Study:
- To investigate the interaction and functional consequences of co-expressing OX1R and CB1R in HEK293 cells.
- To explore the role of receptor heterodimerization in ligand-induced and spontaneous receptor trafficking.
- To determine if ligands targeting one receptor can modulate the function of a co-expressed, unrelated receptor.
Main Methods:
- Inducible and constitutive expression of OX1R and CB1R in HEK293 cells.
- Confocal microscopy and single-cell fluorescence resonance energy transfer (FRET) imaging to assess receptor localization and heterodimerization.
- Pharmacological treatments with receptor agonists and antagonists (SR-141716A for CB1R, SB-674042 for OX1R).
- Measurement of mitogen-activated protein kinase (MAPK) pathway activation (ERK1/2 phosphorylation) to assess receptor function.
Main Results:
- Constitutive CB1R expression led to spontaneous internalization, while OX1R showed agonist-induced internalization.
- Co-expression of OX1R and CB1R resulted in both receptors exhibiting spontaneous internalization, forming intracellular heterodimers/oligomers.
- Antagonists for one receptor (SR-141716A or SB-674042) induced cell surface re-localization of both co-expressed receptors.
- Ligands targeting one receptor altered the potency of the other receptor's agonist in activating ERK1/2 signaling specifically in co-expressing cells.
Conclusions:
- OX1R and CB1R form functional heterodimers that influence each other's trafficking and signaling.
- Ligands can indirectly modulate the function of unrelated receptors through their action on receptor heterodimers.
- This study reveals a novel pharmacological paradigm where receptor heterodimerization mediates cross-talk between distinct receptor systems.
More Related Videos
Related Concept Videos
 01:22
Opioid Receptors: Overview
 01:29
The Two-State Receptor Model
The binding affinity of a drug determines its interaction with one...
 01:31
Transducer Mechanism: Nuclear Receptors
About 48 different soluble family members of nuclear receptors are identified that can be divided into two main classes:
 01:25
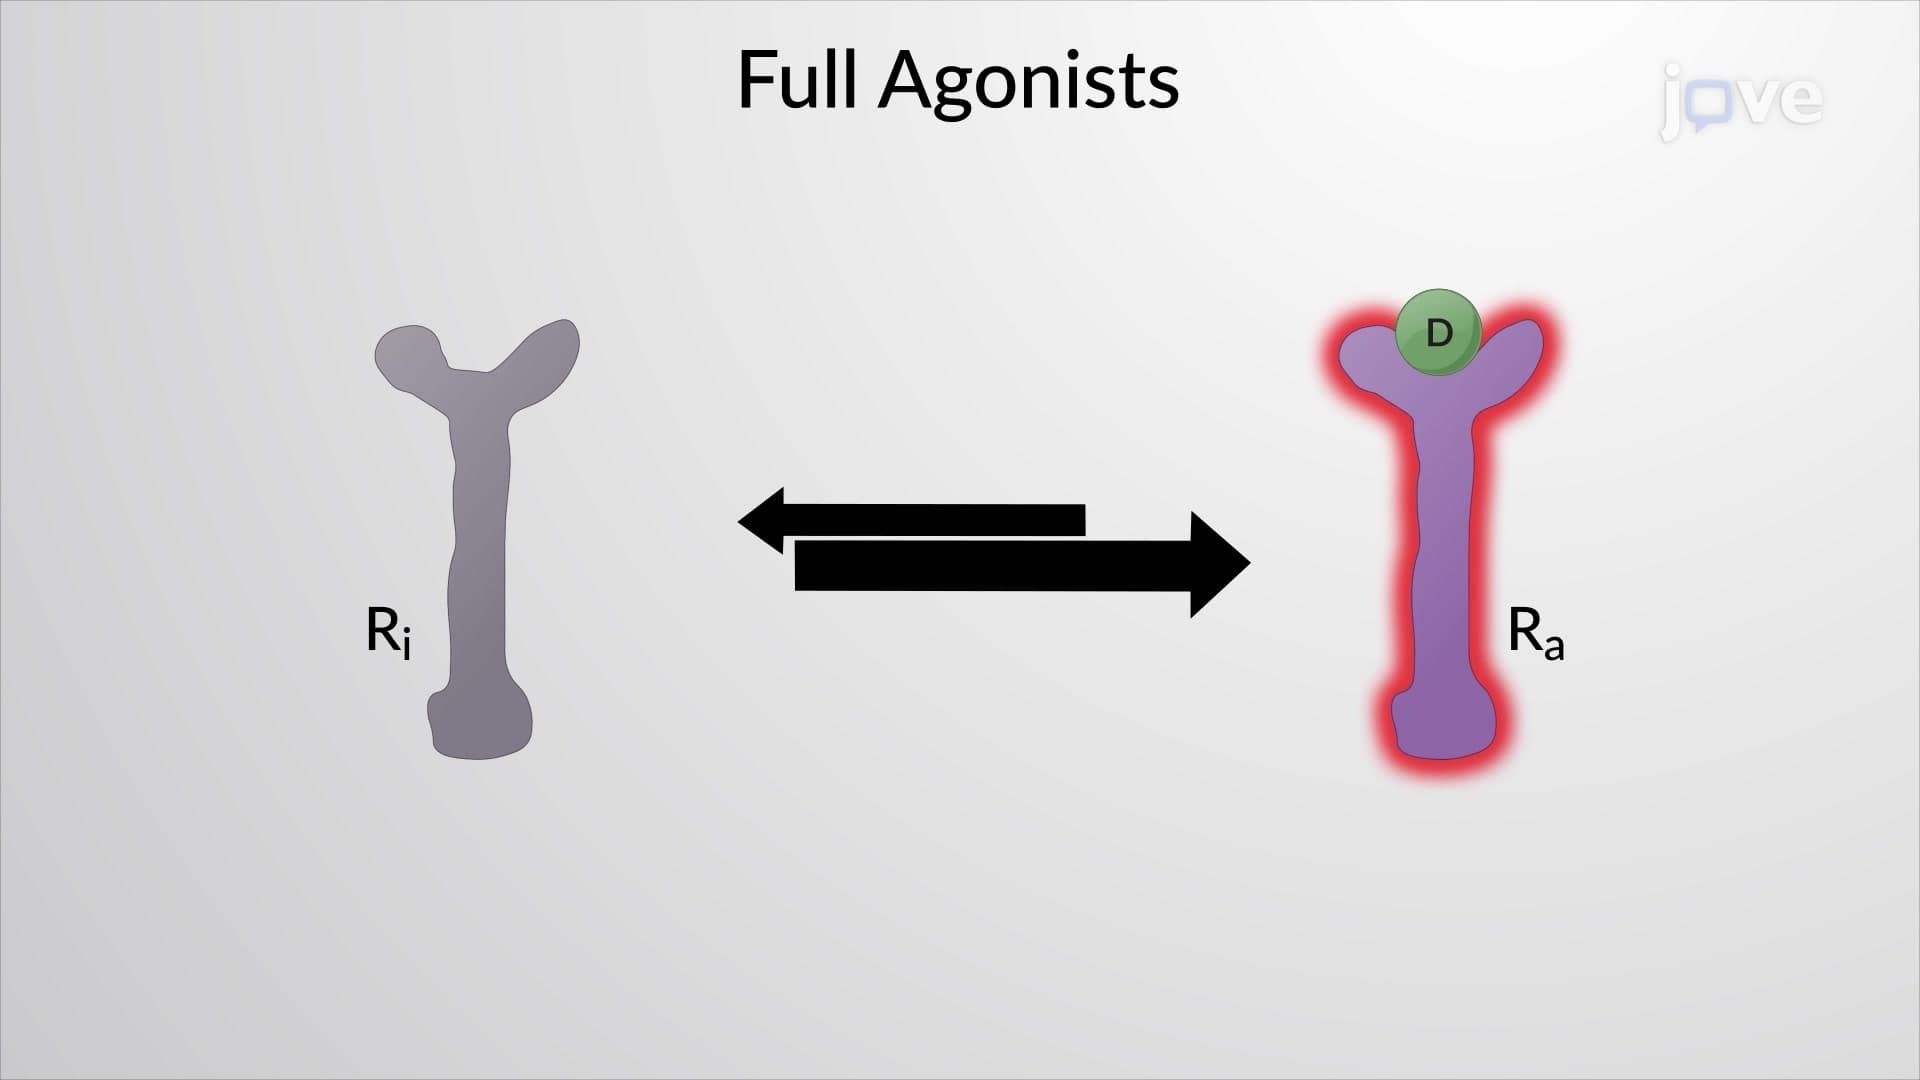
Drug-Receptor Interaction: Agonist
Agonists can bind to receptors in different ways. Some agonists bind directly to the receptor's active site, mimicking the endogenous ligand's action.
 01:12
GPCR Desensitization
 01:30
Quantitative Aspects of Drug-Receptor Interaction