Related Experiment Video
Updated: Jul 19, 2026


Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
W4 theory for computational thermochemistry: In pursuit of confident sub-kJ/mol predictions
Amir Karton1, Elena Rabinovich, Jan M L Martin
1Department of Organic Chemistry, Weizmann Institute of Science, IL-76100 Rehovot, Israel.

Abstract:
In an attempt to improve on our earlier W3 theory [A. D. Boese et al., J. Chem. Phys. 120, 4129 (2004)] we consider such refinements as more accurate estimates for the contribution of connected quadruple excitations (T4), inclusion of connected quintuple excitations (T5), diagonal Born-Oppenheimer corrections (DBOC), and improved basis set extrapolation procedures. Revised experimental data for validation purposes were obtained from the latest version of the Active Thermochemical Tables thermochemical network. The recent CCSDT(Q) method offers a cost-effective way of estimating T4, but is insufficient by itself if the molecule exhibits some nondynamical correlation. The latter considerably slows down basis set convergence for T4, and anomalous basis set convergence in highly polar systems makes two-point extrapolation procedures unusable. However, we found that the CCSDTQ-CCSDT(Q) difference converges quite rapidly with the basis set, and that the formula 1.10[CCSDT(Q)cc-pVTZ+CCSDTQcc-pVDZ-CCSDT(Q)cc-pVDZ] offers a very reliable as well as fairly cost-effective estimate of the basis set limit T4 contribution. The T5 contribution converges very rapidly with the basis set, and even a simple double-zeta basis set appears to be adequate. The largest T5 contribution found in the present work is on the order of 0.5 kcal/mol (for ozone). DBOCs are significant at the 0.1 kcal/mol level in hydride systems. Post-CCSD(T) contributions to the core-valence correlation energy are only significant at that level in systems with severe nondynamical correlation effects. Based on the accumulated experience, a new computational thermochemistry protocol for first- and second-row main-group systems, to be known as W4 theory, is proposed. Its computational cost is not insurmountably higher than that of the earlier W3 theory, while performance is markedly superior. Our W4 atomization energies for a number of key species are in excellent agreement (better than 0.1 kcal/mol on average, 95% confidence intervals narrower than 1 kJ/mol) with the latest experimental data obtained from Active Thermochemical Tables. Lower-cost variants are proposed: the sequence W1-->W2.2-->W3.2-->W4lite-->W4 is proposed as a converging hierarchy of computational thermochemistry methods. A simple a priori estimate for the importance of post-CCSD(T) correlation contributions (and hence a pessimistic estimate for the error in a W2-type calculation) is proposed.


Related Concept Videos
 01:26
Thermodynamic Potentials
 02:55
Thermochemical Equations
 01:24
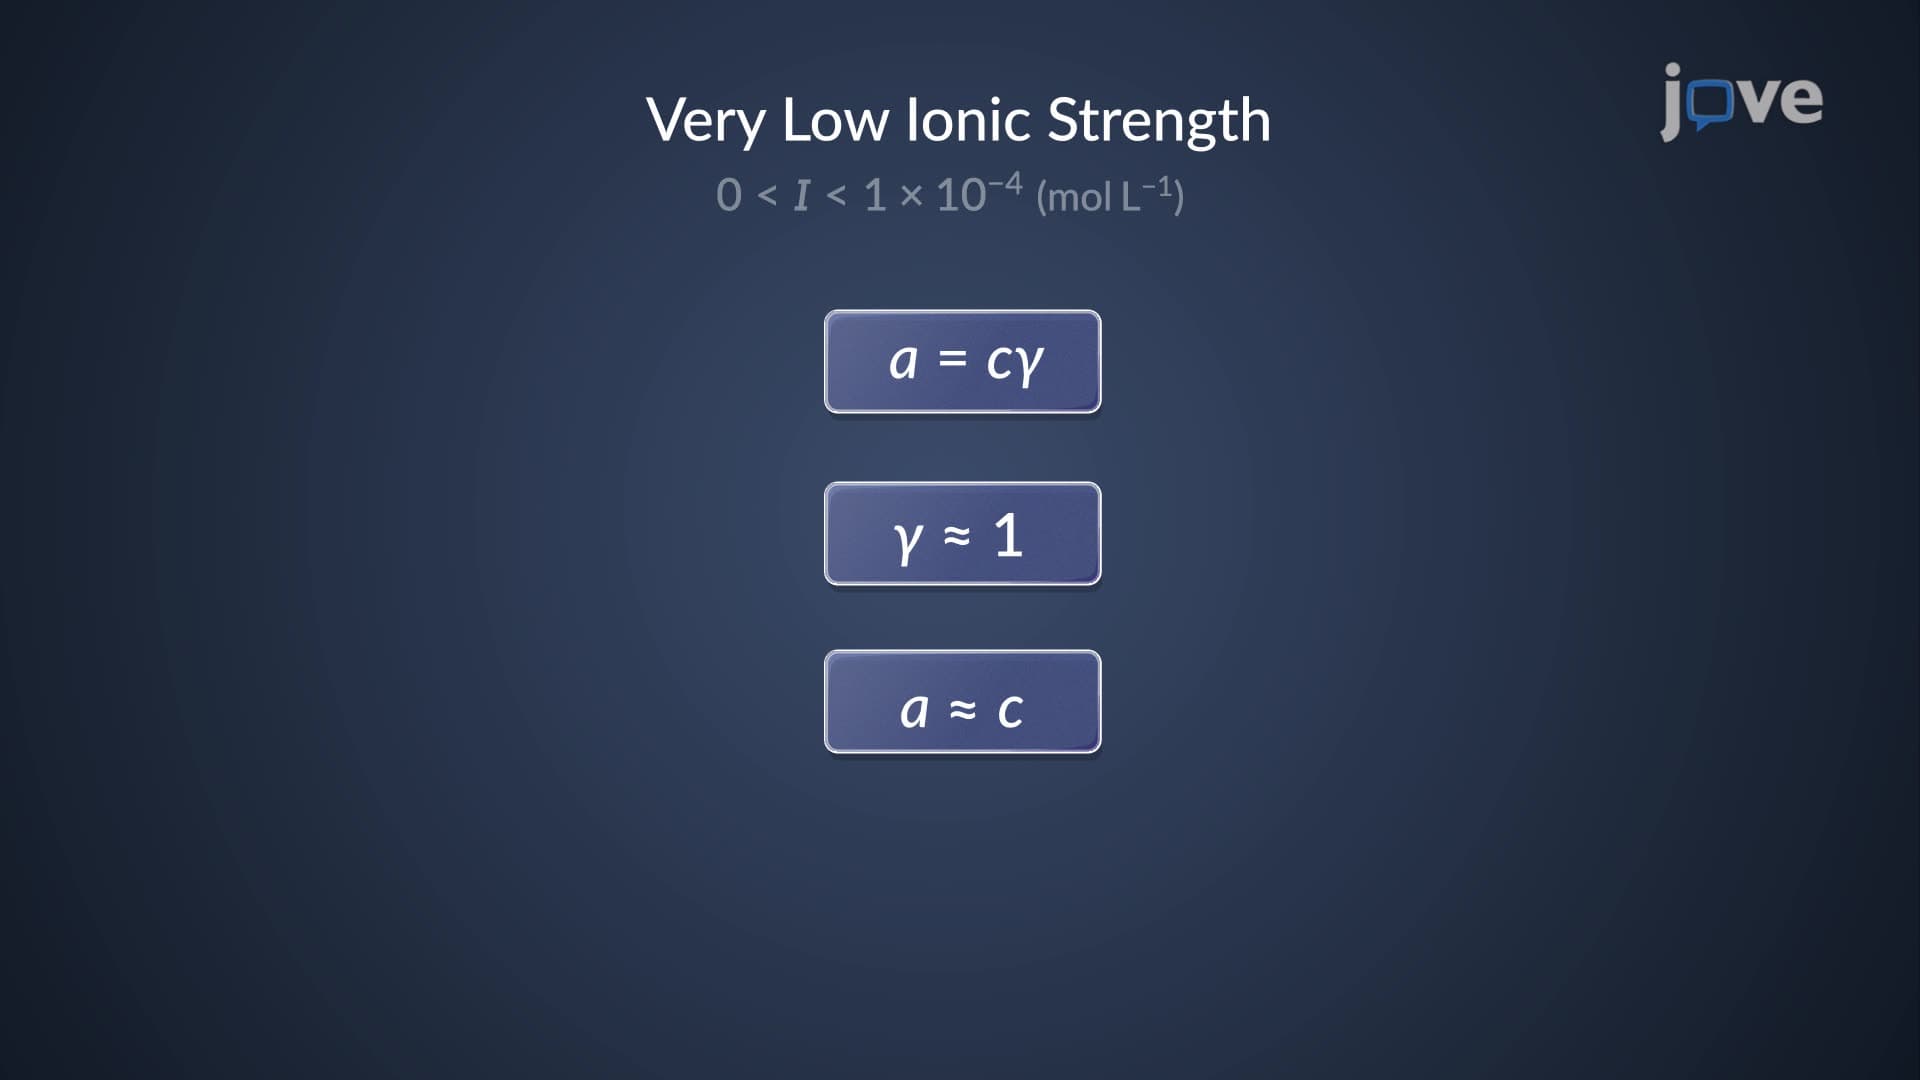
Thermodynamics: Activity Coefficient
The activity coefficient is a measure of the deviation from ideal behavior. When the ionic strength of the solution is minimal, the activity coefficient of an ionic species is close to unity, making...
 01:16
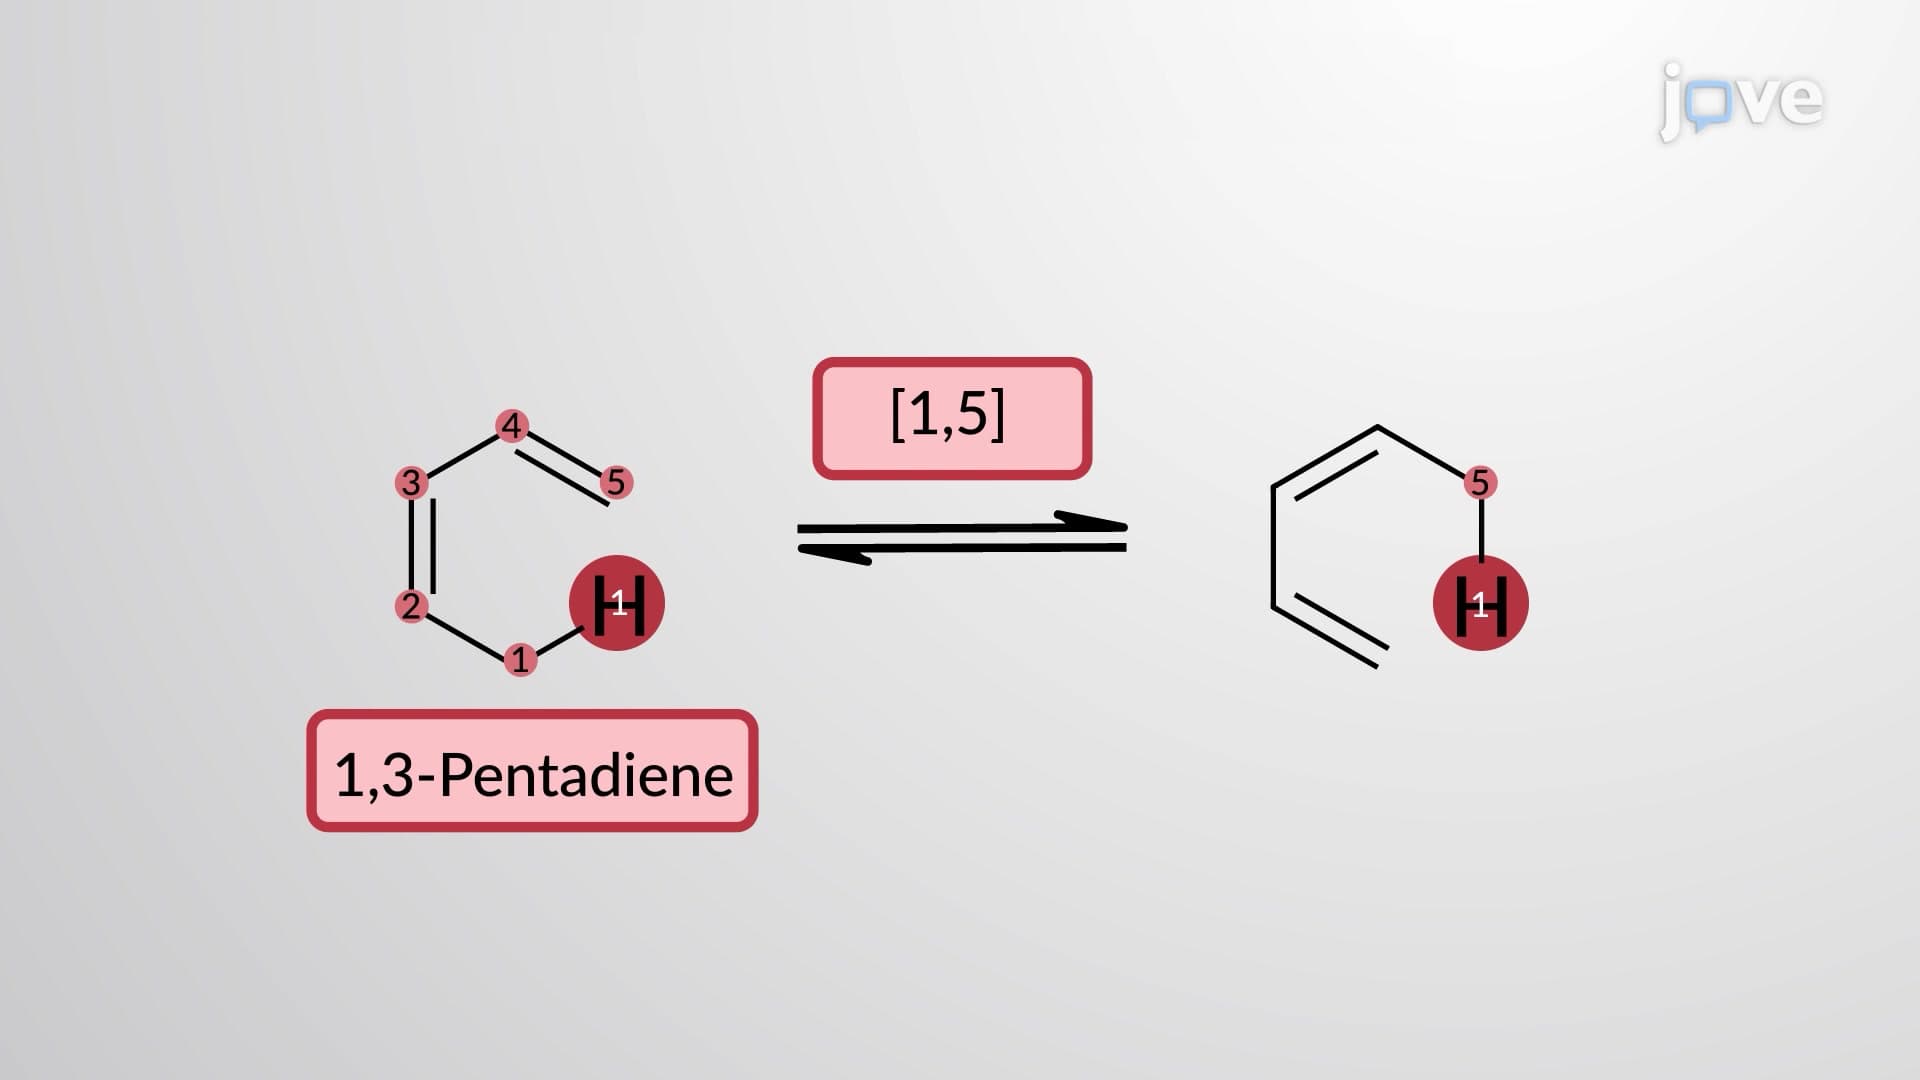
Thermal Sigmatropic Reactions: Overview
Sigmatropic shifts are classified based on an order term [i, j ], where i and j indicate the number of atoms across which each end of the σ bond migrates. Below are examples of a [3,3] sigmatropic shift in 1,5-hexadiene, referred to as...
 01:10
Thermodynamics: Chemical Potential and Activity
The thermodynamic equilibrium constant is more accurately defined in terms of activity rather than concentration.
 02:24
Predicting Reaction Outcomes