Related Experiment Video
Updated: Jul 19, 2026
Line Shape Analysis of Dynamic NMR Spectra for Characterizing Coordination Sphere Rearrangements at a Chiral Rhenium Polyhydride Complex
Published on: July 27, 2022
Characterization by theory of H-transfers and onium reactions of CH3CH2CH2N+H=CH2
Charles E Hudson1, David J McAdoo
1Department of Neuroscience and Cell Biology, University of Texas Medical Branch, Galveston, Texas 77555-1043, USA.

Abstract:
H-transfers by 4-, 5-, and 6-membered ring transition states to the pi-bonded methylene of CH3CH2CH2NH+=CH2 (1) are characterized by theory and compared with the corresponding transfers in cation radicals. Four-membered ring H-transfers converting 1 to CH3CH2CH=N+HCH3 (2) and CH3N+H=CH2 to CH2=NH+CH3 are high-energy processes involving rotation of the source and destination RHC= groups (R = H or C2H5) to near bisection by skeletal planes; migrating hydrogens move near these planes. The H-transfer 1 --> CH3C+HCH2NHCH3 (3) has a higher energy transition-state than 1 --> 2, in marked contrast to the corresponding relative energies of 4- and 5-membered ring H-transfers in cation-radicals. Six-membered ring H-transfer-dissociation (1 --> CH2=CH2 + CH2=N+HCH3) is a closed shell analog of the McLafferty rearrangement. It has a lower energy transition-state than either 1 --> 2 or 1 --> 3, but is still a much higher energy process than 6-membered ring H-transfers in aliphatic cation radicals. In contrast to the stepwise McLafferty rearrangement in cation radicals, H-transfer and CC bond breaking are highly synchronous in 1 --> CH3N+H=CH2 + CH2=CH2. H-transfers in propene elimination from 1 are ion-neutral complex-mediated: 1--> [CH3CH2CH2+ ---NH=CH2] --> [CH3C+HCH3 NH=CH2] --> CH3CH = CH2 + CH2=NH2+. Intrinsic reaction coordinate tracing demonstrated that a slight preference for H-transfer from the methyl containing the carbon from which CH2=NH is cleaved is due to CH2=NH passing nearer this methyl than the other on its way to abstracting H, i.e., some memory of the initial orientation of the partners accompanies this reaction.
More Related Videos
 05:51
Isotopic Effect in Double Proton Transfer Process of Porphycene Investigated by Enhanced QM/MM Method
Published on: July 19, 2019
 06:34
Synthesis of Antiviral Tetrahydrocarbazole Derivatives by Photochemical and Acid-catalyzed C-H Functionalization via Intermediate Peroxides (CHIPS)
Published on: June 20, 2014
Related Concept Videos
 01:11
ortho–para-Directing Activators: –CH3, –OH, –NH2, –OCH3
 01:03
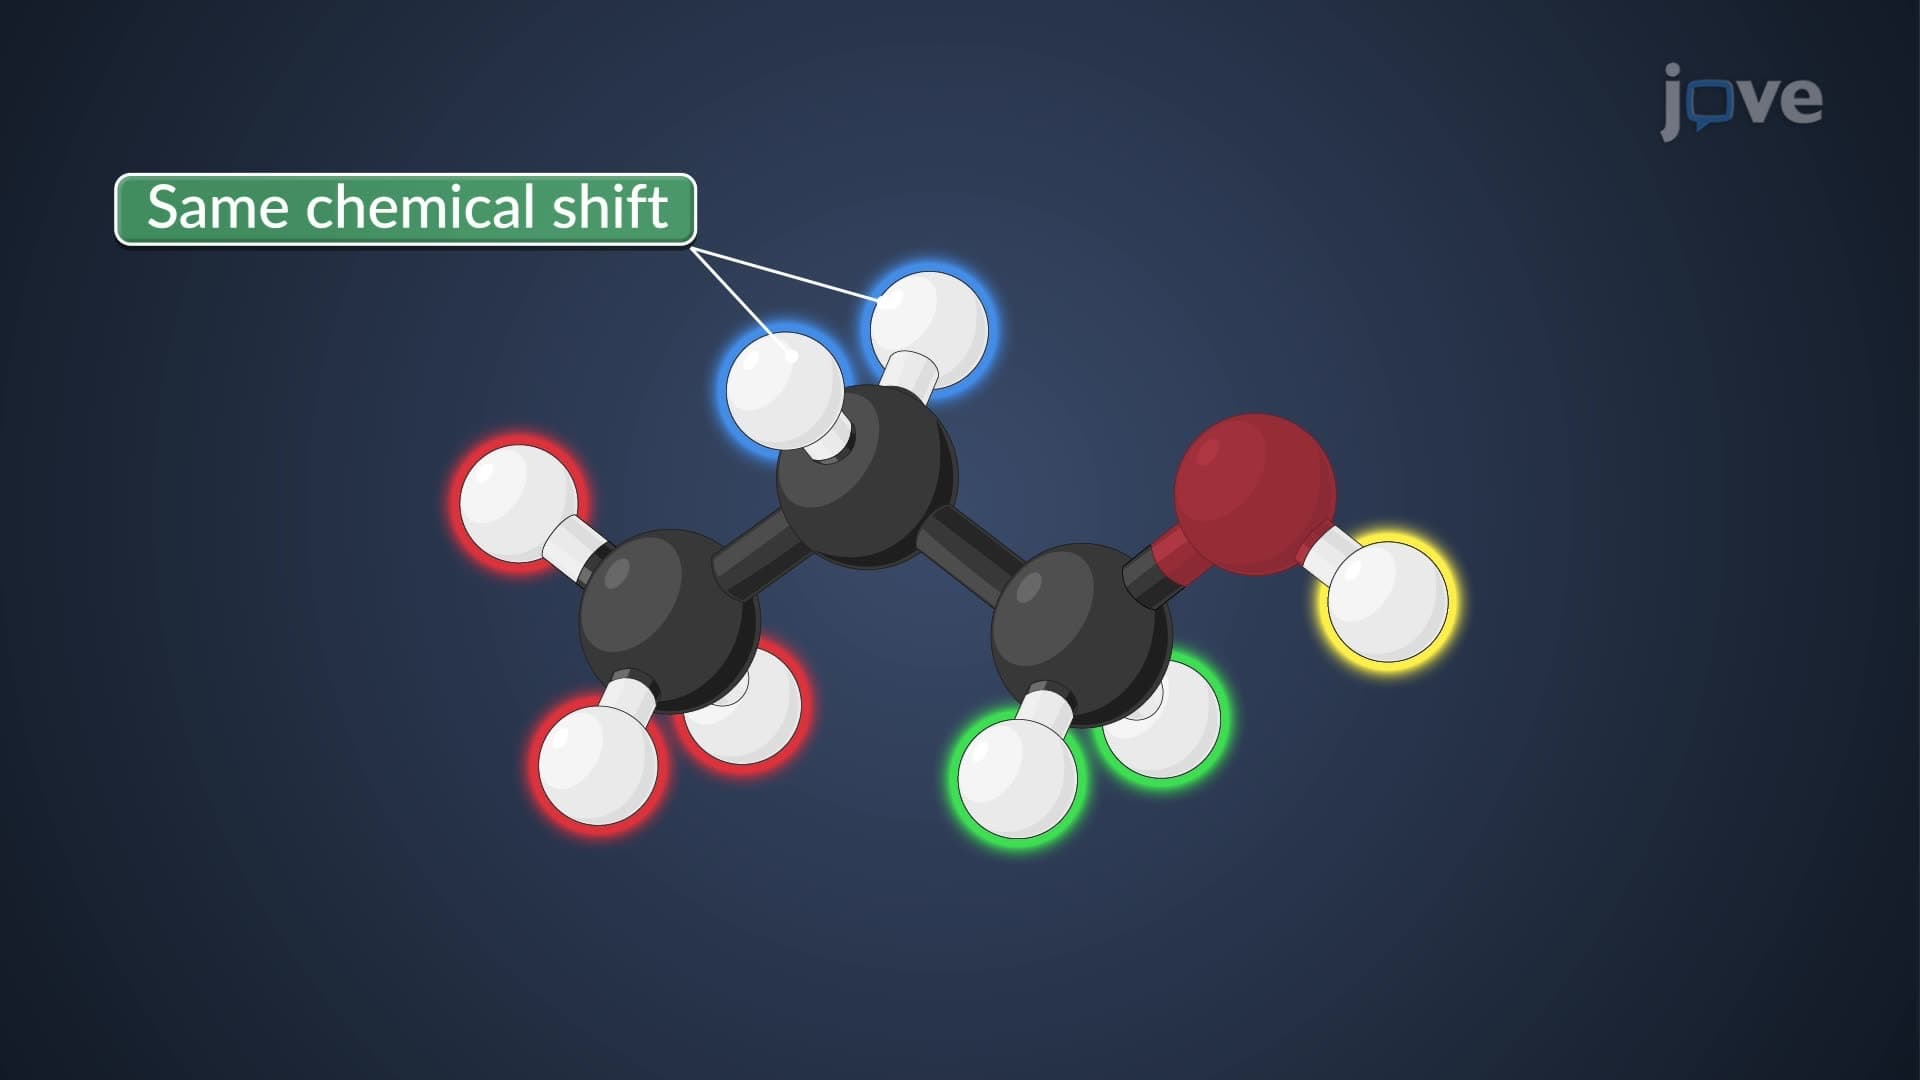
¹H NMR Chemical Shift Equivalence: Homotopic and Heterotopic Protons
 00:58
¹H NMR Chemical Shift Equivalence: Enantiotopic and Diastereotopic Protons
In chiral compounds such as 2-butanol, replacing the methylene hydrogens at C3 produces a pair of...
 02:05
Prochirality
 02:34
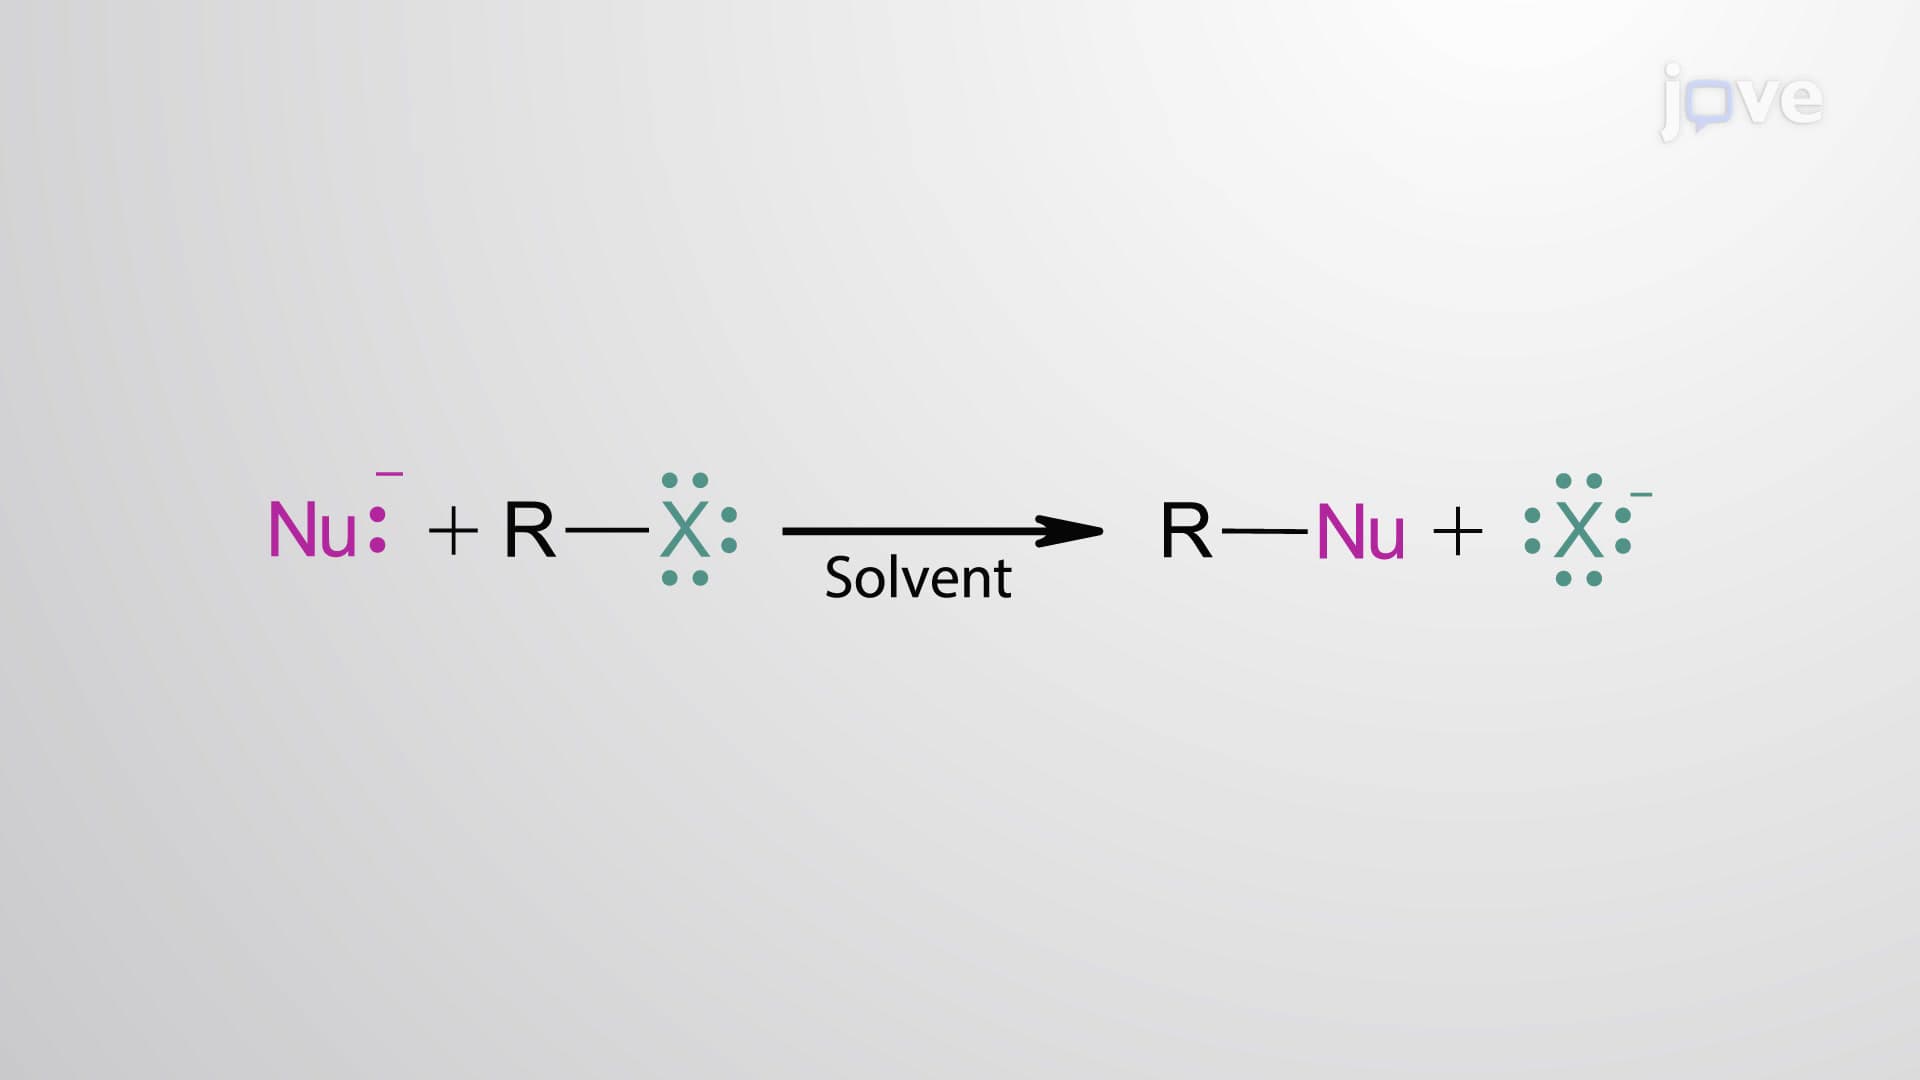
Nucleophilic Substitution Reactions
In 1896, the German chemist Paul Walden discovered that he could interconvert pure enantiomeric (+) and (-) malic acids through a series of reactions. This conversion suggested the involvement of optical inversion during the substitution reaction. Further, in 1930, Sir Christopher Ingold described for the first time two different forms of nucleophilic substitution reactions, which are known as SN1 (nucleophilic substitution unimolecular) and SN2 (nucleophilic substitution...
 02:38
Electrophilic Addition to Alkynes: Halogenation
Halogenation is another class of electrophilic addition reactions where a halogen molecule gets added across a π bond. In alkynes, the presence of two π bonds allows for the addition of two equivalents of halogens (bromine or chlorine). The addition of the first halogen molecule forms a trans-dihaloalkene as the major product and the cis isomer as the minor product. Subsequent addition of the second equivalent yields the tetrahalide.