Related Experiment Video
Updated: Jul 19, 2026
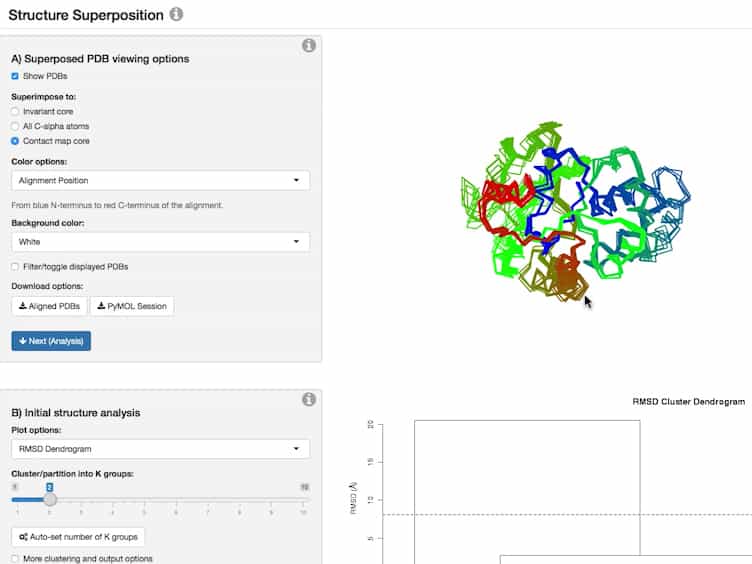

Investigating Protein Sequence-structure-dynamics Relationships with Bio3D-web
Published on: July 16, 2017
Efficient parameterized algorithms for biopolymer structure-sequence alignment
Yinglei Song1, Chunmei Liu, Xiuzhen Huang
1Department of Computer Science, University of Georgia, Athens, GA 30602, USA. song@cs.uga.edu

Abstract:
Computational alignment of a biopolymer sequence (e.g., an RNA or a protein) to a structure is an effective approach to predict and search for the structure of new sequences. To identify the structure of remote homologs, the structure-sequence alignment has to consider not only sequence similarity, but also spatially conserved conformations caused by residue interactions and, consequently, is computationally intractable. It is difficult to cope with the inefficiency without compromising alignment accuracy, especially for structure search in genomes or large databases. This paper introduces a novel method and a parameterized algorithm for structure-sequence alignment. Both the structure and the sequence are represented as graphs, where, in general, the graph for a biopolymer structure has a naturally small tree width. The algorithm constructs an optimal alignment by finding in the sequence graph the maximum valued subgraph isomorphic to the structure graph. It has the computational time complexity O[k(t)N(2)] for the structure of N residues and its tree decomposition of width t. Parameter k, small in nature, is determined by a statistical cutoff for the correspondence between the structure and the sequence. This paper demonstrates a successful application of the algorithm to RNA structure search used for noncoding RNA identification. An application to protein threading is also discussed.


