Ensemble docking of multiple protein structures: considering protein structural variations in molecular docking
Related Experiment Video
Updated: Jul 18, 2026

Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
1Dalton Cardiovascular Research Center and Department of Biochemistry, University of Missouri, Columbia, Missouri 65211, USA.

Abstract:
One approach to incorporate protein flexibility in molecular docking is the use of an ensemble consisting of multiple protein structures. Sequentially docking each ligand into a large number of protein structures is computationally too expensive to allow large-scale database screening. It is challenging to achieve a good balance between docking accuracy and computational efficiency. In this work, we have developed a fast, novel docking algorithm utilizing multiple protein structures, referred to as ensemble docking, to account for protein structural variations. The algorithm can simultaneously dock a ligand into an ensemble of protein structures and automatically select an optimal protein structure that best fits the ligand by optimizing both ligand coordinates and the conformational variable m, where m represents the m-th structure in the protein ensemble. The docking algorithm was validated on 10 protein ensembles containing 105 crystal structures and 87 ligands in terms of binding mode and energy score predictions. A success rate of 93% was obtained with the criterion of root-mean-square deviation <2.5 A if the top five orientations for each ligand were considered, comparable to that of sequential docking in which scores for individual docking are merged into one list by re-ranking, and significantly better than that of single rigid-receptor docking (75% on average). Similar trends were also observed in binding score predictions and enrichment tests of virtual database screening. The ensemble docking algorithm is computationally efficient, with a computational time comparable to that for docking a ligand into a single protein structure. In contrast, the computational time for the sequential docking method increases linearly with the number of protein structures in the ensemble. The algorithm was further evaluated using a more realistic ensemble in which the corresponding bound protein structures of inhibitors were excluded. The results show that ensemble docking successfully predicts the binding modes of the inhibitors, and discriminates the inhibitors from a set of noninhibitors with similar chemical properties. Although multiple experimental structures were used in the present work, our algorithm can be easily applied to multiple protein conformations generated by computational methods, and helps improve the efficiency of other existing multiple protein structure(MPS)-based methods to accommodate protein flexibility.
More Related Videos
 14:34
A Bilingual Computational Workflow for Identifying Potential PLK1 Inhibitors in American Sign Language and English
Published on: April 3, 2026
 05:08
Application of I TASSER, trRosetta, UCSF Chimera, HADDOCK server, and HEX loria for De Novo and In Silico Design of Proteins
Published on: July 8, 2025
Related Concept Videos
 01:24
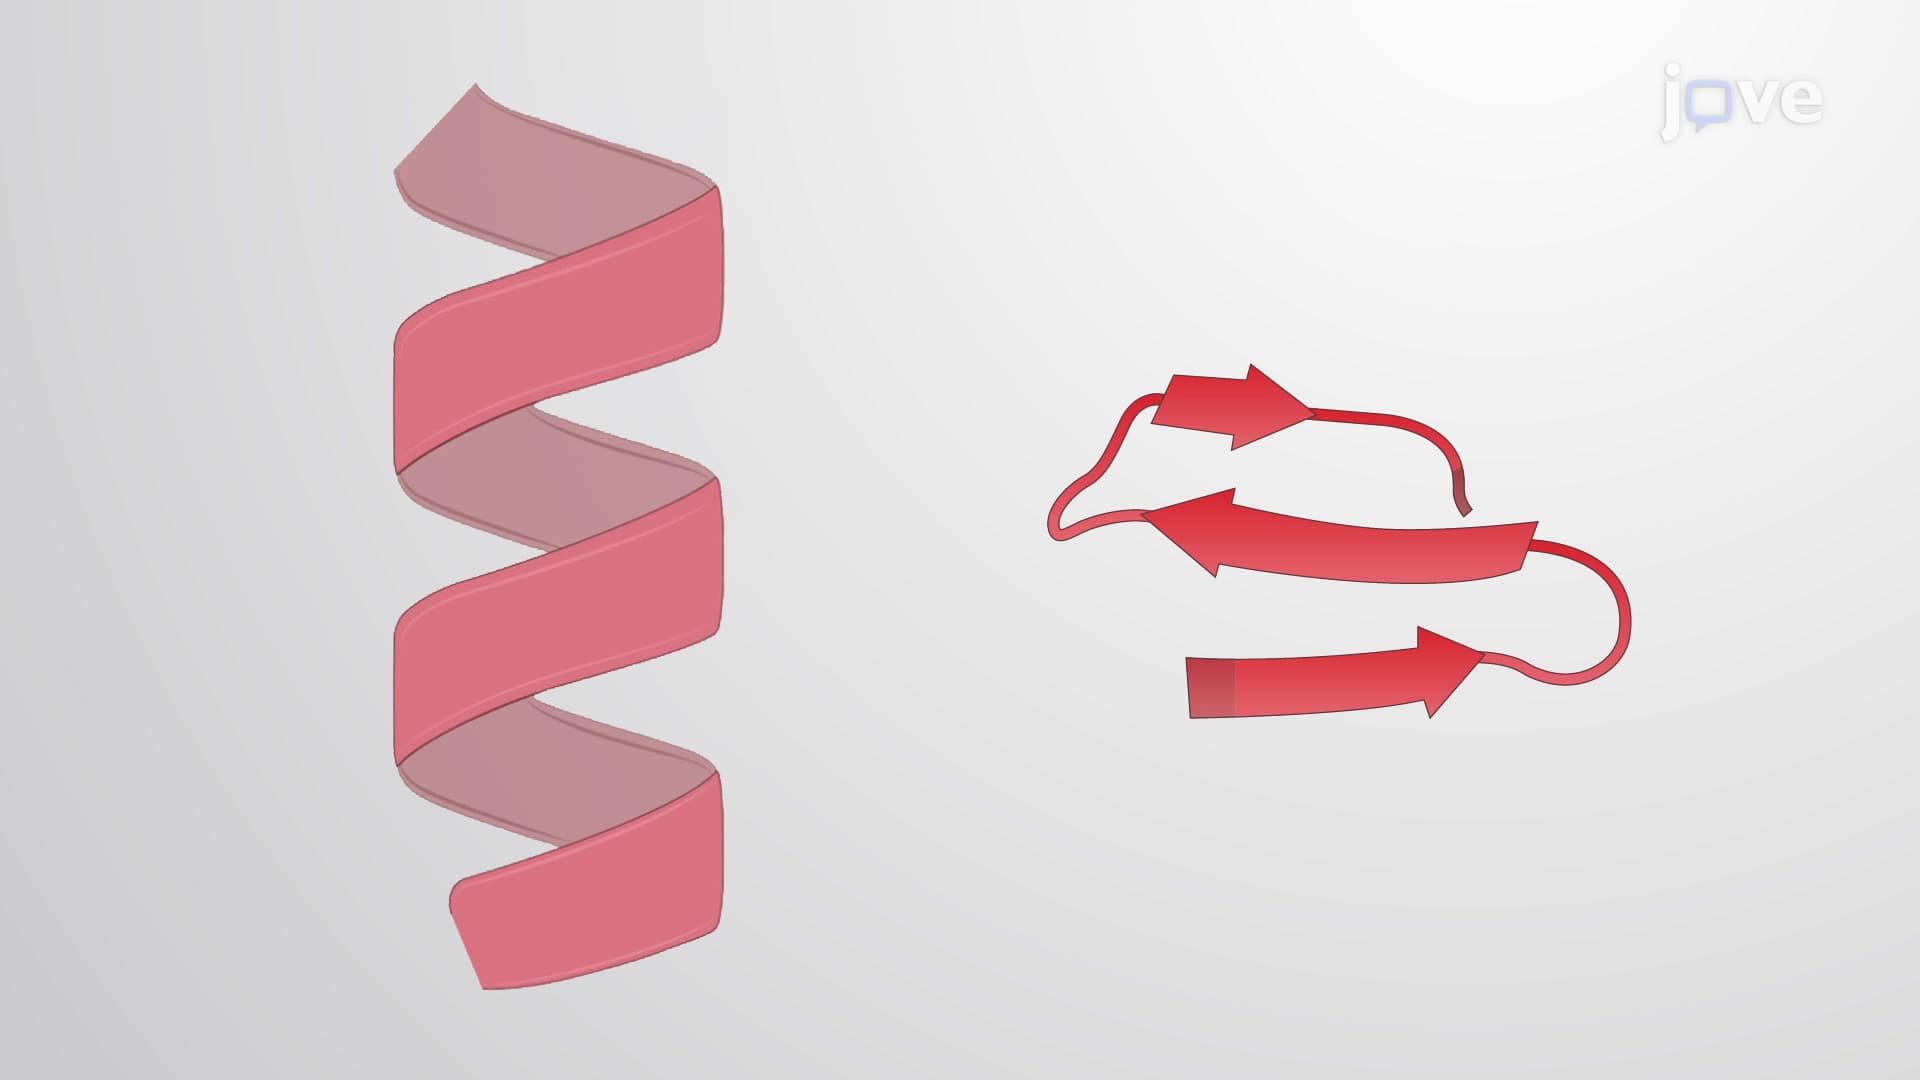
Protein Organization
The primary structure of a protein is its amino acid sequence.
 01:13
Protein Organization
01:24Protein Organization
The primary structure of a protein is its amino acid sequence.
 02:04
Protein-protein Interfaces
02:04Protein-Protein Interfaces
 00:49
Ligand Binding and Linkage