Related Experiment Video
Updated: Jul 18, 2026


Incorporating Target Protein Structure Flexibility and Dynamics in Computational Drug Discovery Using Ensemble-Based Docking Analysis
Published on: June 20, 2025
Synergistic use of compound properties and docking scores in neural network modeling of CYP2D6 binding: predicting
Peter S Bazeley1, Sridevi Prithivi, Craig A Struble
1Department of Mathematics, Statistics, and Computer Science, Marquette University, P.O. Box 1881, Milwaukee, Wisconsin 53233-1881, USA.

Abstract:
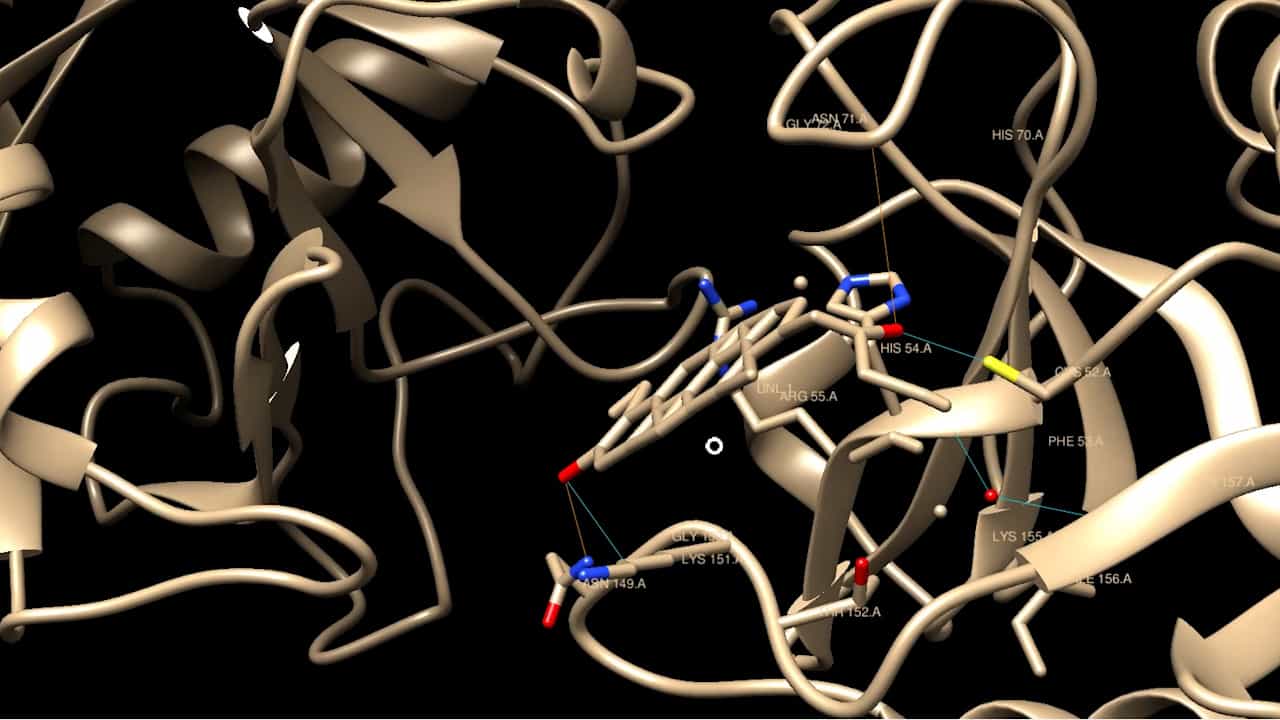
Cytochrome P450 2D6 (CYP2D6) is used to develop an approach for predicting affinity and relevant binding conformation(s) for highly flexible binding sites. The approach combines the use of docking scores and compound properties as attributes in building a neural network (NN) model. It begins by identifying segments of CYP2D6 that are important for binding specificity, based on structural variability among diverse CYP enzymes. A family of distinct, low-energy conformations of CYP2D6 are generated using simulated annealing (SA) and a collection of 82 compounds with known CYP2D6 affinities are docked. Interestingly, docking poses are observed on the backside of the heme as well as in the known active site. Docking scores for the active site binders, along with compound-specific attributes, are used to train a neural network model to properly bin compounds as strong binders, moderate binders, or nonbinders. Attribute selection is used to preselect the most important scores and compound-specific attributes for the model. A prediction accuracy of 85+/-6% is achieved. Dominant attributes include docking scores for three of the 20 conformations in the ensemble as well as the compound's formal charge, number of aromatic rings, and AlogP. Although compound properties were highly predictive attributes (12% improvement over baseline) in the NN-based prediction of CYP2D6 binders, their combined use with docking score attributes is synergistic (net increase of 23% above baseline). Beyond prediction of affinity, attribute selection provides a way to identify the most relevant protein conformation(s), in terms of binding competence. In the case of CYP2D6, three out of the ensemble of 20 SA-generated structures are found to be the most predictive for binding.
Related Concept Videos
 01:49
Conserved Binding Sites
Binding sites are often located in large pockets, and if their location on a protein’s surface is unknown, it can be predicted using various approaches. The energetic method computationally analyses the...
 01:28
Structure-Activity Relationships and Drug Design
SAR studies the intricate relationship between a drug's chemical structure and biological activity. It focuses on understanding how modifications to a drug's structure can influence its...
 02:18
The Equilibrium Binding Constant and Binding Strength
 01:16
Protein-Drug Binding: Mechanism and Kinetics
Various forces drive these interactions, including hydrogen bonds, hydrophobic interactions, ionic bonds, electrostatic interactions, and van der Waals forces. These bonds enable drugs to bind to specific sites on proteins,...
 02:40
Ligand Binding Sites
Protein-ligand interactions are quite specific; even though numerous potential ligands surround a cellular protein at any given time, only a particular ligand can bind to that protein. Moreover, a ligand binds only to a dedicated area on the surface of the protein, known as the...
 01:22
Protein-Drug Binding: Determination Methods
Indirect methods involve isolating the bound drug from its free form in biological samples such as blood, serum, or plasma. These techniques aim to measure the percentage of drugs bound to proteins. Equilibrium dialysis is a commonly used method where the free drug concentration at equilibrium is measured by separating the bound...