Related Experiment Video
Updated: Jul 18, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
Quantum dynamics of the H + O2 --> O + OH reaction on an accurate ab initio potential energy surface

Abstract:
We report exact time-dependent and time-independent quantum mechanical studies of the title reaction on an accurate ab initio potential energy surface of Xu et al. (J. Chem. Phys. 2005, 122, 24305). The J = 0 reaction probabilities for several reactant states show sharp resonance structures superimposed on relatively low backgrounds, and they are remarkably different from existing quantum results on an earlier potential energy surface (DMBE-IV). The new findings reported here suggest that our current understanding of this important reaction might require significant revision.


Related Concept Videos
 01:26
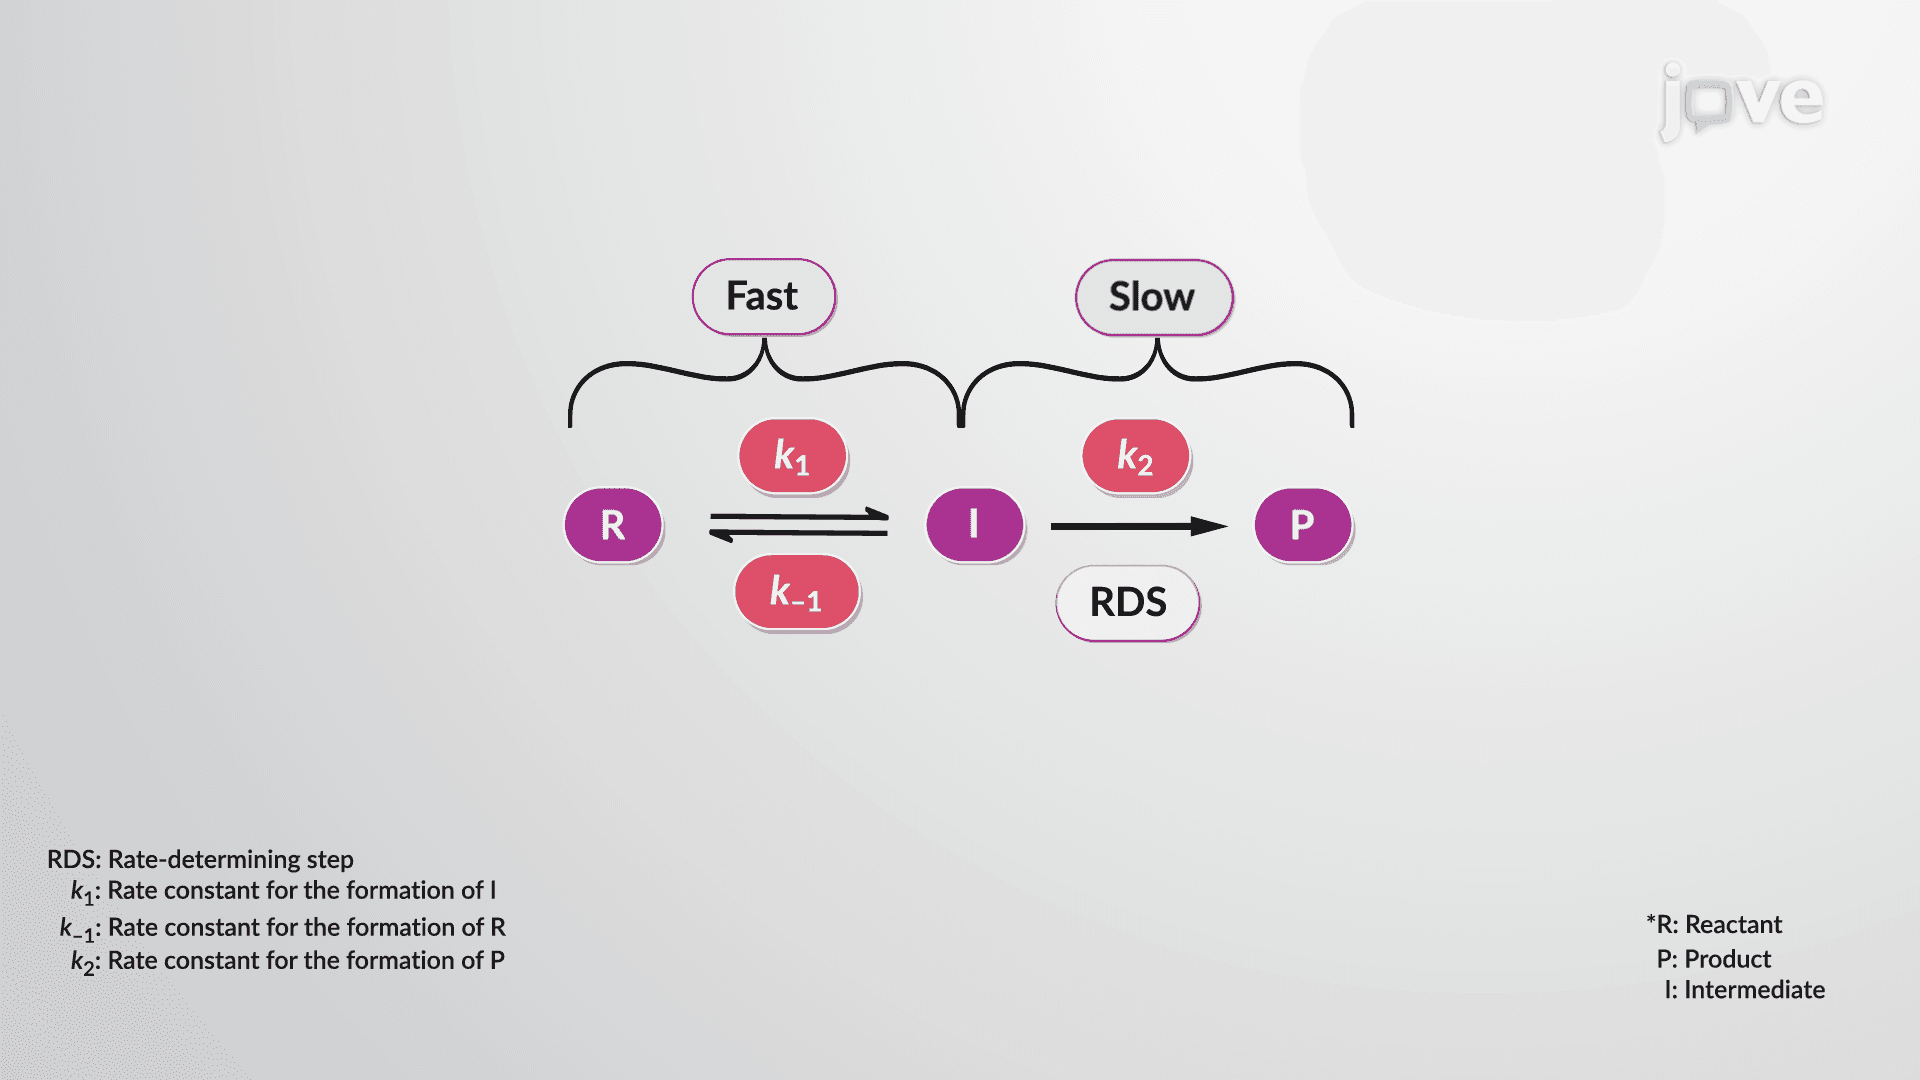
Reaction Mechanisms: The Steady-State Approximation
 02:13
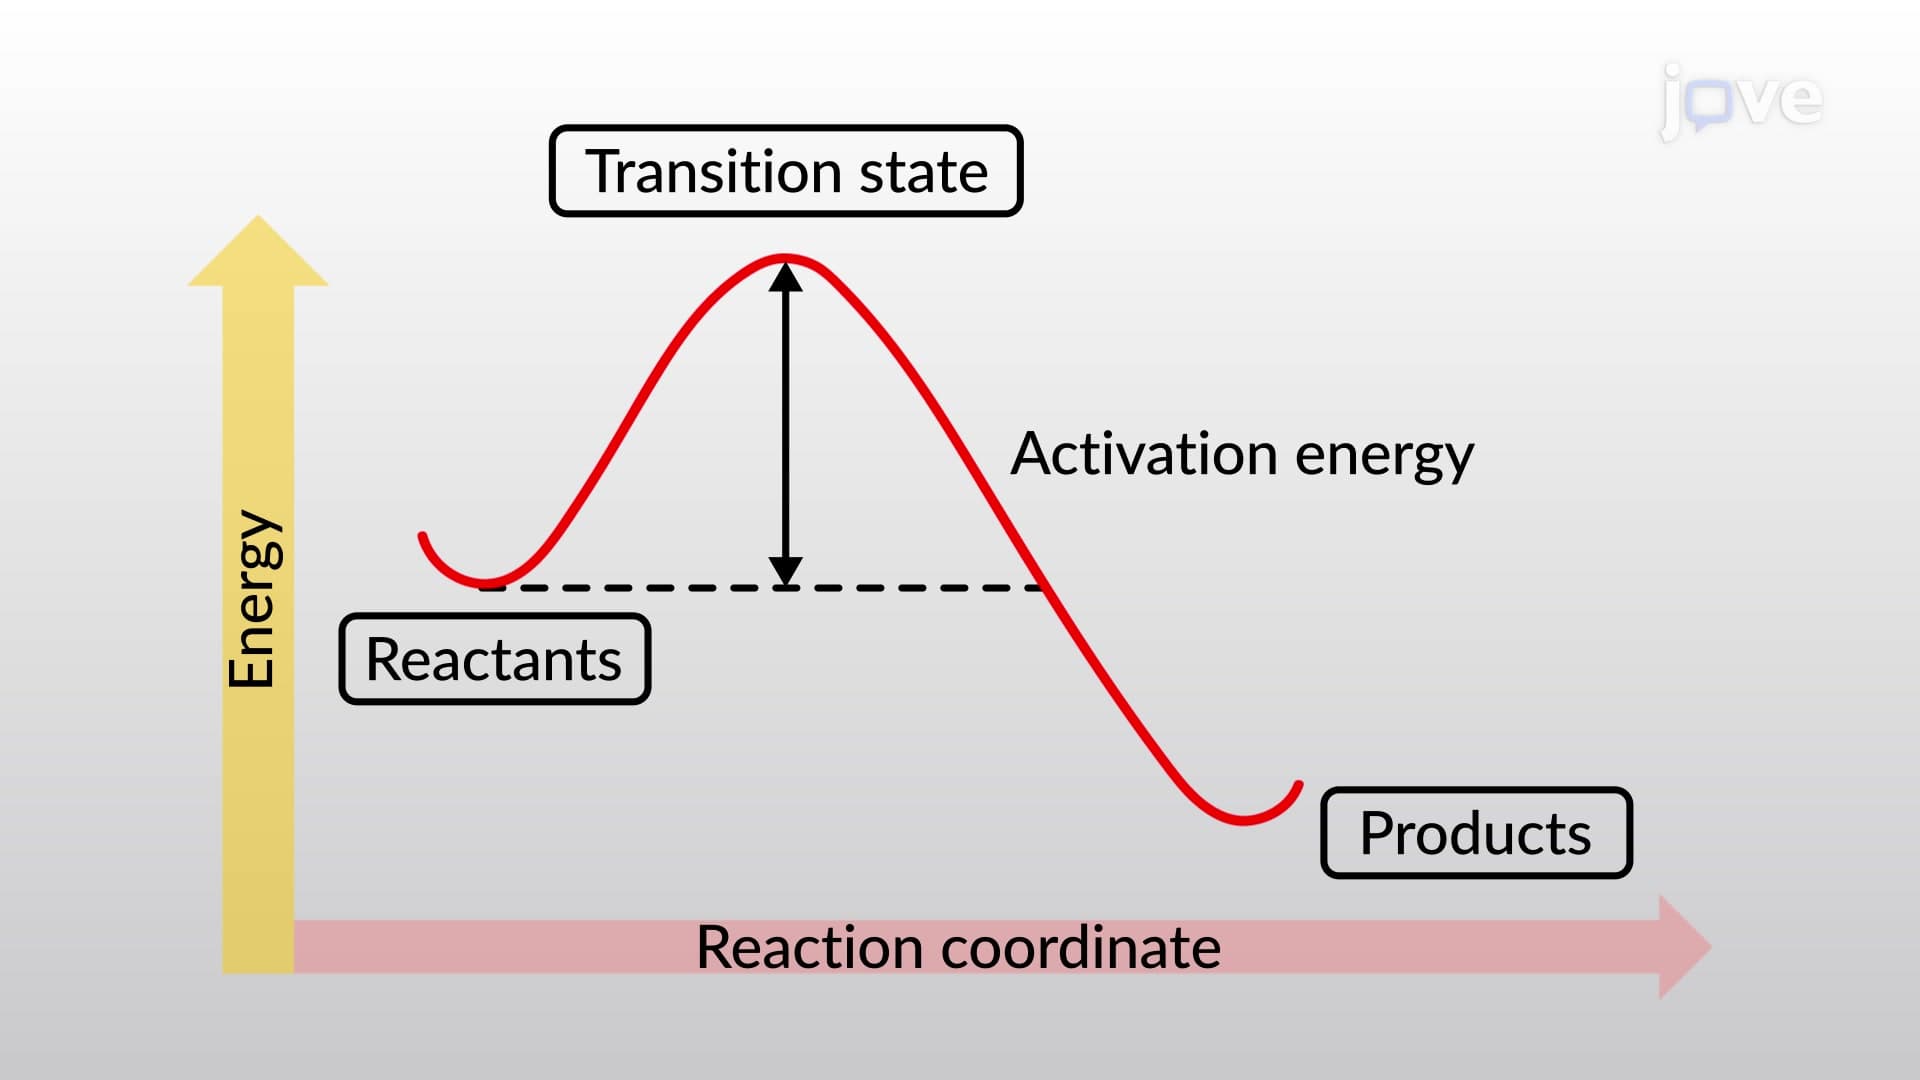
Bond Dissociation Energy and Activation Energy
 03:35
Hybridization of Atomic Orbitals II
 03:51
Molecular Orbital Theory II
 03:24
Hybridization of Atomic Orbitals I
 02:35
Molecular Orbital Theory I