Related Experiment Video
Updated: Jul 18, 2026


Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
A rigorous full-dimensional quantum dynamics calculation of the vibrational energies of H3O-2
1Department of Chemistry, Brookhaven National Laboratory, Upton, NY 11973-5000, USA. hgy@bnl.gov

Abstract:
The vibrational energy levels of the H(3)O-(2) anion have been calculated using a rigorous quantum dynamics method based on an accurate ab initio potential energy surface. The eigenvalue problem is solved using the two-layer Lanczos iterative diagonalization algorithm in a mixed grid/nondirect product basis set, where the system Hamiltonian is expressed in a set of orthogonal polyspherical coordinates. The lowest 312 vibrational energy levels in each inversion symmetry, together with a comparison of fundamental frequencies with previous quantum dynamics calculations, are reported. Finally, a statistical analysis of nearest level spacing distribution is carried out, revealing a strongly chaotic nature.
More Related Videos
 12:11
Computation of Atmospheric Concentrations of Molecular Clusters from ab initio Thermochemistry
Published on: April 8, 2020
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
Related Concept Videos
 01:16
IR Spectroscopy: Hooke's Law Approximation of Molecular Vibration
According to Hooke's law, the vibrational frequency is directly proportional to the...
 01:24
π Molecular Orbitals of 1,3-Butadiene
The simplest conjugated diene is 1,3-butadiene: a four-carbon system where each carbon is sp2-hybridized and has an unhybridized p orbital that contains an unpaired electron. According to molecular orbital theory, atomic orbitals combine to form molecular orbitals such that the number...
 03:51
Molecular Orbital Theory II
 02:11
Conformations of Cyclohexane
The chair form is the most stable and derives its name from its resemblance to the “easy chair.” In the chair conformation, two carbon atoms are arranged out-of-plane — one above and one below, minimizing the torsional strain. In the chair form, the bond angle is very close to the ideal tetrahedral value,...
 02:46
Crystal Field Theory - Tetrahedral and Square Planar Complexes
Crystal field theory (CFT) is applicable to molecules in geometries other than octahedral. In octahedral complexes, the lobes of the dx2−y2 and dz2 orbitals point directly at the ligands. For tetrahedral complexes, the d orbitals remain in place, but with only four ligands located between the axes. None of the orbitals points directly at the tetrahedral ligands. However, the dx2−y2 and dz2 orbitals (along the Cartesian axes) overlap with the ligands less than the dxy,...
 01:10
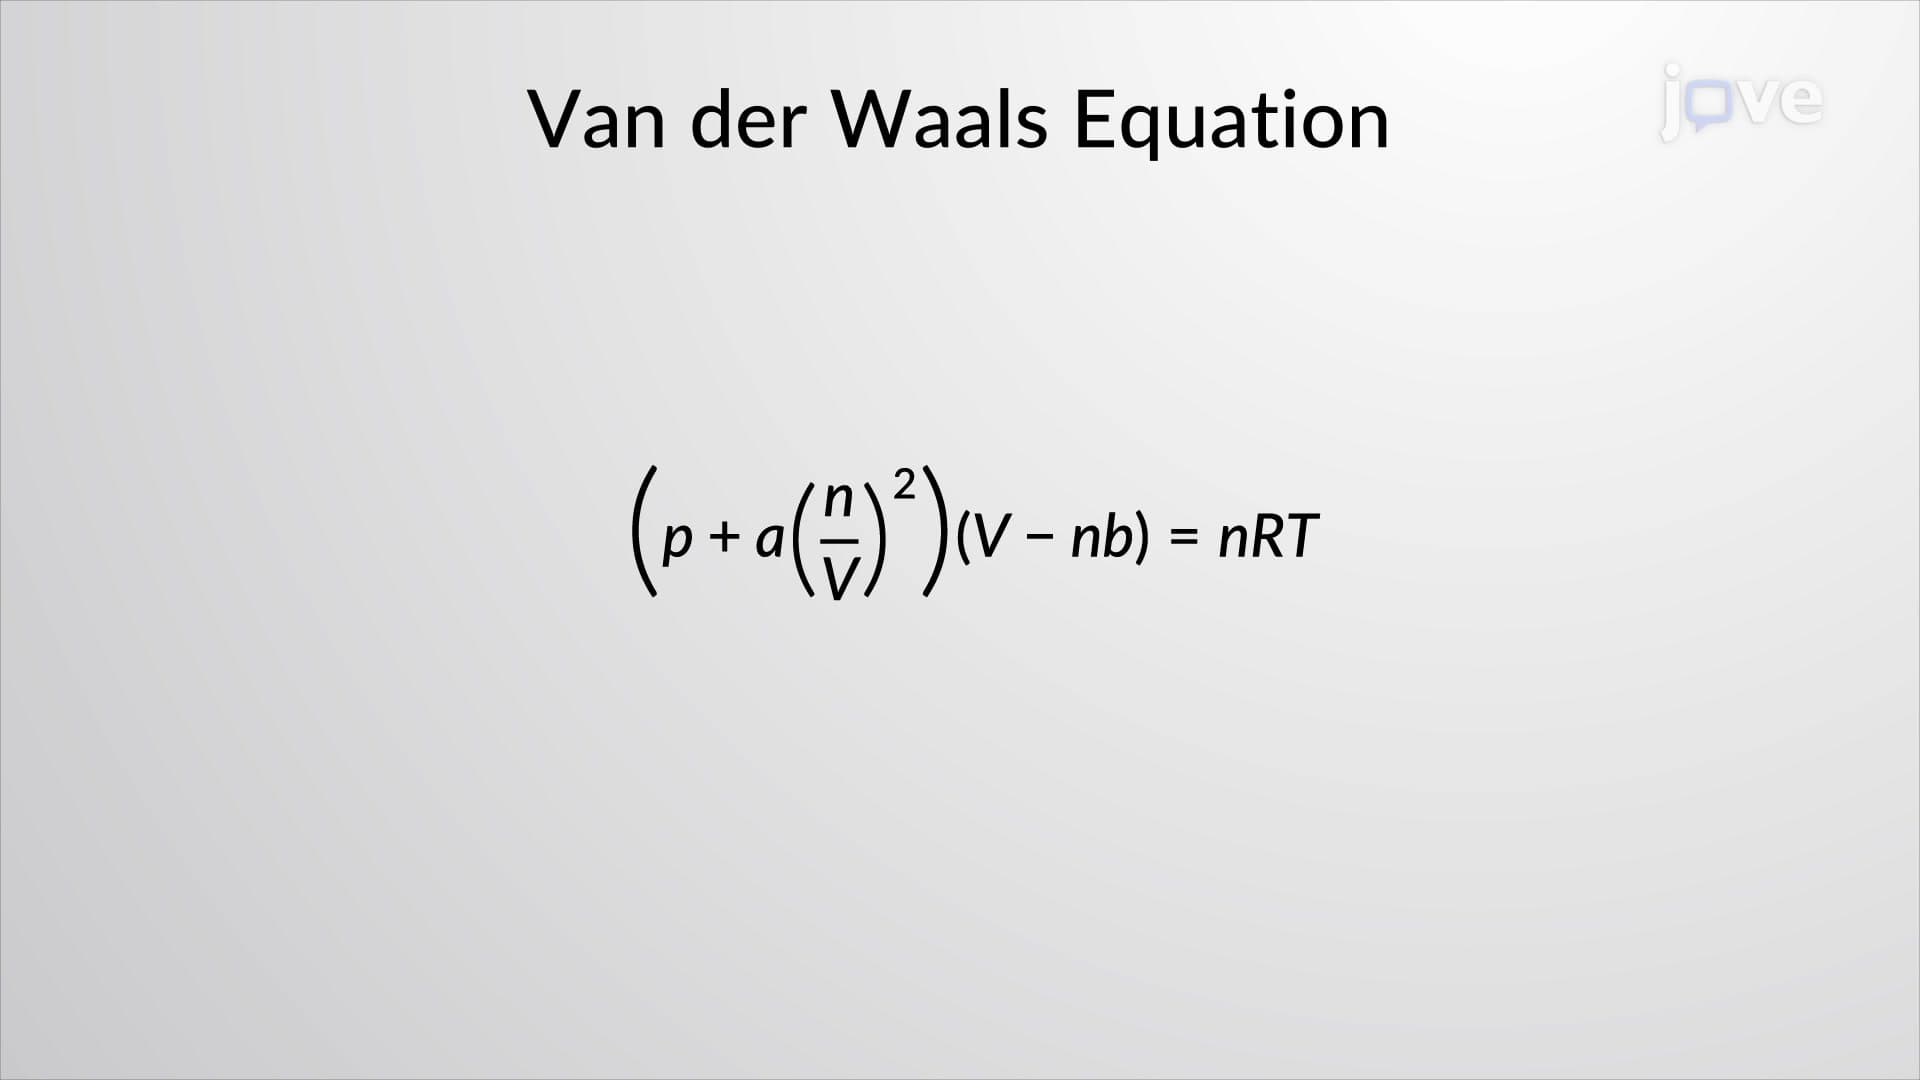
Van der Waals Equation
First, the attractive forces between molecules, which are stronger at higher densities and reduce the pressure, are considered by adding to the pressure a term equal to the square of the molar density multiplied by a positive coefficient a. Second, the volume...