Related Experiment Video
Updated: Jul 18, 2026
Imine Metathesis by Silica-Supported Catalysts Using the Methodology of Surface Organometallic Chemistry
Published on: October 18, 2019
An ab initio and DFT study of structure and vibrational spectra of disiloxane H3SiOSiH3 conformers. Comparison to
C Carteret1, A Labrosse, X Assfeld
1Laboratoire de Chimie Physique et Microbiologie pour l'Environnement, Equipe Chimie et Spectrochimie des Interfaces, UMR 7564 CNRS-Université Henri Poincaré, 405 rue de Vandoeuvre, 54600 Villers-lès-Nancy, France. cedric.carteret@lcpme.cnrs-nancy.fr

Abstract:
The structural and vibrational properties of siloxane monomers may account in the physical and chemical properties of silicone polymers. Because disiloxane (H(3)SiOSiH(3)) is the smallest molecule in the set which runs through small siloxanes like hexamethyldisiloxane (CH(3))3SiOSi(CH(3))3 to silicone polymers, its energetic, structural and vibrational features have been investigated in detail using density functional theory (B3LYP), post Hartree-Fock methods (MP2 and CCSD(T)) and basis sets up to spdfg quality. Five conformations were considered: three bent structures with C2v (double staggered, SS, and double eclipsed, EE, conformations) and Cs symmetries, and two linear forms with D3d and D3h symmetries. At all levels of theory, the relative stability was C2v(SS) approximately C2v(EE)>Cs>D3h>D3d. The difference of energy between the two C2v conformers is lower than 0.04 kcal/mol. At the highest level of theory (CCSD(T)/cc-pVQZ), the barrier to linearisation from C(2v) to D(3h) conformers was calculated at 0.43 kcal/mol, which is extremely low. Most of the structural and vibrational features of the disiloxane do not depend on the conformation of the molecule but are strongly influenced by the SiOSi angle. Anharmonic calculations allowed, without any scaling factor, an exhaustive reinvestigation of the assignments of observed wavenumbers in the infrared and Raman spectra of gaseous disiloxane. Particularly, in the gas phase spectrum, the SiOSi symmetric and antisymmetric stretches have been assigned at 599 and 1105, 596 and 1060, 527 and 1093 cm(-1) for H(3)SiOSiH(3), H(3)Si(18)OSiH(3) and D(3)SiOSiD(3), respectively. The experimental wavenumber splitting of SiOSi symmetric and antisymmetric stretches of H(3)SiOSiH(3) gave an estimation of the SiOSi angle at around 145 degrees . Ab initio methods were revealed more accurate for structural parameters, when DFT/B3LYP was enough for spectral assignments, even at the harmonic level using a single scaling factor.
More Related Videos
 08:54
Vibrational Spectra of a N719-Chromophore/Titania Interface from Empirical-Potential Molecular-Dynamics Simulation, Solvated by a Room Temperature Ionic Liquid
Published on: January 25, 2020
 06:37
Analyzing Melts and Fluids from Ab Initio Molecular Dynamics Simulations with the UMD Package
Published on: September 17, 2021
Related Concept Videos
 01:28
Stability of Conjugated Dienes
A comparison of the enthalpies of hydrogenation of dienes reveals that conjugated dienes release less heat on hydrogenation, rendering them more stable than their nonconjugated analogs.
 02:52
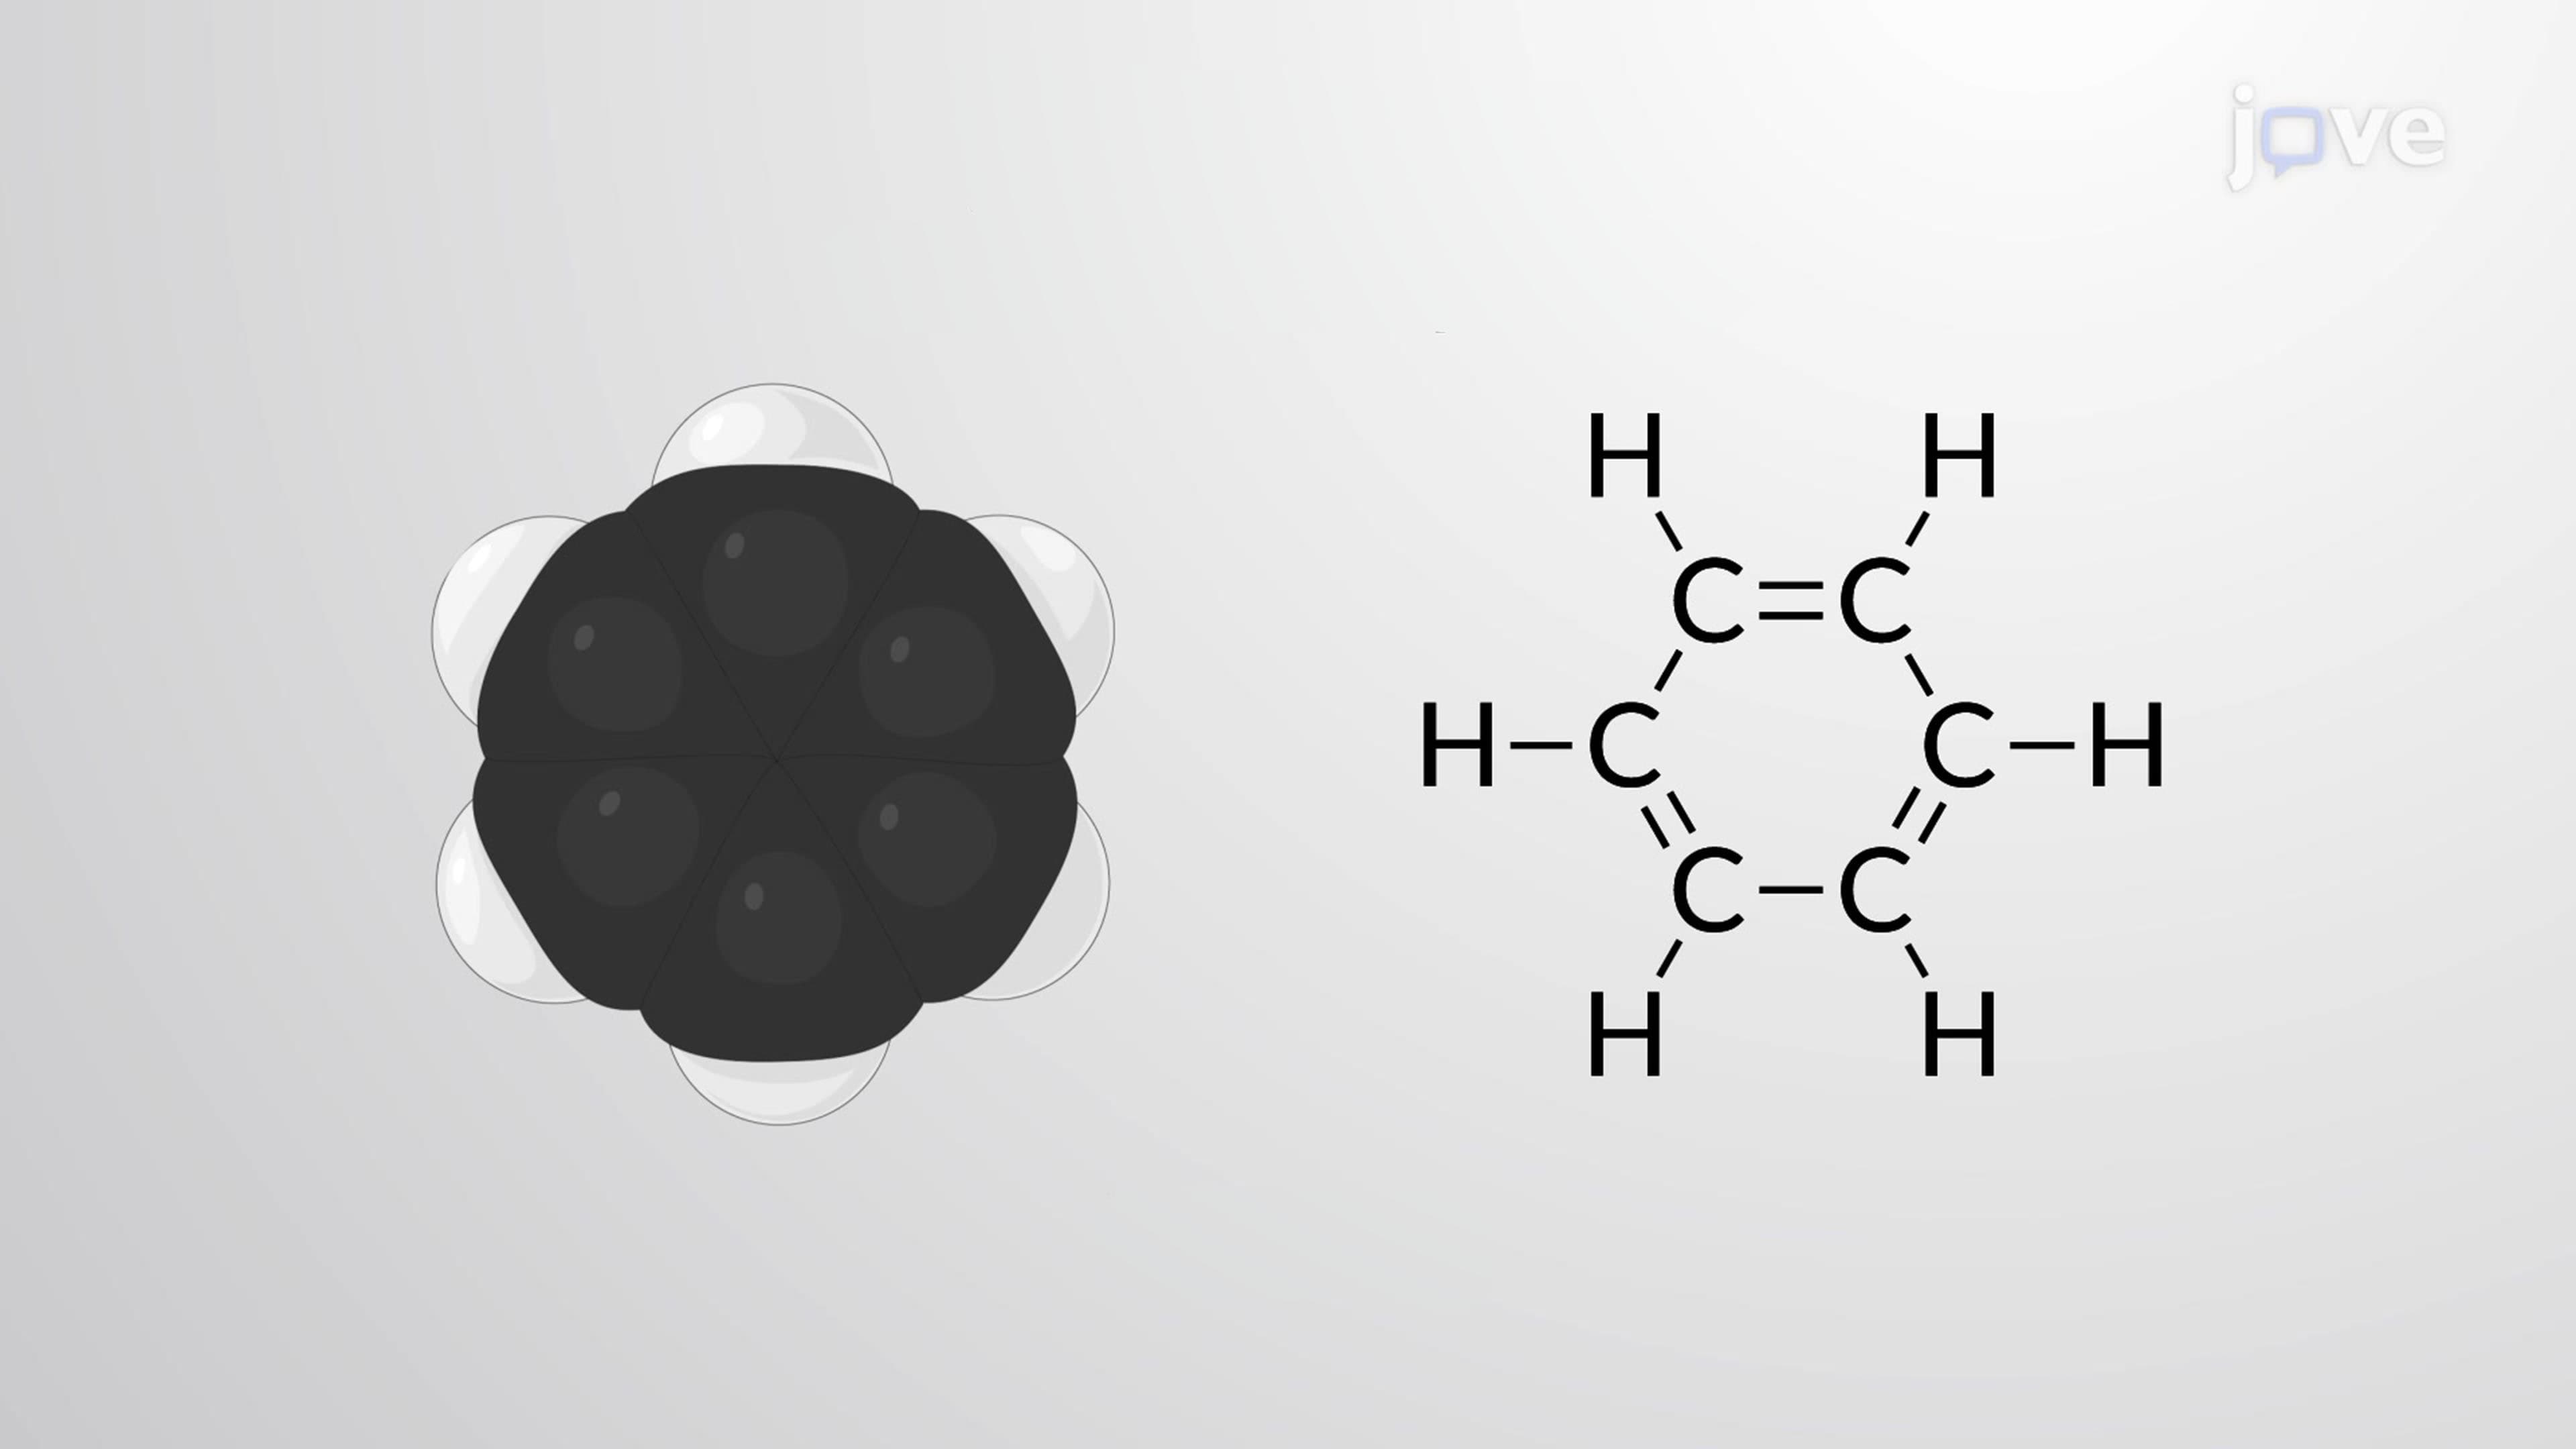
Resonance
 02:54
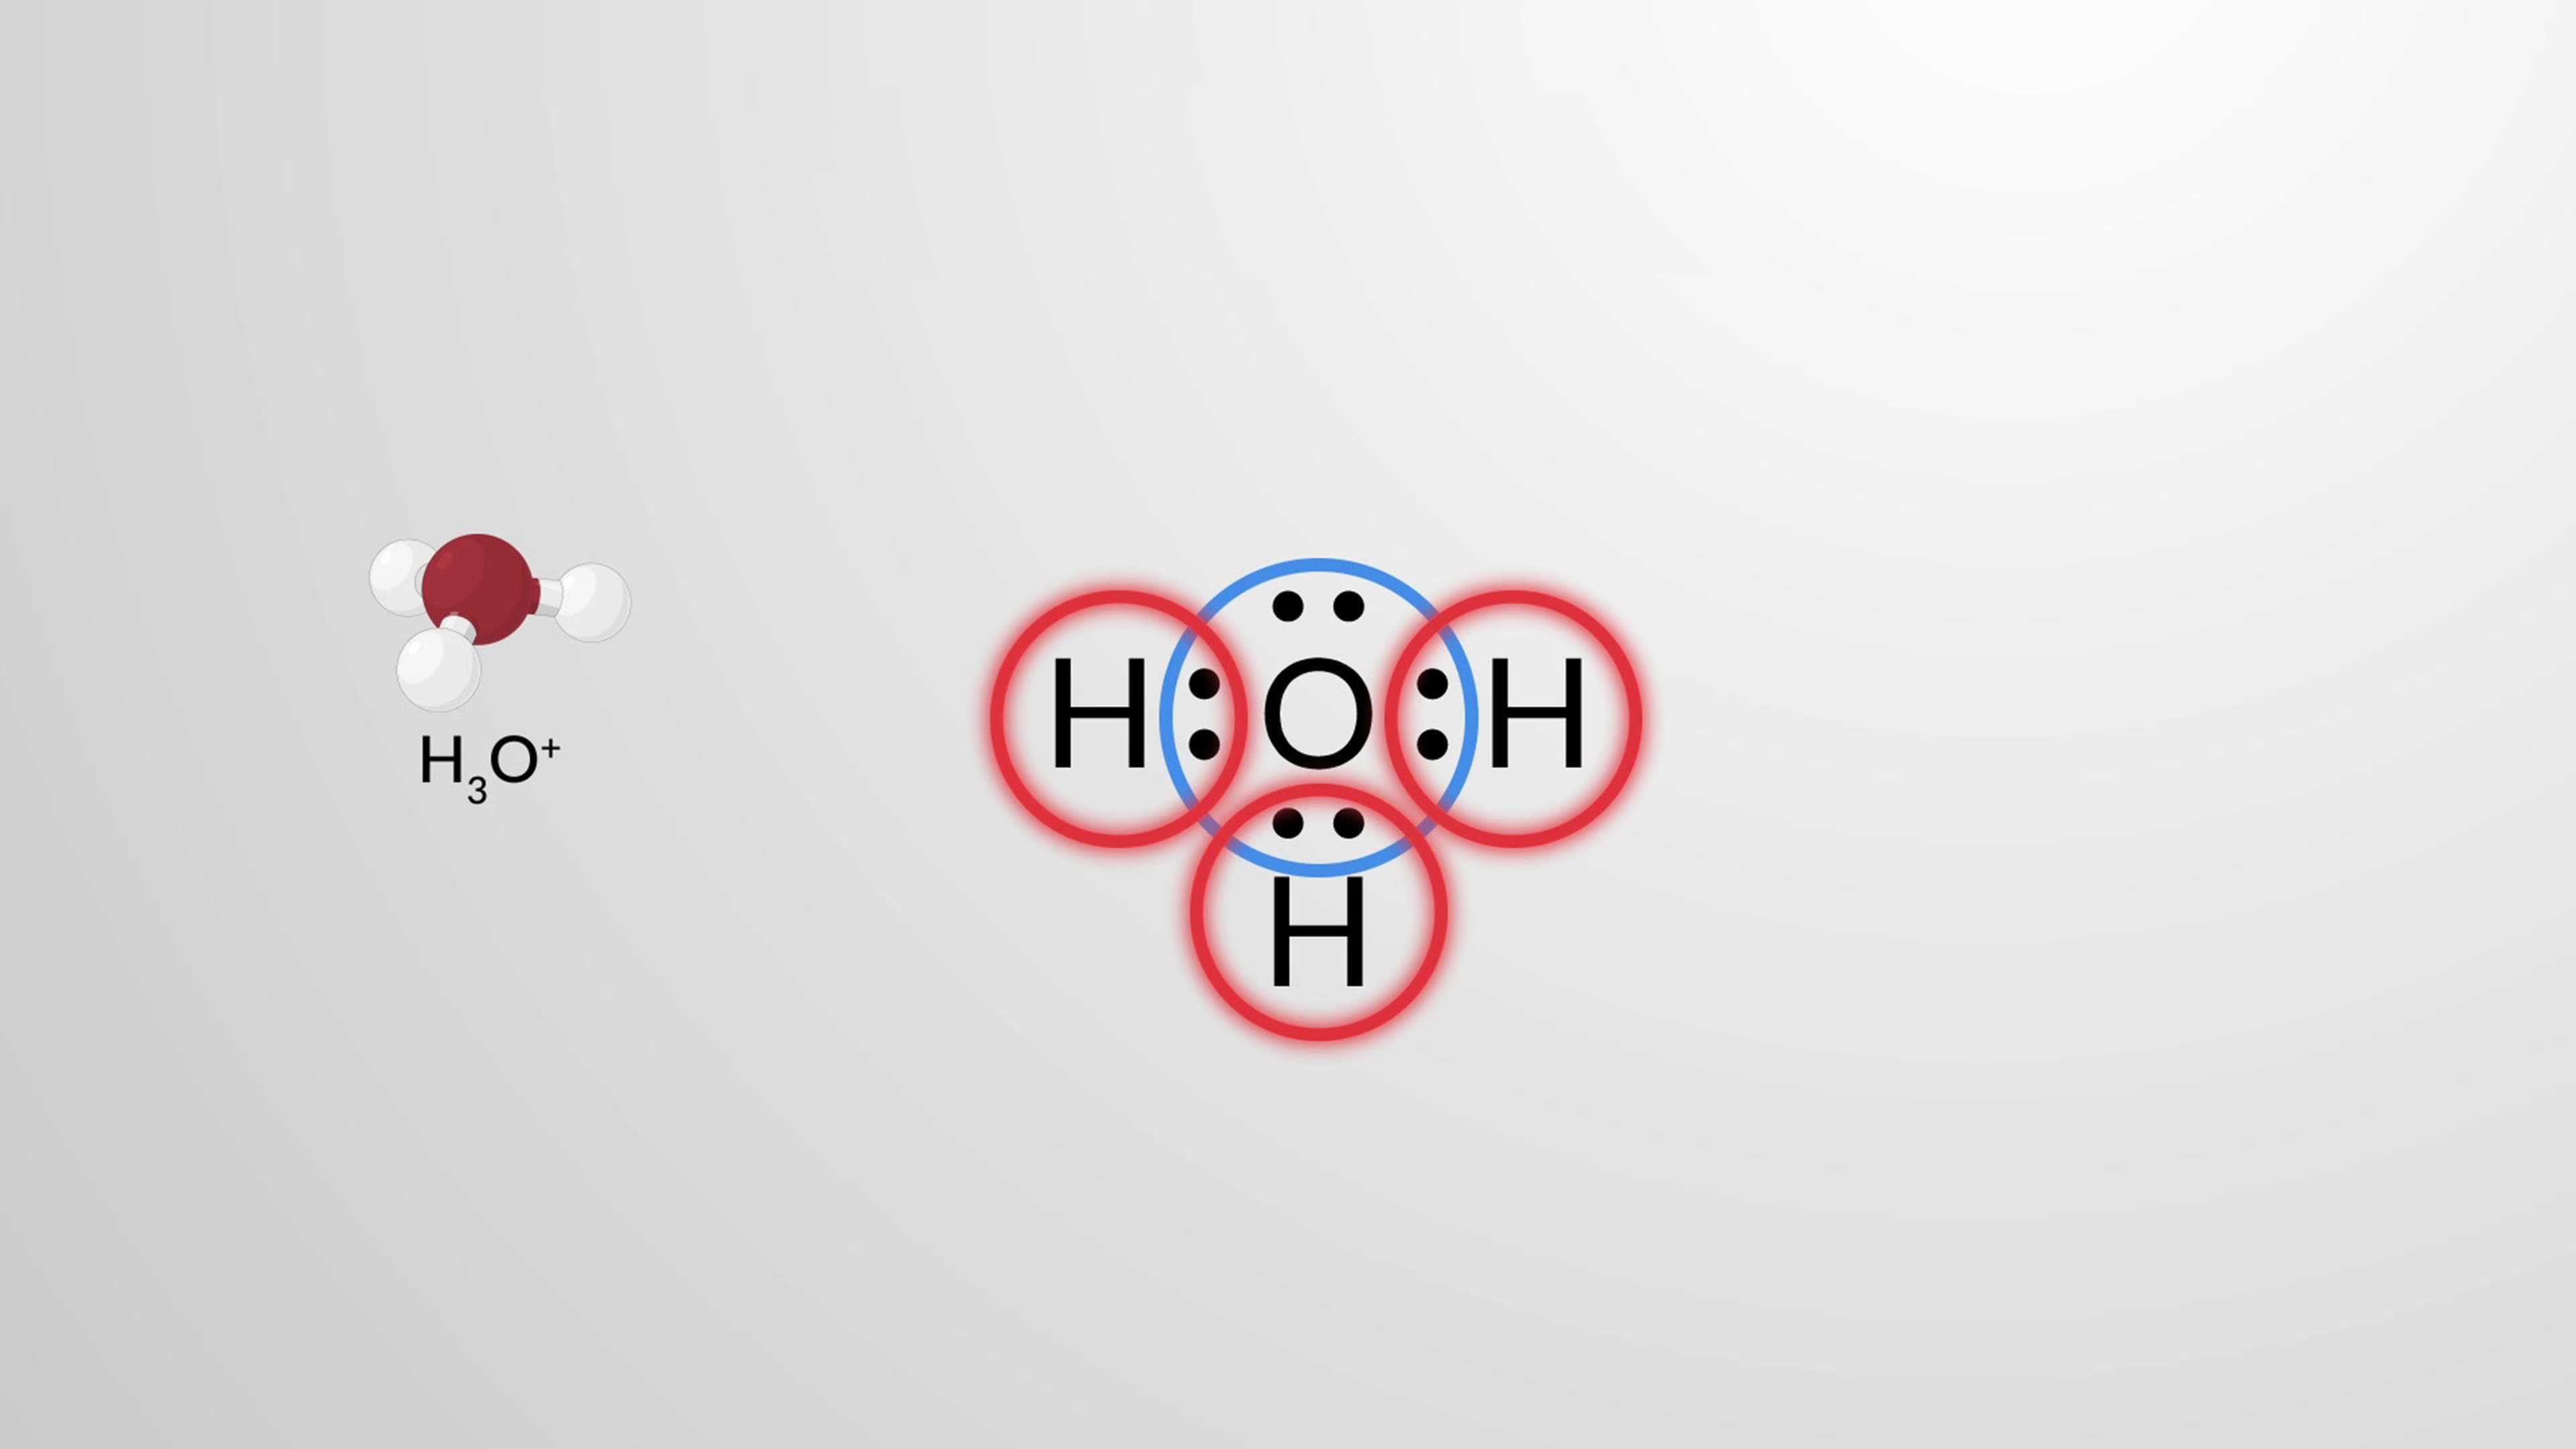
Lewis Structures of Molecular Compounds and Polyatomic Ions
 03:24
Hybridization of Atomic Orbitals I
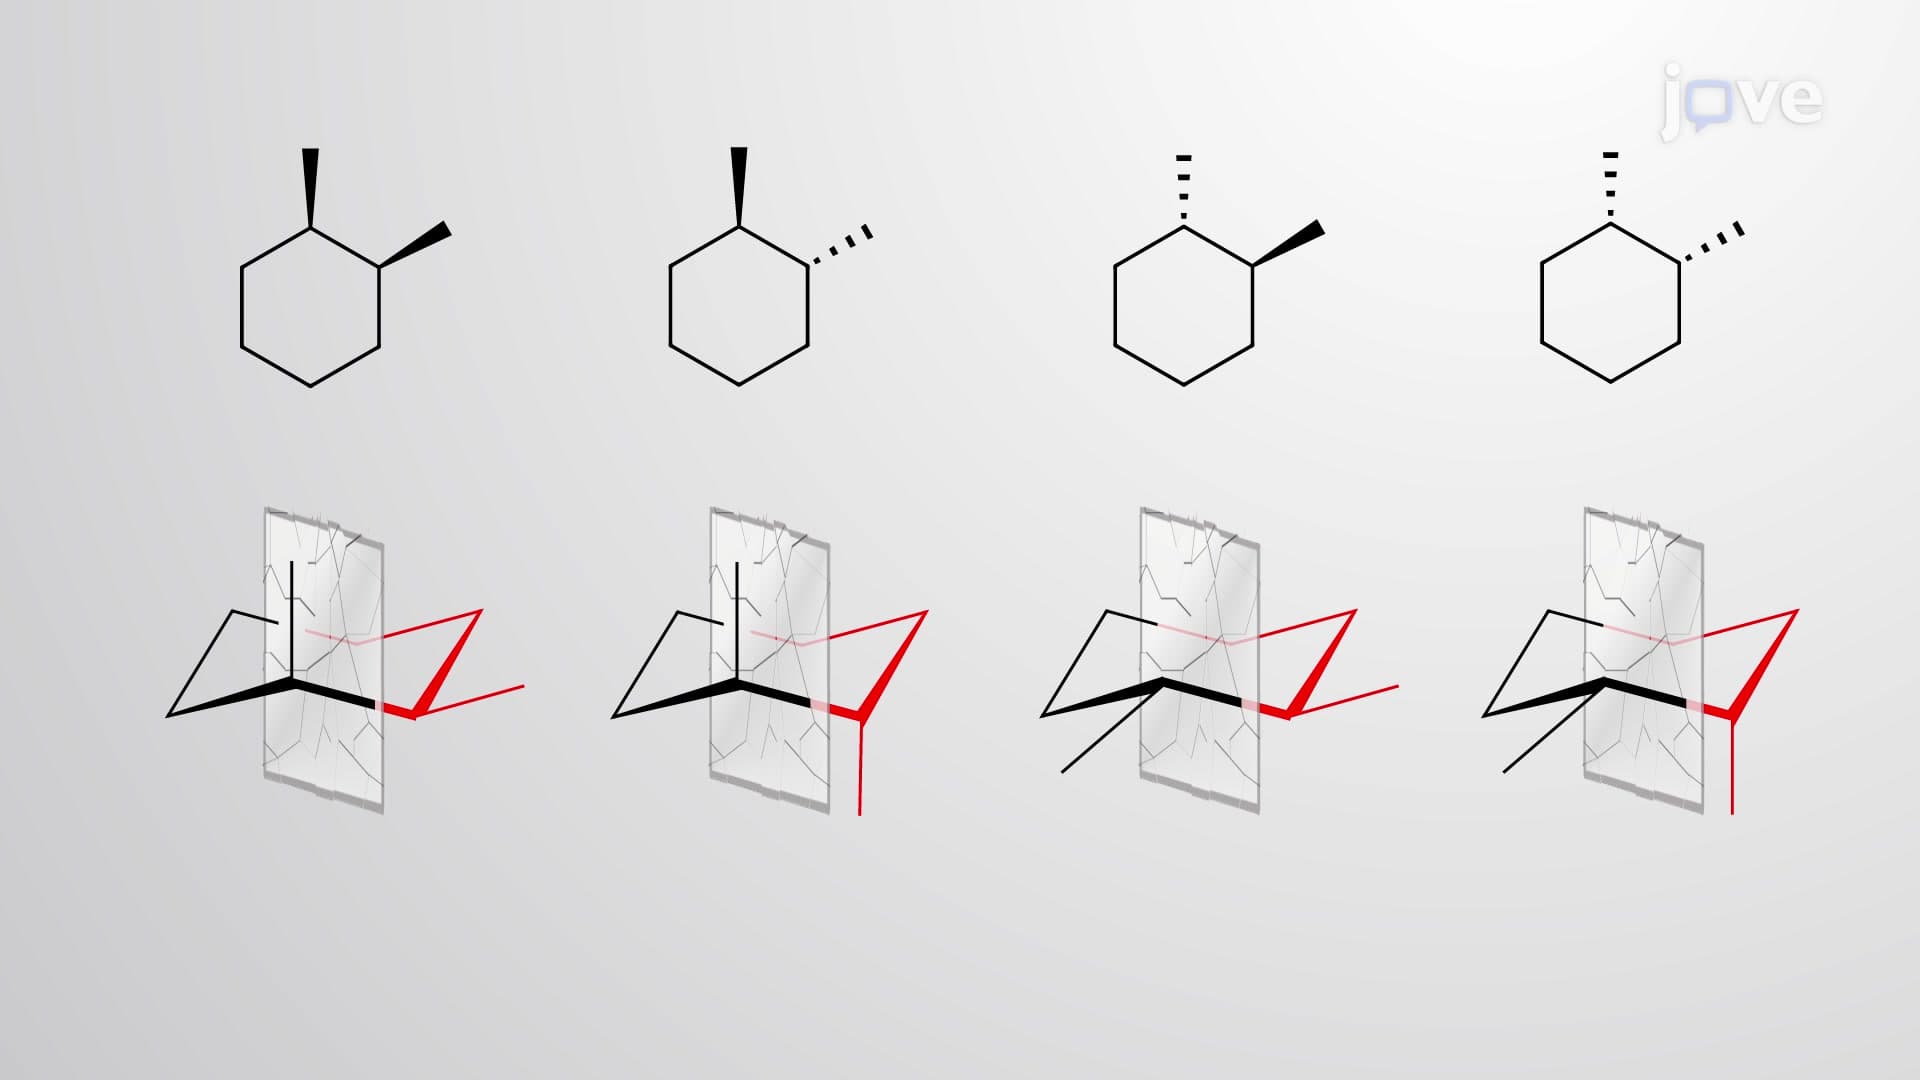 02:33
Stereoisomerism of Cyclic Compounds
 02:30
Stability of Substituted Cyclohexanes
The two chair conformations of cyclohexanes undergo rapid interconversion at room temperature. Both forms have identical energies and stabilities, each comprising equal amounts of the equilibrium mixture. Replacing a hydrogen atom with a functional group makes the two conformations energetically non-equivalent.
For example, in...