Protein dynamics from X-ray crystallography: anisotropic, global motion in diffuse scattering patterns
Related Experiment Video
Updated: Jul 18, 2026
Study of Protein Dynamics via Neutron Spin Echo Spectroscopy
Published on: April 13, 2022
Lars Meinhold1, Jeremy C Smith
1Interdisciplinary Center for Scientific Computing (IWR), Computational Molecular Biophysics, University of Heidelberg, D-69120 Heidelberg, Germany. lars.meinhold@iwr.uni-heidelberg.de

Abstract:
Understanding X-ray crystallographic diffuse scattering is likely to improve our comprehension of equilibrium collective protein dynamics. Here, using molecular dynamics (MD) simulation, a detailed analysis is performed of the origins of diffuse scattering in crystalline Staphylococcal nuclease, for which the complete diffuse scattering pattern has been determined experimentally. The hydrogen-atom contribution and the scattering range over which the scattering can be considered to be a sum of solvent and protein scattering are determined. Two models of correlated protein motion are investigated by calculating the model-derived diffuse scattering and comparing with the scattering calculated directly from MD trajectories. In one model, previously used in diffuse scattering interpretation, the atomic displacement correlations decay isotropically with increasing separation. Model correlation lengths are obtained by refining the model scattering against the simulation-derived scattering pattern, and are found to be significantly different from those correlation lengths derived directly from the MD trajectories. Furthermore, the convergence between the model-derived and MD-derived scattering is poor. The second model, in which the displacement correlations are calculated from the principal components of the MD trajectories, is capable of fully reproducing the MD-derived diffuse scattering if the approximately 50% lowest-frequency modes are included. However, a small number ( approximately 10) of lowest-frequency and largest-amplitude modes dominates the diffuse scattering and thus the correlated protein motions. A detailed analysis of the principal components is performed. In particular, the effective free energy profile associated with each principle mode is analyzed and the eigenfrequency and damping coefficient computed using a model of Brownian dynamics. Those collective modes with effective frequencies below approximately 0.5 THz, including those that determine the diffuse scattering, are overdamped.
More Related Videos
Related Concept Videos
 01:10
X-ray Diffraction of Biological Samples
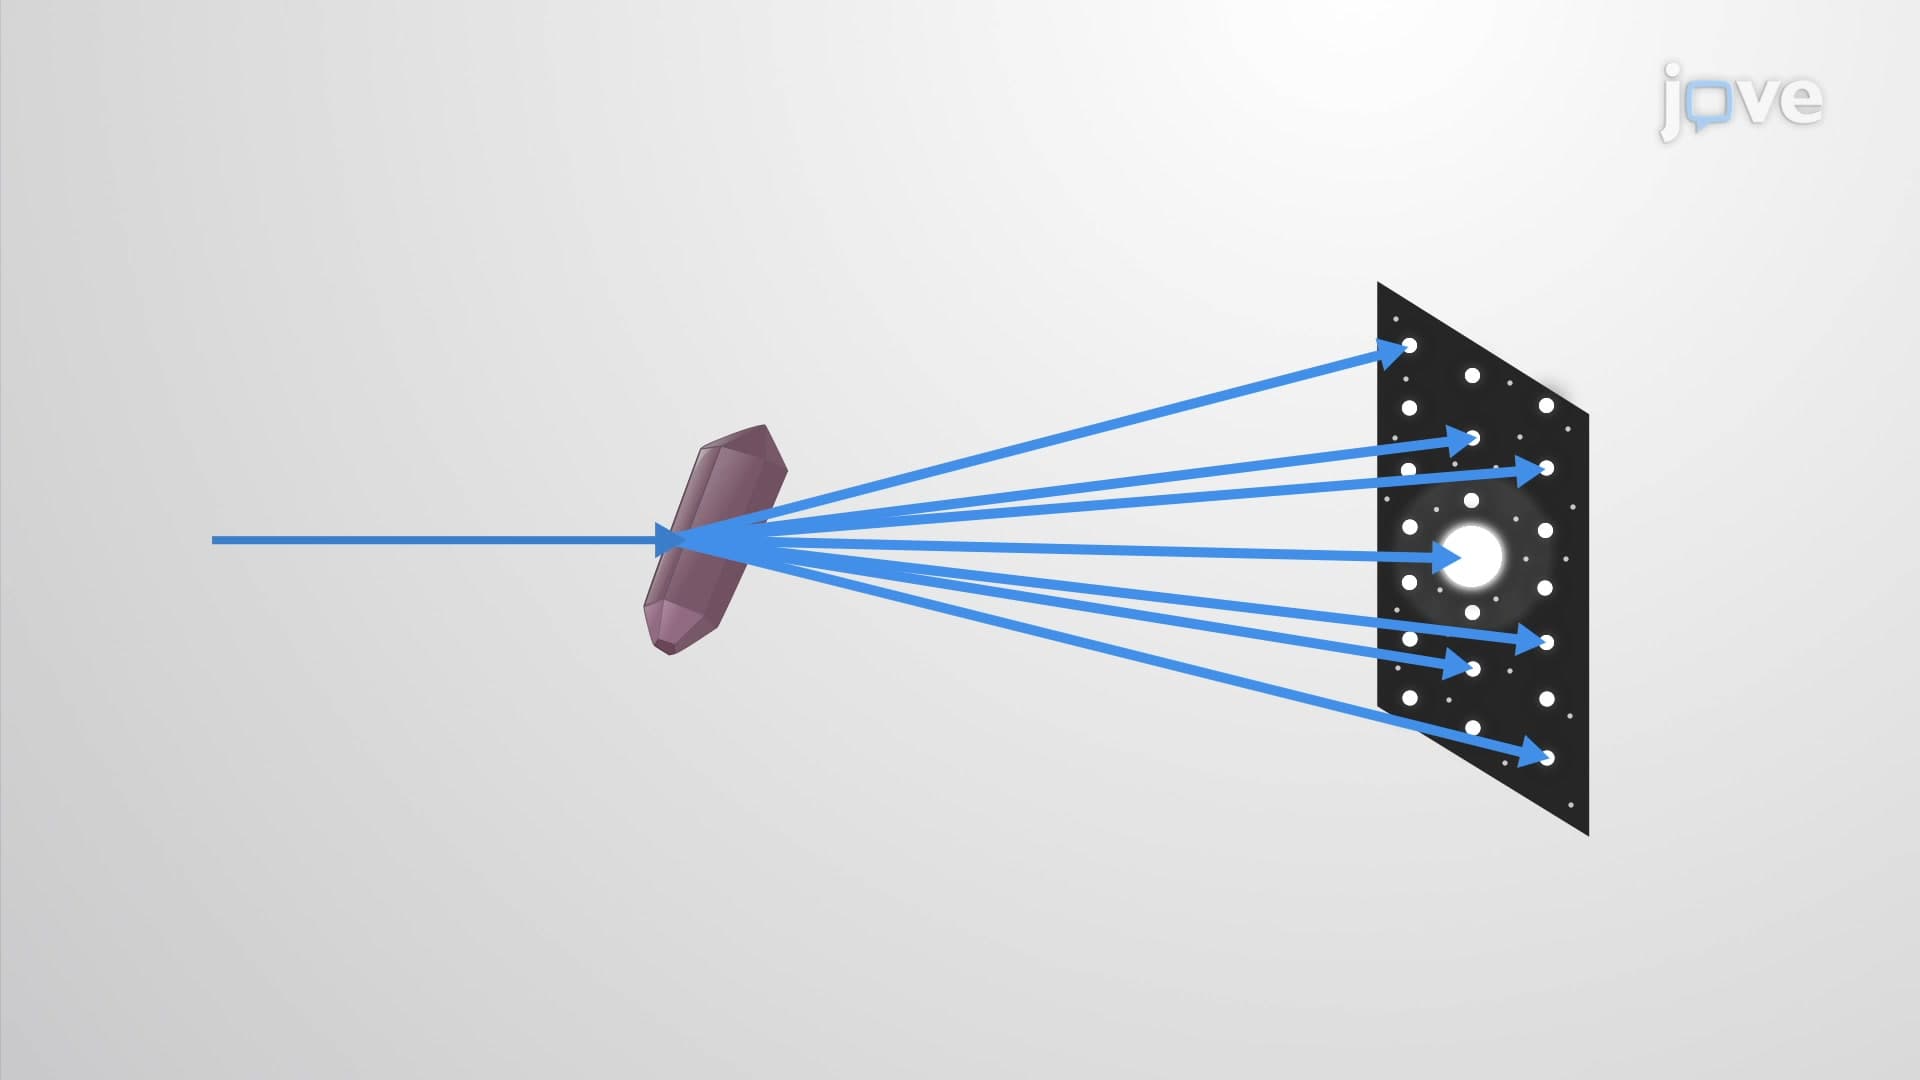
According to Bragg's law, when X-rays strike the sample positioned on a stage, the rays are scattered by the electron clouds around the sample atoms. The X-ray diffraction or scattering is caused by constructive interference of the X-ray waves that reflect off the internal crystal...
 02:18
X-ray Crystallography
Diffraction
Diffraction is the change in the direction of travel experienced by an electromagnetic wave when it encounters a physical barrier whose dimensions are comparable to those of the wavelength of the light. X-rays are electromagnetic radiation with wavelengths about as long as the distance between neighboring...
 01:29
Determination of Crystal Structures
 01:24
Protein Diffusion in the Membrane
 01:19
Protein Dynamics in Living Cells
Fluorescent recovery after photobleaching (FRAP) is a fluorescent-protein-based detection technique used to quantify protein movement rates within the cell. This method exposes a small portion of the cell to an intense laser beam. The laser beam causes permanent photobleaching of the fluorophore-tagged proteins in the exposed region. As the bleached...