Absence of severe recurrent infections in glycogen storage disease type Ib with neutropenia and neutrophil

P D'Eufemia1, R Finocchiaro, M Celli

  • 1Department of Pediatrics, University of Rome La Sapienza, Rome, Italy. patdeufemia@inwind.it

Related Concept Videos

Lysosomal Hydrolases01:22

Lysosomal Hydrolases

Lysosomes are the site for the degradation of macromolecules and biological polymers released during membrane trafficking events such as secretory, endocytic, autophagic, and phagocytic pathways. The membrane-enclosed area of the lysosome, called the lumen, contains hydrolytic enzymes active in an acidic environment. These acid hydrolases are functional at a pH between 4.5 and 5 and are involved in cellular processes such as cell signaling, energy metabolism, restoration of the plasma membrane,...
Immunodeficiency Diseases01:25

Immunodeficiency Diseases

Immunodeficiency disorders are conditions in which the immune system's ability to fight infectious disease and cancer is compromised or entirely absent. The immune system comprises a complex network of cells, tissues, and organs that work together to protect the body from potentially harmful invaders. When this system is deficient or not functioning properly, it leaves the body susceptible to infections, diseases, or other complications.
There are three main causes of immunodeficiency disorders...
Type I Diabetes III: Clinical Manifestations01:19

Type I Diabetes III: Clinical Manifestations
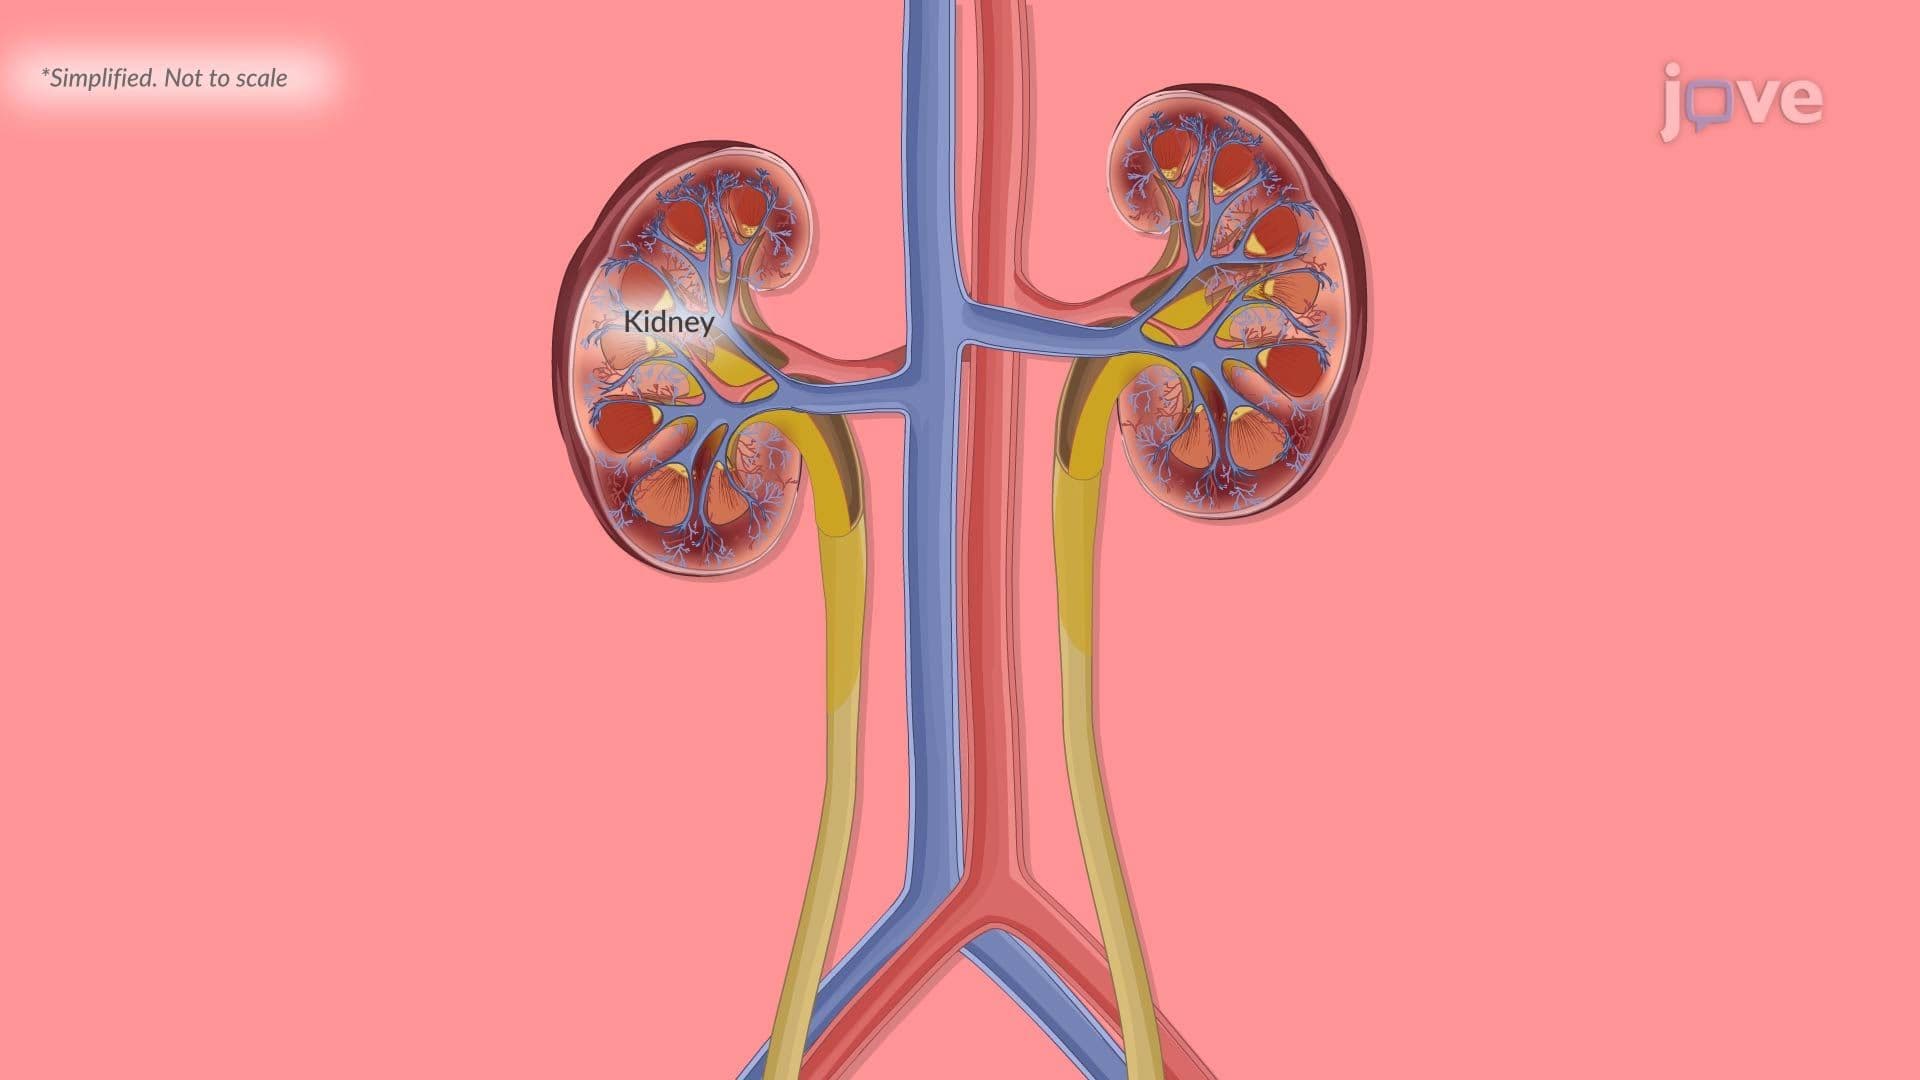
Type 1 diabetes mellitus typically presents with rapid-onset symptoms due to the body’s inability to utilize glucose in the absence of insulin. Since insulin is required for glucose uptake into cells, its deficiency leads to hyperglycemia and cellular energy deprivation, resulting in characteristic clinical features.Polyuria and PolydipsiaOne of the earliest, most prominent symptoms is polyuria (excessive urination). When blood glucose concentrations rise above the renal threshold, the kidneys...
Type I Diabetes II: Pathophysiology01:26

Type I Diabetes II: Pathophysiology

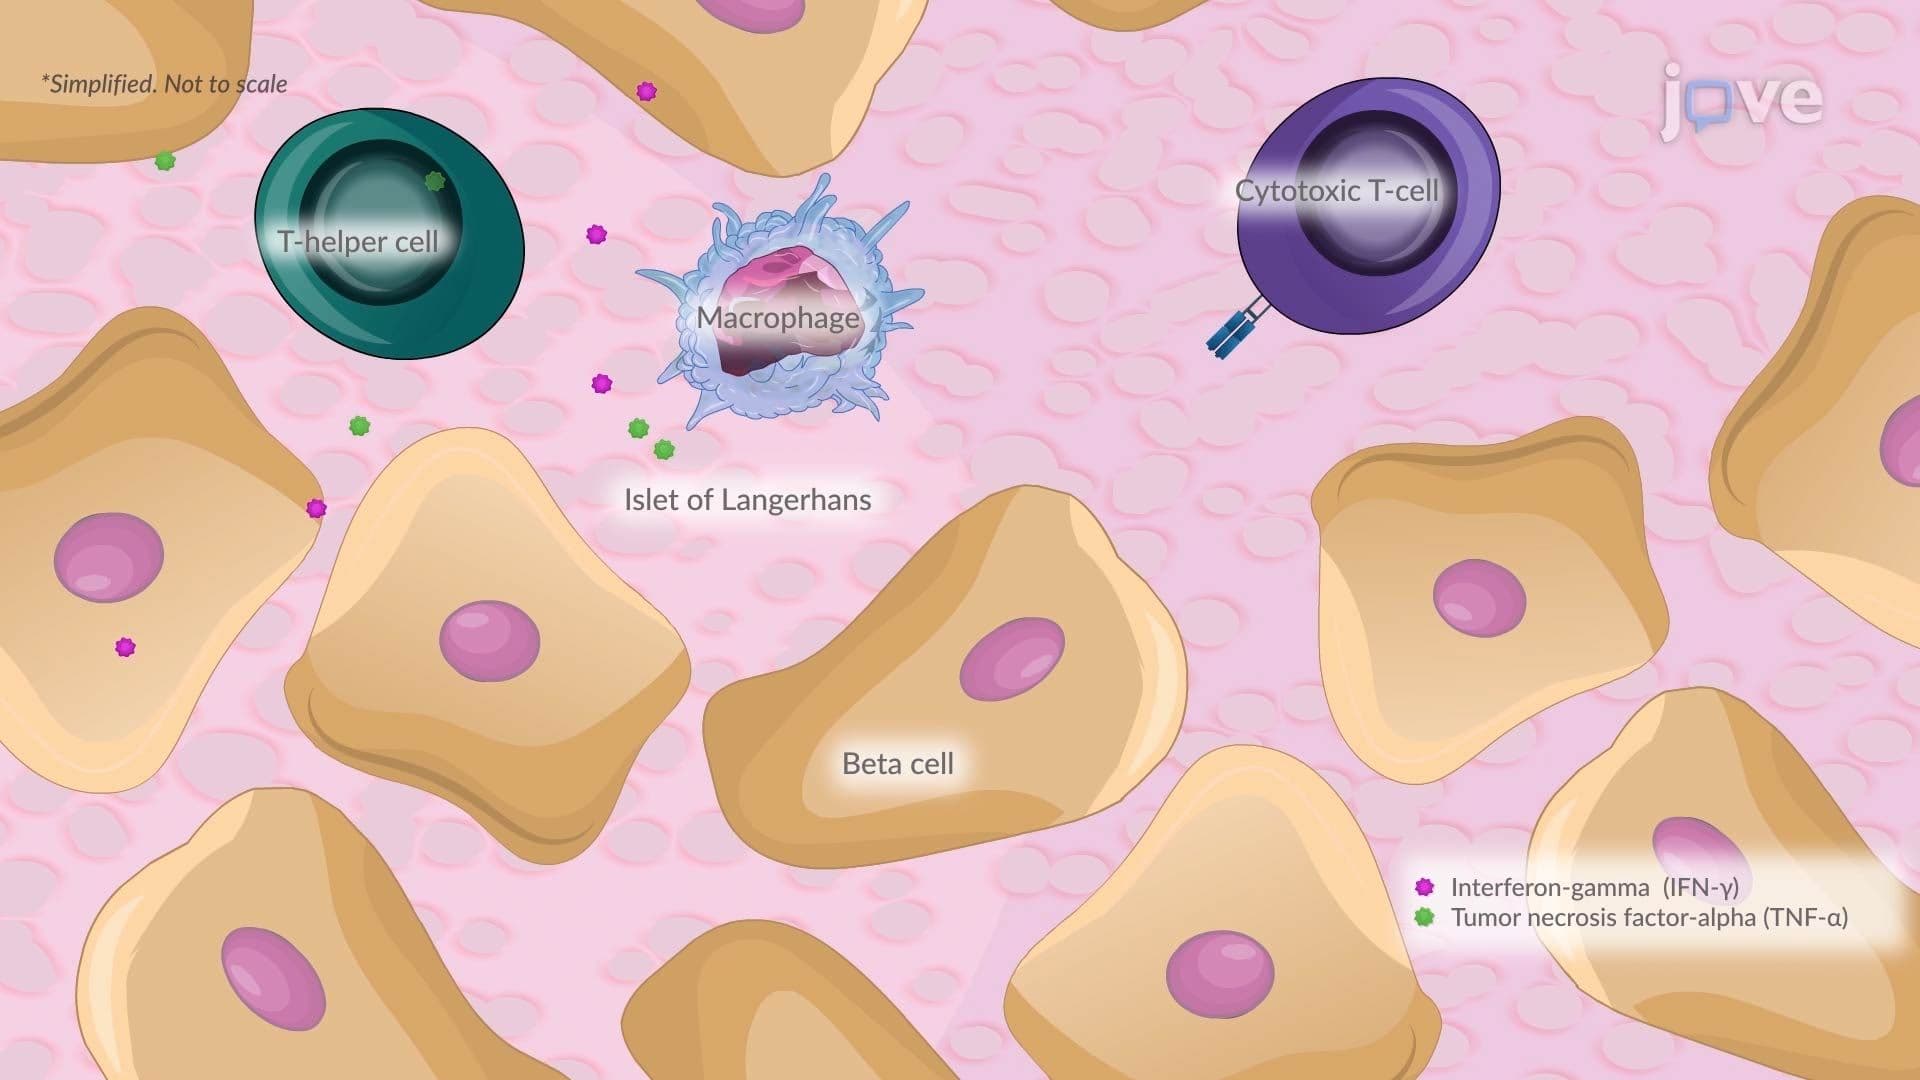
Type 1 diabetes mellitus arises from an immune-mediated destruction of pancreatic β-cells, resulting in an absolute deficiency of insulin. This process develops in genetically susceptible individuals when autoimmunity, environmental exposures, and immunologic dysregulation converge to trigger a targeted attack on the insulin-producing cells of the pancreas. The β-cells are located within the islets of Langerhans and are essential for regulating blood glucose by facilitating cellular uptake of...
Inborn Errors of Metabolism01:20

Inborn Errors of Metabolism

Phenylketonuria (PKU) is a protein metabolism disorder characterized by high blood levels of the amino acid phenylalanine. This results from a mutation in the gene responsible for phenylalanine hydroxylase, an enzyme that converts phenylalanine into tyrosine. When this enzyme is deficient, phenylalanine builds up in the blood, leading to symptoms such as vomiting, rashes, seizures, growth deficiency, and severe mental retardation. An early diagnosis and a diet restricting phenylalanine intake...
Cystic Fibrosis: Pathogenesis01:23

Cystic Fibrosis: Pathogenesis

Cystic fibrosis (CF), an autosomal recessive disorder, significantly affects the function of exocrine glands. This genetically inherited disease is characterized by the production of thick and sticky mucus, which can severely affect various organs and systems in the body.
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...