Related Experiment Video
Updated: May 6, 2026
Hyperinsulinemic-euglycemic Clamps in Conscious, Unrestrained Mice
Published on: November 16, 2011
Glucokinase and IRS-2 are required for compensatory beta cell hyperplasia in response to high-fat diet-induced
Yasuo Terauchi1, Iseki Takamoto, Naoto Kubota
1Department of Metabolic Diseases, Graduate School of Medicine, University of Tokyo, Hongo, Tokyo, Japan.

Abstract:
Glucokinase (Gck) functions as a glucose sensor for insulin secretion, and in mice fed standard chow, haploinsufficiency of beta cell-specific Gck (Gck(+/-)) causes impaired insulin secretion to glucose, although the animals have a normal beta cell mass. When fed a high-fat (HF) diet, wild-type mice showed marked beta cell hyperplasia, whereas Gck(+/-) mice demonstrated decreased beta cell replication and insufficient beta cell hyperplasia despite showing a similar degree of insulin resistance. DNA chip analysis revealed decreased insulin receptor substrate 2 (Irs2) expression in HF diet-fed Gck(+/-) mouse islets compared with wild-type islets. Western blot analyses confirmed upregulated Irs2 expression in the islets of HF diet-fed wild-type mice compared with those fed standard chow and reduced expression in HF diet-fed Gck(+/-) mice compared with those of HF diet-fed wild-type mice. HF diet-fed Irs2(+/-) mice failed to show a sufficient increase in beta cell mass, and overexpression of Irs2 in beta cells of HF diet-fed Gck(+/-) mice partially prevented diabetes by increasing beta cell mass. These results suggest that Gck and Irs2 are critical requirements for beta cell hyperplasia to occur in response to HF diet-induced insulin resistance.
More Related Videos
Related Concept Videos
 01:05
Insulin Secretory Vesicles
 01:16
Hormones Regulating Blood Glucose
In addition to accelerating glucose uptake and utilization, insulin has...
 01:27
Glucose Homeostasis: Pancreatic Islets and Insulin Secretion
Insulin and C-peptide are...
 01:28
Insulin: The Receptor and Signaling Pathways
 01:26
Type I Diabetes II: Pathophysiology
 01:24
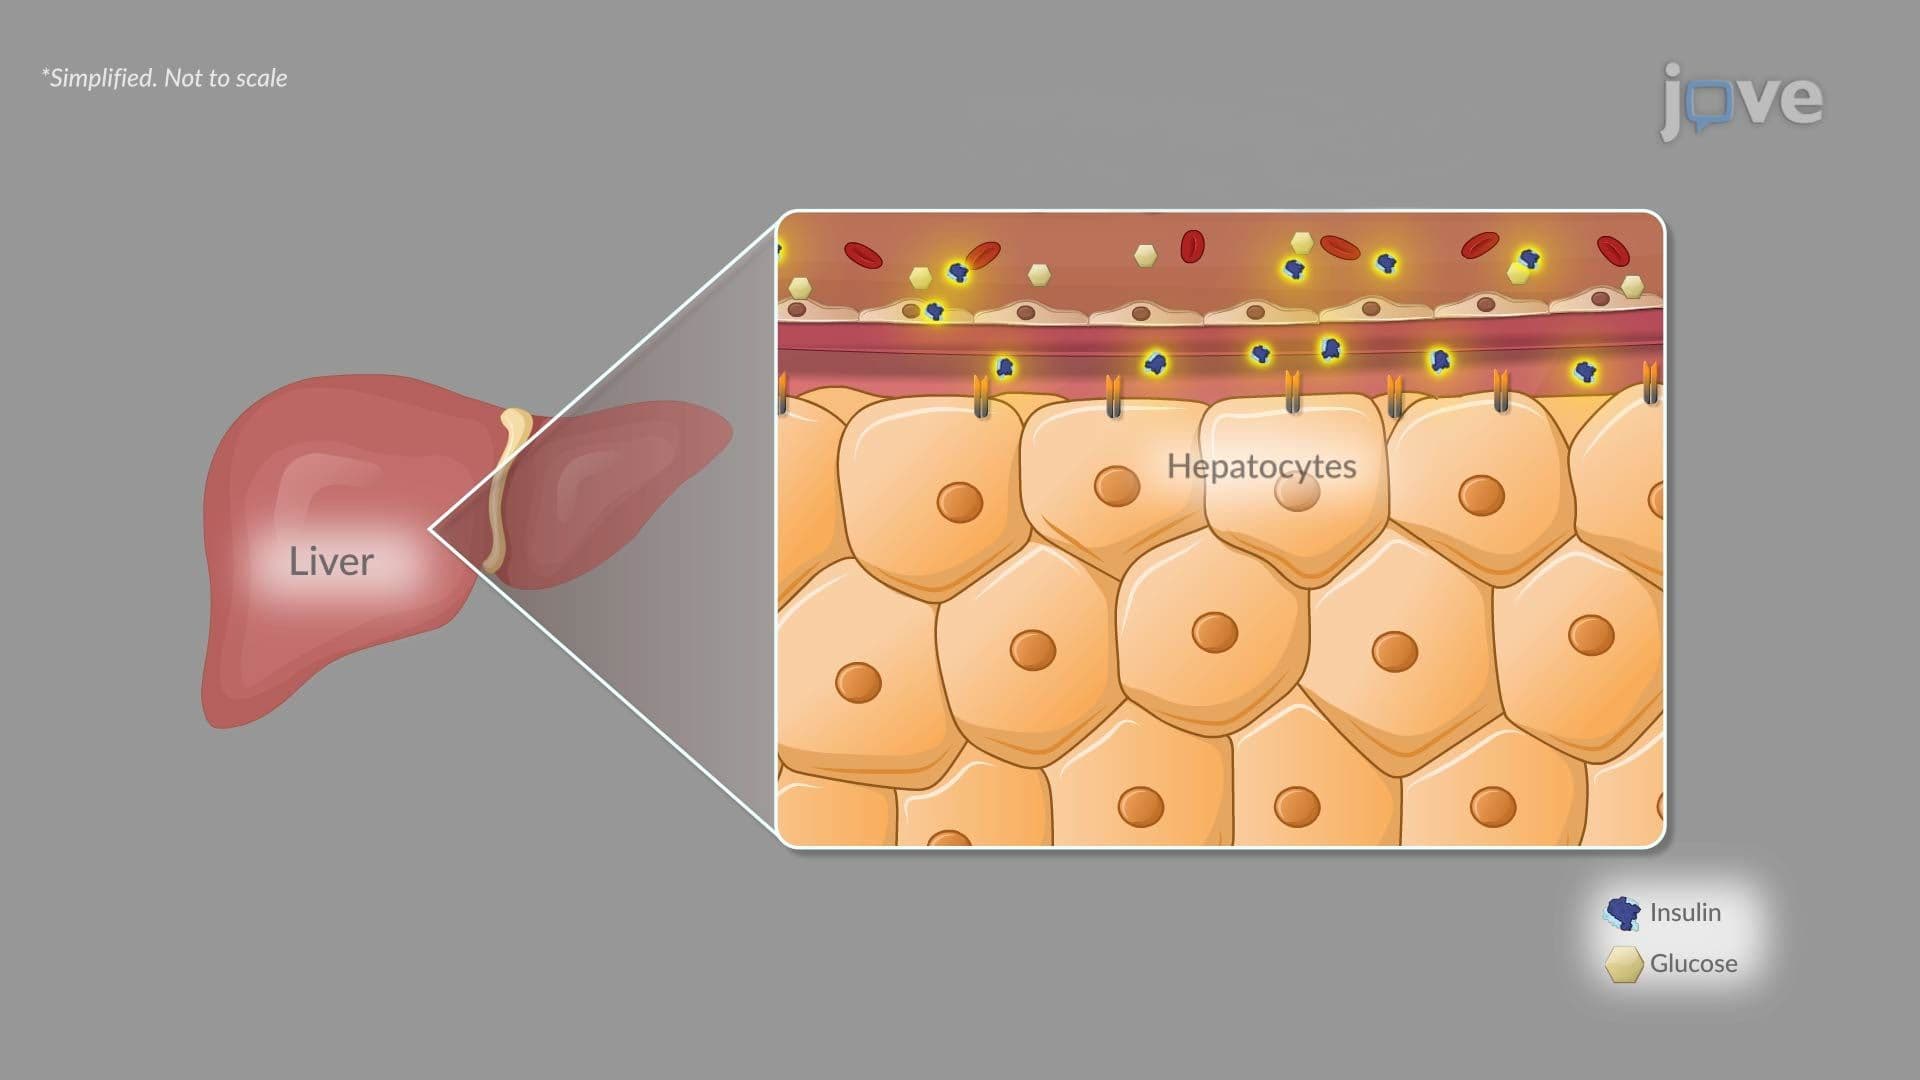
Type II Diabetes II: Pathophysiology