Related Experiment Video
Updated: Jul 17, 2026


In Vitro Enzyme Measurement to Test Pharmacological Chaperone Responsiveness in Fabry and Pompe Disease
Published on: December 20, 2017
Broad spectrum of Pompe disease in patients with the same c.-32-13T->G haplotype
M A Kroos1, R J Pomponio, M L Hagemans
1Department of Clinical Genetics, Erasmus MC, PO Box 2040, 3000 CA, Rotterdam, the Netherlands.

Background:
Pompe disease (acid maltase deficiency, glycogen storage disease type II; OMIM 232300) is an autosomal recessive lysosomal storage disorder characterized by acid alpha-glucosidase deficiency due to mutations in the GAA gene. Progressive skeletal muscle weakness affects motor and respiratory functions and is typical for all forms of Pompe disease. Cardiac hypertrophy is an additional fatal symptom in the classic infantile subtype. c.-32-13T-->G is the most common mutation in adults.
Objective:
To delineate the disease variation among patients with this mutation and to define the c.-32-13T-->G haplotypes in search for genotype-phenotype correlations.
Methods:
We studied 98 compound heterozygotes with a fully deleterious mutation (11 novel mutations are described) and the common c.-32-13T-->G mutation.
Results:
All patients were Caucasian. None had the classic infantile form of Pompe disease. The clinical course varied far more than anticipated (age at diagnosis <1 to 78 years; age at onset: <1 to 52 years). The acid alpha-glucosidase activities in a subset of patients ranged from 4 to 19.9 nmol/mg/h. Twelve different c.-32-13T-->G haplotypes were identified based on 17 single-nucleotide polymorphisms located in the GAA gene. In 76% of the cases, c.-32-13T-->G was encountered in the second most common GAA core haplotype (DHRGEVVT). In only one case was c.-32-13T-->G encountered in the major GAA core haplotype (DRHGEIVT).
Conclusion:
Patients with the same c.-32-13T-->G haplotype (c.q. GAA genotype) may manifest first symptoms at different ages, indicating that secondary factors may substantially influence the clinical course of patients with this mutation.
More Related Videos
 05:58
Digital Polymerase Chain Reaction Assay for the Genetic Variation in a Sporadic Familial Adenomatous Polyposis Patient Using the Chip-in-a-tube Format
Published on: August 20, 2018
 10:44
Chemical Inactivation of the E3 Ubiquitin Ligase Cereblon by Pomalidomide-based Homo-PROTACs
Published on: May 15, 2019
Related Concept Videos
 01:16
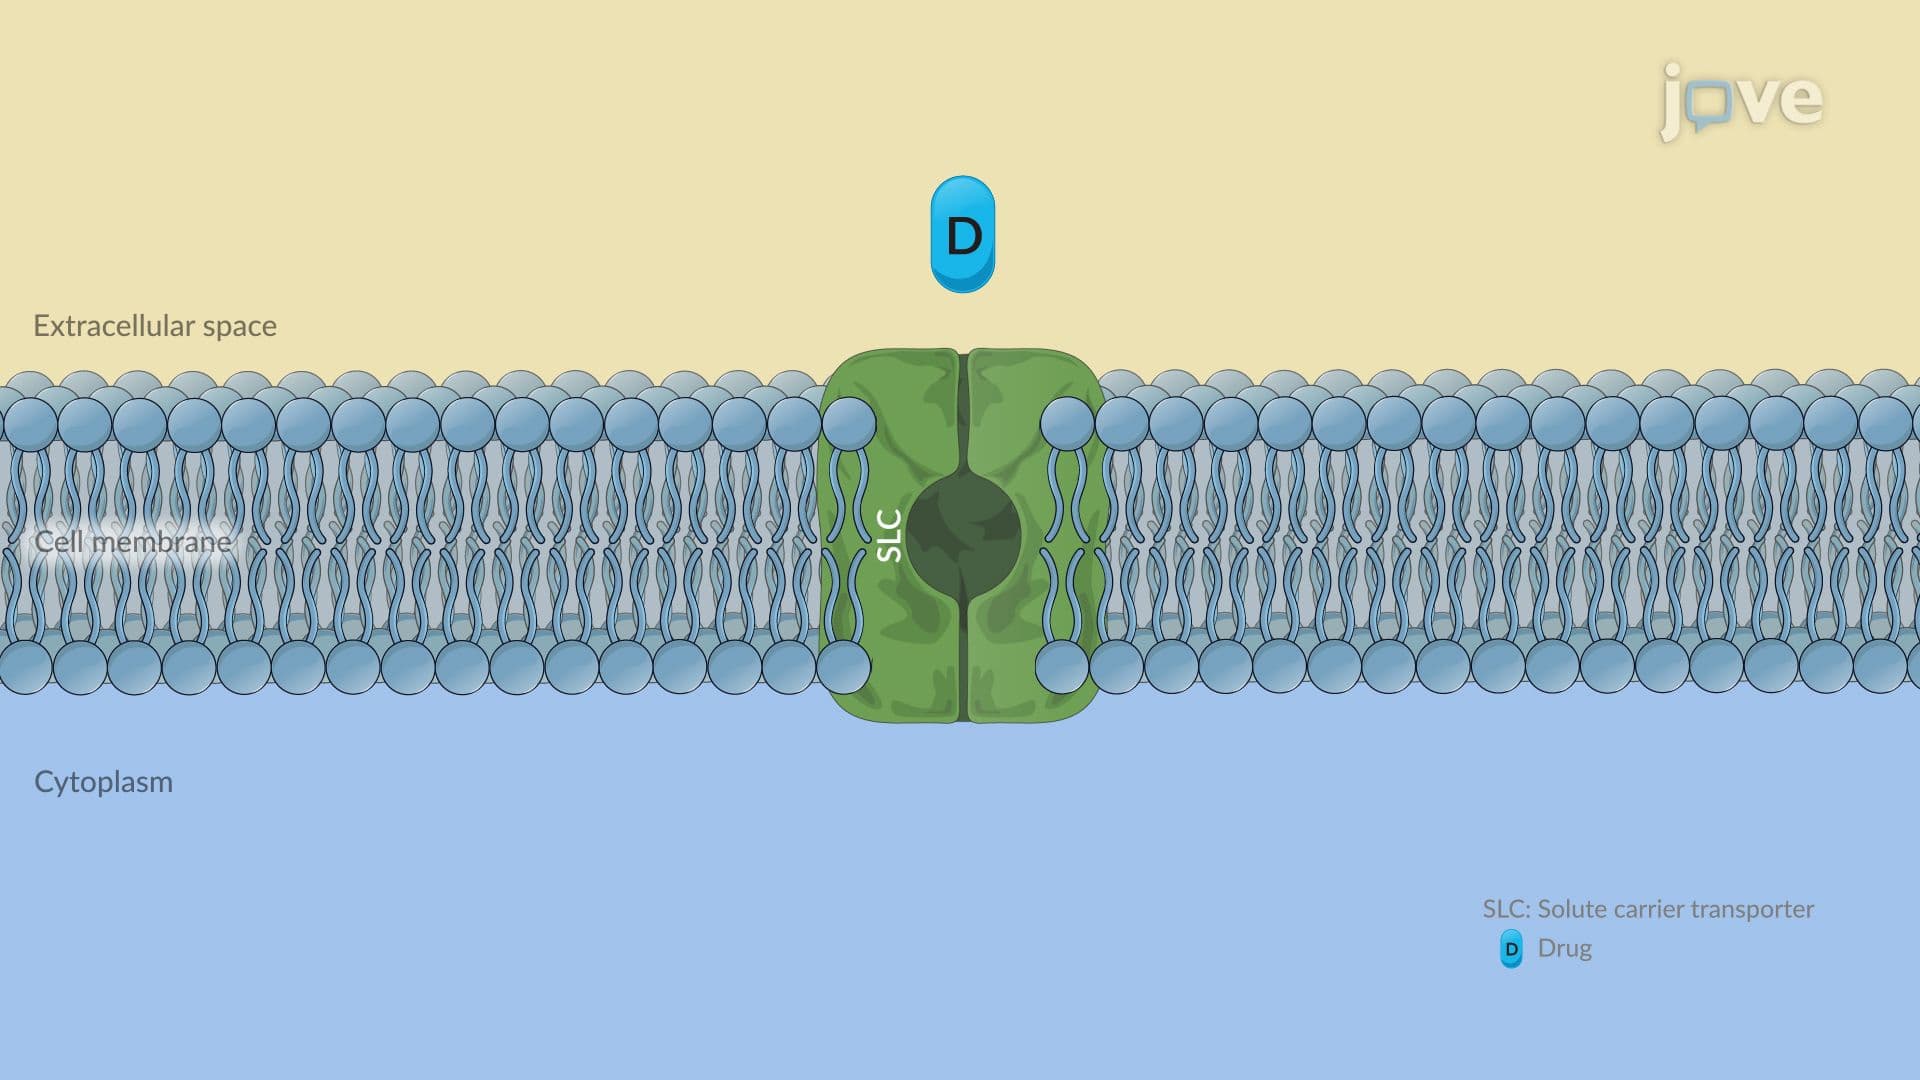
Pharmacogenetics of Drug Transporters: P-Glycoprotein and Solute Carrier Transporters
 01:11
Genome-wide Association Studies-GWAS
GWAS does not require the identification of the target gene involved in...
 01:33
Pleiotropy
 01:22
Lysosomal Hydrolases
 01:21
Huntington Disease l: Introduction
 01:28
Pharmacogenetics of Phase I Enzymes: Cytochrome P450 Isozymes