Related Experiment Video
Updated: Jul 17, 2026
A Plate-Based Assay for the Measurement of Endogenous Monoamine Release in Acute Brain Slices
Published on: August 11, 2021
Trace amine-associated receptor 1 displays species-dependent stereoselectivity for isomers of methamphetamine,
E A Reese1, J R Bunzow, S Arttamangkul
1Department of Physiology and Pharmacology, Oregon Health & Science University, 3181 SW Sam Jackson Park Rd., Portland, OR 97239, USA.

Abstract:
The synthetic amines methamphetamine (METH), amphetamine (AMPH), and their metabolite para-hydroxyamphetamine (POHA) are chemically and structurally related to the catecholamine neurotransmitters and a small group of endogenous biogenic amines collectively referred to as the trace amines (TAs). Recently, it was reported that METH, AMPH, POHA, and the TAs para-tyramine (TYR) and beta-phenylethylamine (PEA) stimulate cAMP production in human embryonic kidney (HEK)-293 cells expressing rat trace amine-associated receptor 1 (rTAAR1). The discovery that METH and AMPH activate the rTAAR1 motivated us to study the effect of these drugs on the mouse TAAR1 (mTAAR1) and a human-rat chimera (hrChTAAR1). Furthermore, because S-(+)-isomers of METH and AMPH are reported to be more potent and efficacious in vivo than R-(-), we determined the enantiomeric selectivity of all three species of TAAR1. In response to METH, AMPH, or POHA exposure, the accumulation of cAMP by HEK-293 cells stably expressing different species of TAAR1 was concentration- and isomer-dependent. EC50 values for S-(+)-METH were 0.89, 0.92, and 4.44 microM for rTAAR1, mTAAR1, and h-rChTAAR1, respectively. PEA was a potent and full agonist at each species of TAAR1, whereas TYR was a full agonist for the rodent TAAR1s but was a partial agonist at h-rChTAAR1. Interestingly, both isomers of METH were full agonists at mTAAR1 and h-rChTAAR1, whereas both were partial agonists at rTAAR1. Taken together, these in vitro results suggest that, in vivo, TAAR1 could be a novel mediator of the effects of these drugs.
Related Concept Videos
 01:16
Adrenergic Agonists: Chemistry and Structure-Activity Relationship
Aromatic ring substitutions: Substituting the aromatic ring with –OH groups at positions 3 and 4 yields catecholamines (e.g., epinephrine), which have a high affinity for adrenoceptors. Hydrogen bonding between –OH groups and receptors enhances adrenergic activity.
Separation of the aromatic...
 01:25
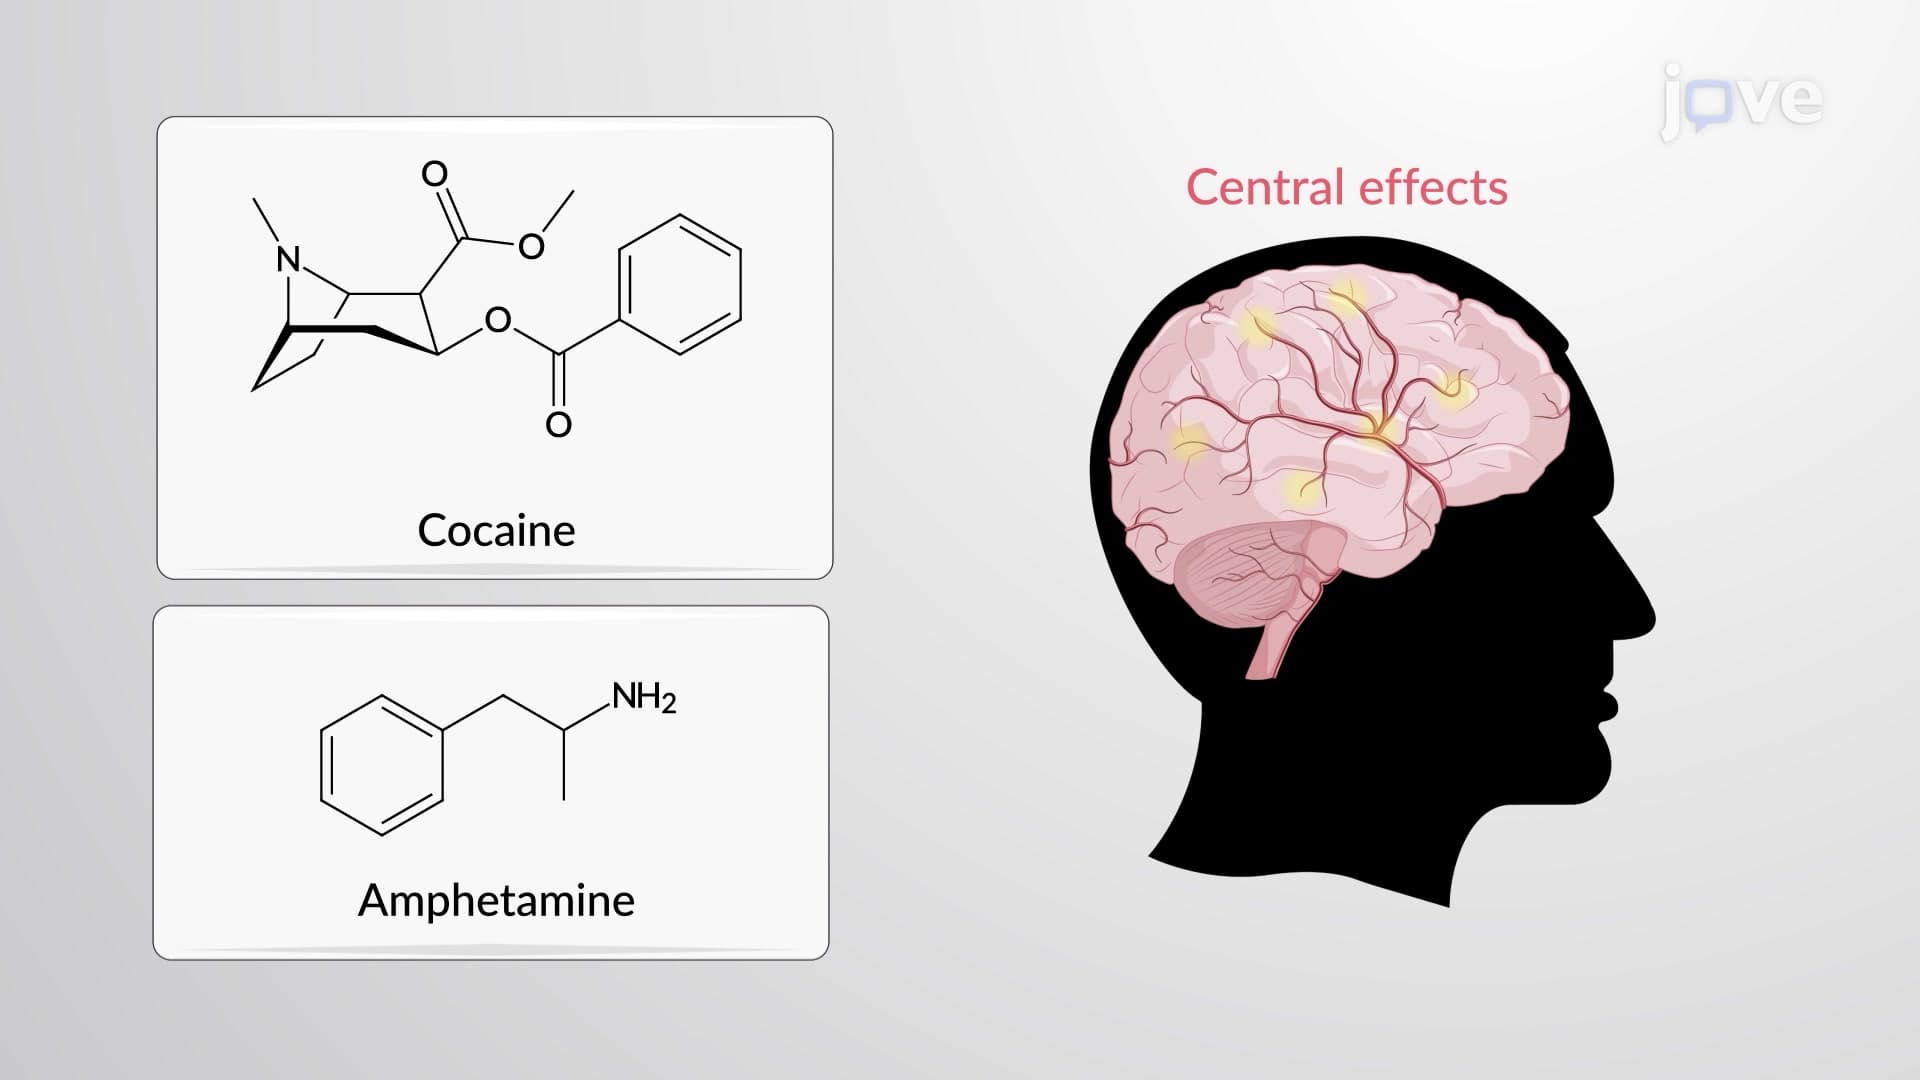
Adrenergic Agonists: Indirect-Acting Agents
One mechanism involves depleting stored catecholamines by displacing them from synaptic vesicles. These agents, known as "displacers," are transported into vesicles at the expense of noradrenaline. Examples include amphetamine and tyramine, which lack a catechol moiety, resulting in prolonged action, improved oral bioavailability, and...
 02:32
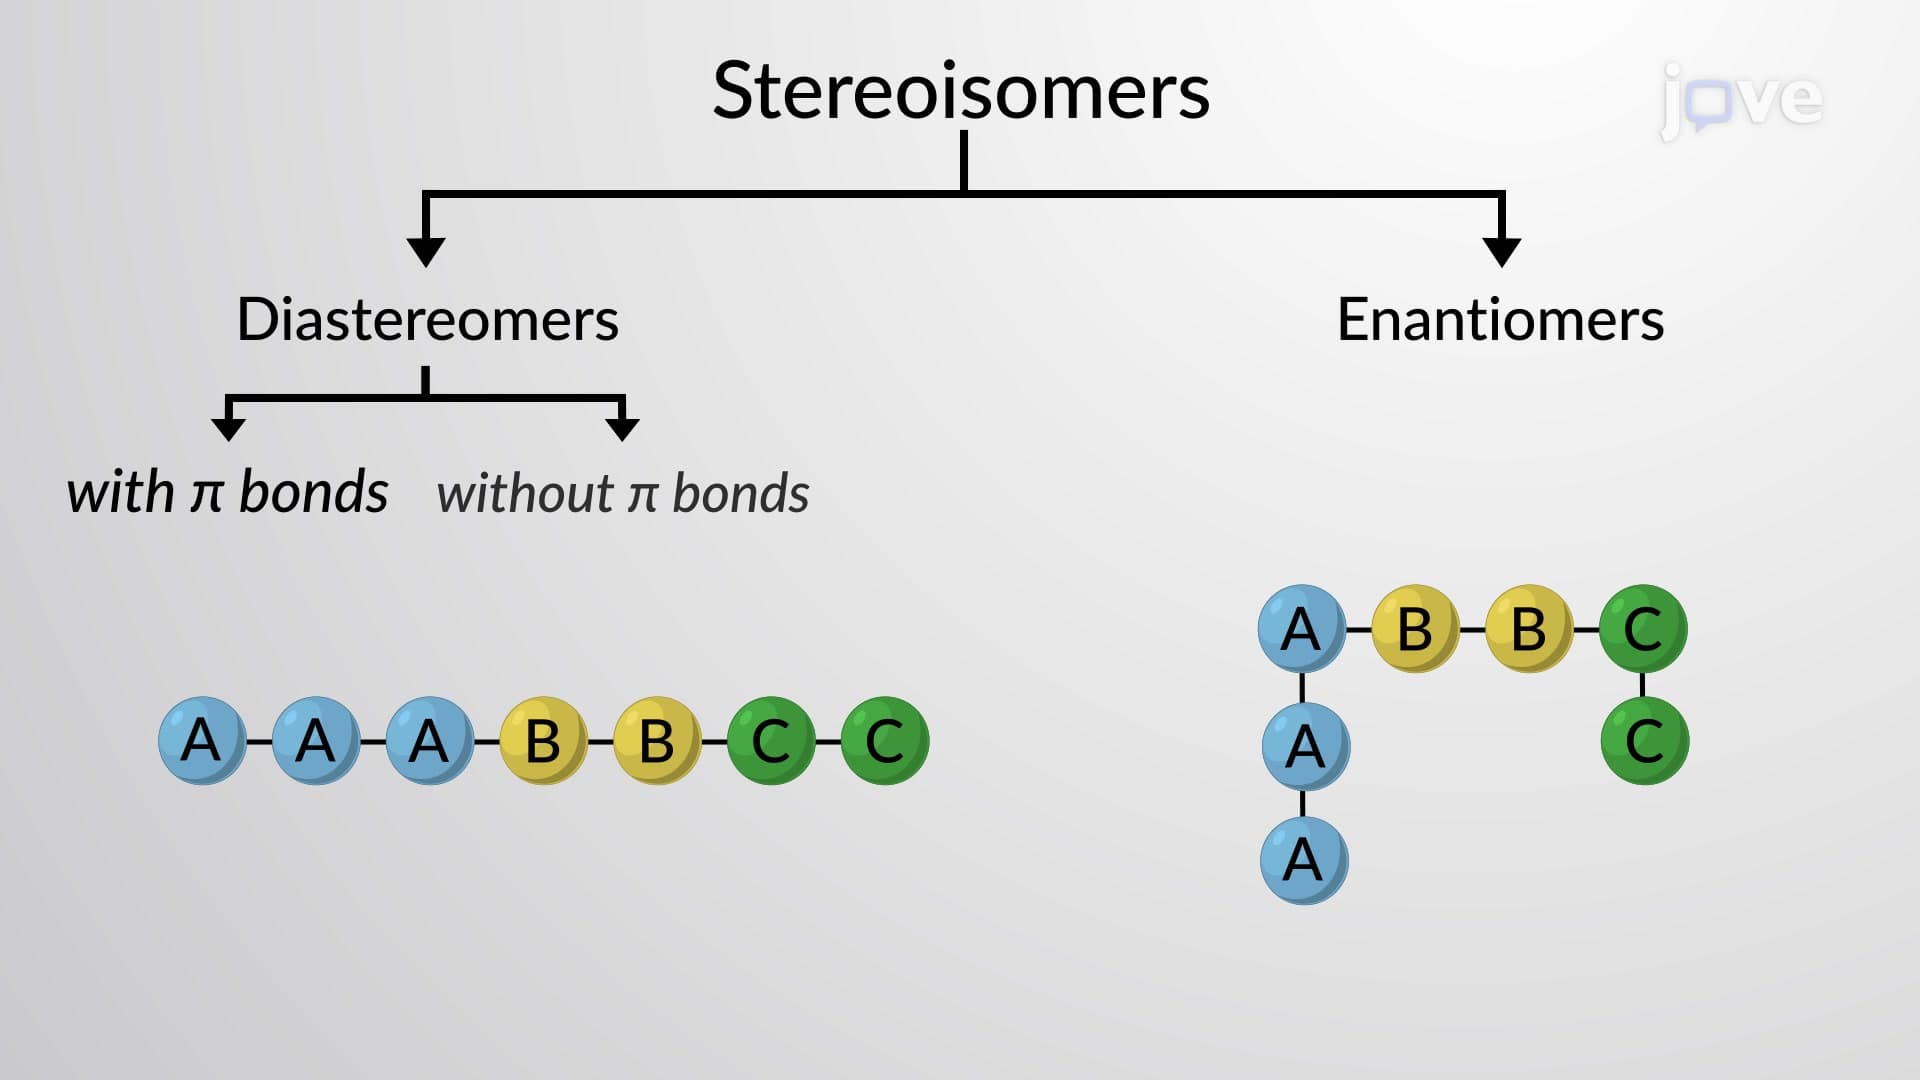
Stereoisomers
 01:17
Adrenergic Antagonists: Chemistry and Classification of ɑ-Receptor Blockers
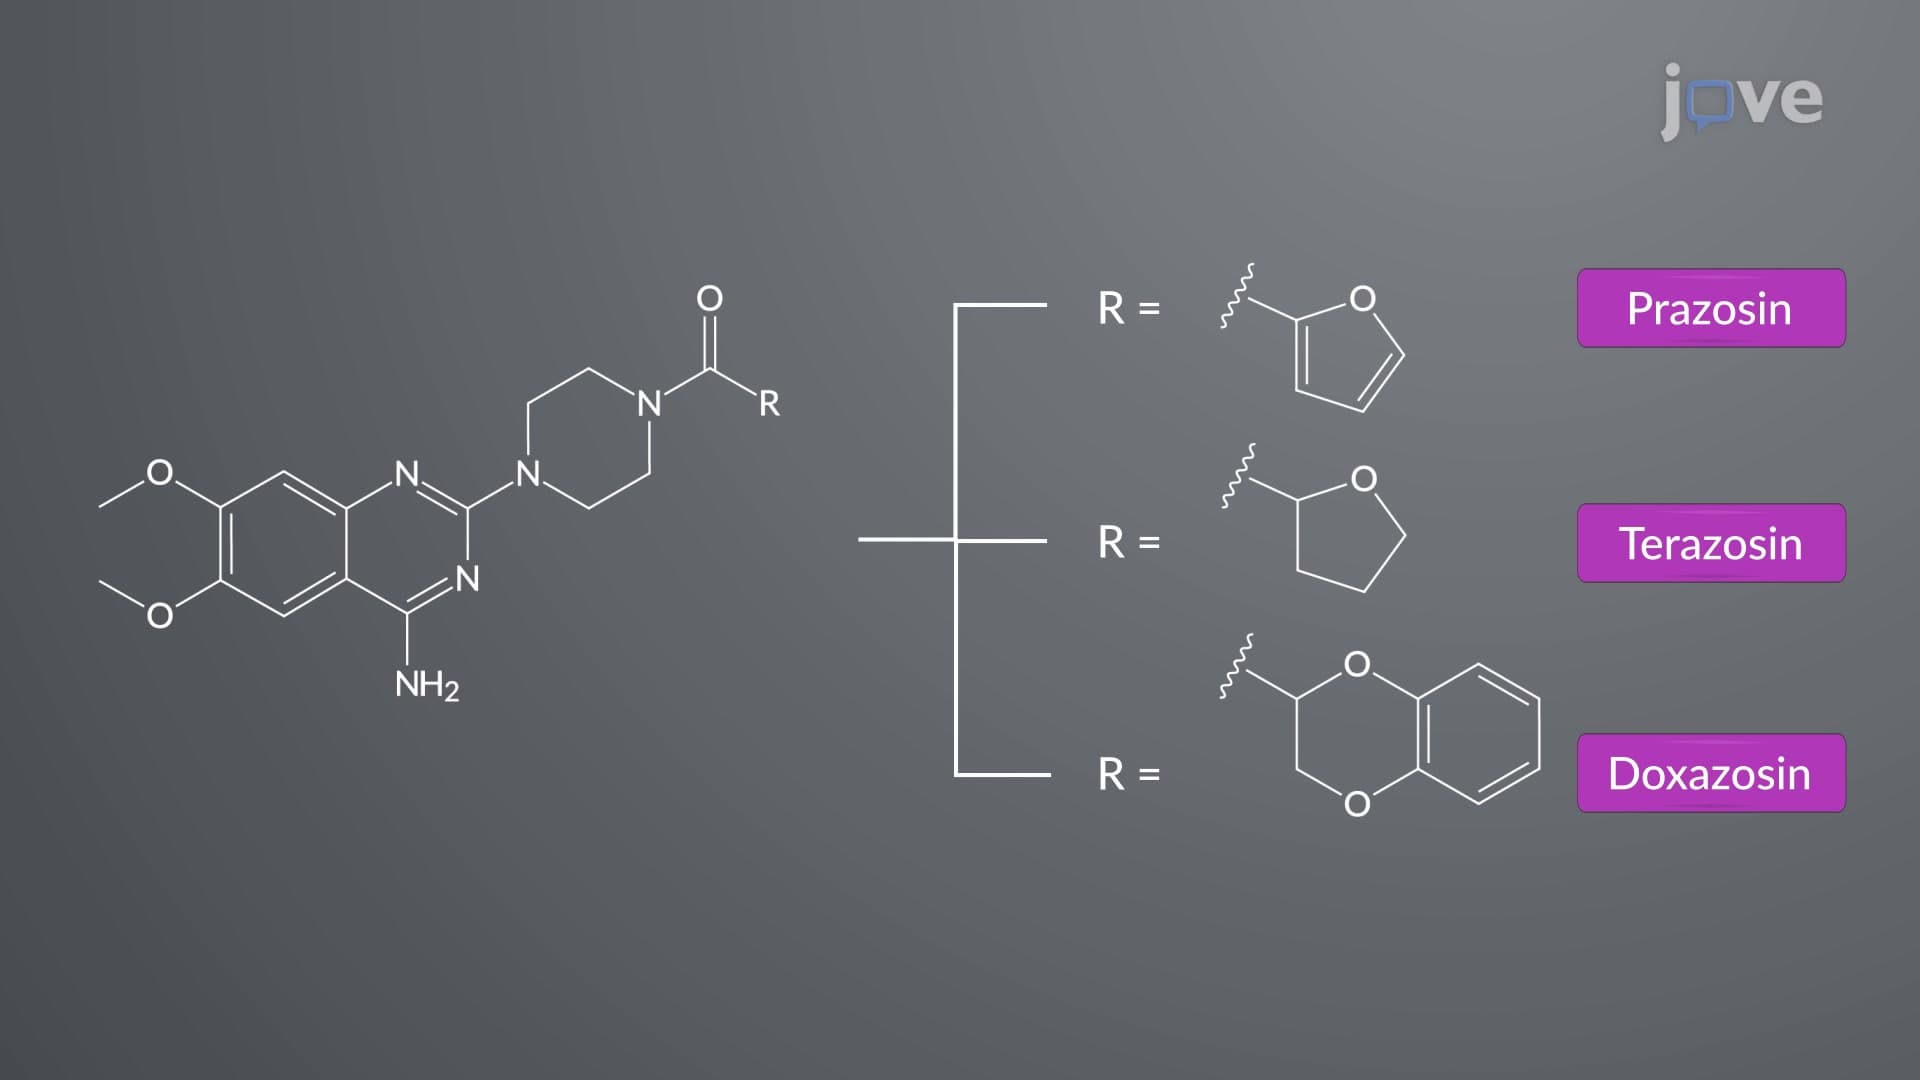
Nonselective α-blockers: Nonselective α-blockers contain haloalkylamine or imidazoline moieties. Phenoxybenzamine, with a haloalkylamine...
 01:21
Drugs Affecting Neurotransmitter Release or Uptake
 01:31
Adrenergic Receptors: ɑ Subtype
Adrenaline ≥ Noradrenaline >> Isoprenaline
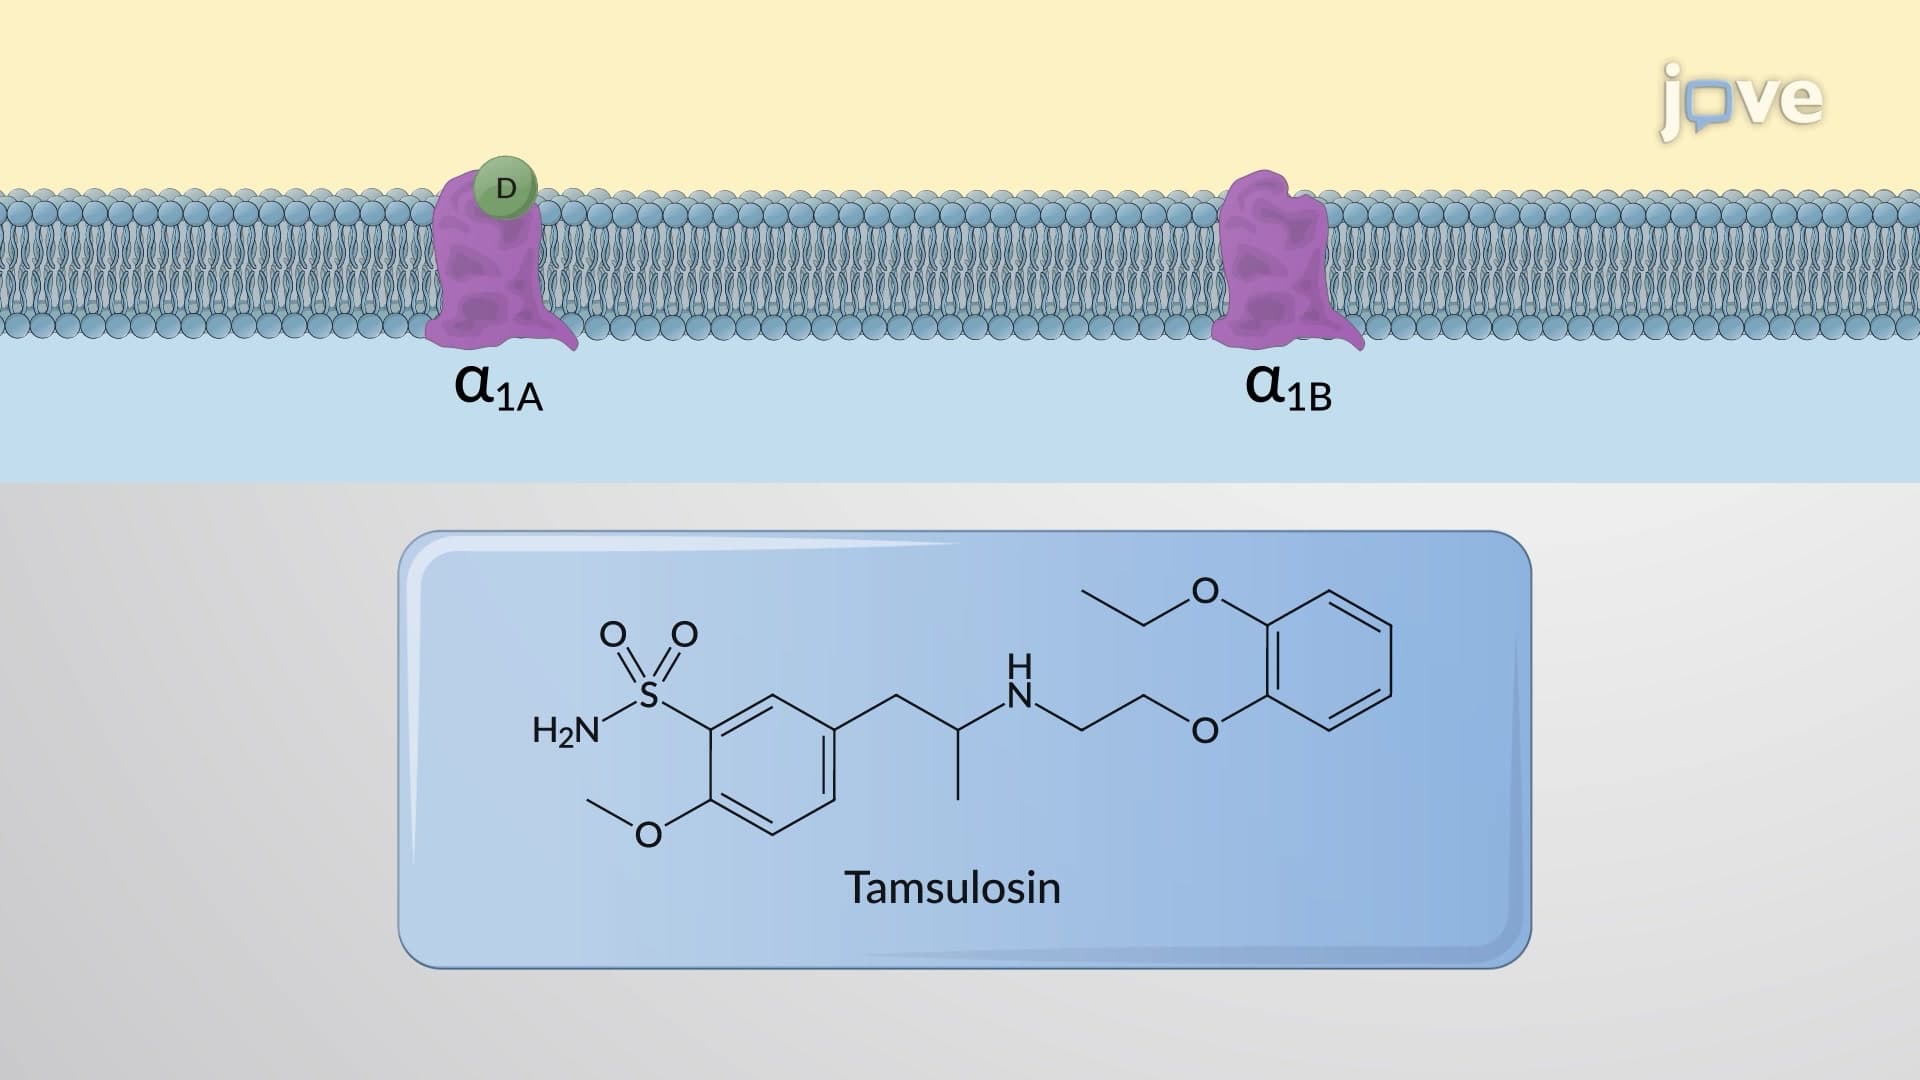
α-adrenoceptors are further divided into α1 and α2-adrenoceptors.
α1-Adrenoceptors: These receptors are located postsynaptically on the effector organs and cause constriction of smooth muscle mediated by activation of phospholipase C—inositol-1,4,5-trisphosphate...