Related Experiment Video
Updated: Jul 17, 2026

In Silico Modeling Method for Computational Aquatic Toxicology of Endocrine Disruptors: A Software-Based Approach Using QSAR Toolbox
Published on: August 28, 2019
Progress in QSAR toxicity screening of pharmaceutical impurities and other FDA regulated products
Naomi L Kruhlak1, Joseph F Contrera, R Daniel Benz
1US Food and Drug Administration, Center for Drug Evaluation and Research, Office of Pharmaceutical Science, Informatics and Computational Safety Analysis Staff, US Food and Drug Administration, 10903 New Hampshire Avenue, Silver Spring, MD 20993, USA.

Abstract:
Active ingredients in pharmaceutical products undergo extensive testing to ensure their safety before being made available to the American public. A consideration during the regulatory review process is the safety of pharmaceutical contaminants and degradents which may be present in the drug product at low levels. Several published guidances are available that outline the criteria for further testing of these impurities to assess their toxic potential, where further testing is in the form of a battery of toxicology assays and the identification of known structural alerts. However, recent advances in the development of computational methods have made available additional resources for safety assessment such as structure similarity searching and quantitative structure-activity relationship (QSAR) models. These methods offer a rapid and cost-effective first-pass screening capability to assess toxicity when conventional toxicology data are limited or lacking, with the potential to identify compounds that would be appropriate for further testing. This article discusses some of the considerations when using computational toxicology methods for regulatory decision support and gives examples of how the technology is currently being applied at the US Food and Drug Administration.
Related Concept Videos
 01:28
Preclinical Development: Overview
 01:25
Drug Regulation
 01:23
Toxicity Testing in Animals
 01:14
Drug Dissolution: Requirements and Profile Comparison
 01:29
Clinically Relevant Drug Product Specifications: Methods of Establishment
 01:20
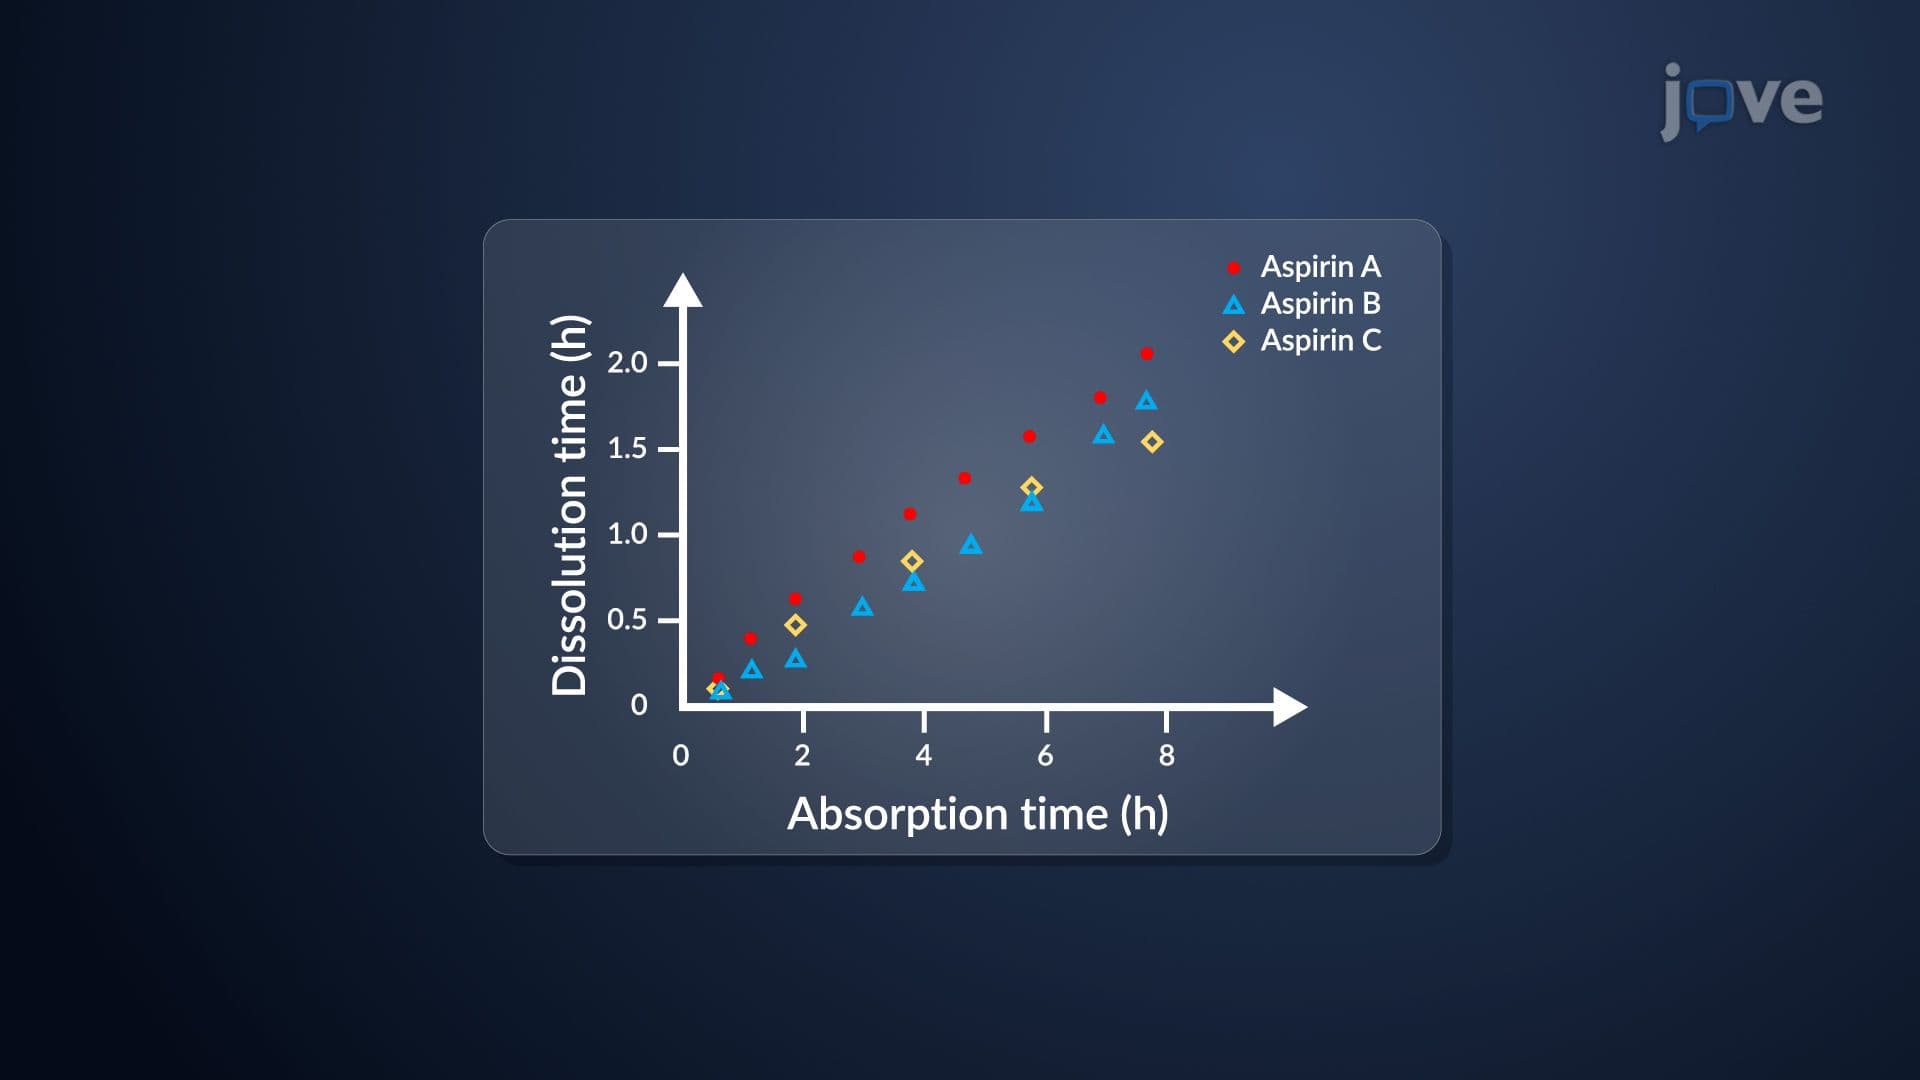
Drug Product Performance: In Vitro–In Vivo Correlation