Related Experiment Video
Updated: Jul 17, 2026

High-Throughput Optical Controlling and Recording Calcium Signal in iPSC-Derived Cardiomyocytes for Toxicity Testing and Phenotypic Drug Screening
Published on: March 31, 2022
Sequential metabolism is responsible for diltiazem-induced time-dependent loss of CYP3A
Ping Zhao1, Caroline A Lee, Kent L Kunze
1Pharmacokinetics, Dynamics and Metabolism, Pfizer Global Research and Development, La Jolla Laboratories, La Jolla, California, USA. pingz@sonuspharma.com

Abstract:
Kinetic parameters (k(inact) and K(I)) obtained in microsomes are often used to predict time-dependent inactivation. We previously reported that microsomal inactivation kinetic parameters of diltiazem underpredicted CYP3A inactivation in hepatocytes. In this study, we evaluated the contributions of inactivation and reversible inhibition of CYP3A by diltiazem and its N-desmethyl (MA) and N,N-didesmethyl (MD) metabolites. In human liver microsomes, MA was a more potent time-dependent inactivator of CYP3A than its parent drug, with apparent k(inact) approximately 4-fold higher than that of diltiazem at a microsomal protein concentration of 0.2 mg/ml. MD did not inactivate CYP3A. Inactivation of CYP3A by diltiazem was dependent on microsomal protein concentration (25, 36, and 41% decrease in CYP3A activity at 0.2, 0.4, and 0.8 mg/ml microsomal protein, respectively, incubated with 10 microM diltiazem over 20 min), whereas inactivation by MA did not seem to be protein concentration-dependent. MA and MD were reversible inhibitors of CYP3A with competitive Ki values of 2.7 and 0.2 microM, respectively. In cryopreserved hepatocytes incubated with diltiazem, time-dependent loss of CYP3A was accompanied by increased formation of MA and MD, with the MA level similar to its K(I) at higher diltiazem concentrations. In addition, the metabolites appeared to be accumulated inside the cells. In summary, time-dependent CYP3A inactivation by MA seems to be the major contributor responsible for the loss of CYP3A in human liver microsomes and human hepatocytes incubated with diltiazem. These findings suggest that prediction of CYP3A loss based solely on microsomal inactivation parameters of parent drug may be inadequate.
Insights
The metabolite MA, not the parent drug diltiazem, is the primary cause of CYP3A inactivation. This highlights the inadequacy of using only parent drug parameters to predict drug metabolism changes in the liver.
Area of Science:
- Pharmacology
- Drug Metabolism
- Enzyme Kinetics
Background:
- Microsomal kinetic parameters are frequently used to predict time-dependent drug inactivation.
- Previous studies indicated that diltiazem's microsomal parameters underestimated CYP3A inactivation in hepatocytes.
Purpose of the Study:
- To investigate the roles of diltiazem and its metabolites (MA and MD) in CYP3A inactivation and reversible inhibition.
- To determine the contribution of these compounds to CYP3A loss in human liver microsomes and hepatocytes.
Main Methods:
- Assessed time-dependent inactivation kinetics (k(inact), K(I)) of diltiazem, MA, and MD on CYP3A in human liver microsomes.
- Evaluated reversible inhibition of CYP3A by MA and MD.
- Incubated diltiazem in cryopreserved human hepatocytes to monitor CYP3A activity, metabolite formation, and intracellular accumulation.
Main Results:
- MA was a more potent CYP3A inactivator than diltiazem in microsomes; MD did not cause inactivation.
- Diltiazem inactivation was microsomal protein-dependent, while MA inactivation was not.
- MA and MD acted as competitive reversible inhibitors of CYP3A.
- In hepatocytes, diltiazem treatment led to MA/MD formation and CYP3A loss, with MA levels near its K(I).
Conclusions:
- Time-dependent inactivation of CYP3A by the metabolite MA is the main driver of CYP3A loss.
- Predicting CYP3A loss solely based on the parent drug's microsomal inactivation parameters is insufficient.
- Metabolite activity and intracellular accumulation are critical factors in drug-induced enzyme inactivation.
Related Concept Videos
 01:23
Nonlinear Pharmacokinetics: Dependence of Elimination Half-Life and Dose Clearance
A study on guinea pigs examined the...
 01:15
Chronopharmacokinetics: Circadian Rhythms and Influence on Drug Response
The time of drug administration is an important factor to consider, as it can influence the toxic dose of a drug. For example, a study conducted by Prins et al. in 1997 examined the effects of the timing of...
 01:25
Pharmacokinetics: Drug–Drug Interactions
 01:18
Pharmacokinetics in Geriatric Patients: Effect of Age on Drug Metabolism
 01:20
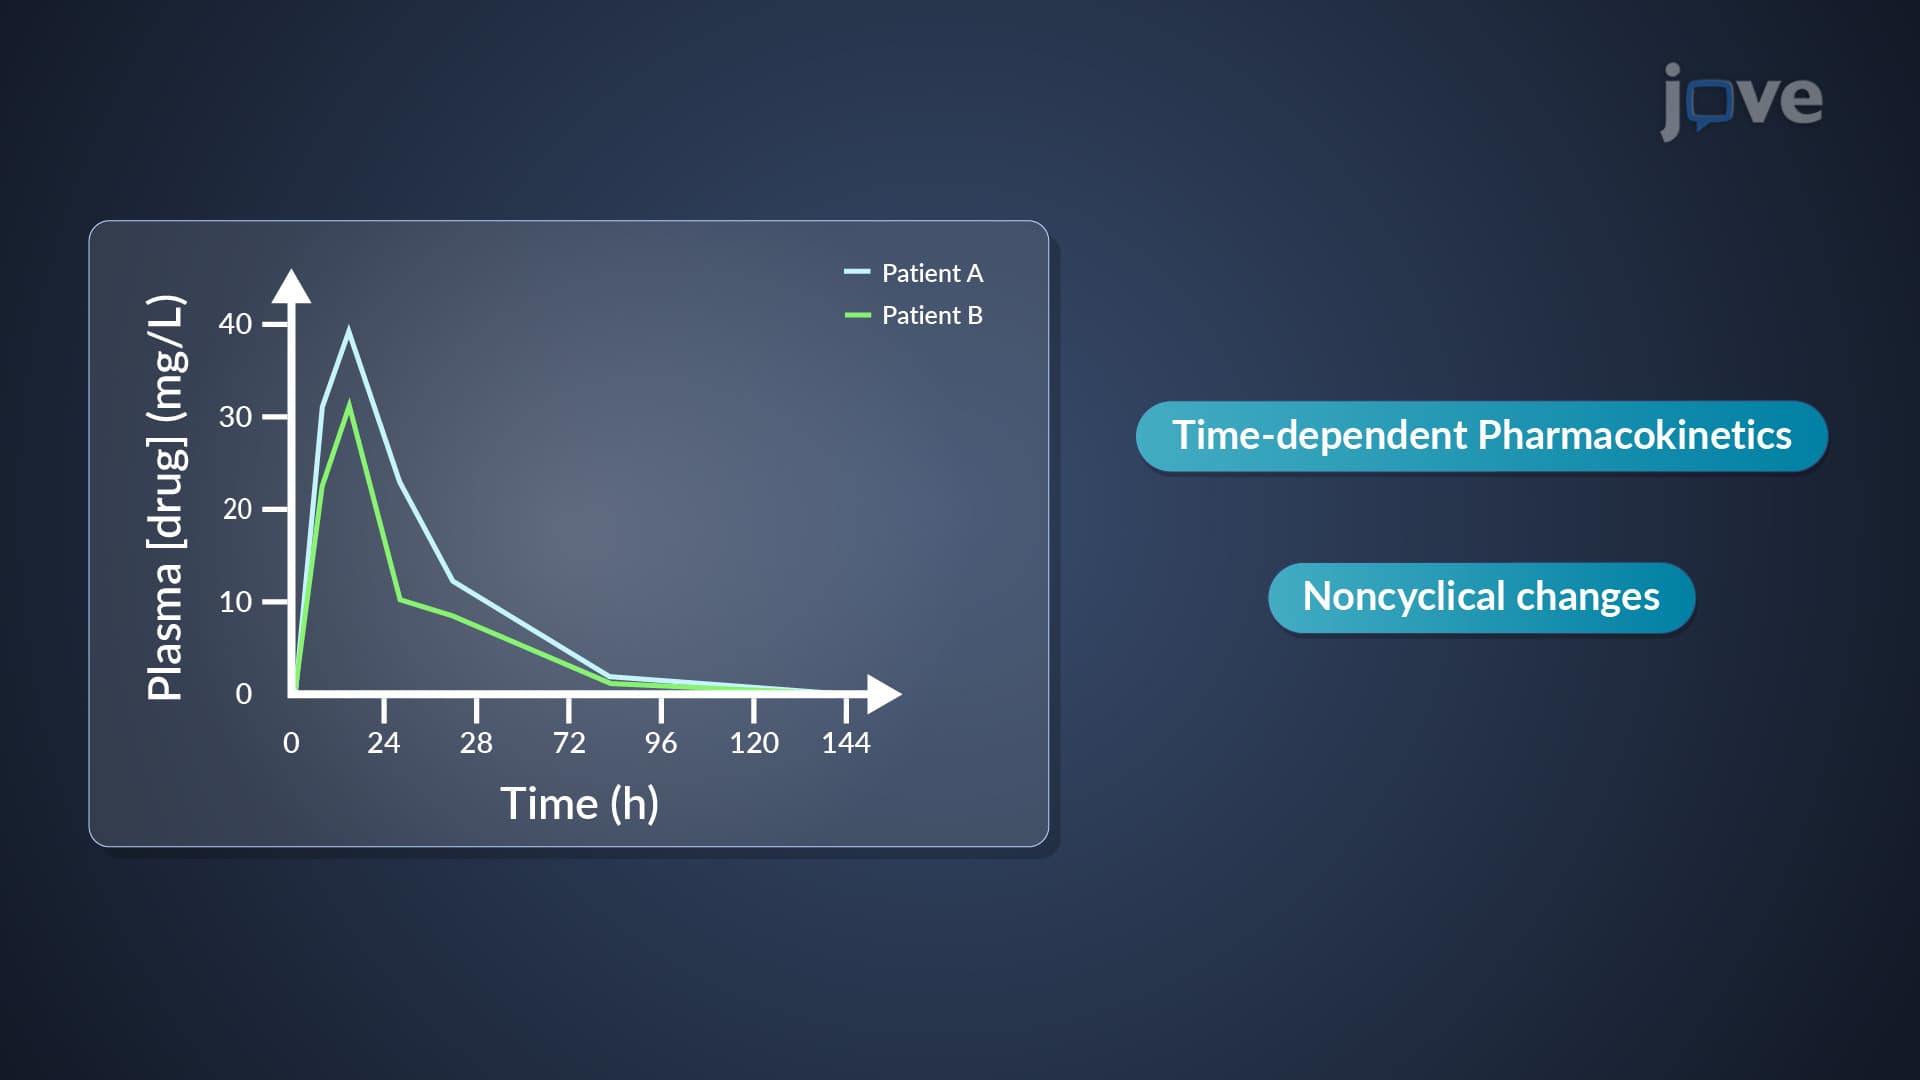
Chronopharmacokinetics: Time-Dependent Pharmacokinetics
Time-dependent pharmacokinetics refers to non-cyclical changes in drug rate processes over a period of time. It can lead to nonlinear pharmacokinetics, where the relationship between drug concentration and time is not proportional. Non-cyclical...
 01:14
Time Course of Drug Effect