Related Experiment Video
Updated: Jun 4, 2026
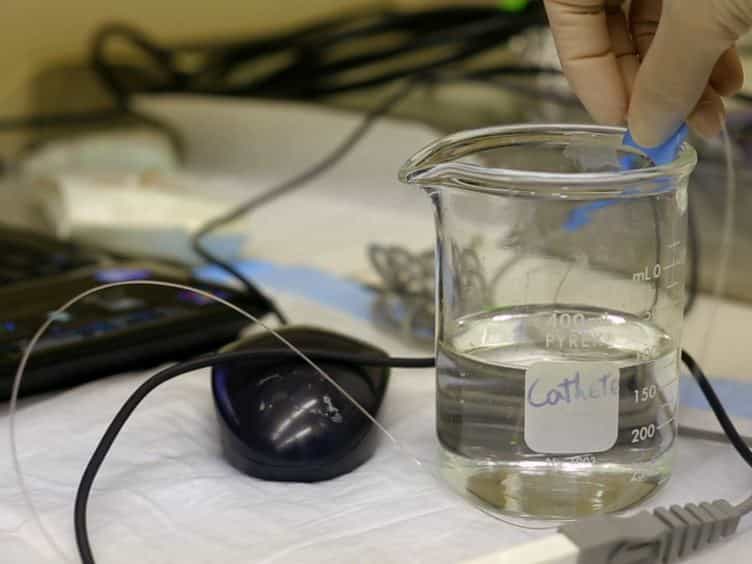

Right Ventricular Systolic Pressure Measurements in Combination with Harvest of Lung and Immune Tissue Samples in Mice
Published on: January 16, 2013
Moderate pulmonary arterial hypertension in male mice lacking the vasoactive intestinal peptide gene
Sami I Said1, Sayyed A Hamidi, Kathleen G Dickman
1Departments of Medicine, State University of New York at Stony Brook, Stony Brook, NY, USA. sami.i.said@stonybrook.edu

Background:
Vasoactive intestinal peptide (VIP), a pulmonary vasodilator and inhibitor of vascular smooth muscle proliferation, has been reported absent in pulmonary arteries from patients with idiopathic pulmonary arterial hypertension (PAH). We have tested the hypothesis that targeted deletion of the VIP gene may lead to PAH with pulmonary vascular remodeling.
Methods And Results:
We examined VIP knockout (VIP-/-) mice for evidence of PAH, right ventricular (RV) hypertrophy, and pulmonary vascular remodeling. Relative to wild-type control mice, VIP-/- mice showed moderate RV hypertension, RV hypertrophy confirmed by increased ratio of RV to left ventricle plus septum weight, and enlarged, thickened pulmonary artery and smaller branches with increased muscularization and narrowed lumen. Lung sections also showed perivascular inflammatory cell infiltrates. No systemic hypertension and no arterial hypoxemia existed to explain the PAH. The condition was associated with increased mortality. Both the vascular remodeling and RV remodeling were attenuated after a 4-week treatment with VIP.
Conclusions:
Deletion of the VIP gene leads to spontaneous expression of moderately severe PAH in mice during air breathing. Although not an exact model of idiopathic PAH, the VIP-/- mouse should be useful for studying molecular mechanisms of PAH and evaluating potential therapeutic agents. VIP replacement therapy holds promise for the treatment of PAH, and mutations of the VIP gene may be a factor in the pathogenesis of idiopathic PAH.
Insights
Vasoactive intestinal peptide (VIP) gene deletion causes pulmonary arterial hypertension (PAH) in mice. VIP replacement therapy shows promise for treating PAH, suggesting VIP gene mutations may contribute to idiopathic PAH.
Area of Science:
- Cardiovascular Research
- Pulmonary Medicine
- Genetics
Background:
- Vasoactive intestinal peptide (VIP) is a pulmonary vasodilator.
- VIP is reportedly absent in pulmonary arteries of idiopathic pulmonary arterial hypertension (PAH) patients.
- VIP inhibits vascular smooth muscle proliferation.
Purpose of the Study:
- To test if VIP gene deletion causes PAH with pulmonary vascular remodeling.
- To investigate the role of VIP in the pathogenesis of PAH.
Main Methods:
- Examined VIP knockout (VIP-/-) mice for PAH, right ventricular (RV) hypertrophy, and pulmonary vascular remodeling.
- Compared VIP-/- mice to wild-type control mice.
- Assessed effects of VIP treatment on vascular and RV remodeling.
Main Results:
- VIP-/- mice developed moderate RV hypertension, RV hypertrophy, and pulmonary vascular remodeling with increased muscularization and narrowed lumen.
- Perivascular inflammatory cell infiltrates were observed in lung sections.
- VIP treatment attenuated both vascular and RV remodeling.
Conclusions:
- VIP gene deletion leads to spontaneous PAH in mice.
- The VIP-/- mouse model is useful for studying PAH mechanisms and therapies.
- VIP replacement therapy is a promising treatment for PAH.
- VIP gene mutations may be involved in idiopathic PAH pathogenesis.
More Related Videos
 04:54
Invasive Hemodynamic Assessment for the Right Ventricular System and Hypoxia-Induced Pulmonary Arterial Hypertension in Mice
Published on: October 24, 2019
 05:31
Murine Model of Thoracic Aortic Dissection Induced by Oral β-Aminopropionitrile and Subcutaneous Angiotensin II Infusion
Published on: May 16, 2025