Related Experiment Video
Updated: Jul 16, 2026

In situ FTIR Spectroscopy as a Tool for Investigation of Gas/Solid Interaction: Water-Enhanced CO2 Adsorption in UiO-66 Metal-Organic Framework
Published on: February 1, 2020
Charge distribution in metal organic framework materials: transferability to a preliminary molecular simulation study
N A Ramsahye1, G Maurin, S Bourrelly
1Laboratoire LPMC UMR CNRS 5617, Université Montpellier II, 34095, Montpellier Cedex 05, France.

Abstract:
Density functional theory calculations have been performed in order to extract the charge distribution in the aluminium-containing MIL-53 structure, to allow further computational studies of adsorption in these materials. Both cluster and periodic methods have been used and the charges calculated for each atom constituting the organic and inorganic part of the material, were discussed. Preliminary grand canonical Monte Carlo simulations, based on a consistent set of potential parameters and this newly derived charge distribution, predicted for enthalpies of adsorption for CO(2) at low coverage in the "large" and "narrow" pore versions of MIL-53 (Al) to be significantly different. These calculated enthalpies reproduced the two distinct ranges of values observed by microcalorimetry on either side of 6 bars quite well. This agreement between experiment and simulation validated our previous assumption, suggesting a structural switching of the hybrid material during the adsorption process. The microscopic mode of interaction between the hybrid porous framework and the CO(2) adsorption was then carefully analysed in both of the MIL-53 (Al) structures.
More Related Videos
 10:52
Multiscale Sampling of a Heterogeneous Water/Metal Catalyst Interface using Density Functional Theory and Force-Field Molecular Dynamics
Published on: April 12, 2019
 06:00
Solvothermal Synthesis of MIL-96 and UiO-66-NH2 on Atomic Layer Deposited Metal Oxide Coatings on Fiber Mats
Published on: June 13, 2018
Related Concept Videos
 01:18
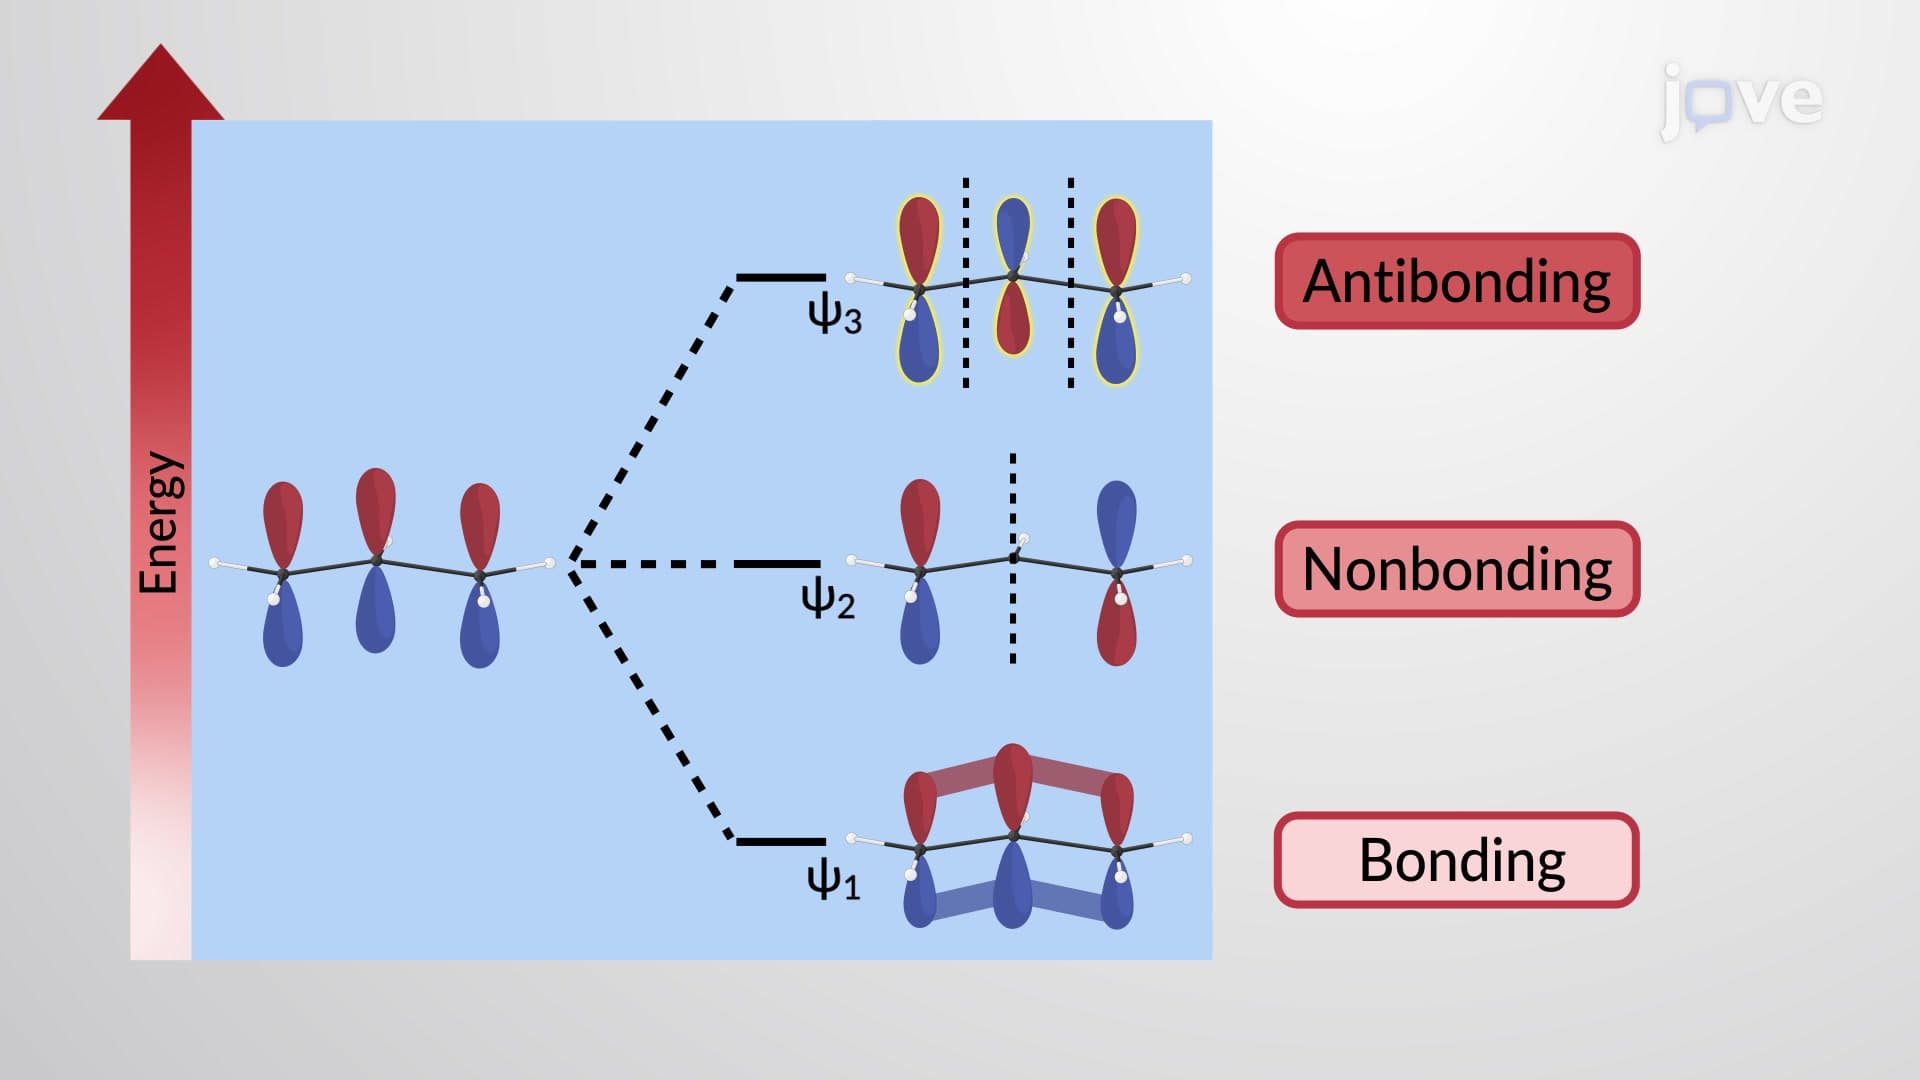
π Molecular Orbitals of the Allyl Cation and Anion
 01:23
Properties of Organometallic Compounds
 02:40
MO Theory and Covalent Bonding