Computational sidechain placement and protein mutagenesis with implicit solvent models
Related Experiment Video
Updated: Jul 16, 2026
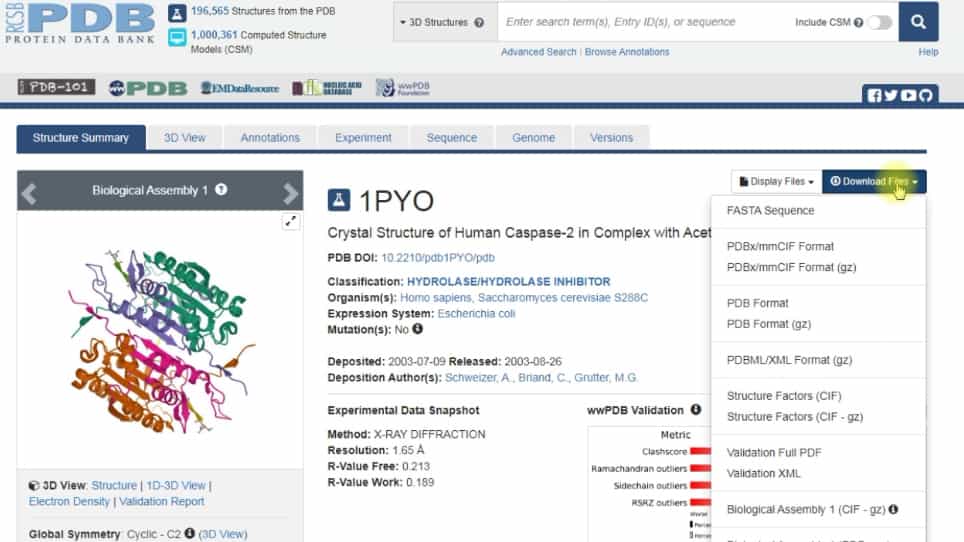
Exploring Caspase Mutations and Post-Translational Modification by Molecular Modeling Approaches
Published on: October 13, 2022
Anne Lopes1, Alexey Alexandrov, Christine Bathelt
1Laboratoire de Biochimie (UMR CNRS 7654), Department of Biology, Ecole Polytechnique, 91128, Palaiseau, France.

Abstract:
Structure prediction and computational protein design should benefit from accurate solvent models. We have applied implicit solvent models to two problems that are central to this area. First, we performed sidechain placement for 29 proteins, using a solvent model that combines a screened Coulomb term with an Accessible Surface Area term (CASA model). With optimized parameters, the prediction quality is comparable with earlier work that omitted electrostatics and solvation altogether. Second, we computed the stability changes associated with point mutations involving ionized sidechains. For over 1000 mutations, including many fully or partly buried positions, we compared CASA and two generalized Born models (GB) with a more accurate model, which solves the Poisson equation of continuum electrostatics numerically. CASA predicts the correct sign and order of magnitude of the stability change for 81% of the mutations, compared to 97% with the best GB. We also considered 140 mutations for which experimental data are available. Comparing to experiment requires additional assumptions about the unfolded protein structure, protein relaxation in response to the mutations, and contributions from the hydrophobic effect. With a simple, commonly-used unfolded state model, the mean unsigned error is 2.1 kcal/mol with both CASA and the best GB. Overall, the electrostatic model is not important for sidechain placement; CASA and GB are equivalent for surface mutations, while GB is far superior for fully or partly buried positions. Thus, for problems like protein design that involve all these aspects, the most recent GB models represent an important step forward. Along with the recent discovery of efficient, pairwise implementations of GB, this will open new possibilities for the computational engineering of proteins.
More Related Videos
 07:08
Optimization of Synthetic Proteins: Identification of Interpositional Dependencies Indicating Structurally and/or Functionally Linked Residues
Published on: July 14, 2015
 15:05
Deciphering the Structural Effects of Activating EGFR Somatic Mutations with Molecular Dynamics Simulation
Published on: May 20, 2020
Related Concept Videos
 01:17
Chemical Shift: Internal References and Solvent Effects
The internal reference compound generally used in NMR spectroscopy is tetramethylsilane (TMS). TMS is preferred because it is chemically inert, soluble in NMR solvents, and easily removable. Also, the highly shielded methyl protons in TMS yield an intense...
 01:16
In vitro Mutagenesis
 02:00
Molecular Models