Related Experiment Video
Updated: Jul 16, 2026
In Vitro Analysis of PDZ-dependent CFTR Macromolecular Signaling Complexes
Published on: August 13, 2012
G551D and G1349D, two CF-associated mutations in the signature sequences of CFTR, exhibit distinct gating defects
Silvia G Bompadre1, Yoshiro Sohma, Min Li
1Department of Medical Pharmacology and Physiology, University of Missouri-Columbia, Columbia, MO 65211, USA.

Abstract:
Mutations in the gene encoding cystic fibrosis transmembrane conductance regulator (CFTR) result in cystic fibrosis (CF). CFTR is a chloride channel that is regulated by phosphorylation and gated by ATP binding and hydrolysis at its nucleotide binding domains (NBDs). G551D-CFTR, the third most common CF-associated mutation, has been characterized as having a lower open probability (Po) than wild-type (WT) channels. Patients carrying the G551D mutation present a severe clinical phenotype. On the other hand, G1349D, also a mutant with gating dysfunction, is associated with a milder clinical phenotype. Residues G551 and G1349 are located at equivalent positions in the highly conserved signature sequence of each NBD. The physiological importance of these residues lies in the fact that the signature sequence of one NBD and the Walker A and B motifs from the other NBD form the ATP-binding pocket (ABP1 and ABP2, named after the location of the Walker A motif) once the two NBDs dimerize. Our studies show distinct gating characteristics for these mutants. The G551D mutation completely eliminates the ability of ATP to increase the channel activity, and the observed activity is approximately 100-fold smaller than WT-CFTR. G551D-CFTR does not respond to ADP, AMP-PNP, or changes in [Mg(2+)]. The low activity of G551D-CFTR likely represents the rare ATP-independent gating events seen with WT channels long after the removal of ATP. G1349D-CFTR maintains ATP dependence, albeit with a Po approximately 10-fold lower than WT. Interestingly, compared to WT results, the ATP dose-response relationship of G1349D-CFTR is less steep and shows a higher apparent affinity for ATP. G1349D data could be well described by a gating model that predicts that binding of ATP at ABP1 hinders channel opening. Thus, our data provide a quantitative explanation at the single-channel level for different phenotypes presented by patients carrying these two mutations. In addition, these results support the idea that CFTR's two ABPs play distinct functional roles in gating.
More Related Videos
Related Concept Videos
 01:23
Cystic Fibrosis: Pathogenesis
CF is primarily caused by a genetic mutation in a chromosome 7 gene coding for the cystic fibrosis transmembrane conductance regulator (CFTR) protein. The most common gene mutation leading to CF is the ΔF508 mutation, but...
 01:09
GPCRs Regulate Adenylyl Cylase Activity
Two...
 01:25
Rab Cascades
 01:30
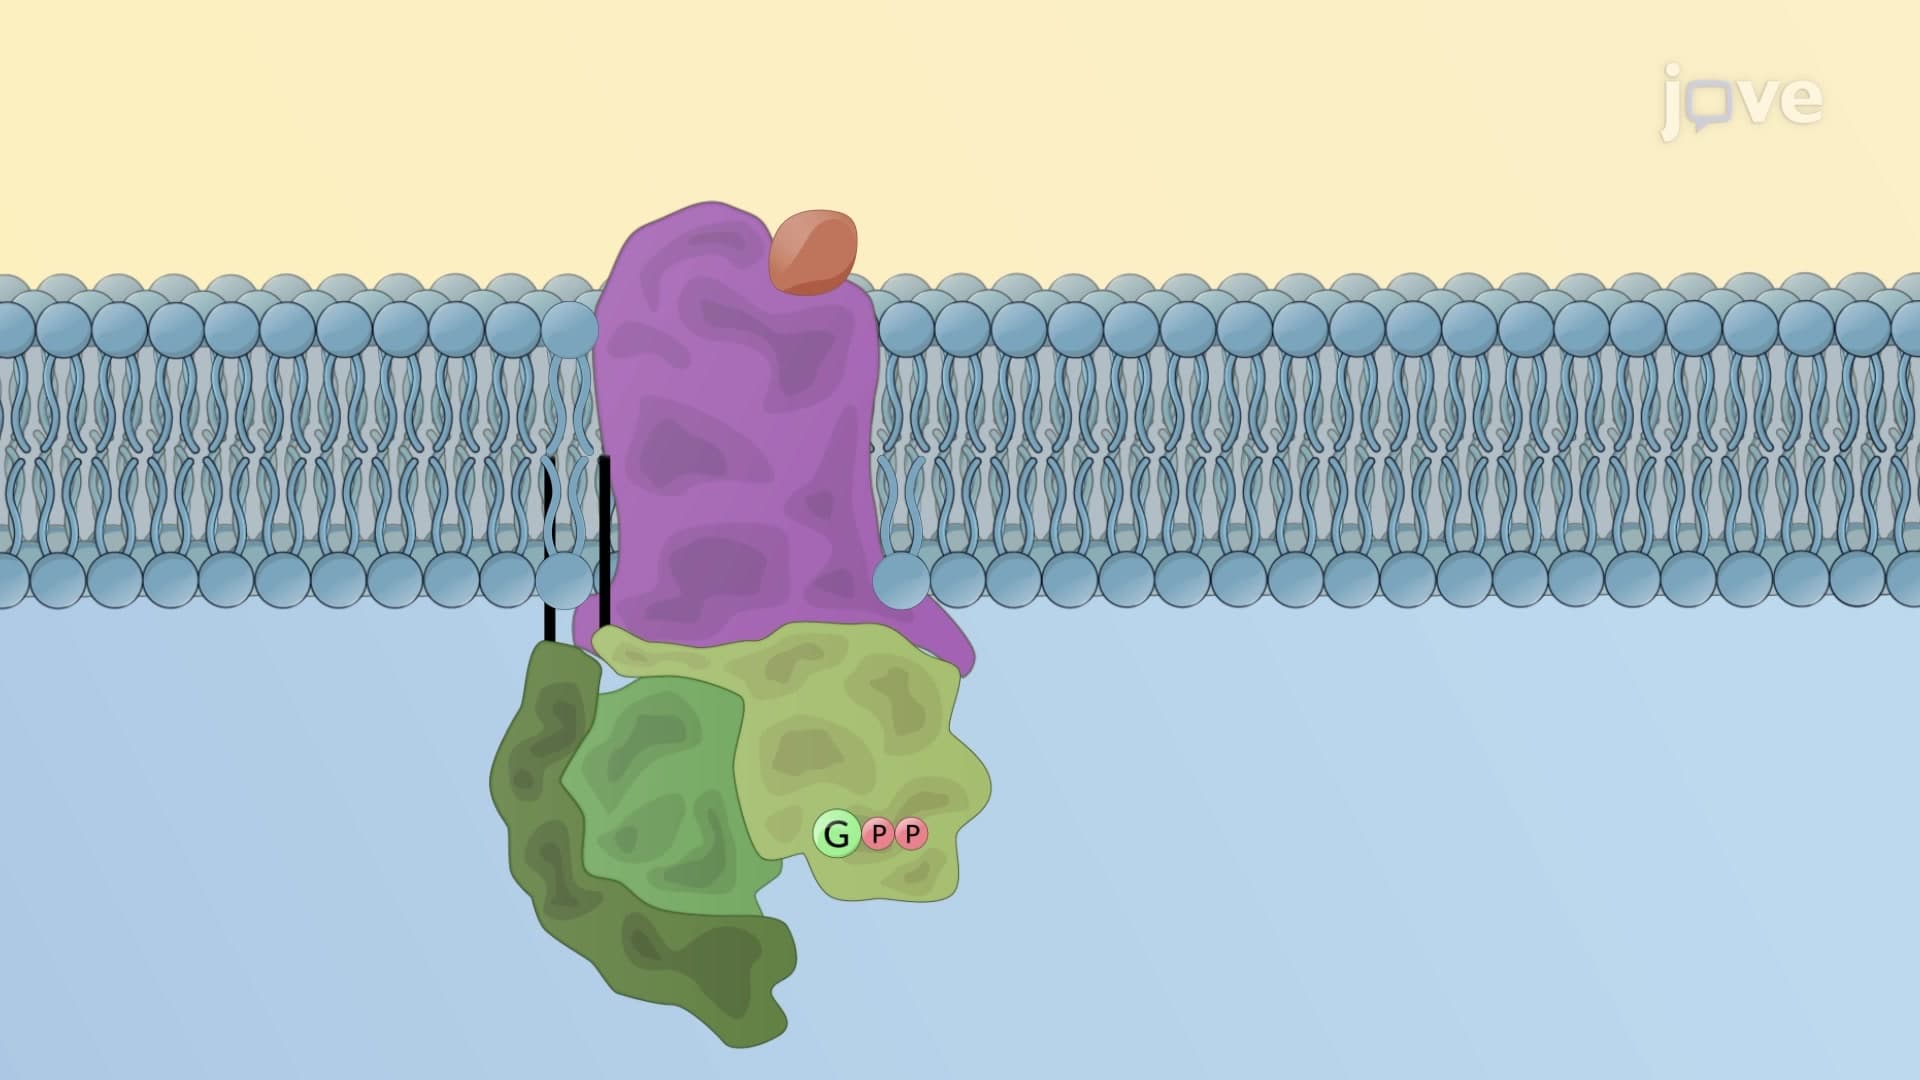
Transducer Mechanism: G Protein–Coupled Receptors
GPCRs are also called heptahelical, 7TM, or...
 01:22
Activation and Inactivation of G Proteins
 01:15
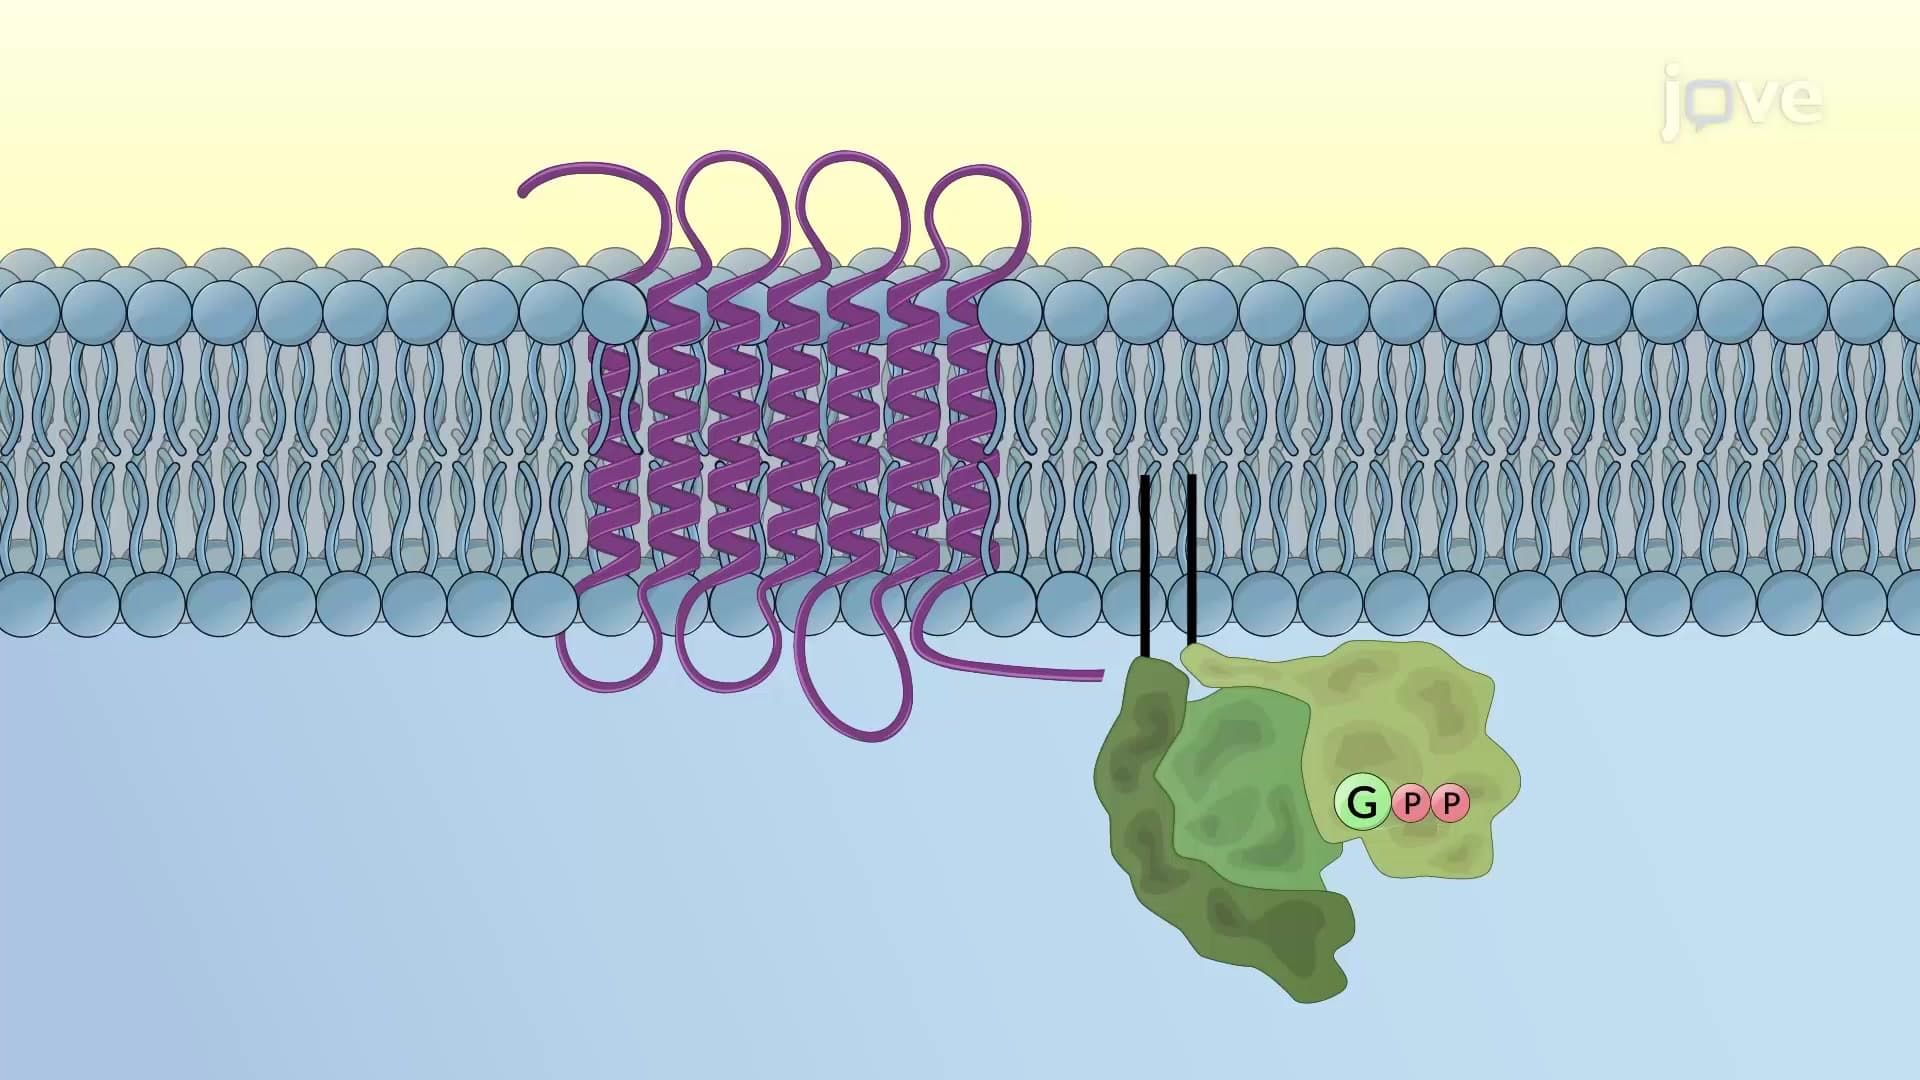
G Protein-coupled Receptors
GPCRs are also called heptahelical, 7TM, or serpentine receptors, and consist of seven (H1-H7) transmembrane alpha-helices that span the bilayer to form a cylindrical core. The transmembrane helices are connected by three extracellular loops and three...