Related Experiment Video
Updated: Jul 16, 2026

Evaluation of LC3-II Release via Extracellular Vesicles in Relation to the Accumulation of Intracellular LC3-positive Vesicles
Published on: October 18, 2024
TDP-43 pathology in familial frontotemporal dementia and motor neuron disease without Progranulin mutations
Harro Seelaar1, H Jurgen Schelhaas, Asma Azmani
1Department of Neurology, Erasmus University Medical Centre, Rotterdam, The Netherlands.

Abstract:
Frontotemporal dementia is accompanied by motor neuron disease (FTD + MND) in approximately 10% of cases. There is accumulating evidence for a clinicopathological overlap between FTD and MND based on observations of familial aggregation and neuropathological findings of ubiquitin-positive neuronal cytoplasmatic inclusions (NCI) in lower motor neurons, hippocampus and neocortex in both conditions. Several familial forms exist with different genetic loci and defects. We investigated the familial aggregation and clinical presentation of FTD + MND cases in a large cohort of 368 FTD patients in The Netherlands. Immunohistochemistry of available brain tissue of deceased patients was investigated using a panel of antibodies including ubiquitin, p62 and TAR DNA-binding protein of 43 kDa antibodies. A total of eight patients coming from six families had a family history positive for FTD + MND (mean age at onset 53.2 +/- 8.4 years). Five patients presented with behavioural changes and cognitive changes followed by motor neuron disease, whereas symptoms of motor neuron disease were the presenting features in the remaining three patients. Other affected relatives in these families showed dementia/FTD, MND or FTD + MND reflecting the clinical interfamilial variation. No mutations were identified in any of the candidate genes, including Superoxide Dismutase 1, dynactin, angiogenin, Microtubule-Associated Protein Tau, valosin-containing protein and progranulin. Available brain tissue of five patients with familial FTD + MND showed NCI in hippocampus, neocortex and spinal cord in all, and neuronal intranuclear inclusions (NII) in two brains. TDP-43 antibody showed robust staining of neuronal inclusions similar in distribution and morphology to NCI and NII. Additionally, TDP-43 antibody also stained ubiquitin-negative glial inclusions in the basal striatum of one case. In conclusion, there exists considerable clinical variation within families with FTD + MND, which may be determined by other genetic or environmental factors. NII are also found in some cases of familial FTD + MND without Progranulin mutations. The observation of glial TDP-43 positive inclusions in one brain is very interesting, although their pathophysiological significance is yet unknown.
Insights
Frontotemporal dementia with motor neuron disease (FTD + MND) shows significant clinical variation within families. Neuropathological analysis revealed TDP-43 protein inclusions in affected brain tissues, suggesting a shared pathology.
Area of Science:
- Neuroscience
- Genetics
- Pathology
Background:
- Frontotemporal dementia (FTD) co-occurs with motor neuron disease (MND) in approximately 10% of cases (FTD + MND).
- Clinicopathological overlap is suggested by familial aggregation and ubiquitin-positive inclusions in FTD and MND.
- Genetic loci and defects vary among familial forms.
Purpose of the Study:
- To investigate the familial aggregation and clinical presentation of FTD + MND in a large Dutch cohort.
- To examine neuropathological findings, including TDP-43 protein, in familial FTD + MND cases.
Main Methods:
- Studied 368 FTD patients in The Netherlands for familial FTD + MND.
- Conducted immunohistochemistry on brain tissue using ubiquitin, p62, and TDP-43 antibodies.
- Screened for mutations in candidate genes (SOD1, dynactin, angiogenin, MAPT, VCP, progranulin).
Main Results:
- Identified eight patients from six families with a history of FTD + MND (mean onset 53.2 years).
- Observed significant interfamilial clinical variation, with presentations including behavioral changes, cognitive decline, and motor neuron disease.
- Found neuronal cytoplasmic inclusions (NCI) and neuronal intranuclear inclusions (NII) in brain tissue, with TDP-43 staining these inclusions.
- No mutations were found in screened candidate genes.
- TDP-43-positive glial inclusions were observed in one case.
Conclusions:
- Familial FTD + MND exhibits considerable clinical variability, potentially influenced by genetic or environmental factors.
- Neuronal intranuclear inclusions are present in some familial FTD + MND cases, even without progranulin mutations.
- The significance of glial TDP-43 inclusions requires further investigation.
More Related Videos
 11:03
Use of Capillary Electrophoresis Immunoassay to Search for Potential Biomarkers of Amyotrophic Lateral Sclerosis in Human Platelets
Published on: February 10, 2020
 13:31
Novel Atomic Force Microscopy Based Biopanning for Isolation of Morphology Specific Reagents against TDP-43 Variants in Amyotrophic Lateral Sclerosis
Published on: February 12, 2015
Related Concept Videos
 01:24
Parkinson Disease ll: Pathophysiology
 01:23
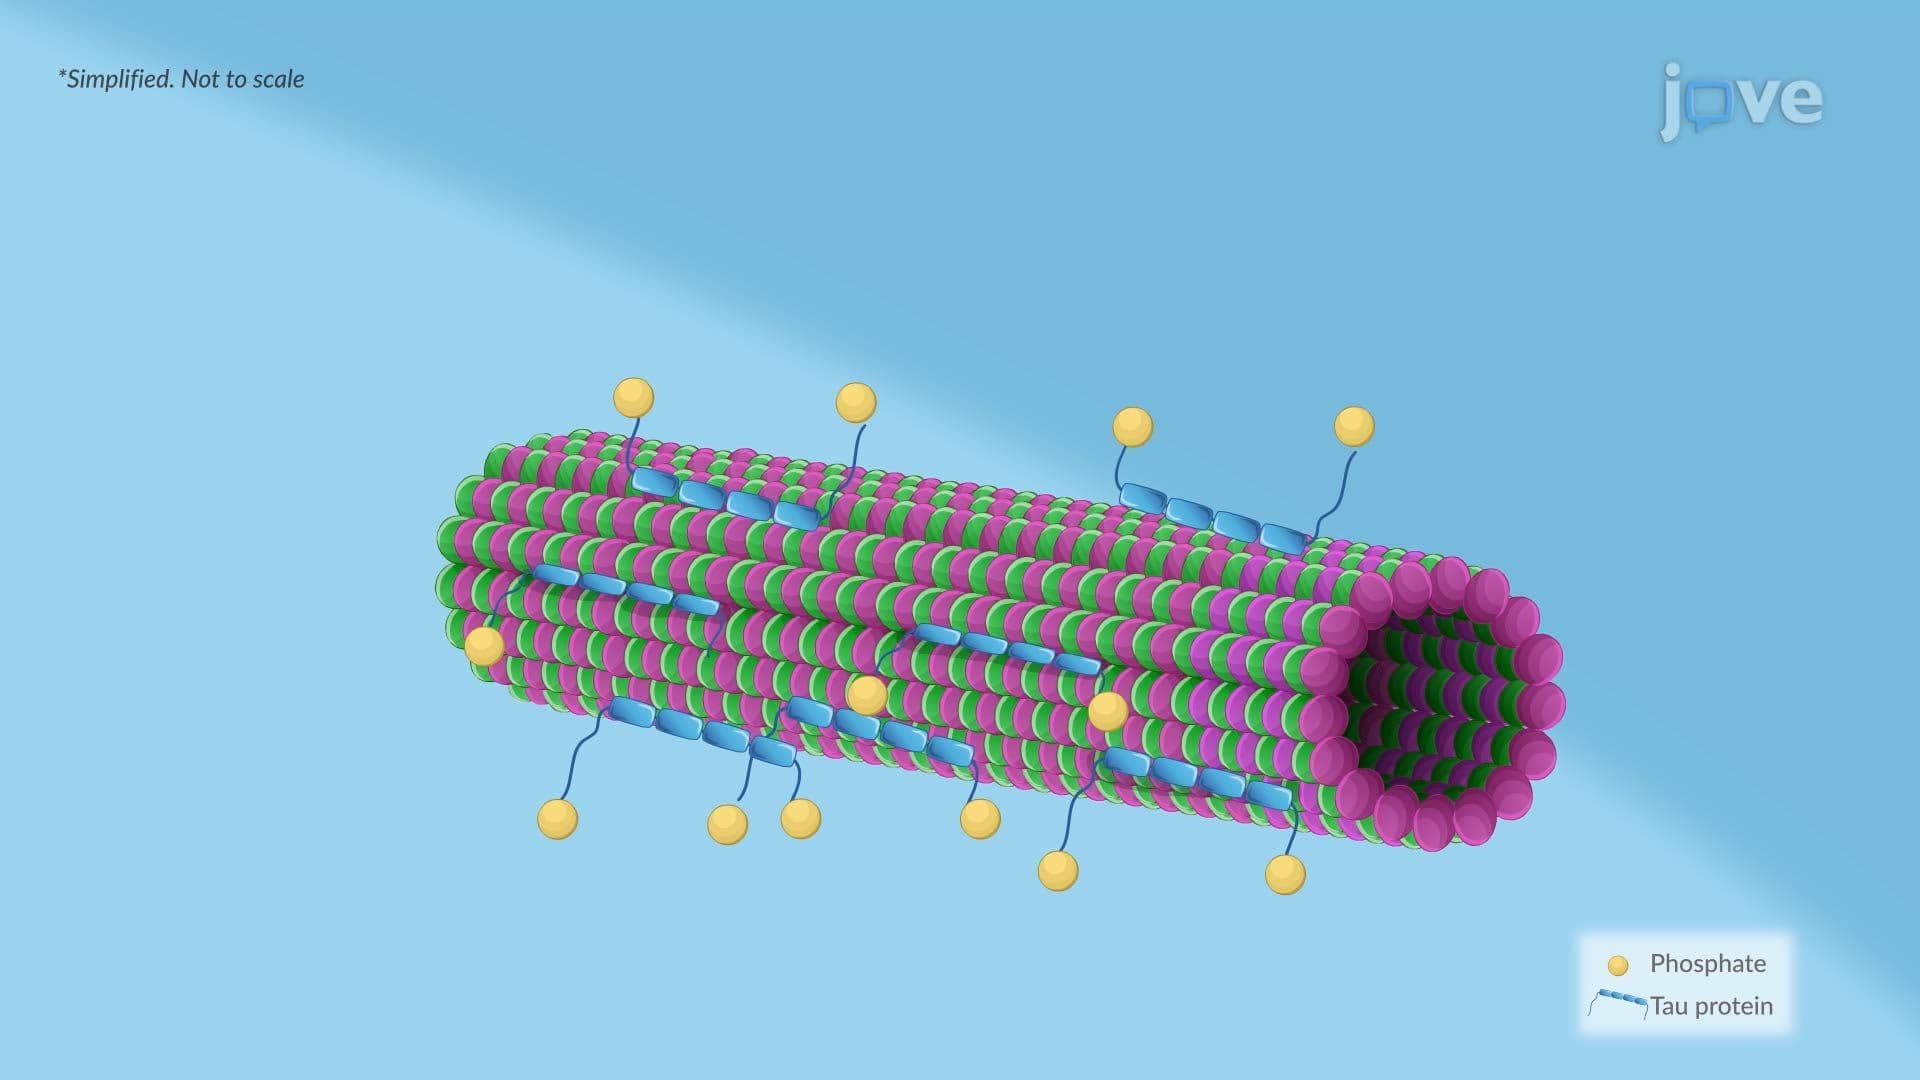
Alzheimer Disease ll: Pathophysiology
 01:29
Alzheimer Disease l: Introduction
 01:21
Huntington Disease l: Introduction
 01:24
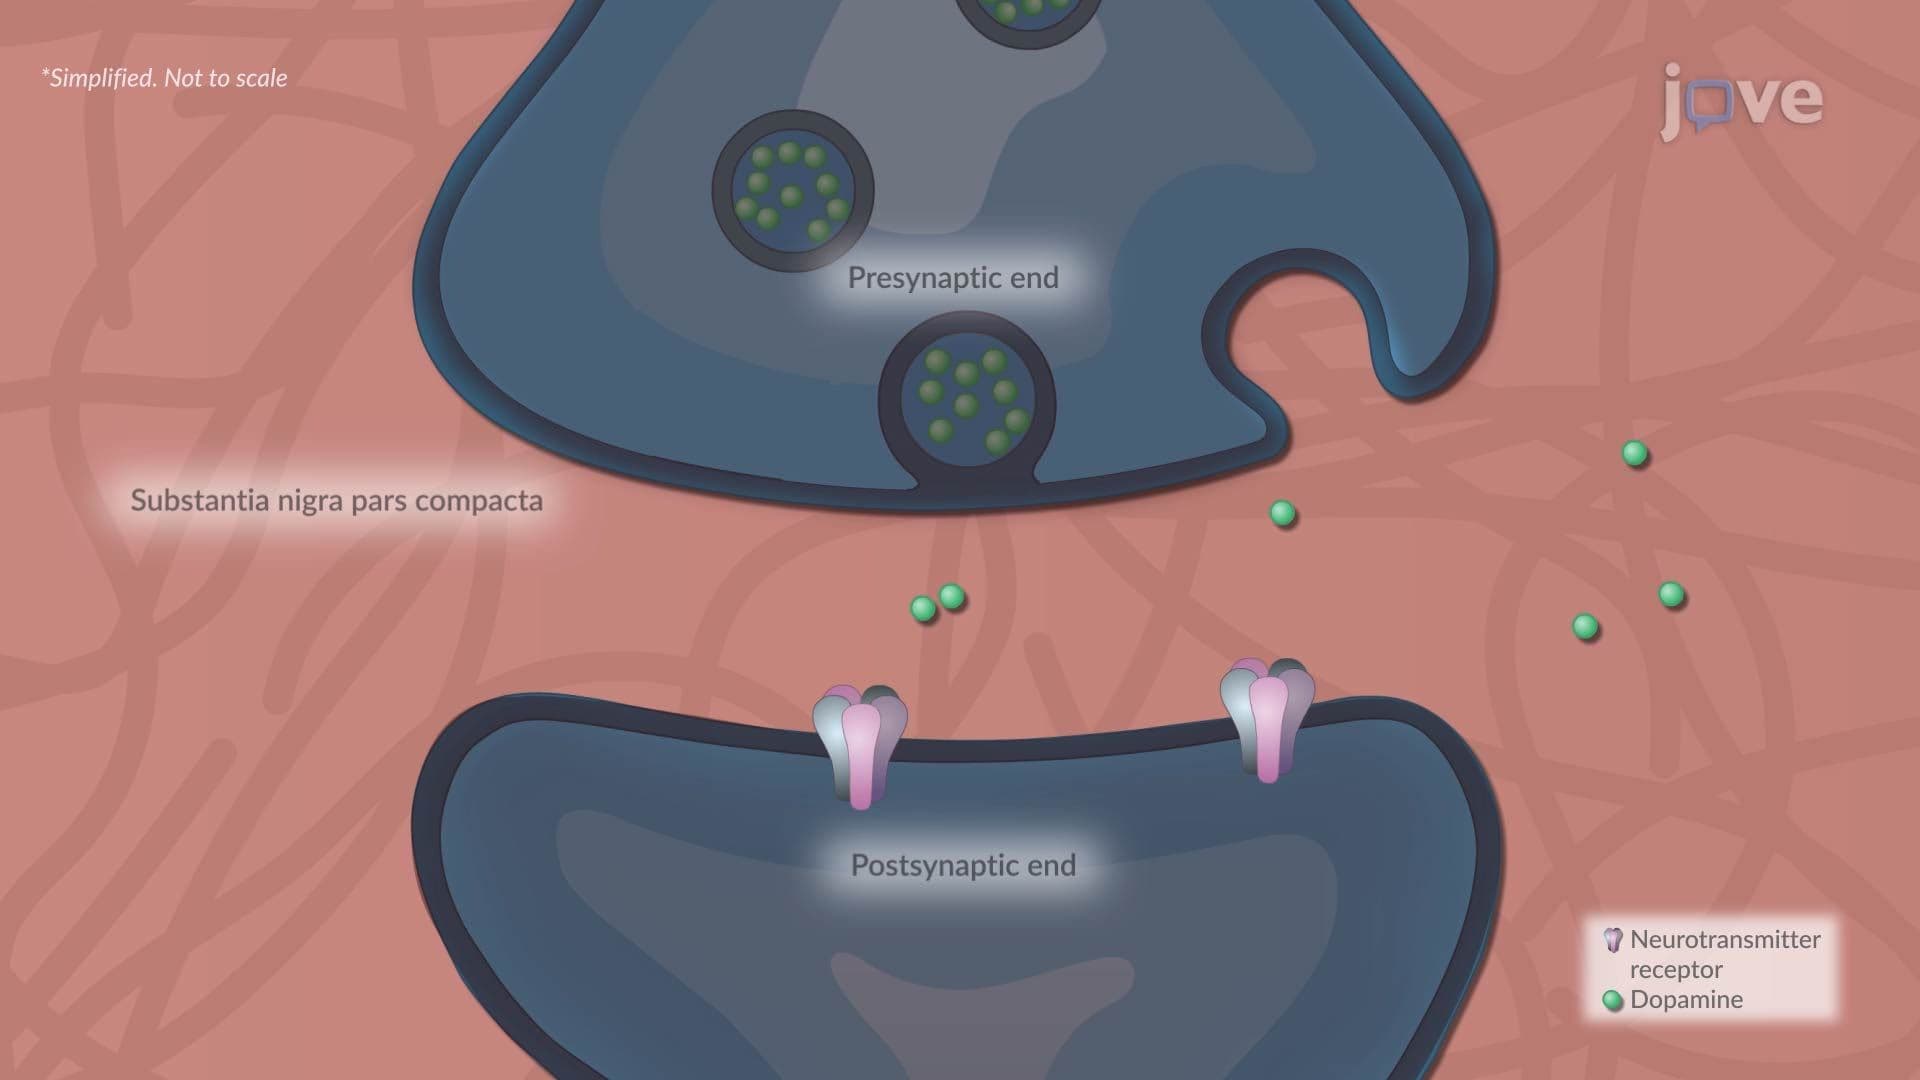
Parkinson Disease l: Introduction
 01:15
Parkinson's Disease: Overview